So You Need to Run a DNA Profile. Here Is What Actually Happens.
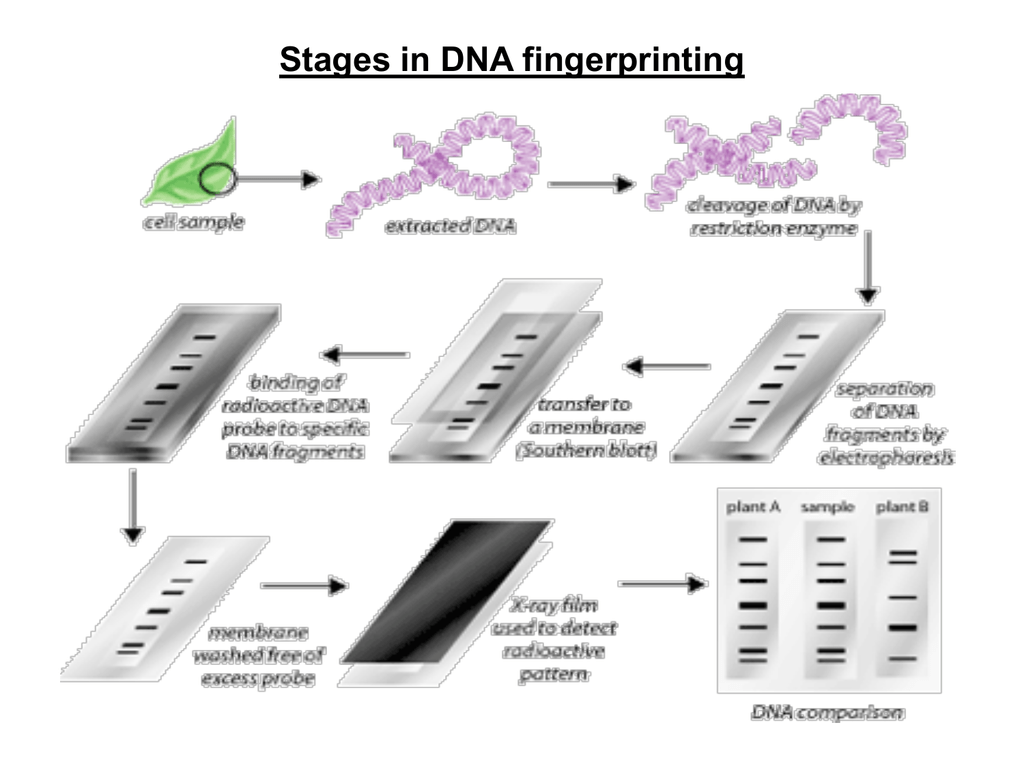
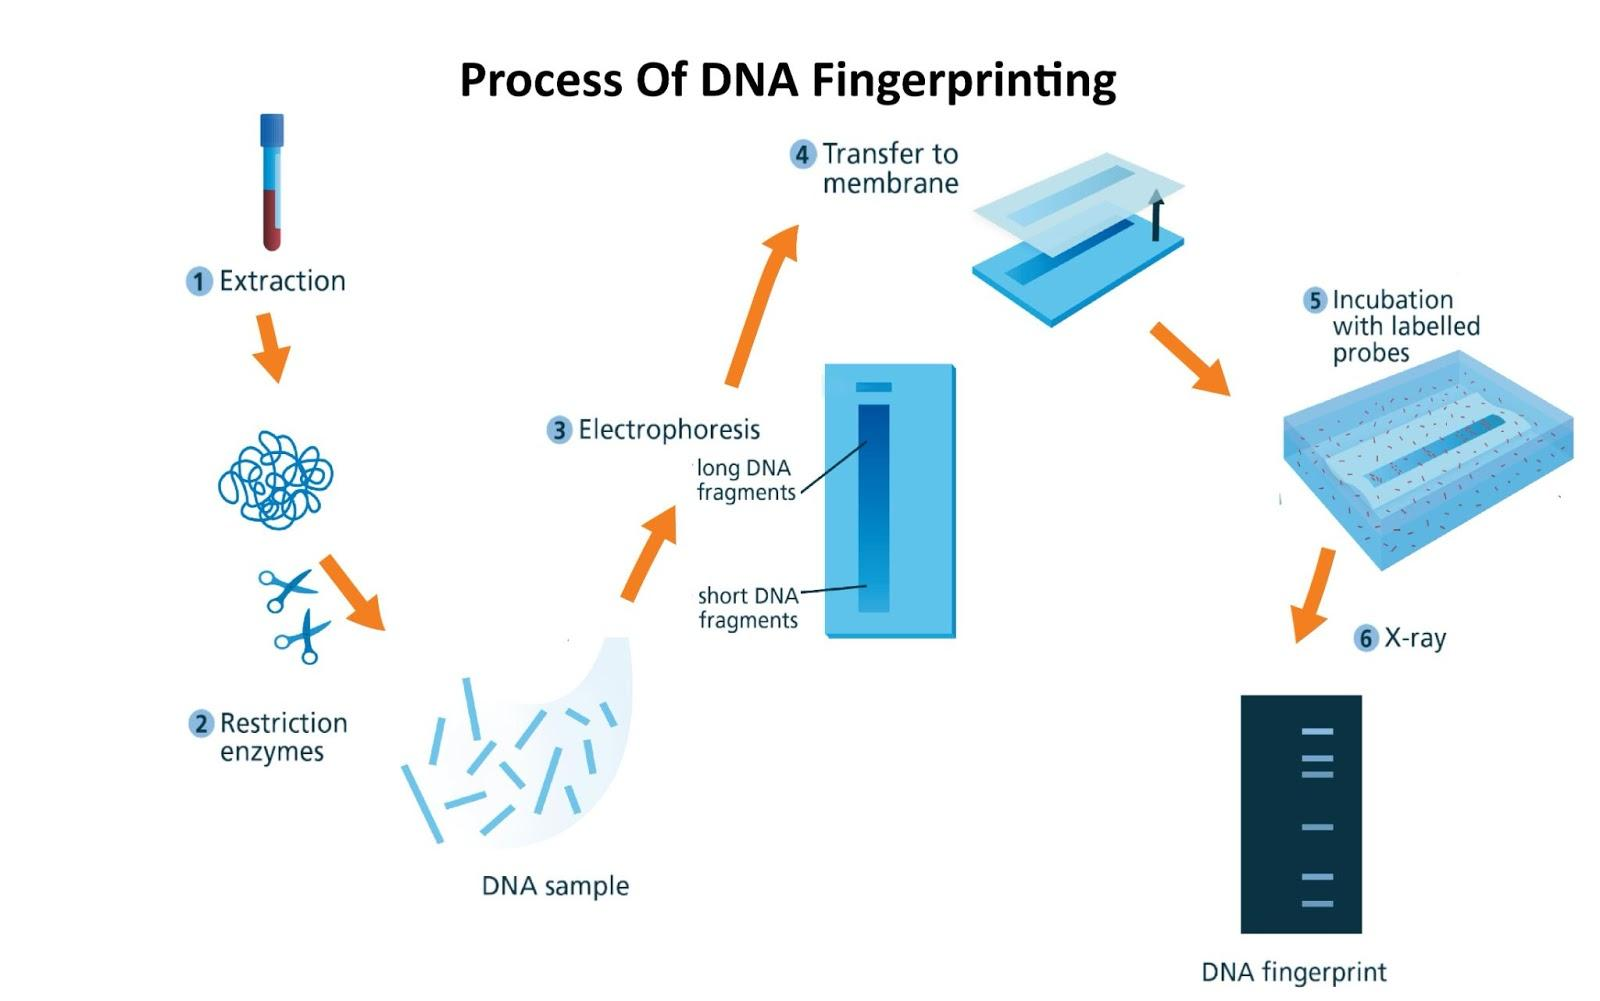
Dna Profiling Dna Fingerprinting is not a single test. It is a chain of molecular steps, and if any one link is weak you will get a result that looks clean but is fundamentally useless. I have watched people throw away weeks of work because they treated the extraction step as something that "just works." It does not. The basic workflow goes like this. You collect a biological sample, lyse the cells, clean up the nucleic acids, amplify specific loci through PCR, separate the fragments by size, and interpret the electropherogram. That is the textbook version. The reality involves more pipetting, more QC checks, and more moments where you stare at a blank screen wondering why your positive control failed.
Dna Profiling Dna Fingerprinting
At its core, this method targets short tandem repeats. These are repeating sequences scattered throughout the genome at known positions. The number of repeats varies between individuals, which is what makes them useful for identification. The standard panels used in forensics look at around 20 loci, including the CODIS core markers. Some labs run additional STRs or switch to SNP panels when the DNA is highly degraded. I want to talk about the part most tutorials skip, which is the extraction choice and why it matters far more than people admit. I spent about three years running casework samples from various substrates. One type of sample that consistently gave me trouble was old bloodstains on porous fabric that had been exposed to heat and moisture. The DNA was fragmented enough that standard amplification would produce partial profiles with drop-out at larger loci. The workaround I ended up using was switching to a mini-STR kit. These primers target shorter amplicons, usually under 200 base pairs, and they recovered usable profiles from samples that were completely flatlining with standard primers. Not every lab has those kits on hand, and they cost more per sample, but it is a real difference between a dead end and a lead. Another counter-intuitive thing is the relationship between DNA quantity and profile quality. More DNA does not automatically mean a better profile. I have seen samples with high template concentrations produce messy electropherograms with artificial peaks, stutter artifacts that obscure real alleles, and allelic imbalance that makes heterozygous calls unreliable. The fix is usually dilution. You run a quantification first, then dilute into the optimal range rather than amplifying straight from the extract. Most kits have a recommended input range, and staying within it prevents more problems than you might expect.
When it comes to choosing a method, there are three main options people consider. Traditional capillary electrophoresis with fluorescently labeled primers is still the workhorse for forensic and paternity testing. It is well validated, widely accepted in courts, and the infrastructure is established. Next-generation sequencing is growing, especially for degraded samples and mixed profiles, but it requires different validation, different bioinformatics, and it is nowhere near as cheap or accessible yet. RFLP analysis is essentially obsolete outside of teaching labs and historical contexts. If you are setting up a lab from scratch, the realistic path is capillary electrophoresis with a commercial STR kit. The major manufacturers all offer multiplex kits that cover the standard loci. You will need a thermal cycler, a centrifuge, a pipette set, a capillary electrophoresis instrument, and software for interpretation. The upfront cost is significant. You should also budget for continuous calibrators, positive and negative controls, and regular proficiency testing. Labs that skip proficiency testing end up with drift they do not notice until a case comes back and someone asks questions. Here is a practical breakdown of the main steps without pretending it is straightforward.
Get the Full Details
Extraction. You can use silica-based spin columns, magnetic beads, or precipitation methods. Silica columns are fast and consistent for clean samples. Magnetic beads scale better for batch processing and integrate well into automated workflows. Precipitation is cheaper but slower and more variable. If your sample has inhibitors like humic acid from soil or heme from blood on certain fabrics, you may need an additional cleanup step or a different extraction chemistry. I once spent four hours troubleshooting a failed PCR only to discover the inhibitor was coming from the filter paper the sample was dried on. Switching to a different collection medium fixed it. Quantification. Real-time PCR with species-specific and human-specific targets tells you how much DNA you have and whether it is human. This step is non-negotiable if you want a reliable profile. Skipping it means you are guessing at input amounts, and guessing leads to suboptimal amplification. Amplification. The PCR reaction mixes primers for multiple loci with fluorescent tags, polymerase, buffer, and nucleotides. A typical run takes about four hours depending on the thermal cycler and the kit protocol. The cycling conditions are strict, and deviating from them, even slightly, can cause allele drop-out or increased stutter. I have seen people change the annealing temperature by two degrees and wonder why their heterozygote balance collapsed.
Separation and detection. Capillary electrophoresis separates the amplified fragments by size. The instrument injects the sample, applies voltage, and detects fluorescence as fragments pass a laser. Run times vary but are usually between 20 and 40 minutes per sample. You need size standards in every run for calibration, and you should include internal lane controls to catch injection or separation issues. Interpretation. The software generates an electropherogram with peaks representing alleles. You assign allele calls based on the size standard. This is where experience matters. Stutter peaks can be mistaken for real alleles, especially in heterozygous pairs where the stutter from one allele lands near the other. Peak height thresholds, stutter filters, and mixture thresholds all affect the call. If you are new to this, spend time looking at trace peaks and learning what normal artifact patterns look like before you start interpreting casework. I learned this the hard way by calling a minor component in a two-person mixture and wasting days chasing a suspect who was not involved. The peak turned out to be noise. There are limitations you need to understand. DNA profiling cannot tell you where someone was or what they did. It can only tell you that biological material matching a profile was found at a location. Transfer is a real issue. Primary transfer happens when DNA is deposited directly. Secondary transfer occurs when material moves from one surface to another via an intermediary. I have seen cases where a low-level profile was attributed to direct contact when the most likely explanation was indirect transfer through a shared object. Touch DNA is sensitive but fragile, and the amount of DNA you recover does not correlate cleanly with the duration or intensity of contact.
Mixed samples are another source of trouble. When two or more people contribute DNA, the peaks can overlap in ways that make deconvolution difficult. Low-template mixtures are particularly problematic because stochastic effects like allele drop-out and drop-in become significant. If you are working with mixtures and the total DNA is below 100 picograms, treat the results with extra caution and consider a likelihood ratio framework rather than a simple qualitative call. Some labs now use probabilistic genotyping software for these cases, which helps but requires validation and a solid understanding of what the model is actually doing. Contamination is the ever-present risk. If you handle multiple samples in the same workspace without proper separation, cross-contamination is almost inevitable. I have encountered situations where a positive control leaked into a nearby sample well because a pipette tip touched the rim and carried a tiny amount of liquid. Single-use tips, filtered tips, and strict unidirectional workflow practices reduce this risk significantly. You should also run extraction blanks and negative controls with every batch. If a blank shows peaks, you have a contamination issue and you need to figure out where it came from before processing any more samples. If you are looking for a way to get started without building a full lab, some research-grade kits and services allow you to send samples out for profiling. This is common in academic and small investigative contexts. The turnaround is usually one to three weeks depending on the provider and the batch size. You lose some control over the process, but the data quality from a validated reference lab is generally reliable.
For home or hobbyist use, the options are very limited and most commercial direct-to-consumer genetic tests do not produce forensic-quality profiles. They give you ancestry information and some health data, but the marker panels are entirely different from STR profiling. If you want actual DNA fingerprinting data, you need a lab or a validated kit designed for that purpose. The field is moving toward more sensitive methods and better mixture interpretation, but the fundamentals remain the same. Good sampling, clean extraction, proper quantification, controlled amplification, careful separation, and cautious interpretation. Rush any of those and the profile you get will not save you time later. It will create work you did not need to do and answers you cannot trust.