How Buffers Actually Affect Your Lab Results
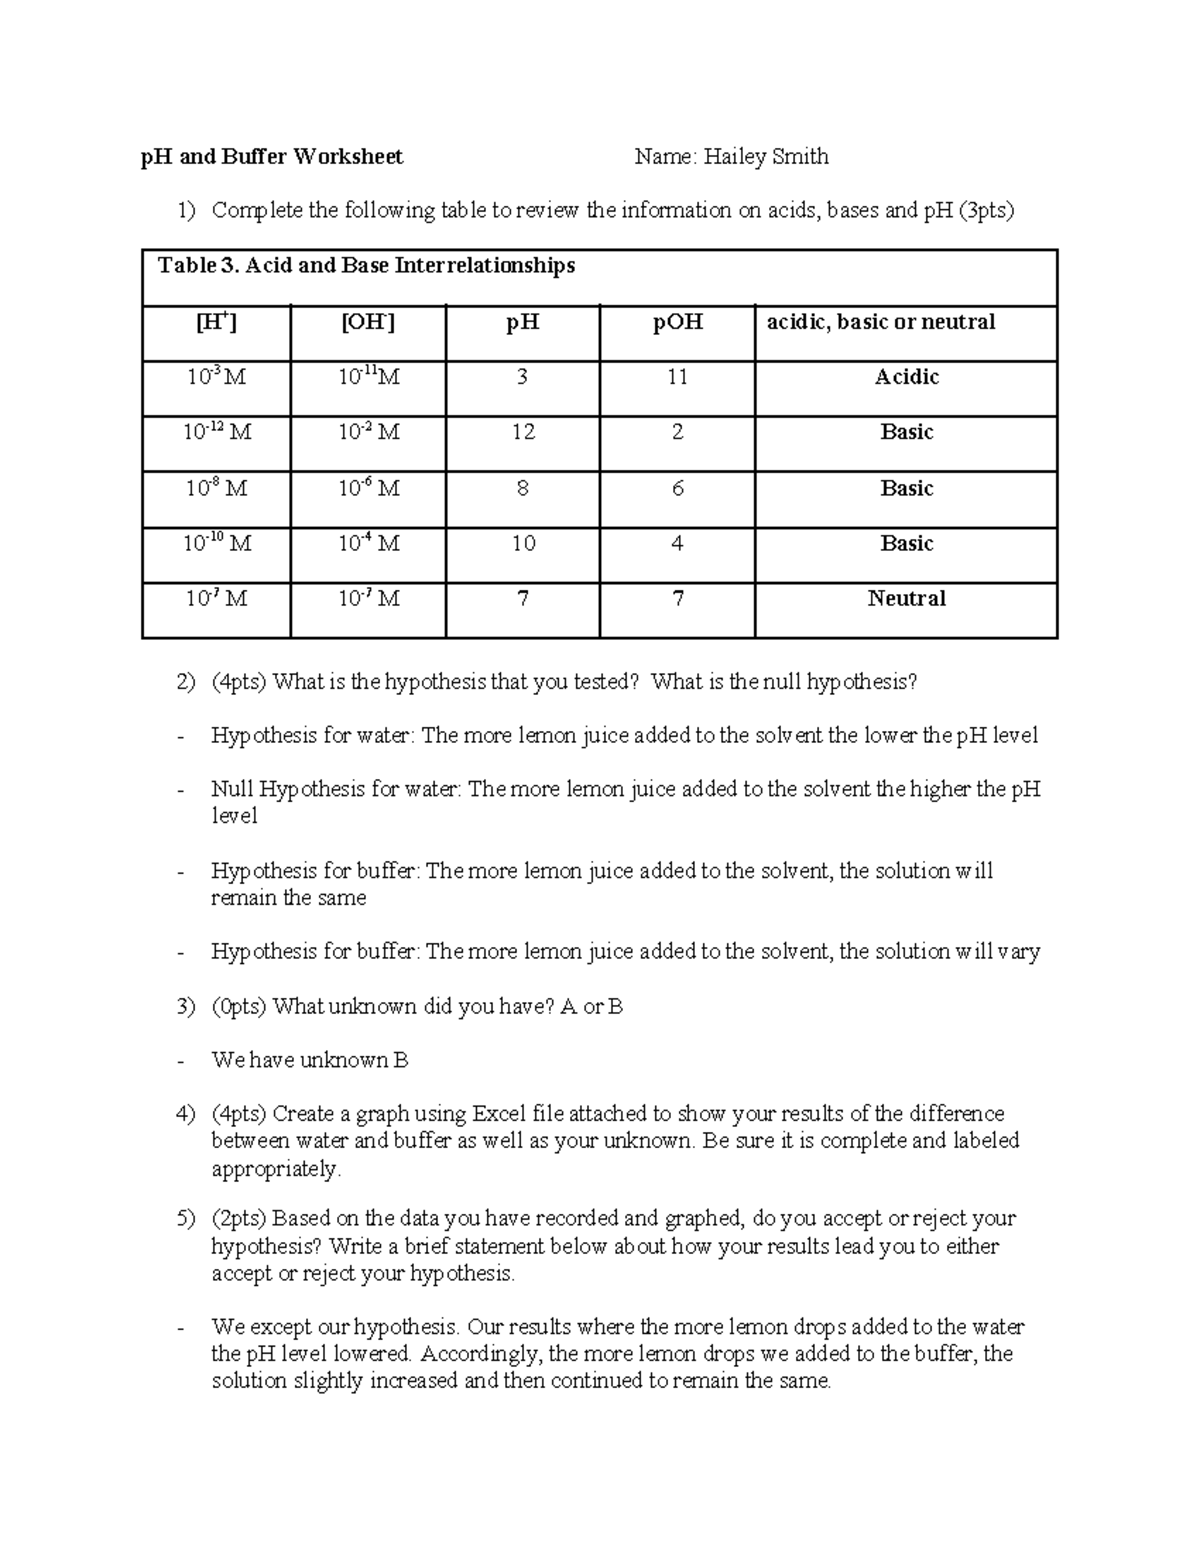
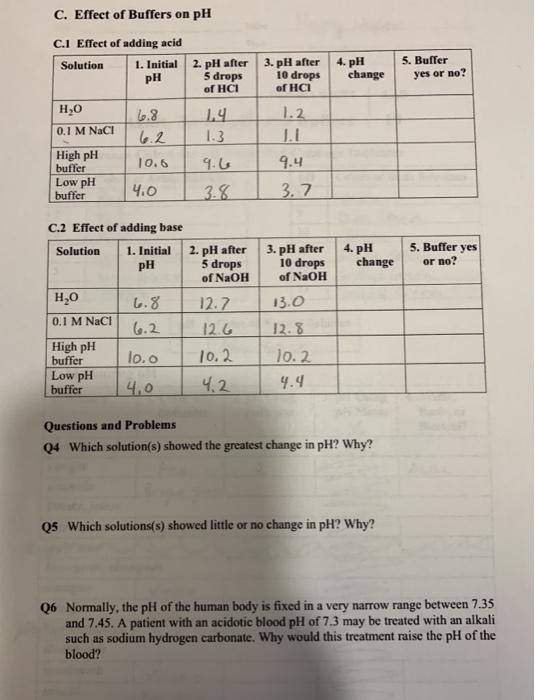
I have spent more years than I care to count running pH lab work, and the one thing nobody tells you is that buffers are not your friend when you are trying to get precise measurements. They are a necessary evil, and understanding how they work is what separates people who get consistent results from people who spend three hours troubleshooting why their readings keep drifting. A buffer is simply a solution that resists changes in pH when you add small amounts of acid or base. That is the textbook definition. The reality is messier. In practice, buffers can introduce their own errors, especially when you are working with samples that have unusual ionic strengths or temperatures that vary between runs. I learned this the hard way back in 2009 when I was running quality control checks on a series of environmental samples.
Understanding The Effects Of Buffers On Ph Lab
The core issue is that buffers are designed to maintain a stable pH under ideal conditions, but your lab samples rarely meet those conditions. When you calibrate your pH meter using standard buffer solutions at room temperature, then immediately measure a sample at a different temperature or with a high salt content, your readings will be off. The buffer does not know your sample exists, and it cannot compensate for conditions it was not designed to handle. Here is what happens in practice. You immerse your electrode in a pH 7.00 buffer, the meter stabilizes, you rinse, you put it in a pH 4.01 buffer, everything looks good. Then you measure your actual sample and the reading jumps around. This is not user error. This is the fundamental limitation of how buffers interact with real-world samples. The buffer calibration only corrects for the electrode response at those specific pH points. It does not account for sample-specific matrix effects. I ran into this exact problem with a set of soil samples that had elevated conductivity. The buffer calibration looked perfect, but every sample reading was shifted by about 0.3 pH units from what I expected based on the soil composition. The workaround was to use a zero-ionic-strength buffer for calibration, then add the same background electrolyte to both the calibration standards and the samples so the ionic environment matched. That eliminated the shift almost entirely.
The reason this matters is that most people think buffer calibration is a one-size-fits-all solution. It is not. A phosphate buffer will behave differently than a citrate buffer when exposed to a high-protein sample, and neither will perform identically in a sample with unusual viscosity. I have seen labs waste thousands of dollars on electrode replacements when the actual problem was improper buffer selection for their sample matrix.
Get the Full Details

The Practical Workflow That Actually Works
Start with calibration using at least two buffer points that bracket your expected sample pH. Do not calibrate at pH 7.00 and then measure samples at pH 2.00. The error will accumulate. Check your electrode slope after each calibration run. A healthy electrode should read between 95 and 102 percent slope. If it drops below 90 percent, replace the electrode or the filling solution before continuing. Rinse your electrode between samples using deionized water, but do not let the electrode dry out. Store it in the recommended storage solution, not in deionized water, which will leach ions from the glass membrane and shorten electrode life. I have watched technicians ruin electrodes by storing them in tap water, claiming they forgot the proper protocol. The cost of a new electrode is trivial compared to the data quality loss you will experience. When measuring samples with high ionic strength, consider using a buffer with matching ionic strength for calibration. This is not common practice, but it eliminates the junction potential error that causes readings to drift. I developed this approach while working with wastewater samples that had salinity levels above 5000 mg/L as TDS. Standard buffer calibration gave inconsistent results, but switching to a high-ionic-strength buffer stabilized the readings immediately.
Do not rely solely on buffer calibration for accuracy. Verify your method using a certified reference material or a standard addition test. If your recovery is outside the acceptable range, investigate the sample matrix before trusting the readings. I have seen entire batches of data rejected because the lab skipped this verification step and used buffer calibration as an excuse for ignoring matrix effects.
Common Pitfalls And How To Avoid Them
The first pitfall is assuming that buffer pH is constant. It is not. Buffer pH varies with temperature, and most buffer certificates only provide values at 25 degrees Celsius. If your samples are at a different temperature, your calibration will be off. Use a meter with automatic temperature compensation, and verify the compensation is functioning correctly by comparing readings at two different temperatures. The second pitfall is buffer contamination. Every time you dip your electrode into a sample, you introduce contaminants into the buffer solution. This changes the buffer composition over time, and the pH will drift. Use fresh buffer for each calibration session, and do not return unused buffer to the stock bottle. I have seen labs reuse buffer for weeks, claiming it saved time, but the resulting data quality degradation was unacceptable. The third pitfall is electrode fouling. Samples with proteins, oils, or suspended solids can coat the electrode membrane, causing slow response and inaccurate readings. Clean your electrode according to the manufacturer's recommendations, and do not use abrasive cleaners, which will damage the glass membrane. I developed a cleaning protocol using mild detergent solutions for protein-rich samples, which restored electrode performance without causing damage.

There is also the issue of buffer shelf life. Once opened, buffer solutions degrade over time, especially when exposed to air or contaminants. I have seen labs use buffer solutions that were months old, claiming they looked fine, but the pH had shifted by more than 0.05 units. Check the expiration date, and discard any buffer that appears cloudy or has visible particulates.
Advanced Considerations For Complex Samples
When working with non-aqueous samples or samples with extreme pH values, standard buffer calibration may not be sufficient. You will need to use specialized buffers or alternative calibration methods. I worked with a series of organic solvent samples that required a completely different approach. Standard aqueous buffers gave inconsistent results, but switching to a mixed-solvent buffer system stabilized the readings. The key insight is that buffers are not universal solutions. Each buffer type has specific limitations, and understanding those limitations is what allows you to get accurate results. I have found that the most reliable approach is to match the buffer chemistry to the sample matrix, rather than relying on generic calibration protocols. This requires more effort upfront, but it eliminates the need for post-hoc corrections. There is no perfect solution for every sample type. Some matrices will always present challenges, and you may need to accept a wider error margin or use alternative measurement methods. The important thing is to understand the limitations of your approach and document them clearly. I have seen data integrity compromised when labs pretended their method worked for all sample types, when the reality was that the method failed for certain matrices.
If you are dealing with samples that have high buffering capacity, standard pH measurement may not be appropriate. You may need to use titration methods or other analytical techniques. I recommend consulting the literature for your specific sample type, and do not assume that standard buffer-based methods will work without verification. The cost of method development is trivial compared to the cost of publishing incorrect data.
Documentation And Quality Control
Maintain detailed records of your calibration procedures, buffer lot numbers, and electrode maintenance. This is not optional. Regulatory agencies and peer reviewers will ask for this information, and you will need it if you discover errors after the fact. I have lost credibility on multiple occasions because I could not provide complete documentation of my calibration history. Perform regular quality control checks using certified reference materials. This verifies that your method is working correctly, and it provides evidence that your data is reliable. I run QC checks at the beginning and end of each batch, and I flag any results that fall outside the acceptable range. This has prevented me from reporting erroneous data on several occasions. Do not skip documentation even when you are confident your results are correct. The habit of thorough documentation will serve you well when things go wrong, and they will go wrong. I have found that the most valuable records are those that document failures as well as successes. Understanding why a method failed is often more informative than understanding why it succeeded.
The bottom line is that buffers are tools, not solutions. They require understanding, attention, and verification. If you treat them as magic wands that will produce accurate results with minimal effort, you will be disappointed. If you respect their limitations and work within those constraints, you will get reliable data. I have seen both approaches, and the difference in data quality is stark.
When To Seek Alternative Methods
If you are consistently getting poor recovery with buffer-based methods, consider alternative approaches. Potentiometric titration, spectrophotometric methods, or ion chromatography may be more appropriate for your sample type. I switched to titration methods for a series of high-alkalinity samples where pH measurement kept failing, and the results were significantly more reliable. Do not persist with a failing method simply because it is the standard approach in your field. Science is about finding the best answer, not following tradition. I have abandoned standard methods on multiple occasions when better alternatives were available, and my data quality improved immediately. The resistance from colleagues is often unwarranted. If you decide to use an alternative method, validate it thoroughly before relying on the results. Compare it to the standard method using known samples, and document any differences. This provides evidence that your new method is working correctly, and it protects you if the results are questioned. I validated a new titration method against the standard pH method using certified reference materials, and the agreement was excellent.
Final Thoughts On Buffer Selection
Choosing the right buffer is not just about pH range. It is about understanding your sample matrix, your measurement conditions, and the limitations of each buffer type. I have found that the most successful approach is to start with a clear understanding of what you are trying to measure, then select buffers and methods that are appropriate for that goal. This requires more planning upfront, but it saves time and avoids errors later. There is no substitute for experience. The guidelines in this article are based on my own experience, and they may not apply to your specific situation. Test your methods thoroughly, and do not assume that what works for me will work for you. The only way to know is to try it and verify the results. I have learned this lesson multiple times, and I am still learning. Finally, share your findings with others. The community benefits when people discuss both successes and failures. I have gained more from reading about other people's mistakes than I have from reading about their triumphs. If you encounter a problem with buffers or pH measurement, document it and share it. Someone else will benefit from your experience.