Working with Standard Enthalpies of Formation in Practice
Most people pull up a table, look up values, and plug them into H°rxn = nH°f(products) mH°f(reactants). That part is straightforward enough. The table itself is just a list of numbers, usually organized by compound name, state of matter, and temperature, typically 298.15 K. The real work starts when you realize how many traps are sitting between looking up a value and getting a correct answer.
Where to Find a Reliable Enthalpy Of Formation Table
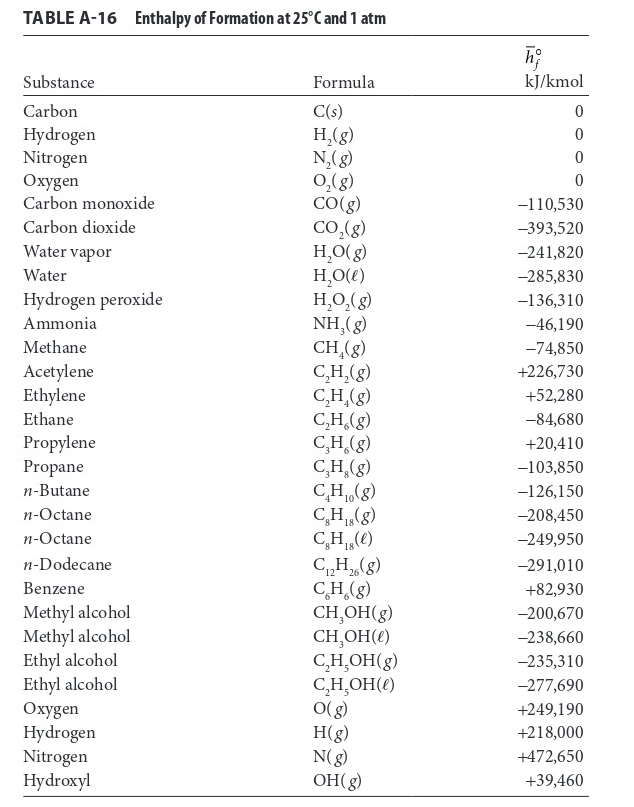
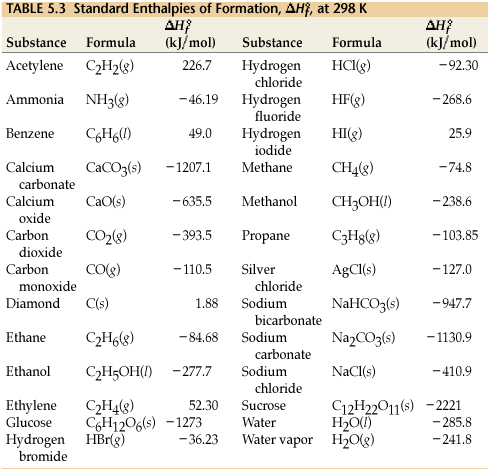
I use the NIST Chemistry WebBook as my primary source. It's free, well-curated, and the data goes back decades. The CRC Handbook of Chemistry and Physics is also solid if you're working offline. Avoid random educational sites that reprint tables without citing sources — I've caught at least three errors across two different "free download" pages where water's H°f was listed as 241.8 kJ/mol instead of 285.8 kJ/mol, and nobody noticed because they didn't check which phase the value corresponded to.
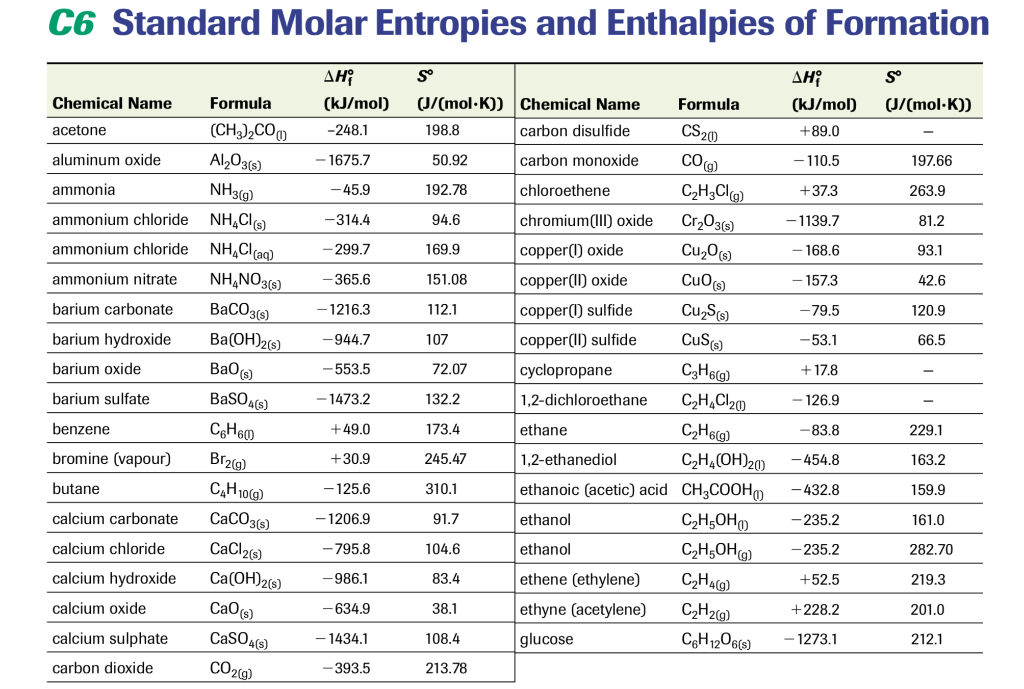
Here's the thing most students miss: the enthalpy of formation for an element in its standard state is zero, but only if it's actually in its standard state. Oxygen as O(g) is zero. Ozone, O(g), is +142.7 kJ/mol. Carbon as graphite is zero. Carbon as diamond is +1.9 kJ/mol. If your problem uses diamond and you pull the graphite value, your answer will be wrong by however many moles of carbon you're working with, and you won't know it happened.
I ran into this exact issue last year on a combustion problem involving carbon nanotubes. The question didn't specify allotrope, and I just pulled the graphite value out of habit. The answer came out about 3% off from the reference solution. It took me twenty minutes to realize what happened — the problem was implicitly referencing the nanotube structure, which has a small but nonzero heat of formation relative to graphite. I had to estimate it using group contribution methods as a workaround since no single measured value existed. If you're ever in that situation, the Joback method or the Benson group additivity scheme can give you a reasonable estimate within ±5 kJ/mol, which is usually good enough for homework-level work.
Phase matters enormously. Water as a liquid has H°f = 285.8 kJ/mol. Water as a gas is 241.8 kJ/mol. That's a 44 kJ/mol difference, which is exactly the enthalpy of vaporization at 298 K. If a reaction produces water vapor and you use the liquid value, your H°rxn will be off by 44 times the number of moles of water produced. This is the single most common error I see, and it's not even close.
Hess's Law Calculations Step by Step
Write the balanced equation first. Make sure it's balanced — I know that sounds obvious, but I've seen people use a 2:1 ratio when the actual stoichiometry was 1:1, cascading every subsequent calculation wrong. Look up each H°f value with the correct state notation. Multiply each by its stoichiometric coefficient. Sum the products. Sum the reactants. Subtract. The sign convention trips people up less now than it used to, but it still happens: products minus reactants, not the other way around.
Let me walk through a concrete example. Combustion of methane:
CH(g) + 2O(g) CO(g) + 2HO(l)
The values from NIST at 298.15 K:
- CH(g): 74.8 kJ/mol
- O(g): 0 kJ/mol
- CO(g): 393.5 kJ/mol
- HO(l): 285.8 kJ/mol
products = (1 × 393.5) + (2 × 285.8) = 965.1 kJ
reactants = (1 × 74.8) + (2 × 0) = 74.8 kJ
H°rxn = 965.1 (74.8) = 890.3 kJ/mol
That matches the literature value. If you'd used HO(g) instead, you'd get 802.3 kJ/mol, which is a substantial difference and completely wrong for liquid water as the product.
I once had a student who got 802 kJ/mol on this exact problem and couldn't figure out why the textbook said 890. She'd looked up water correctly but grabbed the gas value without thinking about the phase. We spent ten minutes on it. The table was right there in front of her — the gas and liquid entries are usually listed separately, often right next to each other. But under time pressure, people read what they expect to see.
Temperature Dependence and When the Table Fails You
Standard tables are almost always at 298.15 K. If your reaction runs at 500 K or 1000 K, those values don't apply directly. You need to integrate the heat capacity over the temperature range. The formula is:
H°(T) = H°(298) + [Cp(products) Cp(reactants)] dT from 298 to T
Cp values are typically given as polynomial functions of temperature — usually a Shomate equation or a simple polynomial like Cp = a + bT + cT² + dT³. NIST provides these coefficients for most common compounds. The integration itself is mechanical but tedious to do by hand, which is why I just write a quick Python script or throw it into a spreadsheet. The actual calculation takes about thirty seconds once the coefficients are in front of you.
There are cases where this approach breaks down entirely. Phase transitions are the main culprit. If a reactant or product melts or boils between 298 K and your target temperature, you need to add the latent heat of transition as a discrete term. Forgetting that term is another classic error, and it shows up in exam problems with surprising regularity. I've seen questions where water goes from liquid to gas between 298 K and 400 K, and students who skip the vaporization enthalpy end up off by 44 kJ/mol again.
Another limitation: the standard tables don't cover unstable intermediates or transient species well. If you're working on a mechanism involving something like a radical or a high-energy intermediate, the H°f value might not exist in any published table. In those cases, you're forced to use computational chemistry — DFT calculations at the B3LYP/6-31G* level or higher, sometimes coupled with isodesmic or isomerization reactions to cancel out systematic errors. This adds hours or days to what should be a quick lookup, and the results carry their own uncertainties, typically ±5 to ±15 kJ/mol depending on the method and the system size.
Common Pitfalls to Watch For
Solution-phase values vs. gas-phase values. Some tables list H°f for aqueous ions, others for neutral gas-phase molecules. If your reaction is in solution and you pull a gas-phase value, you've ignored the solvation enthalpy, which can be substantial. Hydrochloric acid is a good example: H°f for HCl(g) is 92.3 kJ/mol, but for H(aq) + Cl(aq) it's 167.2 kJ/mol combined. That's a 75 kJ/mol difference just from dissolving the gas.
Sign errors in the subtraction step. The formula is products minus reactants, and the reactant sum often includes negative numbers. When you subtract a negative sum, you're adding. I still catch people writing 393.5 + (74.8) instead of 393.5 (74.8). It's a arithmetic slip, but it ruins the entire answer.
Using average bond enthalpies when you have formation data. Average bond enthalpies are approximate — they're averaged across many different molecules, so they carry typical uncertainties of ±10 to ±20 kJ/mol per bond. If you have H°f values available, use them. Bond enthalpies should be a fallback, not a primary method. I see this mistake constantly in introductory courses where both methods are taught, and students pick the wrong one because it seems faster. It is faster, but the accuracy cost is real.
Not checking units. Some tables use kJ/mol, some use kcal/mol, and a few older sources still use cal/mol. If you mix units without converting, your answer will be wrong by a factor of 4.184 or more. I once graded a set of problem sets where half the class used a table in kcal and the other half in kJ, and the instructor hadn't noticed because the numerical answers happened to look plausible. It took me two hours to trace back where the discrepancy originated.
Building Your Own Reference File
If you're doing this work regularly, copying values from NIST every time is inefficient. I keep a personal CSV file with the most common compounds — about 150 entries covering hydrocarbons, common inorganics, aqueous ions, and a handful of intermediates. Each row has the compound name, formula, state, H°f, G°f, and Cp coefficients. The file takes about three hours to build the first time, but after that, I can run through a multi-step thermochemistry problem in under five minutes because I'm not hunting through webpages between each lookup.
A spreadsheet with named ranges or a simple lookup table is sufficient. I've also seen people write small utilities in Python using the Cantera package, which loads thermodynamic data directly from NASA polynomials and handles temperature corrections automatically. That's overkill for occasional use but cuts calculation time to near zero for routine work once it's set up.
The Enthalpy Of Formation Table is fundamentally just a starting point. The actual skill is knowing which value to trust, when to adjust for temperature, and how to catch the mistakes that creep in during the transcription from table to calculation. Most errors aren't conceptual — they're transcription errors, phase mismatches, or sign mistakes. Slow down on those steps and the rest follows naturally.