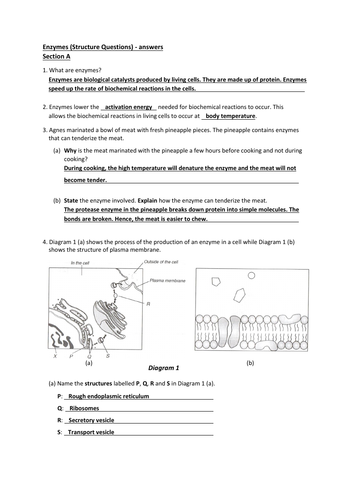
Working With Enzymes In Practice
I have spent more years than I want to admit troubleshooting enzyme kinetics assays in diagnostic labs and research settings. The theory is straightforward—proteins that catalyze reactions—but the practical side has plenty of rough edges. People ask me about enzymes constantly, usually because they are stuck on why their assay data looks wrong or how to interpret Vmax and Km values from a messy experiment. The first question I get is usually about enzyme specificity. Enzymes are not just catalysts that speed everything up; each one has a precise active site geometry that determines which substrates it will bind and react with. I remember running a phosphatase assay once where the phosphate buffer completely inhibited the reaction. Not a partial inhibition—a total shutdown. I had used standard lab phosphate buffer without thinking about it. Switching to Tris buffer fixed the problem immediately. That is the kind of thing you learn through painful trial and error. Then there is the question of how enzymes actually work at the molecular level. They lower the activation energy of a reaction by stabilizing the transition state. The substrate binds to the active site, forming an enzyme-substrate complex, and the reaction proceeds faster than it would on its own. After the product releases, the enzyme is unchanged and ready for another cycle. This is standard biochemistry, but the details matter when you are designing an actual experiment.
Kinetic Parameters and What They Mean
Km and Vmax are the numbers you will see everywhere. Km is the substrate concentration at which the reaction runs at half its maximum velocity. It is often mistaken for a measure of affinity, but that is only approximately true under certain conditions. A low Km generally indicates tighter binding, but allosteric enzymes and multi-substrate reactions complicate that picture significantly. Vmax is the maximum rate when every enzyme molecule is saturated with substrate. In practice, you rarely reach true Vmax because substrate solubility limits or enzyme instability intervene first. I have seen researchers extrapolate Vmax from data that barely reached steady state, and the resultingKm values were nowhere near accurate. The workaround is to ensure you are measuring initial rates under conditions where substrate depletion is less than five percent over the measurement period.
Common Inhibition Types and How to Distinguish Them
Competitive inhibition is the simplest case. The inhibitor competes with substrate for the active site, increasing apparent Km without changing Vmax. You can overcome it by adding more substrate. This is the mechanism behind many drugs, including statins and various kinase inhibitors used in cancer therapy. Noncompetitive inhibition is trickier. The inhibitor binds at a different site and reduces Vmax without affecting Km. Adding more substrate does nothing to reverse it. Mixed inhibition sits somewhere in between, affecting both parameters. I once spent three days trying to figure out why my inhibitor data did not fit any standard Lineweaver-Burk plot. The problem turned out to be that the compound was aggregating at higher concentrations, creating artifactual inhibition patterns. Running dynamic light scattering before the kinetics experiment would have saved me a week.
Get the Full Details
Practical Issues With Enzyme Assays
Enzyme stability is a constant headache. Many enzymes lose activity within hours if you keep them on ice without stabilizers. I typically add ten percent glycerol and work at four degrees Celsius whenever possible. Some enzymes require specific cofactors or metal ions to remain active. Magnesium is essential for most kinases, and zinc appears in the active site of carbonic anhydrase and many proteases. Forgetting to include the right cofactor in your buffer is an easy way to get zero activity and waste several days of work. pH sensitivity is another factor I wish people appreciated more. Enzymes have narrow optimal pH ranges, and even small deviations can dramatically reduce activity. The histidine residues in many active sites change protonation state around pH six to seven, which explains why many enzymes drop off sharply outside their optimal range. I always check pH at the working temperature, not room temperature, because pH changes with temperature for most buffer systems.
Understanding Michaelis-Menten Kinetics Proper
The Michaelis-Menten equation describes hyperbolic saturation kinetics for single-substrate reactions under steady-state assumptions. It assumes the enzyme-substrate complex reaches a constant concentration and that product formation is irreversible during the initial rate measurement. Those assumptions break down in several common scenarios. Pre-steady-state kinetics, where you observe the burst phase before equilibrium, requires stopped-flow equipment and gives you information about individual catalytic steps rather than overall parameters. Products that inhibit the enzyme violate the irreversibility assumption. Many dehydrogenases are subject to product inhibition by NADH, which is why coupled enzyme assays exist—to keep product concentrations low by immediately converting them to another measurable reaction. I rely on coupled assays whenever possible because they extend the linear range and reduce noise from product accumulation.
When Enzyme Data Looks Wrong
If your progress curve is not linear during the initial rate phase, check for substrate depletion, enzyme instability, or product inhibition. The rule of thumb is to measure rates during the first five to ten percent of substrate conversion. Beyond that, things start changing enough to distort the kinetics. Another frequent issue is enzyme adsorption to tube walls. At low enzyme concentrations, a significant fraction can stick to plastic surfaces, reducing the effective enzyme concentration in solution. Using silicone-coated tubes or adding a small amount of detergent like 0.01 percent Tween-20 can solve this problem. I learned this the hard way when my enzyme activity appeared to decrease linearly with dilution, which should never happen for a true catalyst.

Allosteric Enzymes and Cooperative Binding
Not all enzymes follow Michaelis-Menten kinetics. Allosteric enzymes show sigmoidal saturation curves because binding at one subunit affects the affinity of other subunits. Hemoglobin is the classic example, though it is not technically an enzyme. Aspartate transcarbamoylase and phosphofructokinase are enzyme examples that display cooperativity. The Hill coefficient quantifies the degree of cooperativity. A value greater than one indicates positive cooperativity, while less than one suggests negative cooperativity. Fitting Hill equations to experimental data requires careful consideration of confidence intervals, because the Hill model is phenomenological rather than mechanistic. It describes the shape of the curve without explaining why the cooperativity exists at the molecular level.
Enzyme Purification and Activity Measurements
Specific activity, measured in micromoles of product per minute per milligram of protein, is the standard metric for enzyme purity. If your specific activity does not increase through purification steps, you are either not removing contaminants or you are losing enzyme along with the impurities. I track yield and specific activity at every step, because the fold-purification number alone can hide catastrophic losses. Protein concentration determination introduces its own errors. Bradford assays interact differently with various proteins depending on their aromatic amino acid content. Absorbance at 280 nanometers requires knowledge of the extinction coefficient, which you can calculate from the amino acid sequence but may differ from the theoretical value if the protein has unusual disulfide bonds or post-translational modifications. I typically cross-check concentration measurements with at least two methods before trusting the data.
Modern Applications and Assay Development
High-throughput screening has changed how enzymes are used in drug discovery. Automated liquid handlers, plate readers, and miniaturized reactions allow thousands of compounds to be tested against a target enzyme in a single day. The challenge is maintaining kinetic relevance at small volumes and short incubation times. Surface plasmon resonance and isothermal titration calorimetry provide binding data that complements kinetic measurements but requires different instrumentation and expertise. Directed evolution has become a powerful tool for improving enzyme properties. By introducing random mutations and screening for desired activities, researchers have created enzymes with improved stability, altered substrate specificity, and enhanced catalytic efficiency. Frances Arnold won the Nobel Prize for this work, and the techniques are now standard in many industrial biotechnology applications. The process is iterative and labor-intensive, but modern automation has reduced the time required from months to weeks for well-designed screening campaigns.

Enzymes Questions And Answers for Beginners
The most important thing to understand is that enzyme kinetics is experimental, not theoretical. Equations describe idealized behavior, but real enzymes behave according to the conditions you create. Buffer composition, temperature, ionic strength, and even the quality of water used to prepare reagents can affect your results. I always include appropriate controls and repeat experiments because enzyme preparations vary between batches in ways that are difficult to predict. If you are just starting with enzyme work, begin with well-characterized systems like alkaline phosphatase, luciferase, or horseradish peroxidase. These have robust assays, published protocols, and predictable behavior. Once you understand the basics, move to your protein of interest and invest time in optimizing conditions before generating kinetic data. The time you spend on optimization will save you weeks of troubleshooting later.