The Actual Process Nobody Warns You About
Separating layers in a separatory funnel sounds simple until you're standing there at 11pm with a cloudy interface and no idea which layer is which. I've wasted enough solvent and product to know that this is where most people hit problems, not during the extraction itself but during the washing and final separation. Here's what actually happens when you run through a standard liquid-liquid extraction with organic and aqueous layers, and where things go sideways.
Extraction And Washing Organic Vs Aqueous Layer
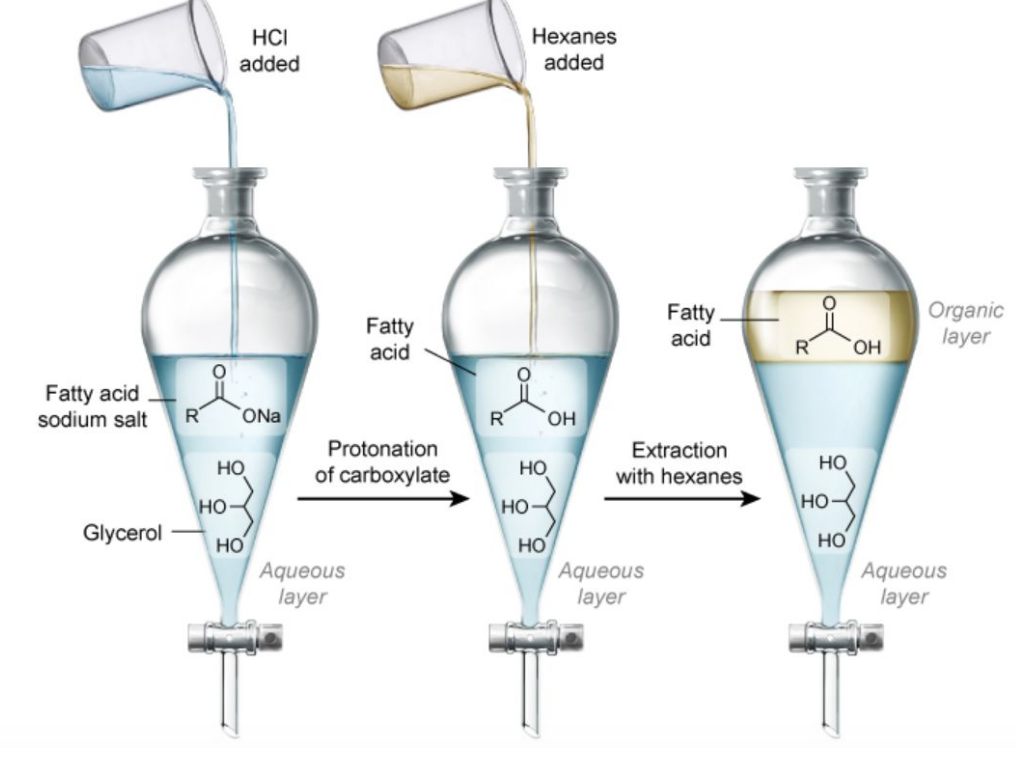
You start by choosing your solvent system. This is the part that matters most and the part people rush. If your target compound is polar, you might use water and ethyl acetate. Non-polar compounds often pair with hexanes or dichloromethane and water. The rule of thumb is simple: like dissolves like, but the devil is in the details of your specific molecule's logP value. Mix the layers thoroughly. Close the funnel, invert it, and vent immediately. Gas pressure builds up fast, especially with volatile solvents like DCM or ethyl acetate. I learned this the hard way after I had solvent spraying into my fume hood because I didn't vent within the first five seconds of shaking. Open the stopcock carefully to release pressure, close it, shake again, vent again. Repeat about four or five times for a standard extraction. Don't over-shake. Vigorous agitation creates emulsions that can take hours to separate. Let the layers settle. This is where patience pays off. A clean separation usually takes two to five minutes depending on your solvent pair. DCM and water separate faster than ethyl acetate and water because of the density difference. If you're seeing a cloudy middle layer or droplets suspended in one phase, that's an emulsion forming. Don't panic. Add a small amount of saturated sodium chloride solution and swirl gently. The salt helps break the emulsion by increasing the ionic strength of the aqueous phase and reducing the solubility of organic compounds in water. This alone has saved me from losing material more times than I can count.
Now for the critical moment: identifying which layer is which. This is where beginners lose product. You need to know the densities. DCM is denser than water, so it sits on the bottom. Ethyl acetate is less dense than water, so it floats on top. Hexanes definitely float. If you're using something unusual or a solvent mixture, test it. Add a few drops of water to the funnel and watch where they go. If the drops dissolve into the top layer, the bottom is your organic phase. If they fall through and dissolve into the bottom layer, the top is organic. This takes ten seconds and prevents catastrophic mistakes. Drain the bottom layer into a clean flask. Keep the top layer in the funnel if you need to do a second extraction, which you should. Run the process again with fresh solvent. Two extractions with half the volume each are more efficient than one extraction with the full volume. This is basic partition coefficient math, but people still do single extractions and wonder why their yield is lower than expected. Combine your organic extracts. Now you wash them. Washing removes residual aqueous-soluble impurities. Common washes include saturated sodium bicarbonate to neutralize acids, brine to help with phase separation and remove water, and sometimes dilute acid if you need to remove basic impurities. Each wash goes through the same process: add the wash solution, shake gently with venting, let separate, drain layers. Use roughly equal volumes to your organic layer for each wash step.
Get the Full Details

After your final wash, dry the organic layer over anhydrous sodium sulfate or magnesium sulfate. Swirl it around and let it sit for about ten minutes. The drying agent should flow freely like sand. If it's clumped together, you need more. Filter or decant the dried organic solution into a round-bottom flask and rotate evaporate to remove the solvent.
Edge Cases That Actually Matter
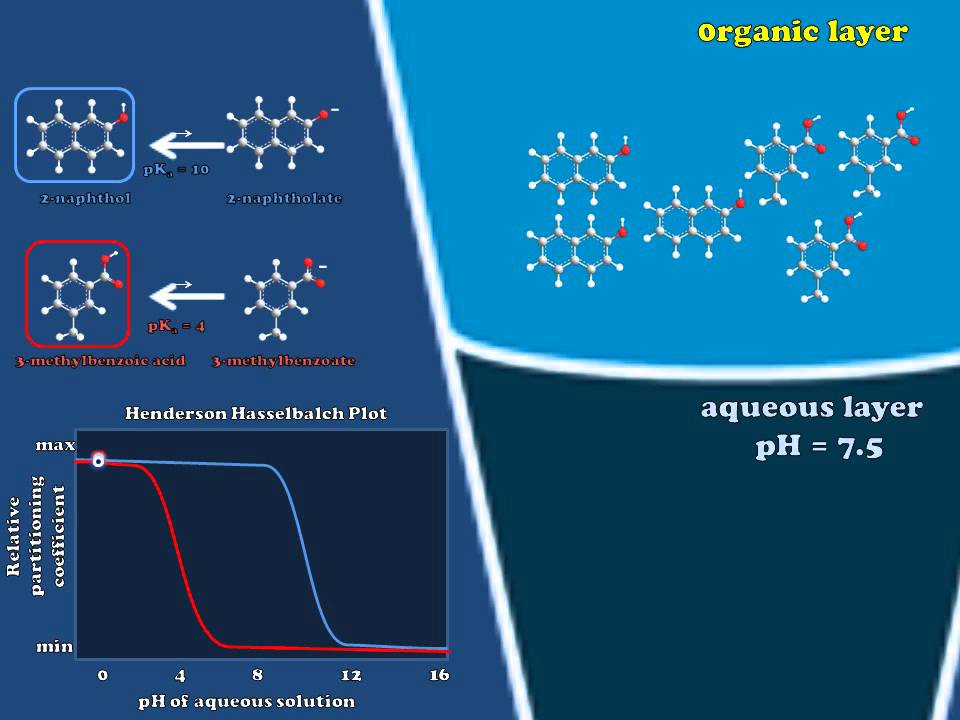
Here's something you won't find in most lab manuals. When extracting basic compounds, the pH of your aqueous layer controls everything. If you're working with an amine and your aqueous phase isn't basic enough, your compound will stay protonated in the water and never move into the organic layer. I once spent three hours trying to extract a product that was stuck in the aqueous phase because I'd used the wrong pH adjustment. Adding a few drops of concentrated sodium hydroxide to the aqueous layer and re-extracting pulled the product out cleanly. The fix was trivial, but I wouldn't have known to look there without wasting half a day first. Another issue that catches people off guard: some compounds have similar solubility in both layers regardless of what you do. If you're consistently getting poor partitioning, consider changing your solvent system entirely rather than doing more extractions. Switching from ethyl acetate to methyl tert-butyl ether or adjusting the pH can dramatically improve your recovery. There's no point in running ten extractions if your distribution coefficient is near one. The drying step deserves more attention than it gets. Over-drying with too much sodium sulfate can actually adsorb your product onto the solid surface, especially for polar compounds. Use just enough to see free-flowing powder, not a excess pile of drying agent. A quick gravity filtration through a small plug of cotton or a pipette tip packed with glass wool works fine for small scale work and loses less material than filter paper.
And one more thing about emulsions. If brine doesn't work and you're stuck with a stubborn emulsion, try adding a single drop of a surfactant-free antifoam agent or simply letting it sit overnight in the refrigerator. Cold temperatures reduce solubility and surface activity, and the emulsion usually collapses on its own. I've recovered material from emulsions that looked completely irrecoverable just by walking away and coming back the next morning. The whole process from start to finish on a standard 50 millimole scale takes about twenty to thirty minutes if you're working smoothly. The bottleneck is always waiting for layers to separate. You can reduce that time by centrifuging small-scale extractions at low speed for a minute, which forces phase separation instantly. It's not always available, but when it is, it's worth using.