Working With First Order Kinetics in the Lab
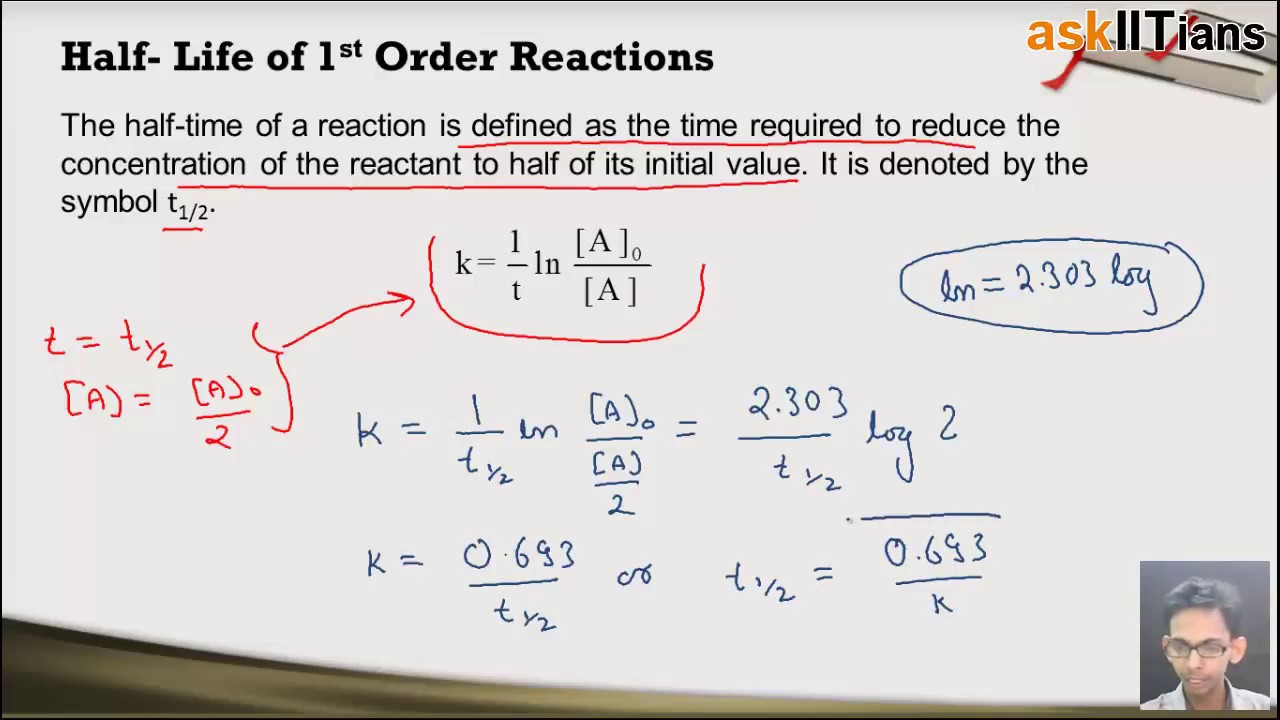
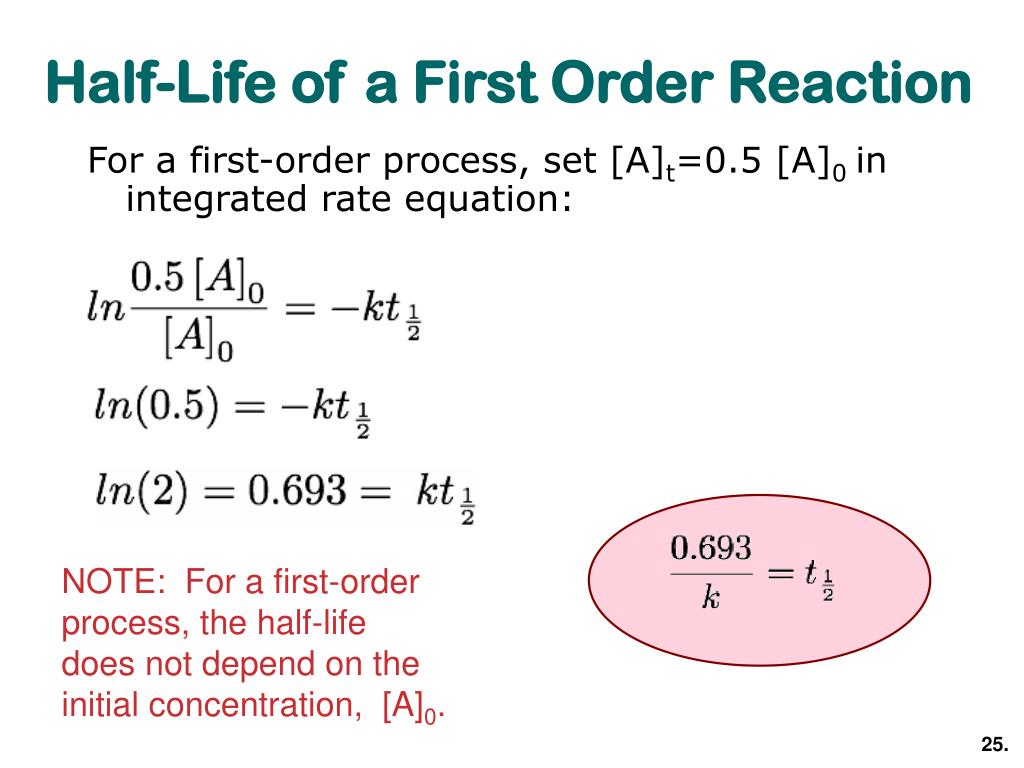
The half-life formula for a first-order reaction is t1/2 = ln(2) / k, or about 0.693 divided by the rate constant. That's it. But using it correctly in practice is where things get messy, and most people who are just learning kinetics mess it up on the first problem set. I need to nail down the method before I define anything, because the way you apply this equation tells you whether you actually understand first-order kinetics or if you're just plugging numbers into a template.
First Order Half Life Equation
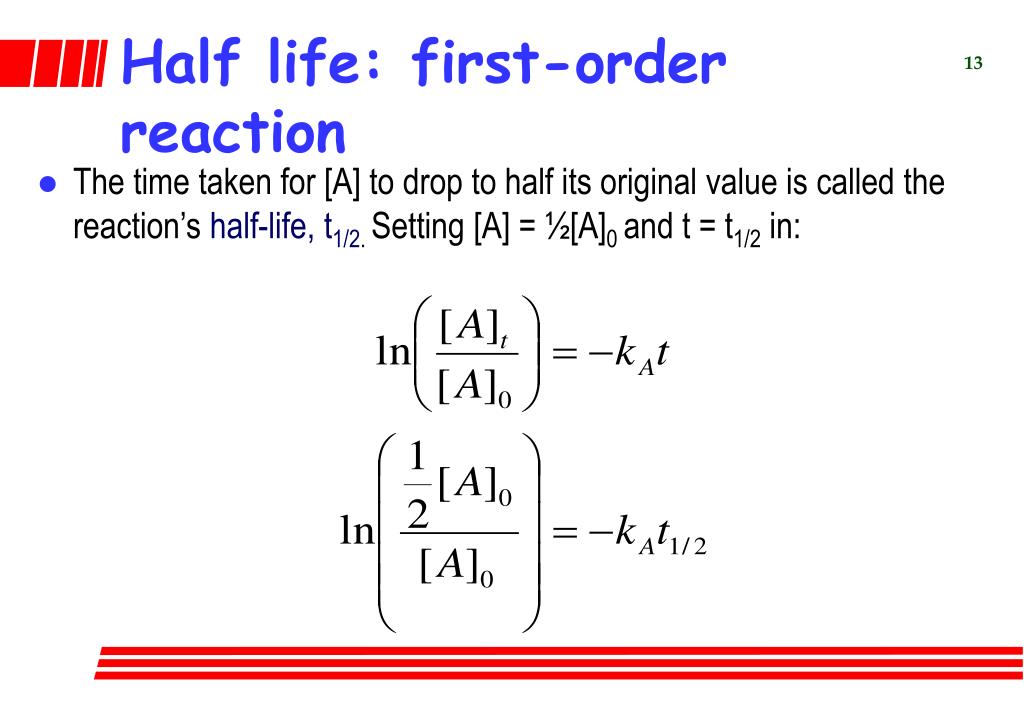
Here's how I use it. You start with your concentration data at two different time points. Let's say you have [A] at time zero and [A] at some later time t. You take the natural log of the ratio [A] at t divided by [A] at zero, and that gives you minus k times t. Rearrange for k, then drop that k into the half-life equation. Done. The equation itself doesn't care about units as long as you're consistent, which sounds obvious but causes more errors than I can count. I've seen people plug in minutes for t and then expect the half-life to come out in hours without doing the conversion. It doesn't work that way. k carries the inverse time unit, so if your time is in seconds, your half-life is in seconds. One thing beginners consistently get wrong is assuming the half-life depends on the starting concentration. For first-order reactions it doesn't. Zero-order half-lives change depending on where you start, but first-order is constant across all concentrations. That's actually the defining feature. If your experimental half-lives shift as concentration drops, the reaction isn't first-order, period. You need to rethink your mechanism or check your data.
I ran into this a few years ago with a degradation study on a pharmaceutical compound. The textbook said the reaction should be first-order, and every calculation I did suggested it was. But when I plotted the half-lives calculated from different initial concentrations, they drifted by about twelve percent across the range. I spent two days recalculating before I realized the problem was in my sampling protocol, not the chemistry. The HPLC injector was carrying over a small amount between runs, and at low concentrations that carryover was skewing the readings enough to make the kinetics look concentration-dependent. I switched to running blank injections between samples and the drift disappeared. The reaction was first-order the whole time. Another practical issue is that ln(2) is approximately 0.693147, and most people round it to 0.693. That's fine for homework. In a regulatory setting where you're reporting half-lives for drug clearance, even that small rounding can matter if you're propagating errors through multiple calculations. I keep it as ln(2) in my spreadsheets and only round at the very end. You can also work backwards from a known half-life to find k, which is what you'll do most of the time in pharmacokinetics. A drug with a six-hour half-life has a k of about 0.1155 per hour. Simple division, but again, the units are where people trip up. Make sure you write them down explicitly.
Get the Full Details

The integrated rate law for first-order reactions is ln[A] = -kt + ln[A]0. This is the equation you plot when you need to verify linearity. If you graph ln of concentration versus time and it's not a straight line, your reaction order assumption is wrong. I've seen this used as a shortcut to avoid calculating half-lives at all, but the plot tells you much more than a single half-life number ever will. It shows you whether the model fits the data across the entire range, not just at one point. There are situations where this equation breaks down completely. Enzyme kinetics following Michaelis-Menten don't follow first-order across all substrate concentrations. At high substrate levels the reaction approaches zero-order. If you're working with a system like that and you apply the first-order half-life equation blindly, you'll get answers that look reasonable but are fundamentally wrong. The same goes for autocatalytic reactions or any system where the rate constant changes over time due to product inhibition or catalyst deactivation. I also want to mention a common pitfall with radioactive decay. The first-order half-life equation works perfectly for that because nuclear decay is inherently first-order, but people sometimes try to apply it to chemical reactions that only approximate first-order behavior under specific conditions. If you're working in a regime where the reaction is pseudo-first-order because one reactant is in large excess, the half-life you calculate is only valid for those conditions. Change the excess concentration and your effective rate constant changes. It's not a true half-life, it's a conditional one.
For anyone doing this work regularly, I'd recommend keeping a reference sheet with the common conversions and the rearranged forms of the equation. The basic form t1/2 = 0.693/k is useful, but you'll also need k = 0.693/t1/2 and the concentration-time relationship [A] = [A]0 × e-kt constantly. Having them all in one place saves you from derivations that eat time and introduce rounding errors. The main thing to take away is that the equation is trivial. Understanding when it applies and when it doesn't is what actually matters. Check your linearity, verify your units, and don't trust the half-life if the plot isn't clean.