So You Need to Actually Run an HPLC Method
I have spent more years than I want to admit troubleshooting HPLC systems that were behaving as if they had a personal vendetta against me. The reality is that most people treat High Performance Liquid Chromatography like it is something you just turn on and walk away from. That approach usually works until it does not. I am going to walk through what actually matters, starting with the method setup because that is where the most problems originate. You begin with a column. Most labs default to a C18 reverse-phase column, 150 by 4.6 millimeters, five-micron particles. This is the bread-and-butter configuration for polar and moderately non-polar analytes. It handles roughly eighty percent of what people throw at it without requiring any exotic conditions. The mobile phase is typically water with some form of acid modifier and an organic modifier like acetonitrile or methanol. Let us talk about your gradient. If you are running an isocratic method and your run times are exceeding twenty minutes, you are probably doing something wrong or your sample has too many compounds with widely varying retention factors. A shallow gradient from ten percent to sixty percent organic over twelve minutes with a post-run re-equilibration of eight minutes is a reasonable starting point for an unknown mixture. That gives you a chromatogram you can actually interpret rather than a solid blob that looks like a skyline from a city center.
The flow rate for a standard four-six millimeter column is one milliliter per minute. If you move to a narrower column like a two-point-one millimeter internal diameter, you drop the flow to two hundred to three hundred microliters per minute. The detector cell volume matters here. A standard flow cell holds maybe eight microliters. At low flow rates with narrow columns, the band broadening from the cell itself becomes a real problem. I lost a resolution of about two baseline peaks once because I did not account for the extra-column volume of my detector during a method transfer from a four-six column to a two-one column.
Things That Break Methods Before You Even Start
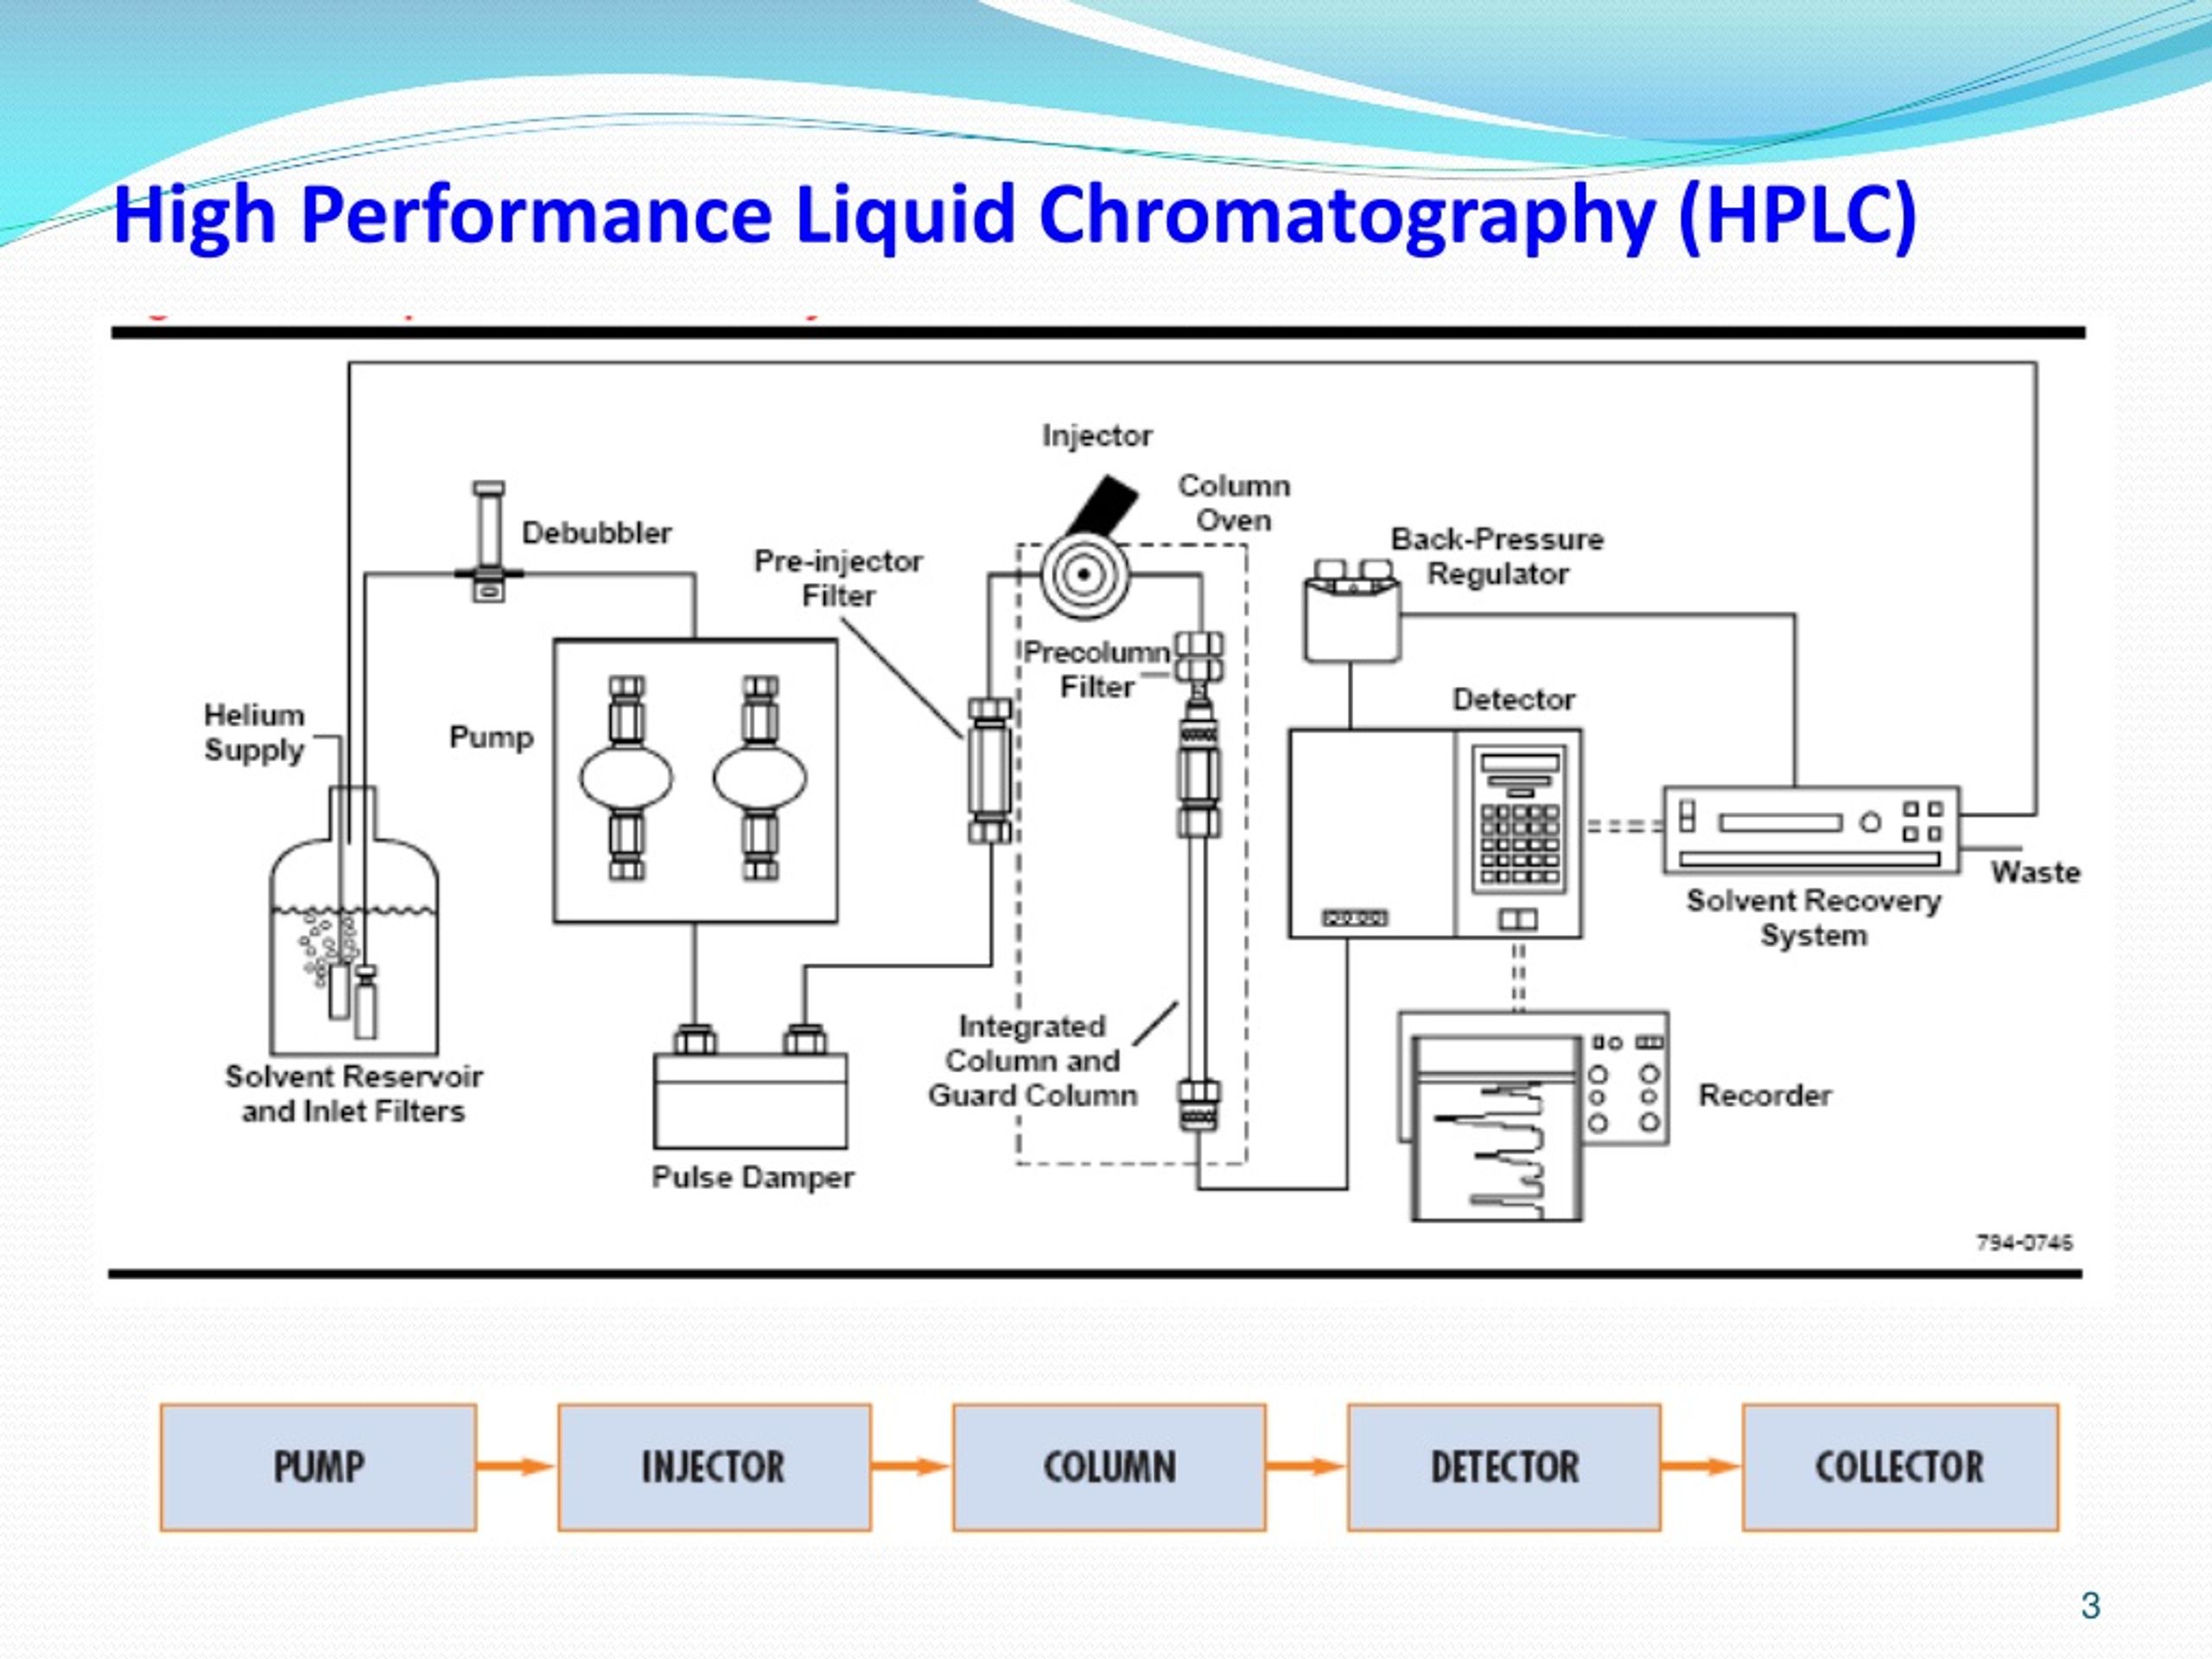
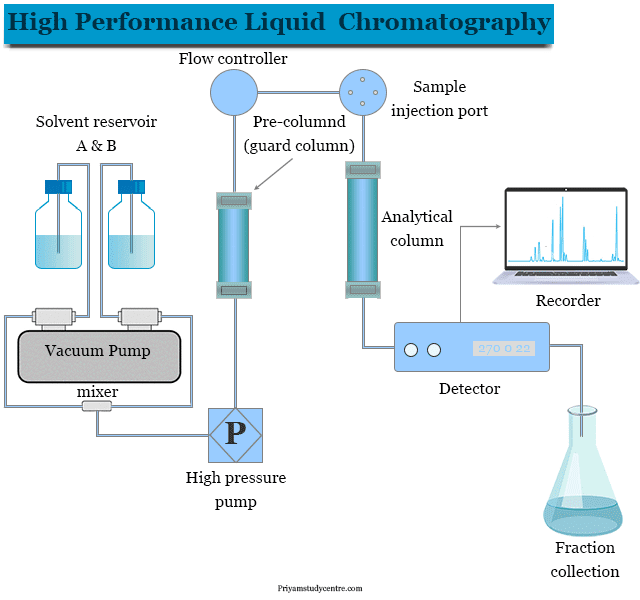
Filter your mobile phases. Three reasons. Particulates will destroy your injector seals over time. Dissolved gases will come out of solution when the pressure drops at the column inlet and create bubbles in your flow cell. And trace contamination from the solvent bottles themselves will eventually ghost onto your chromatogram. I use nylon filters for aqueous phases and PTFE for organic ones. Some people swear by PVDF. Pick one and stick with it. Using mismatched filter materials with certain solvent combinations can lead to filter degradation and the filter itself becomes your contamination source. Degassing is non-negotiable if you are running gradients. A simple inline vacuum degasser on the pump assembly solves most bubble-related issues. If you skip this step, you will see noisy baselines and shifting retention times that make no analytical sense. The noise increases because light scattering from microbubbles in the flow cell changes the absorbance reading unpredictably. The retention time shifts happen because your actual flow rate becomes inconsistent when gas pockets move through the system. Column temperature control is another thing most people ignore until their data looks sloppy. Running a column at ambient lab temperature means your retention times will drift with the seasons. A column oven set to thirty-five degrees Celsius stabilizes your method and usually improves peak shape because mass transfer kinetics are faster at elevated temperature. I once spent three days trying to figure out why my retention times were drifting by nearly a minute every morning before I realized the HVAC in the lab cycled on and off and the column was getting hit with cold draft air from a vent directly above the instrument.
Get the Full Details

A Real Problem and What Actually Fixed It
Here is a specific case. I was analyzing a pharmaceutical compound that showed severe peak tailing on a fresh C18 column. The tailing factor was around 2.1, which is unacceptable for quantitative work. The sample was dissolved in mobile phase, the pH was in the recommended range for the column, and the injection volume was only ten microliters at a concentration that should not have overloaded the column. Everything looked correct on paper. The issue turned out to be secondary interactions between the basic analyte and residual silanol groups on the silica surface of the column. This is a classic problem that does not show up in any beginner tutorial. The workaround was straightforward but not obvious if you do not have experience with it. I added 10 millimolar ammonium acetate to the mobile phase and adjusted the pH to 4.5 using formic acid. The buffered conditions suppressed the ionization of the residual silanols and the peak tailing dropped to a tailing factor of 1.2 within the first few injections. The key insight here is that adding a volatile buffer serves two purposes. It controls the ionization state of your analyte and it passivates the silanol sites on the stationary phase. The buffer needs to be volatile because you are often coupling HPLC to mass spectrometry downstream, and non-volatile buffers like phosphate will destroy your MS source in about an hour of running.
Understanding What the Chromatogram Is Actually Telling You
Resolution is the metric that matters. Two peaks need a resolution of at least 1.5 to be considered baseline separated. The equation for resolution involves the selectivity factor, the efficiency of the column, and the retention factor. Most beginners focus on efficiency, which is why they keep buying more expensive columns instead of adjusting the mobile phase composition. Selectivity is far more powerful than efficiency for separating closely eluting peaks. Changing the organic modifier from acetonitrile to methanol can shift selectivity dramatically even though the column and flow rate stay exactly the same. I resolved a pair of isomers that co-eluted perfectly on acetonitrile simply by switching to methanol as the organic modifier. The run time increased by about four minutes but the separation was clean and the quantification became reliable. Plate count measurements are useful for checking column health but they are not the whole story. A column might have 12,000 plates per meter and still produce garbage peaks if the stationary phase is degraded or if there is void formation at the column head. I check plate count monthly as a maintenance log entry but I do not let it be the sole determinant of whether a column is still good. Peak asymmetry and the retention time stability of a standard mix over a week tell me more about the actual performance of the column.
Common Pitfalls That Waste Everyone's Time
Solvent miscibility is a problem people encounter constantly. Water and acetonitrile mix cleanly. Water and methanol mix cleanly. Water and hexane do not mix cleanly. If you ever decide to run a normal-phase method or use an unusual solvent system, make sure the components are actually miscible across the entire gradient range. Phase separation in the mixed manifold of your pump causes all kinds of chaotic behavior including pressure fluctuations and erratic retention times. I once had a method that appeared to work fine until I tried to include a high organic wash step and the baseline went completely unstable. Checking the binary pump mixing chamber revealed a cloudy layer of immiscible solvent. It took three hours of flushing with isopropanol to clear it out. Sample solvent strength is another area where people routinely sabotage their own separations. If you dissolve your sample in pure acetonitrile and inject it onto a column running at ten percent organic, the strong solvent effect will cause peak distortion and splitting. The sample plug is essentially being deposited into a solvent that is much weaker than where it was dissolved, and the analyte can precipitate or interact strangely at the column head. The rule of thumb is that your sample solvent should match the initial mobile phase composition or be weaker. If you need to dissolve a hydrophobic compound, use a small percentage of organic solvent in your sample diluent rather than pure organic. A fifty fifty mixture of water and methanol usually works as a universal sample solvent for reverse-phase methods. Maintenance schedules matter more than most people admit. Replace the guard column every time you notice a pressure increase of more than five hundred psi above your baseline or when peak shapes begin to degrade. The guard column is cheap insurance. Swapping it takes about thirty seconds and it protects the main analytical column from particulates and strongly retained contaminants. The main column replacement costs around two hundred to four hundred dollars. I replaced a main column once because I had been ignoring a clogged guard for months and the backpressure had been climbing gradually. The column was packed with precipitated sample material and the seals were starting to fail. It was a painful lesson in preventive maintenance.

When HPLC Is the Wrong Tool
Reverse-phase HPLC has hard limitations. Thermally labile compounds can degrade in the injector or on-column, especially at elevated temperatures. Highly polar compounds that do not retain on C18 at any reasonable pH will simply flow through in the void volume and you will get nothing useful. For those compounds you need hydrophilic interaction chromatography or ion-pairing chromatography. Ion-pairing agents like tributylamine or heptafluorobutyric acid can help retain polar analytes but they require extensive column equilibration times of up to an hour between runs and they contaminate the system if you ever need to switch back to standard reversed-phase work. The contamination is persistent and cleaning it out takes a full day of washing with strong organic solvent. For compounds that are volatile and thermally stable, gas chromatography may be a better choice. GC provides higher peak capacity and faster analysis times for the right analyte class. HPLC is the better choice when your compounds are non-volatile, polar, or thermally unstable. Knowing which category your analyte falls into prevents you from wasting weeks trying to force a method that was never going to work on an instrument designed for the wrong application.
Practical Method Development Workflow
Start with a quick scouting gradient. Thirty minutes from five percent to ninety-five percent organic at one milliliter per minute on a C18 column with UV detection at two-hundred nanometers. Look at where your analytes elute. If everything comes out between five and fifteen percent organic, you have a polarity problem and need a different column chemistry or a weaker organic modifier. If everything elutes after eighty percent organic, your column might be saturated with strongly retained matrix components or your analytes are too non-polar for standard C18 and you need a C8 or phenyl column. Once you have identified the approximate elution window, narrow the gradient to cover only that range. A gradient spanning twenty percent organic around your analytes will give you far better resolution than a broad sweep. Then optimize the flow rate and column temperature. Lower flow rates increase resolution but extend run time. Higher temperatures reduce backpressure and improve mass transfer but may compromise selectivity. The optimal settings depend entirely on your specific analytes and there is no universal answer. Validate the method after development. Check linearity across your expected concentration range, precision at three concentration levels, accuracy with spiked samples, and stability of the standard and sample solutions. A properly validated method typically requires around two weeks of focused work depending on how many concentrations and replicates you need. If you are working under regulatory conditions such as FDA compliance, you will need to include specificity, limit of detection, limit of quantitation, and robustness testing as well. That adds another week or two minimum.
The instrument itself is reliable when maintained properly. The issues are almost always method-related or sample-preparation-related. If your chromatography looks bad, check the method parameters before you start replacing hardware. I have seen technicians order new columns, pumps, and detectors for problems that were caused by an incorrect pH adjustment or a degraded mobile phase. The mobile phase should be prepared fresh and used within forty-eight hours unless you have validated longer shelf stability. Buffer solutions in particular will support microbial growth over time and the resulting particulates and pH shifts will ruin your method quietly.