Delta H Calculations Aren't as Clean as Textbooks Make Them Look
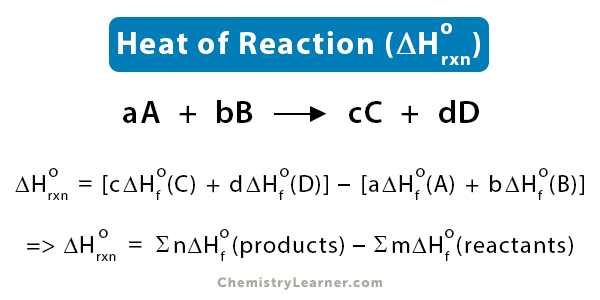
You want the change in enthalpy for a reaction. The standard approach is straightforward enough, but the details are where people get tripped up. Most students and even some early-career engineers just grab a table of standard enthalpies of formation and start subtracting. That works until it doesn't, and by then you've already submitted the wrong answer on an exam or built a process model with a five percent error margin. The baseline method is to take the sum of the standard enthalpies of formation for the products and subtract the sum for the reactants. You look each value up in a reference table, multiply by the stoichiometric coefficient from your balanced equation, and do the arithmetic. Standard state means 25 degrees Celsius and one atmosphere of pressure. That assumption alone will trip you up if your reaction runs anywhere near those conditions or involves phases that shift at standard temperature. I spent a few semesters grading introductory chemistry exams and the same mistakes show up year after year. People forget that elements in their standard states have a delta H of formation equal to zero, but only in that specific form. Oxygen as O2 gas is zero. Oxygen as ozone, O3, is not zero, and using the wrong value for it is a common way to get the wrong answer. Phosphorus is another one — white phosphorus is the standard state, not red phosphorus. Carbon is graphite, not diamond. These aren't minor details, they're the difference between getting the sign right and getting it wrong.
When the Table Values Don't Cut It
Standard enthalpies of formation cover a lot of common compounds, but there are plenty of situations where the data simply isn't available or isn't accurate enough. Transition metals in unusual oxidation states, exotic organometallics, and certain intermediates in catalytic cycles often don't have reliable tabulated values. When that happens, you need to build the enthalpy change from other measurements instead of looking it up. Hess's Law is the workhorse here. If you can break your target reaction into a series of steps whose enthalpies you already know, you add them up. The path doesn't matter, only the initial and final states. This is usually more useful than people give it credit for because it turns a lookup problem into a bookkeeping problem. You're not calculating anything new, just combining existing data in the right way. The trick is identifying which intermediate reactions connect cleanly to your target. I once needed the enthalpy of formation for a metal hydride that had no tabulated value. The compound was unstable at room temperature, so direct measurement wasn't practical. I found the heat of solution for the parent oxide in acid, the heat of solution for the hydride in the same acid, and a third reaction linking the oxide and the element. Subtracting the right combination gave me the formation enthalpy within a few kilojoules of what calorimetry later confirmed. The whole thing took maybe an hour if I knew where to look for the constants.
Calorimetry Is the Direct Route
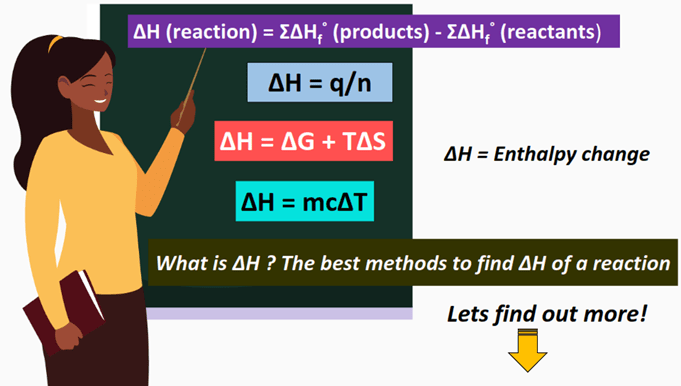
If you have the reaction in front of you, measuring the heat directly is often the most reliable option. A coffee-cup calorimeter works for aqueous reactions at modest temperatures. You mix the reactants, record the temperature change, and multiply by the mass of the solution and its specific heat capacity. For gas-phase or high-temperature reactions, a bomb calorimeter is the standard instrument. It handles the combustion reactions that organic and biochem students spend weeks calculating instead of actually measuring. The practical issue with calorimetry is that everything leaks heat. No calorimeter is perfectly adiabatic, and the correction for heat loss during the measurement period can be larger than people expect. The standard workaround is to run a blank calibration with a known electrical input or a standard reaction, establish the heat capacity of the whole setup, and then apply that calibration to your actual experiment. Without calibration, your delta H values can drift by ten to fifteen percent depending on how quickly you're taking readings and how well insulated your apparatus is.
Get the Full Details
Bond Enthalpies Are an Approximation, Not a Precision Tool
Bond enthalpy tables let you estimate delta H by adding up the energies required to break bonds in the reactants and subtracting the energies released when new bonds form in the products. This is faster than Hess's Law for a rough check, but the numbers are averages pulled from many different molecules. A C-H bond in methane doesn't have the exact same enthalpy as a C-H bond in ethanol, but the table will give you the same value for both. The cumulative error across a reaction with dozens of bonds can easily reach twenty or thirty kilojoules per mole. Use bond enthalpies when you need a quick sanity check or when no better data exists. Don't use them when the assignment or project demands precision. I've seen people treat average bond energies like they're fundamental constants, which is a category error that costs points on exams and confidence on real calculations.
Phase Changes and Temperature Corrections Matter More Than You'd Expect
Standard enthalpies are reported at 298 Kelvin, but reactions don't always happen at 298 Kelvin. If your process runs at 500 K or your products include water vapor instead of liquid water, the tabulated value for liquid water is the wrong number to plug in. The enthalpy of vaporization for water is about forty-four kilojoules per mole, and using the liquid value when the product is actually a gas introduces a significant error. Same thing with heating or cooling substances away from standard temperature — you need heat capacity data and you integrate it over the temperature range. The Kirchhoff equation handles the temperature dependence. You take the standard enthalpy at 298 K and add the integral of the change in heat capacity over the temperature interval. In practice, if the heat capacities don't vary much over your range, you can approximate them as constant and the calculation becomes a simple multiplication. For rough engineering estimates that's usually good enough. For anything where the accuracy matters, you'll want temperature-dependent Cp polynomials from a database like NIST.
A Few Practical Reminders
Make sure your equation is balanced before you touch any numbers. An unbalanced equation gives you wrong stoichiometric coefficients, which scales the entire delta H incorrectly. It sounds obvious and it surprises me how often it's missed. Check your units too. Most tables report kilojoules per mole, but a few older sources use kilocalories, and mixing the two without converting is a fast way to produce garbage results. Pay attention to the physical state notation. H2O liquid versus H2O gas in your reaction equation isn't just a detail, it changes the enthalpy value substantially. If someone hands you an equation without phase labels, assume standard states unless the context clearly indicates otherwise, and note that assumption in your work. That habit will save you from confusion when the answer doesn't match the key. One more thing that comes up in practice: the sign convention. Delta H negative means the reaction releases heat, which is exothermic. Delta H positive means it absorbs heat, endothermic. Students flip these all the time because the math sometimes produces a result that feels backwards depending on how they set up the subtraction. If you consistently use products minus reactants, the sign will come out correctly. There's no ambiguity once you pick a convention and stick to it.


