How Hybridization and MO Theory Actually Work When You Stop Memorizing
Most students hit a wall somewhere around sophomore organic chemistry or physical chemistry. They can draw sp2 and sp3 orbitals from memory. They can sketch a benzene ring with the right number of pi bonds. But the moment you ask them to explain why ethylene's C-C stretch frequency differs from ethane's, or why O2 is paramagnetic despite Lewis structures suggesting otherwise, they go blank. That disconnect between memorized diagrams and actual chemical behavior is exactly what hybridization and MO theory were meant to close.
The two frameworks serve different purposes and they're frequently taught as if they're competing explanations. They aren't. Hybridization is a mathematical convenience for building localized bonding models. MO theory is a quantum mechanical treatment that describes electrons as delocalized over the entire molecule. Both are approximations. Both have real predictive power. Neither is the final answer.
Practical Hybridization And Mo Theory: Where to Start
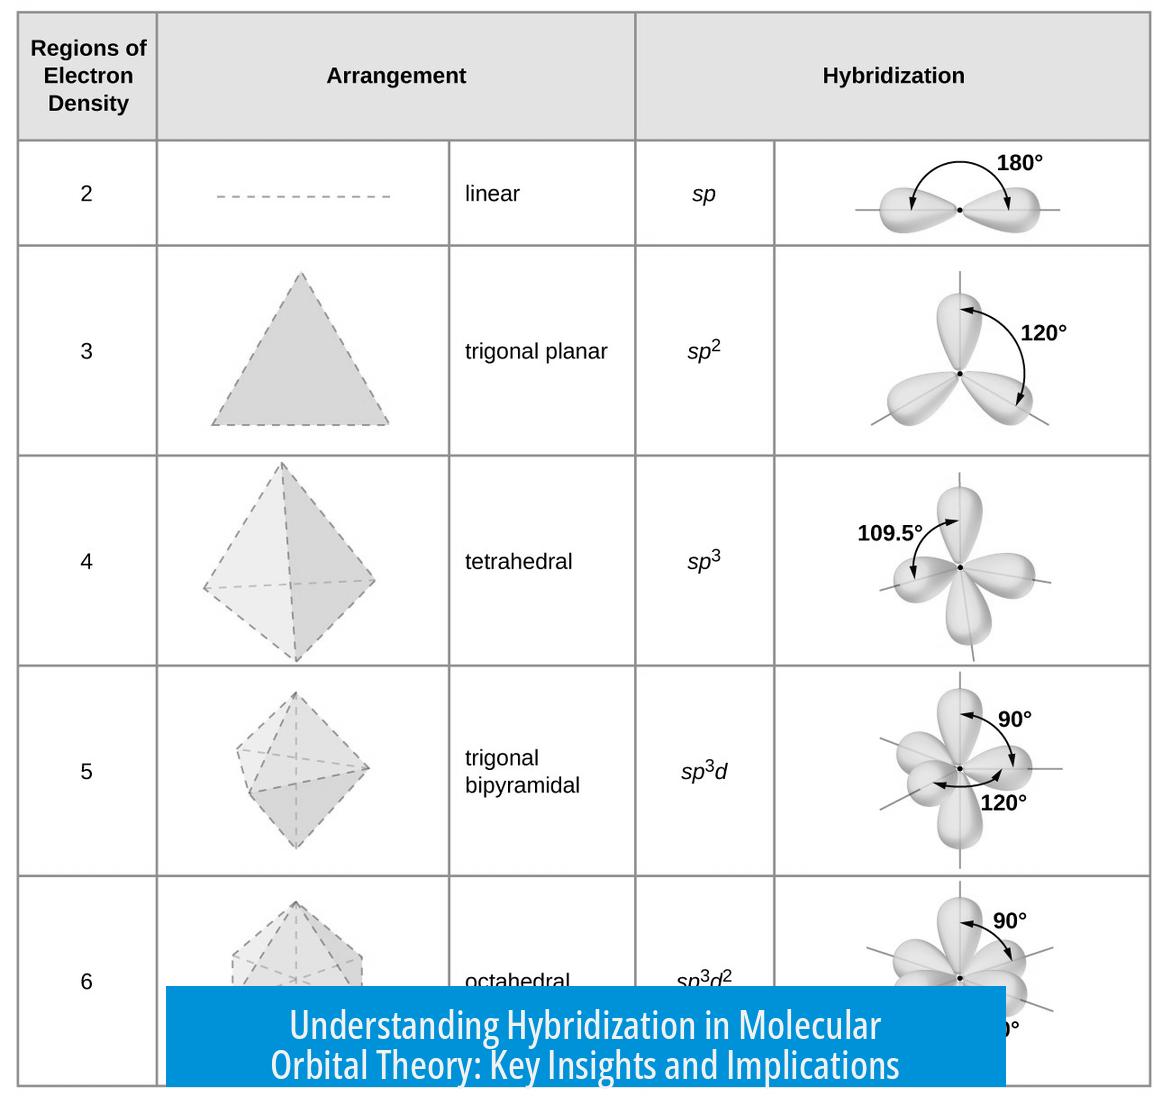
Hybridization starts with atomic orbitals on individual atoms. You take s and p (and sometimes d) orbitals and mix them linearly to produce hybrid orbitals oriented in specific geometries. Carbon in methane uses four sp3 hybrids arranged tetrahedrally. Carbon in ethylene mixes one s and two p orbitals into three sp2 hybrids in a trigonal plane, leaving one unhybridized p orbital perpendicular to that plane for pi bonding.
The practical utility comes from geometry prediction and bond angle estimation. If your central atom has four electron domains, you assign sp3. Three domains gives sp2. Two domains gives sp. This works reliably for main group elements in their common oxidation states. It breaks down noticeably with transition metals, heavy p-block elements where relativistic effects matter, and systems with significant electron correlation.
MO theory requires you to think in terms of molecular orbitals formed from linear combinations of atomic orbitals across the whole molecule. You combine atomic orbitals of similar energy and symmetry to form bonding and antibonding combinations. The bonding MOs sit lower in energy than the parent atomic orbitals. The antibonding MOs sit higher. Electrons fill from the bottom up according to the Aufbau principle, Hund's rule, and the Pauli exclusion principle.
The key difference from hybridization is that MOs are delocalized. In benzene, for example, the six pi electrons occupy three bonding MOs spread across all six carbons rather than existing as three localized double bonds between specific carbon pairs. This explains why all C-C bond lengths in benzene are identical at 1.39 angstroms, somewhere between a single and double bond.
The Diagramming Process for Simple Molecules
Take water as a straightforward example. Oxygen has the valence configuration 2s2 2p4. In the hybridization picture, you treat oxygen as sp3 hybridized with two bonding pairs and two lone pairs, giving a bent geometry with a bond angle near 104.5 degrees. The deviation from the ideal 109.5 degrees comes from lone pair repulsion, which VSEPR handles qualitatively.
In the MO picture for water, you combine the oxygen 2s and 2p orbitals with the hydrogen 1s orbitals. The a1 symmetry combination involves oxygen 2s and 2pz mixing with the symmetric hydrogen combination. The b2 combination involves oxygen 2px mixing with the antisymmetric hydrogen combination. This produces bonding and nonbonding orbitals that account for the two O-H bonds and the two lone pairs. The calculated H-O-H angle from an MO calculation comes out close to the experimental value without invoking VSEPR separately.
For diatomic molecules, MO theory becomes especially powerful. O2 has 12 valence electrons. Filling the MO diagram from lowest to highest energy gives the configuration (sigma2s)2(sigma*2s)2(sigma2pz)2(pi2px)2(pi2py)2(pi*2px)1(pi*2py)1. The two unpaired electrons in the degenerate pi* orbitals explain O2's paramagnetism, something no Lewis structure or simple hybridization model can reproduce. This is the textbook example for a reason.
Common Pitfalls and What I've Learned the Hard Way
Students routinely mix up which orbitals participate in hybridization versus which remain as pure p orbitals for pi bonding. The rule is straightforward once it clicks: the number of hybrid orbitals equals the number of atomic orbitals mixed. For sp2, you mix one s and two p orbitals to get three sp2 hybrids. One p orbital is left untouched. You cannot hybridize all three p orbitals and still have a p orbital available for pi bonding, so sp3d hybrids cannot simultaneously serve as pi bond donors in the simple valence bond picture.
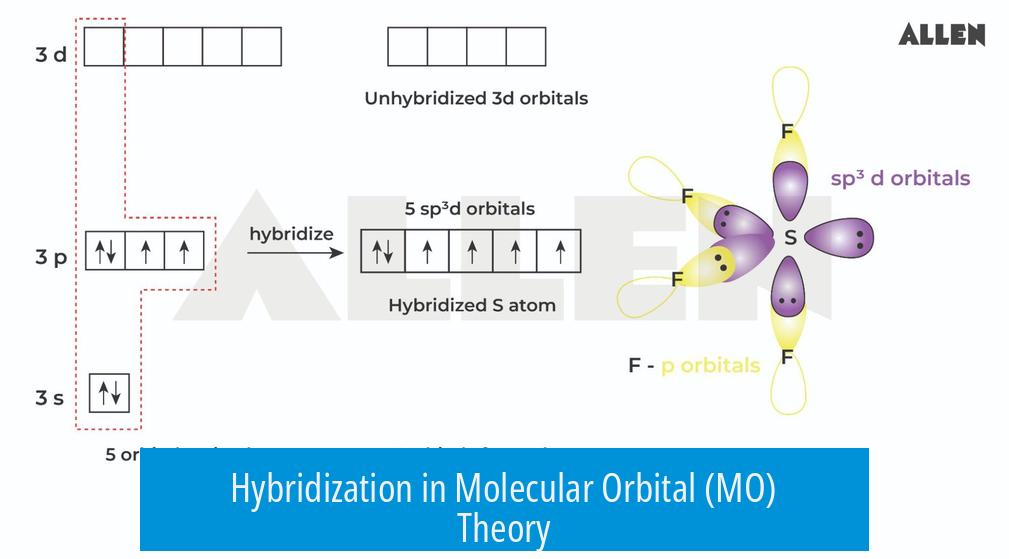
Another frequent error is applying sp3d or sp3d2 hybridization to main group compounds as a blanket explanation for five or six electron domains. This works numerically for things like PF5 and SF6, but it glosses over the fact that d-orbital participation in bonding for third-period elements is minimal. Modern computational chemistry shows that hypervalent molecules are better described using three-center four-electron bonds rather than invoking d-orbital hybridization. The sp3d model gives the right geometry but the wrong physical picture.
I encountered a specific problem when modeling the bonding in NO, a radical with 11 valence electrons. The simple MO diagram suggests a bond order of 2.5, which matches the observed short bond length. But when I tried to fit this into a hybridization framework for use in a teaching lab manual, every reasonable hybrid assignment produced a structure with an incomplete octet on nitrogen or an awkward single-electron orbital that looked terrible on paper. The workaround was straightforward: I dropped the hybridization attempt entirely for NO and used the MO diagram directly, noting to students that hybridization is a localized model and radicals with odd electron counts often resist localization without distortion.
Where These Methods Break Down Completely
Transition metal complexes expose the limitations of both approaches most clearly. The crystal field theory model ignores covalency entirely and treats ligands as point charges. The ligand field theory version incorporates some covalent mixing but still relies on a simplified orbital picture. Neither handles the near-degeneracy problems and static correlation that arise in open d-shell systems with multiple low-lying electronic states. For something like Cr2, which has a formal quadruple bond involving sigma, two pi, and a delta component from d-orbital overlap, neither basic hybridization nor introductory MO theory captures the full bonding situation. You need multiconfigurational methods like CASSCF to get anything resembling the correct electronic structure.
Even for organic molecules, there are edge cases. Fulvene has a six-membered ring with an exocyclic double bond. A naive hybridization analysis would assign sp2 to all carbons and predict uniform aromatic-like delocalization. But the dipole moment and reactivity patterns show that the pi electrons are not evenly distributed. The MO picture handles this better by showing that the exocyclic double bond mixes with the ring pi system in a way that breaks the simple benzene analogy, but even that requires looking at the actual orbital coefficients rather than relying on resonance drawings alone.
Conjugated polyenes present another limitation. The simple Hückel MO method gives reasonable qualitative results for linear chains like butadiene and hexatriene, predicting the correct trend in HOMO-LUMO gaps and explaining why 1,3,5-hexatriene absorbs at longer wavelengths than butadiene. But it fails quantitatively for substituted systems where electron-donating or electron-withdrawing groups shift orbital energies significantly. You need extended Hückel or semiempirical methods like AM1 or PM3 for that level of accuracy, and those introduce their own parameterization dependencies.
Practical Workflow for Solving Bonding Problems
When you're given an unfamiliar molecule and need to determine its bonding characteristics, start by counting valence electrons. This single step catches more mistakes than any other. Then draw the Lewis structure to identify sigma bonds, lone pairs, and pi bonds. The Lewis structure tells you the connectivity and approximate geometry.
Next, assign hybridization based on the sigma bond and lone pair count around each central atom. This gives you the local geometry and identifies which p orbitals are available for pi bonding. Be careful here with atoms that have expanded octets or unusual oxidation states.
Then construct the MO diagram if delocalization or magnetic properties are in question. For small molecules, you can build this by hand using group theory to determine which atomic orbitals have the correct symmetry to combine. The character table for the molecule's point group tells you which irreducible representations apply. This step is non-negotiable if you want to predict spectroscopic properties or explain phenomena like the Jahn-Teller distortion.
For larger molecules, you're going to use computational software. Gaussian, ORCA, or GAMESS will handle the self-consistent field calculation and output the MO energies, coefficients, and occupancy. The default DFT functional like B3LYP or wB97X-D gives results that are qualitatively correct for most organic and main group systems at a computational cost that fits on a modern laptop. For transition metals or systems with strong correlation, you'll need a different functional or a multireference method, and the runtime increases by roughly an order of magnitude.
Counter-Intuitive Details Most Textbooks Skip
Hybridization is not an observable physical quantity. You cannot measure an sp3 orbital directly. It is a mathematical construct that provides a convenient basis set for describing electron density in a molecule. The actual electron density comes from the wavefunction, which in Hartree-Fock theory is a single Slater determinant of spin-orbitals. Hybrid orbitals are just one possible linear transformation of the canonical MOs. Different transformations give different hybrid sets, but they all describe the same electron density. This is why you sometimes see different hybridization assignments for the same molecule in different textbooks.
Another detail that rarely gets emphasized is that hybridization energy costs are real. Mixing s and p character into hybrids requires promotion energy that is partially recovered through better overlap in bonding. For second-row elements, the s-p promotion penalty is significant enough that sp3 hybridization in methane is only favorable because the four C-H bonds formed compensate for it. In heavier elements like silicon, the s-p gap is larger relative to the bond energies, which is one reason why Si prefers different coordination geometries and why Si=Si double bonds are much more reactive than C=C bonds.
MO theory also has a subtlety that trips people up: the ordering of sigma and pi orbitals changes depending on the atomic numbers involved. For B2, C2, and N2, the pi2p orbitals lie below the sigma2p orbital due to reduced s-p mixing. For O2 and F2, the sigma2p drops below the pi2p because increased nuclear charge stabilizes the sigma orbital more. This crossover means you cannot memorize a single MO diagram and apply it universally. The diagram must be adjusted for each row of the periodic table.
When to Use Each Model and When to Move On
Hybridization works well for predicting molecular geometry, estimating bond angles, and understanding the directional character of sigma bonds in organic molecules. It's fast, visual, and sufficient for most undergraduate-level problems. Use it when you need a quick answer about shape or when teaching introductory concepts.
MO theory is necessary when you need to explain magnetic properties, electronic spectra, bond orders that aren't integers, or delocalization effects. It's also essential for predicting reaction mechanisms that involve frontier orbital interactions, like Diels-Alder reactions where HOMO-LUMO symmetry matching determines whether a reaction is thermally allowed.
Neither model is adequate for quantitative predictions of reaction energies, barrier heights, or spectroscopic frequencies without computational support. If you need numbers that match experiment within a few kcal/mol, you're past both models and into the realm of ab initio or DFT calculations with proper basis sets and correlation treatment. For most practical purposes in industry, DFT with a triple-zeta basis set and a dispersion correction runs in minutes to hours on a standard workstation and gives results that are reliable enough for drug design and materials screening.
The bottom line is that hybridization and MO theory are complementary tools, not competing truths. Hybridization gives you intuition about local bonding geometry. MO theory gives you a framework for understanding global electronic structure. Knowing when each breaks down and what to use instead is what separates someone who can draw diagrams from someone who can actually predict how molecules behave.
Gallery Hybridization And Mo Theory
CHEM 1411 L33 Review of Hybridization and MO Theory 2024 - YouTube
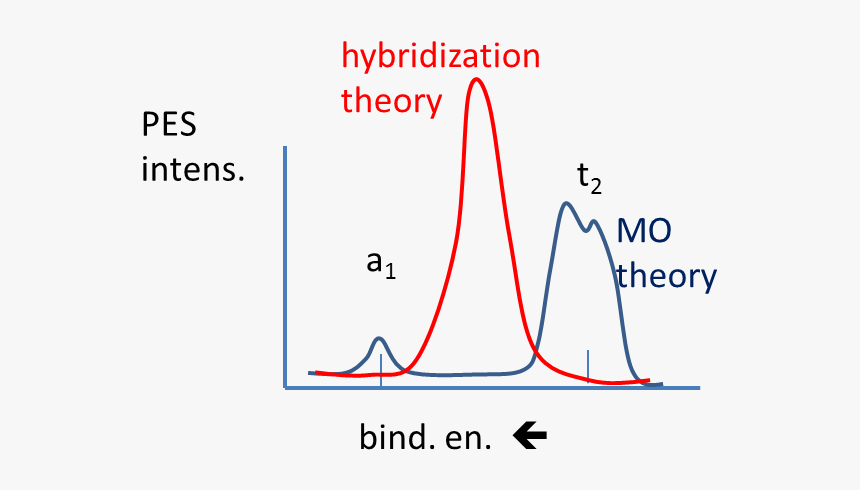
Mo Versus Hybridization Theory For Methane Pes - Methane Pes, HD Png ...
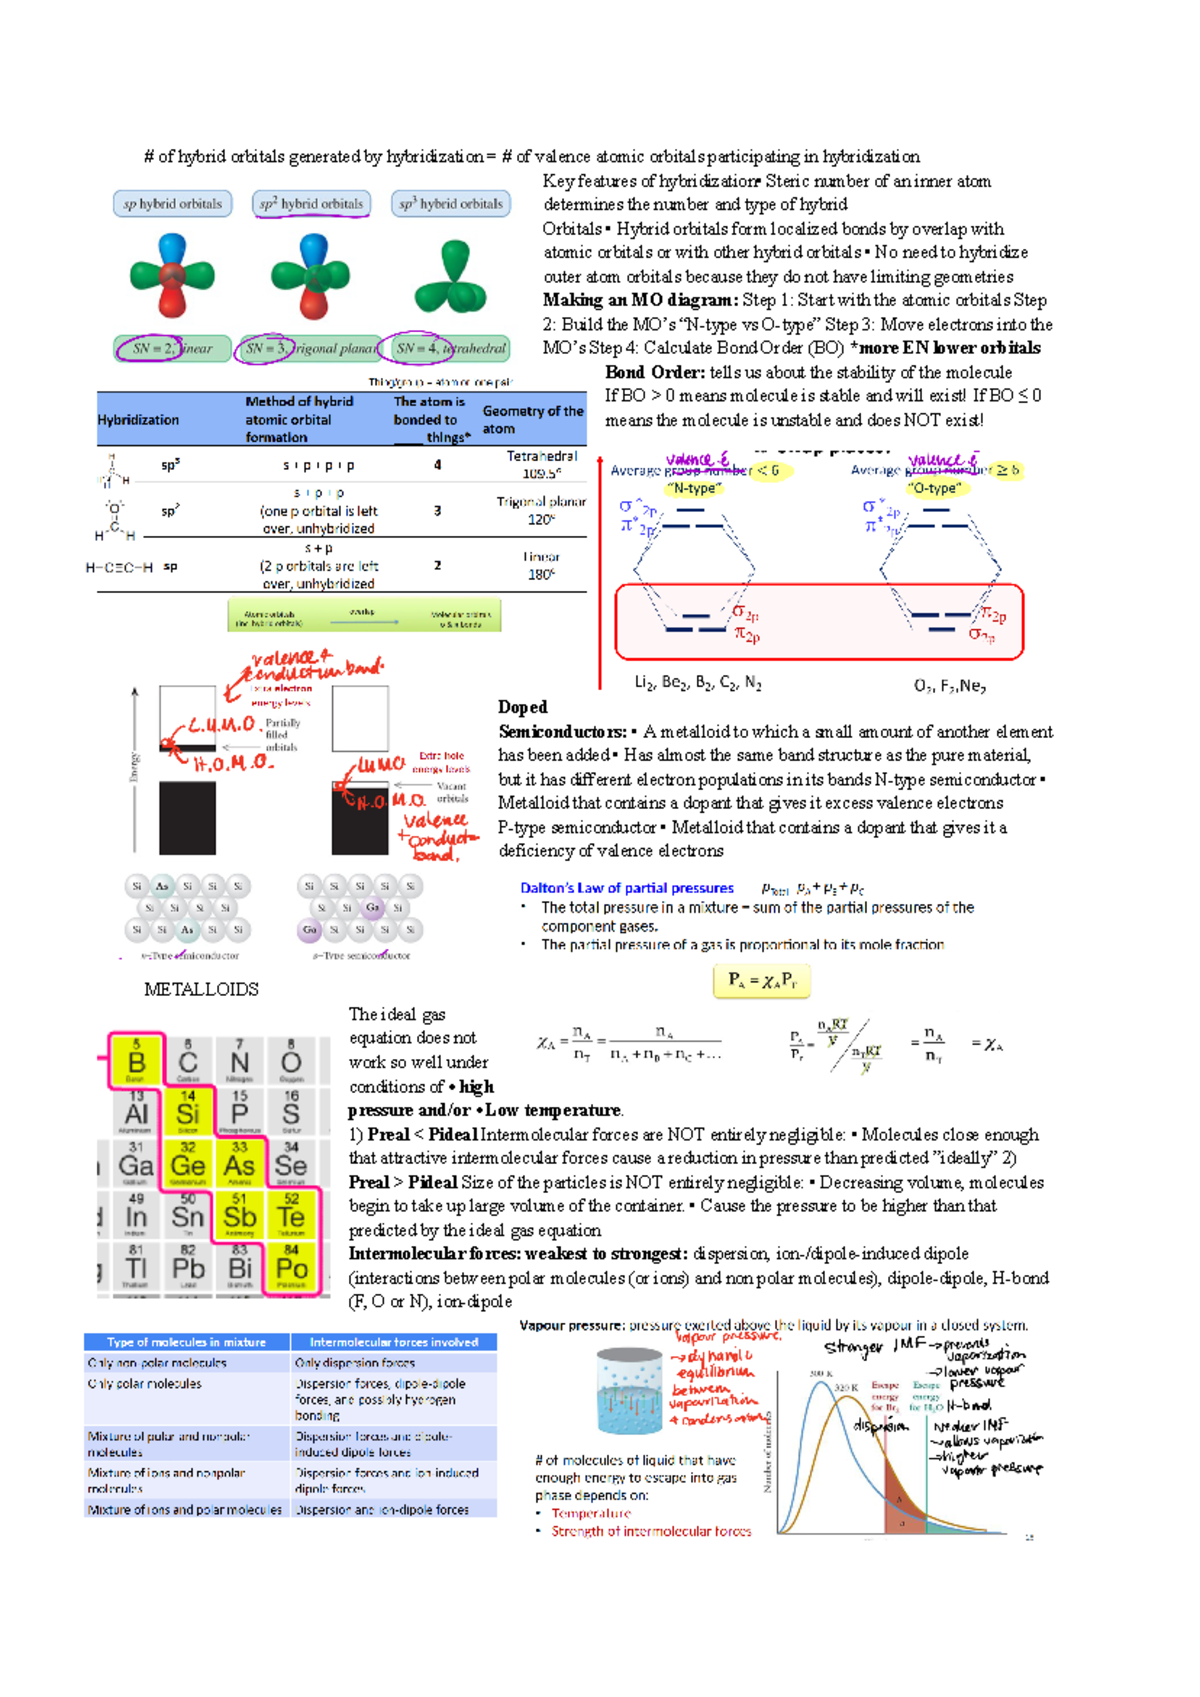
Chem 101 Hybridization and Molecular Orbital Theory Summary - Studocu
Understanding Hybridization in Molecular Orbital Theory: Key Insights ...
Understanding Hybridization in Molecular Orbital Theory: Key Insights ...