Practical Guide to the Kinetic Theory Of A Gas
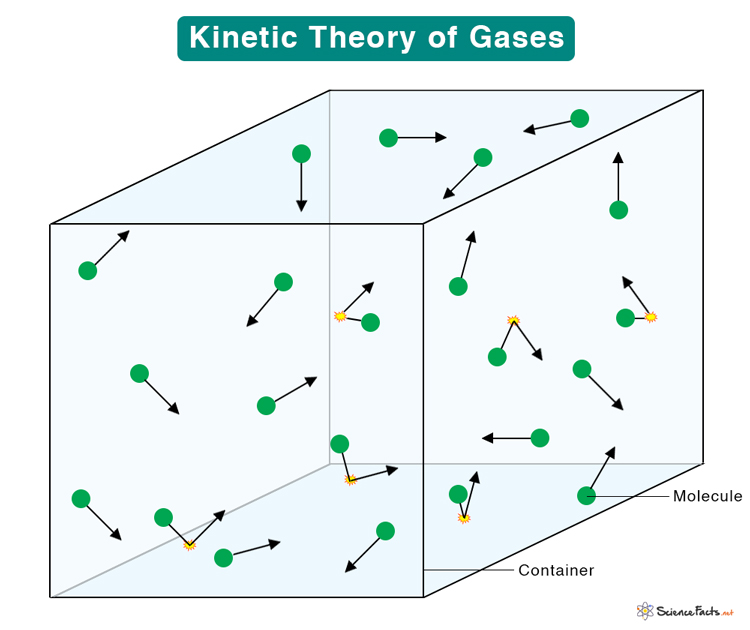
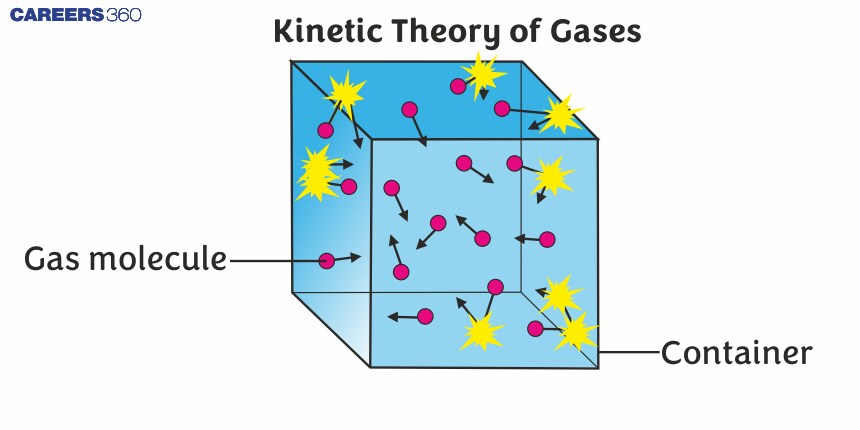
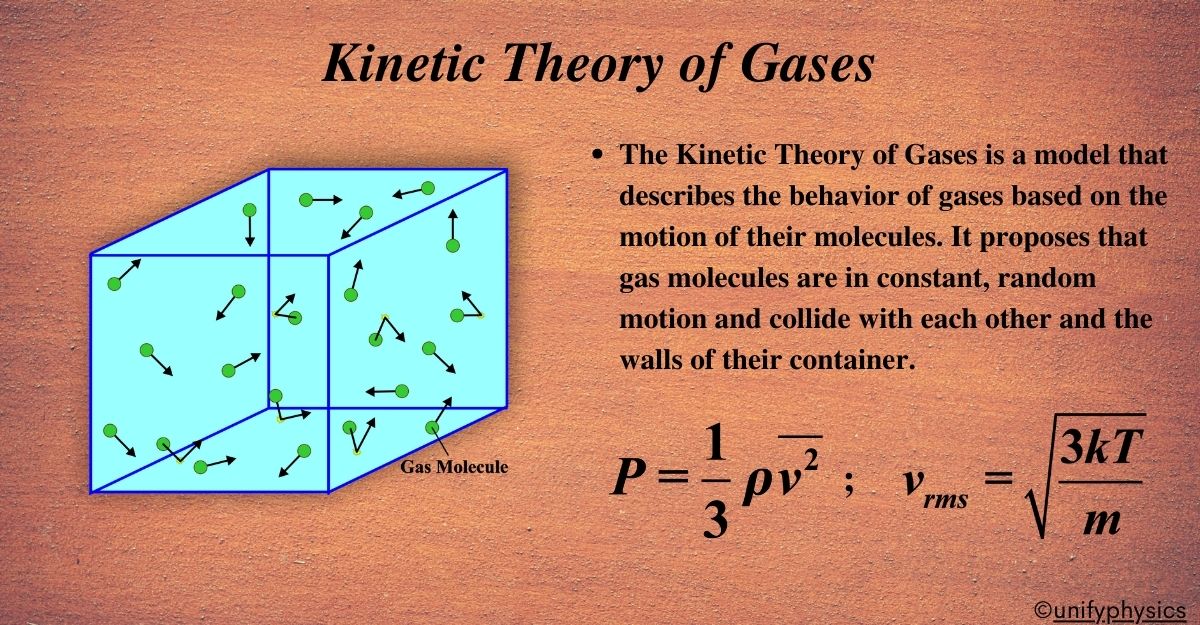
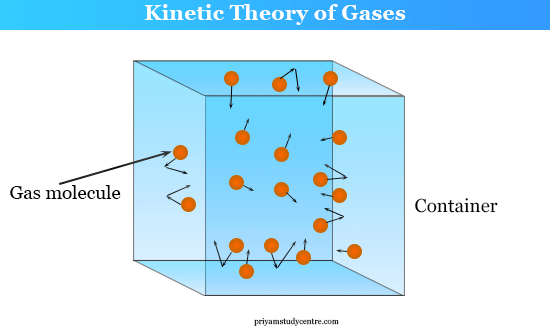
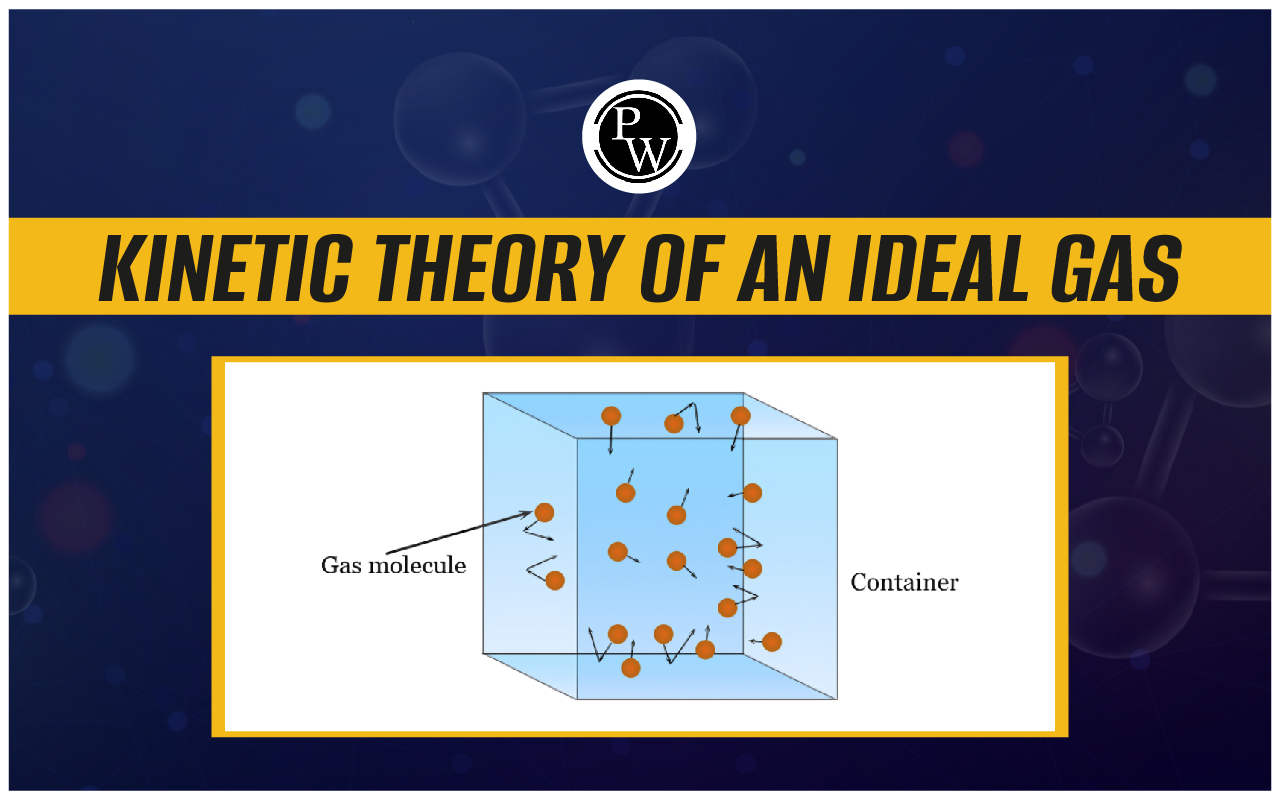
Start with the simplest version. You have a sealed container with gas molecules bouncing around. They're tiny, they move fast, and they don't interact except when they collide. That's basically it. The theory connects what you can measure — pressure, volume, temperature — to what you can't see, which is the motion of individual molecules. Here's how I approach it when I need to solve something real, not just pass an exam. First, pick your gas and assume it behaves ideally unless you have a good reason not to. For nitrogen at room temperature and atmospheric pressure, ideal behavior gets you within about one percent. Above 10 atmospheres or below 200 Kelvin, that assumption starts breaking down and you need something like the van der Waals equation instead. The core equation you'll use most is pV = nRT, but that's just the shortcut. The deeper version comes from working out the pressure from molecular collisions. It looks like this:
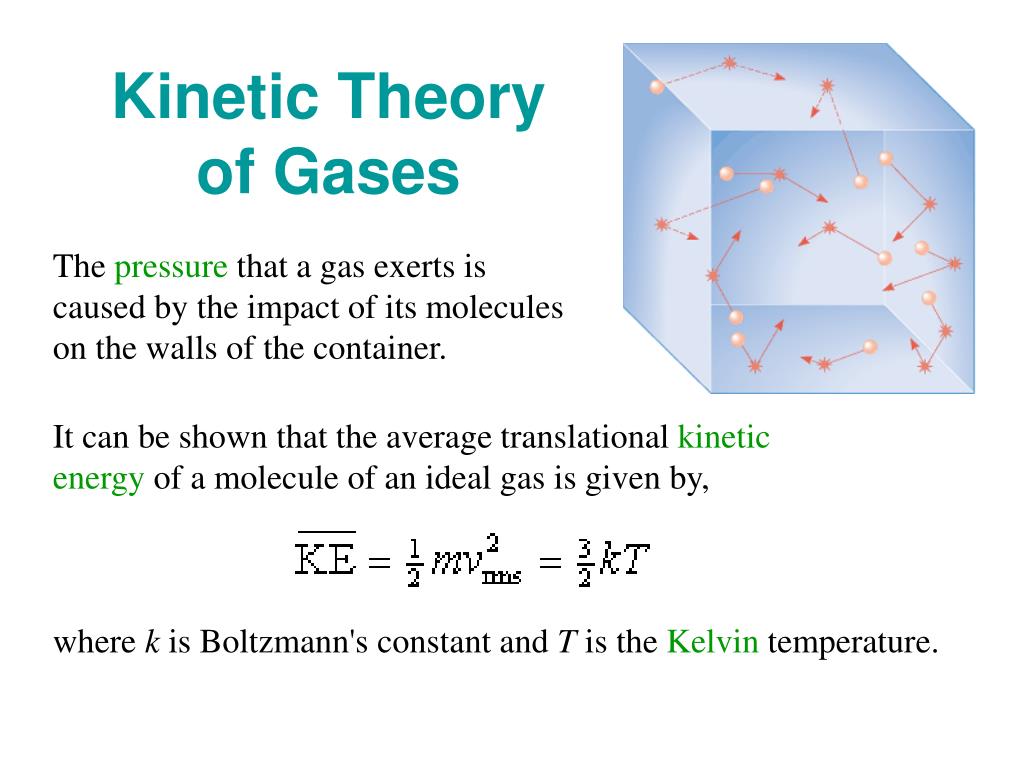
p = (1/3) × (N/V) × m × v²(rms) Where N is the number of molecules, V is volume, m is molecular mass, and v(rms) is the root mean square speed. This tells you that pressure is literally the result of molecules slamming into the walls. Temperature is proportional to the average kinetic energy of those molecules. Specifically, (3/2)kT = (1/2)m × v²(rms), where k is Boltzmann's constant. When I'm calculating RMS speed for a real problem, I usually start by finding the molar mass in kilograms per mole, then plug it into v(rms) = sqrt(3RT/M). Don't skip the unit conversion. I've seen people use grams per mole and get answers off by a factor of sqrt(1000), which is about 31.6. That's a common mistake.
Let me walk through a specific case. Say you have a 2-liter container of helium at 300 K and 1 atmosphere. You want the RMS speed of the helium atoms. The molar mass of helium is 0.004 kg/mol. So v(rms) = sqrt(3 × 8.314 × 300 / 0.004) = sqrt(1,870,650) 1368 m/s. That's about 1.4 kilometers per second. The atoms are moving incredibly fast, but the container only has about 0.08 moles, or roughly 5 × 10²² atoms, so the total mass is tiny and the momentum per collision is small, which is why you only feel 1 atmosphere of pressure. One thing textbooks don't always emphasize: the distribution of speeds matters. Not every molecule moves at the RMS speed. The Maxwell-Boltzmann distribution describes the spread. At 300 K, some helium atoms are moving at 500 m/s, others at 2500 m/s, and the RMS value sits somewhere in between. If you're doing anything involving reaction rates or escape velocity, the tail of that distribution is where the action is. The average speed is actually different from the RMS speed — about 92% of it. For most engineering calculations the difference doesn't matter, but if you're modeling evaporation or atmospheric loss over geological timescales, it does. Here's the problem I ran into that almost cost me a day. I was modeling a noble gas mixture in a vacuum chamber at around 10³ torr and needed to calculate collision frequencies for a sputtering deposition process. The standard kinetic theory formulas assume a dense gas where the mean free path is tiny compared to the container. In that vacuum regime, the mean free path was about 5 centimeters, but my chamber was only 10 centimeters across. The molecules were hitting the walls more often than they were hitting each other. The standard p = (1/3)(N/V)m × v²(rms) formula still gives the right pressure, but any calculation involving collision rates was completely wrong. I had to switch to free-molecular flow equations and treat wall collisions separately from intermolecular collisions. It changed the effective collision frequency by roughly two orders of magnitude.
Get the Full Details
Another nuance that trips people up: kinetic theory works beautifully for monatomic gases. Diatomic and polyatomic gases add rotational and vibrational degrees of freedom, which means the simple (3/2)kT per molecule only accounts for translational energy. At room temperature, nitrogen has five active degrees of freedom — three translational and two rotational. That means the internal energy is (5/2)nRT per mole, not (3/2)nRT. The pressure equation stays the same because pressure only depends on translational motion, but if you're calculating heat capacity or energy changes, missing those extra degrees of freedom throws off your Cp and Cv values. The ratio = Cp/Cv drops from 5/3 for monatomic gases to 7/5 for diatomic gases at room temperature. That's the difference between 1.67 and 1.40, and it matters a lot if you're working with adiabatic processes like compression in an engine cylinder. For practical problem-solving, here's the order I recommend. Identify whether the gas is ideal or not. Check the pressure and temperature against typical deviation points. Calculate what you need using the basic kinetic equations. If collision rates matter, verify that your mean free path is actually much smaller than your system dimensions. If it isn't, use the appropriate rarefied gas corrections. Double-check your units, especially when converting between molar mass and molecular mass, and between kilojoules and joules in energy calculations. The limitations are real. Kinetic theory assumes point particles with elastic collisions and no intermolecular forces except during collisions. Real gases have finite molecular size and attractive forces. At high pressures, the volume of the molecules themselves becomes significant relative to the container volume. At low temperatures, attraction between molecules matters and you get condensation. The theory also doesn't handle quantum effects — at very low temperatures or with light gases like hydrogen and helium, you need quantum statistical mechanics instead of classical kinetic theory. For most introductory and intermediate work, these limitations don't matter much, but they become critical in areas like cryogenics, high-pressure physics, and stellar interiors.
If you're working with non-ideal conditions regularly, look into the virial expansion or the van der Waals equation as practical alternatives. They add correction terms for molecular volume and intermolecular attraction while keeping the same basic framework. For rough estimates they're usually sufficient. For precision work, you'd move toward more sophisticated equations of state like Redlich-Kwong or Peng-Robinson, which are standard in chemical engineering.