Why Your Chemistry Teacher Was Lying To You About Atoms
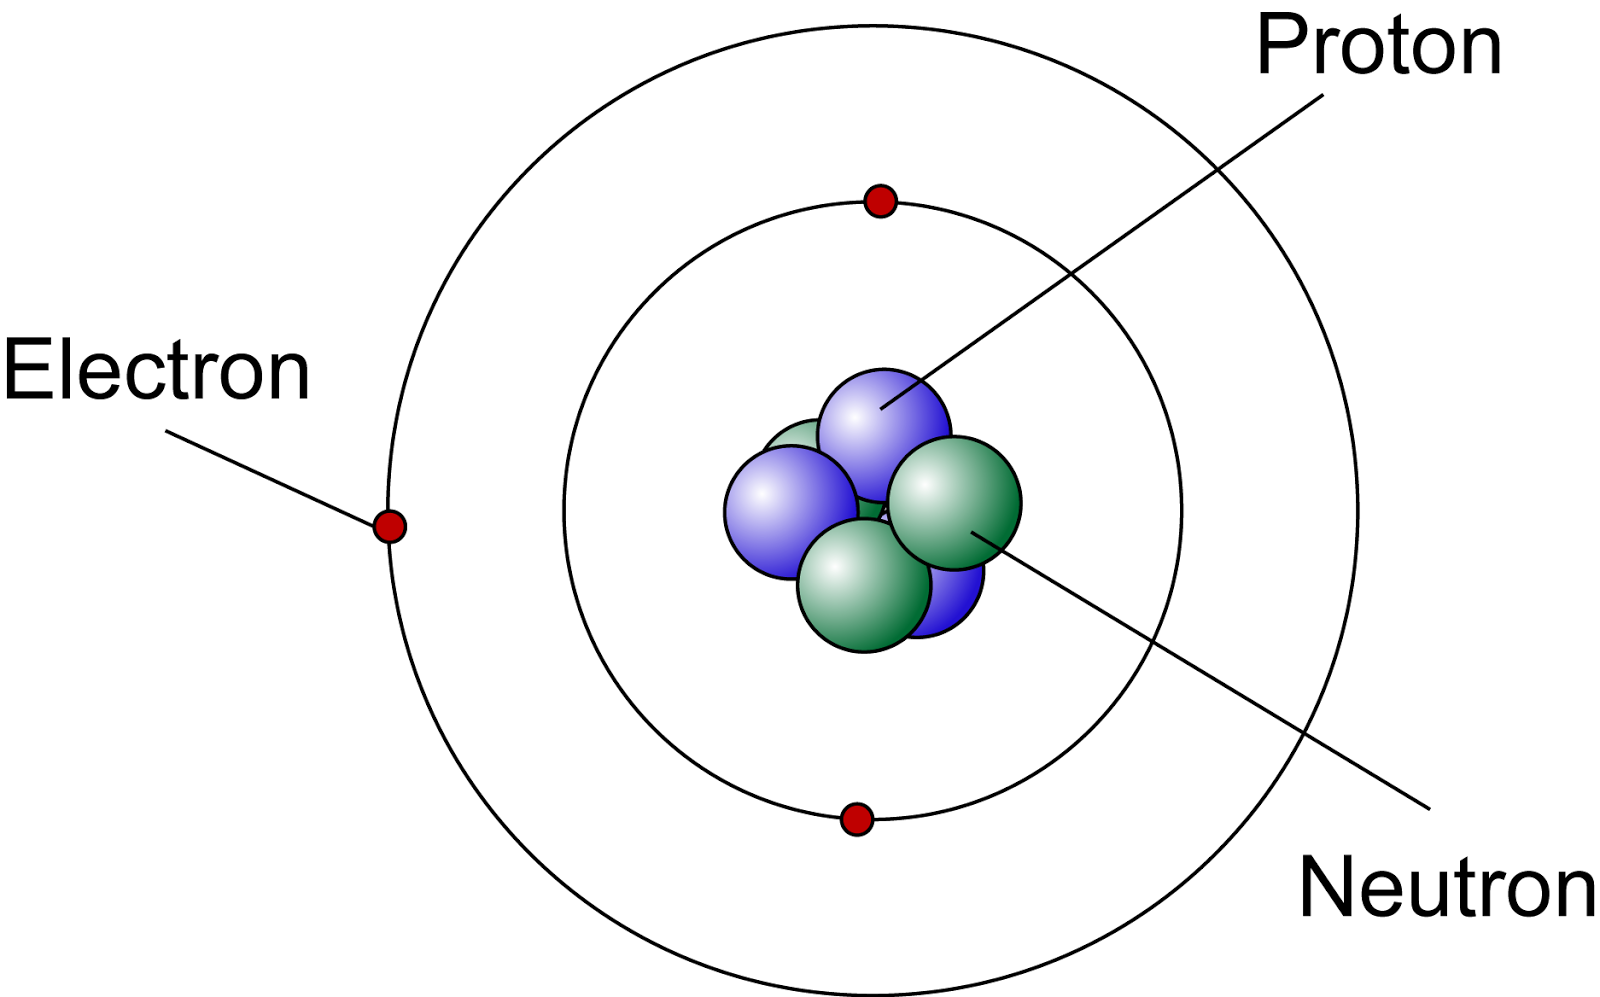
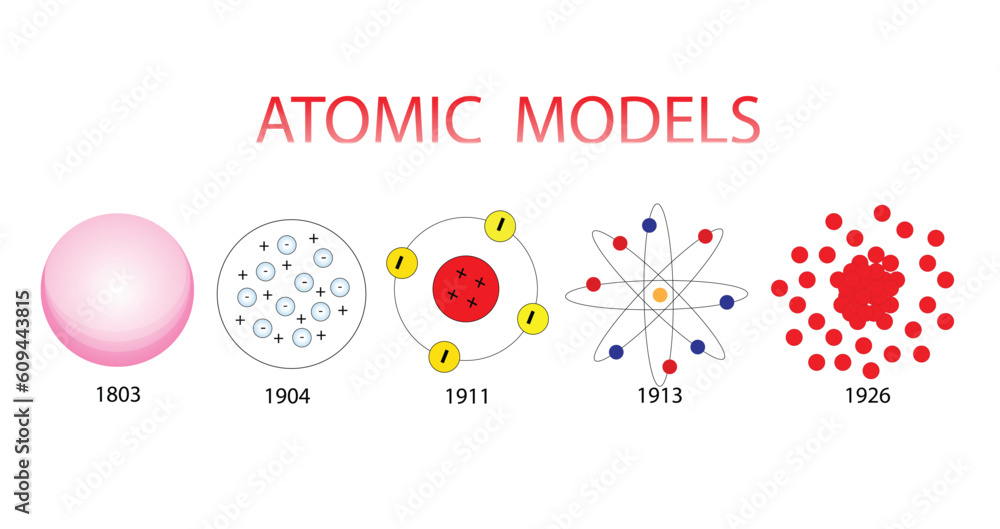
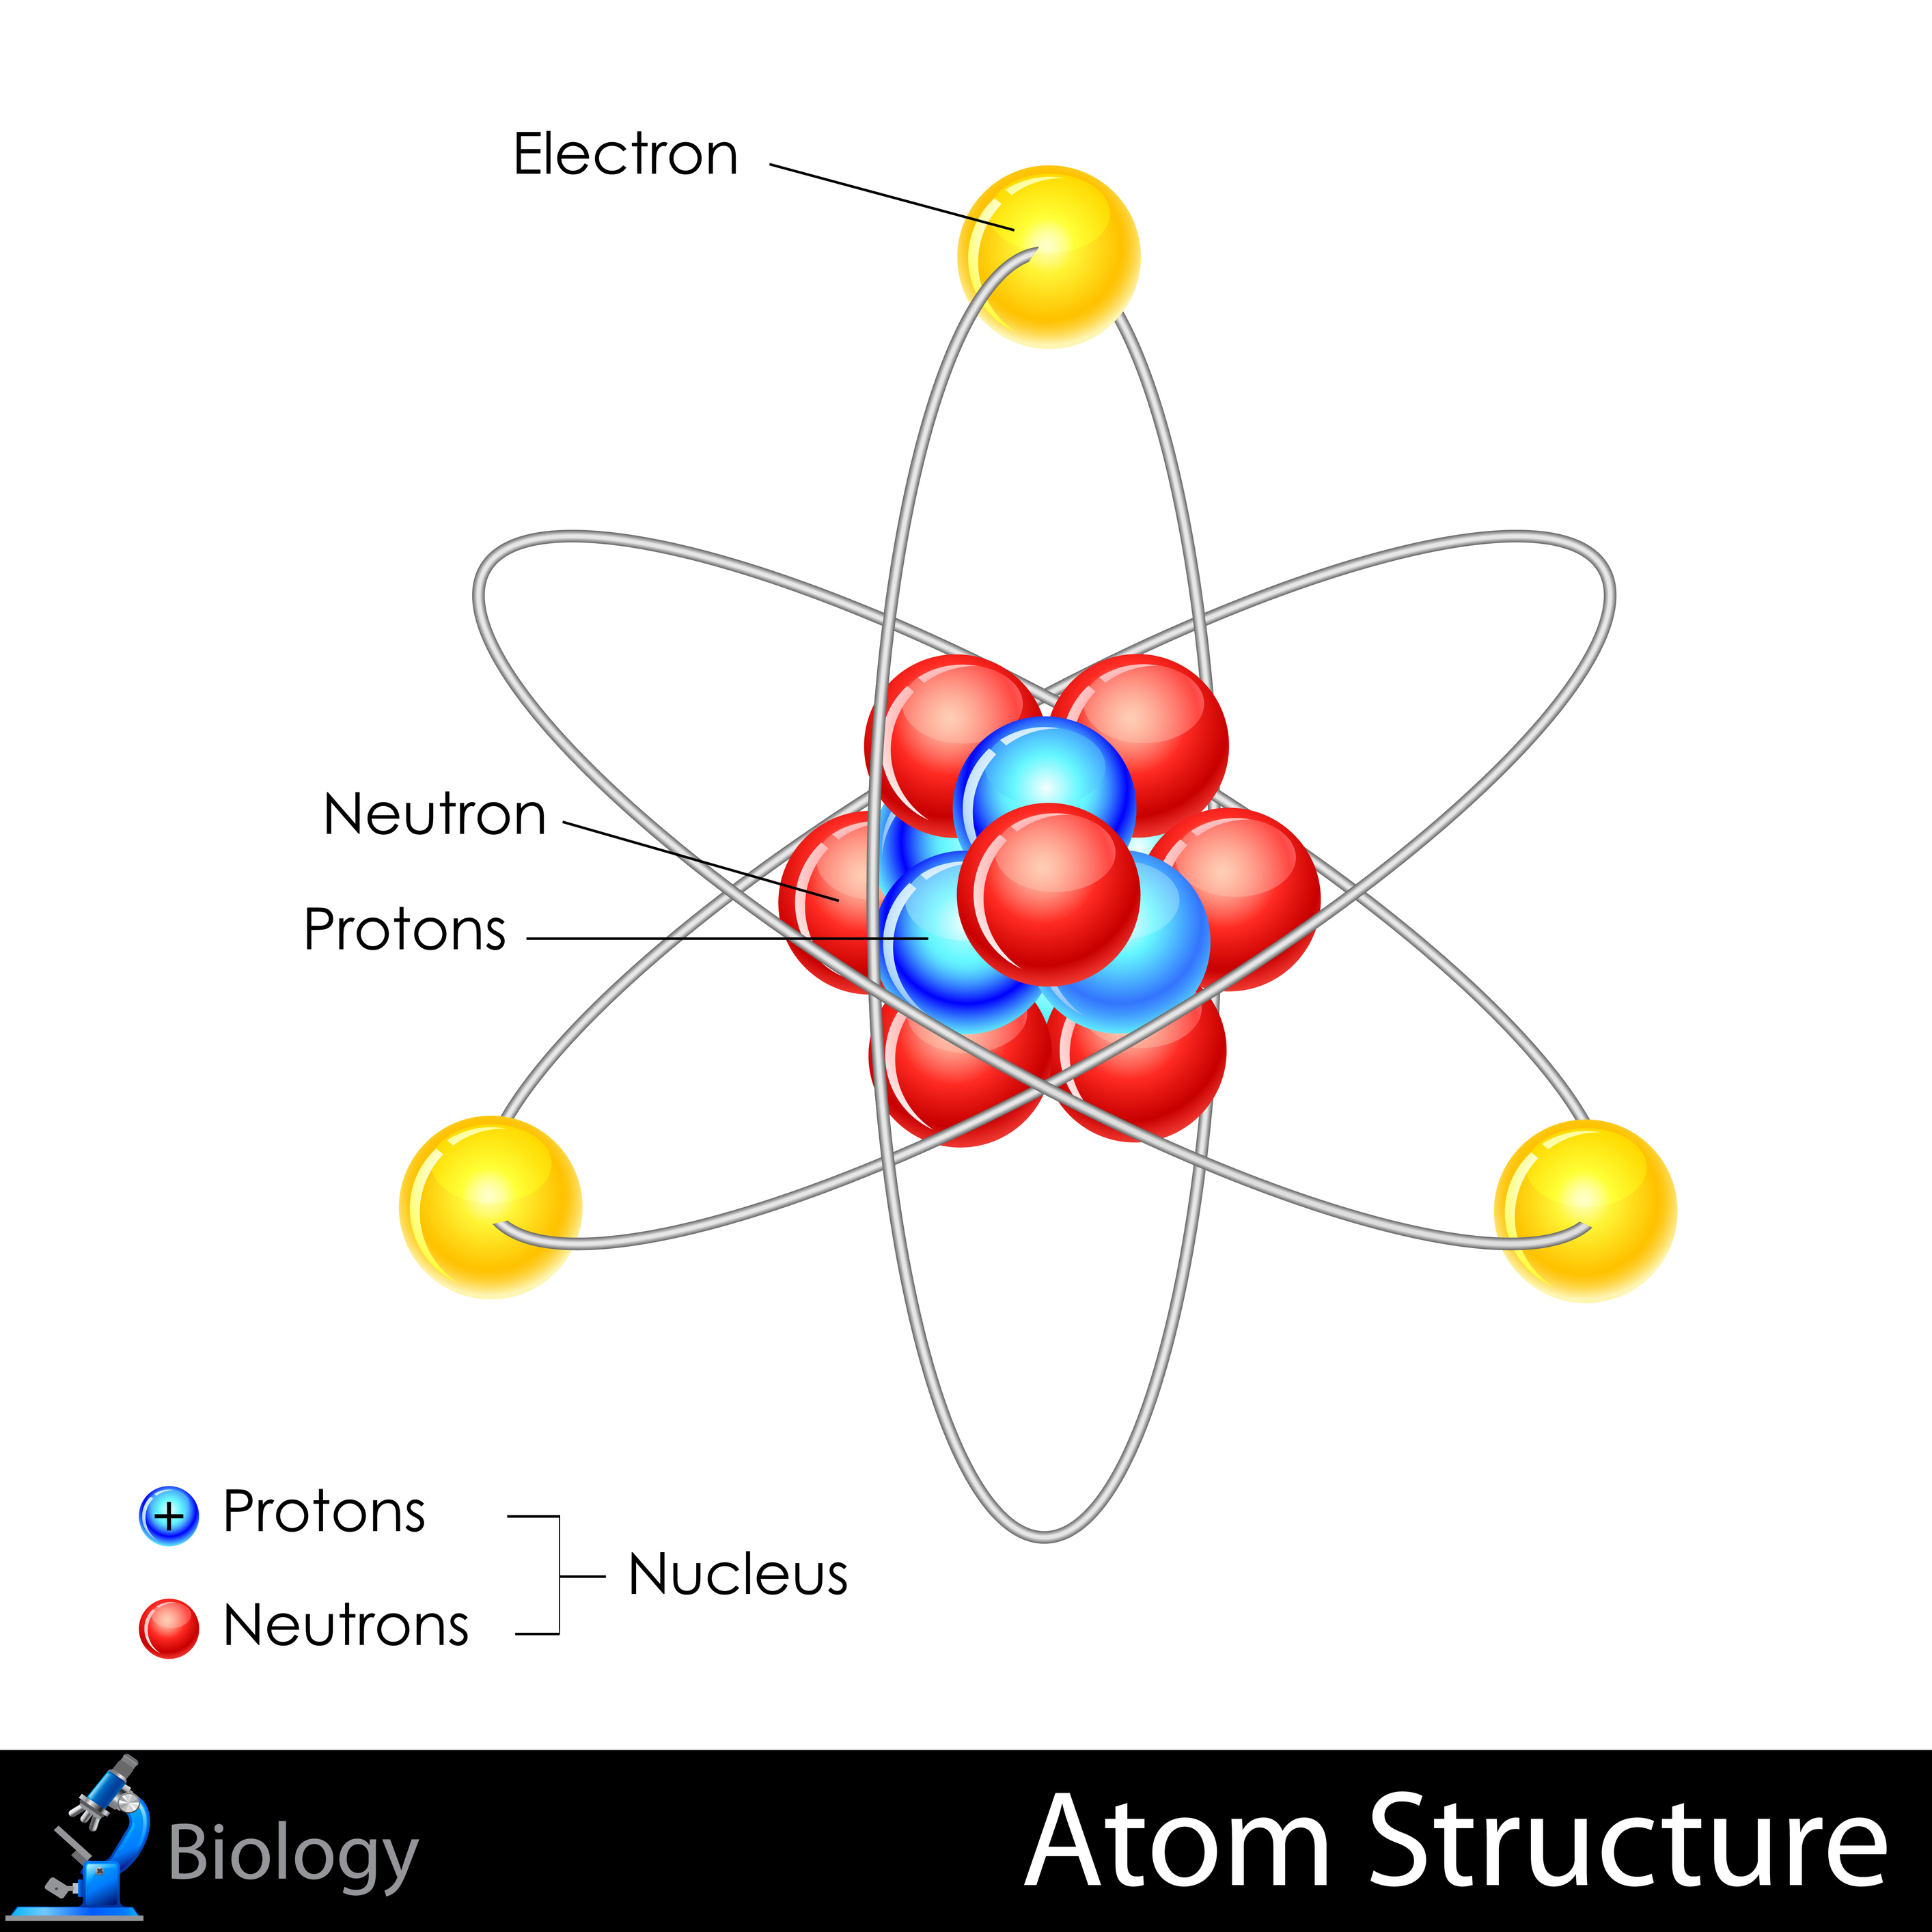
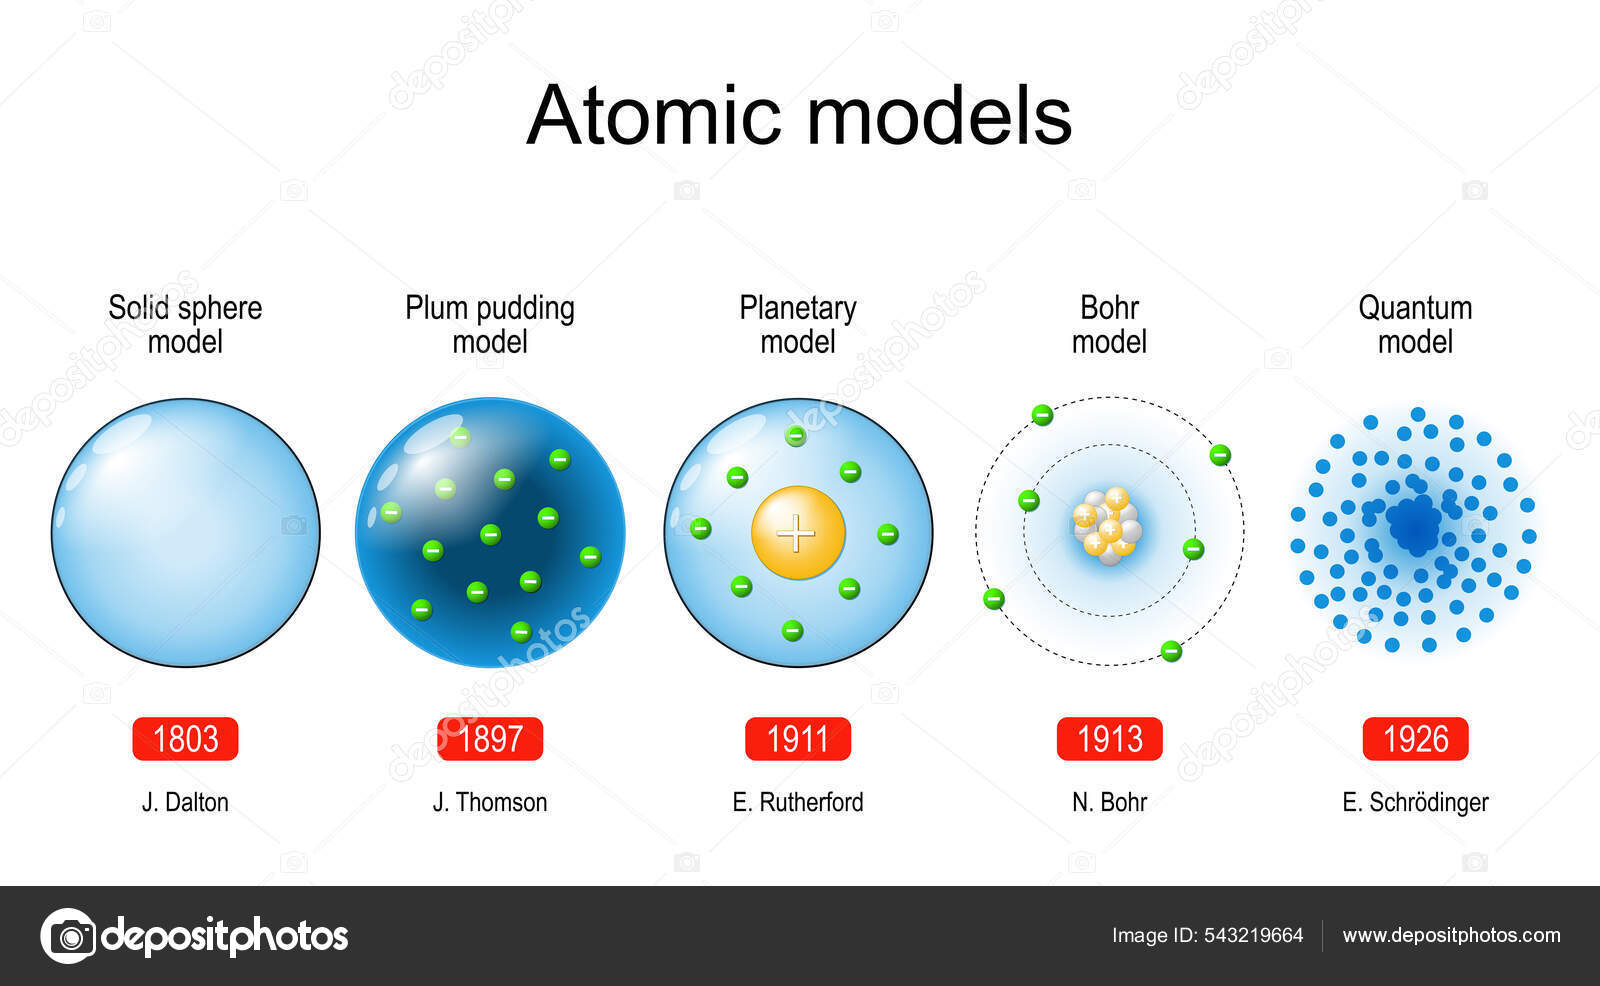
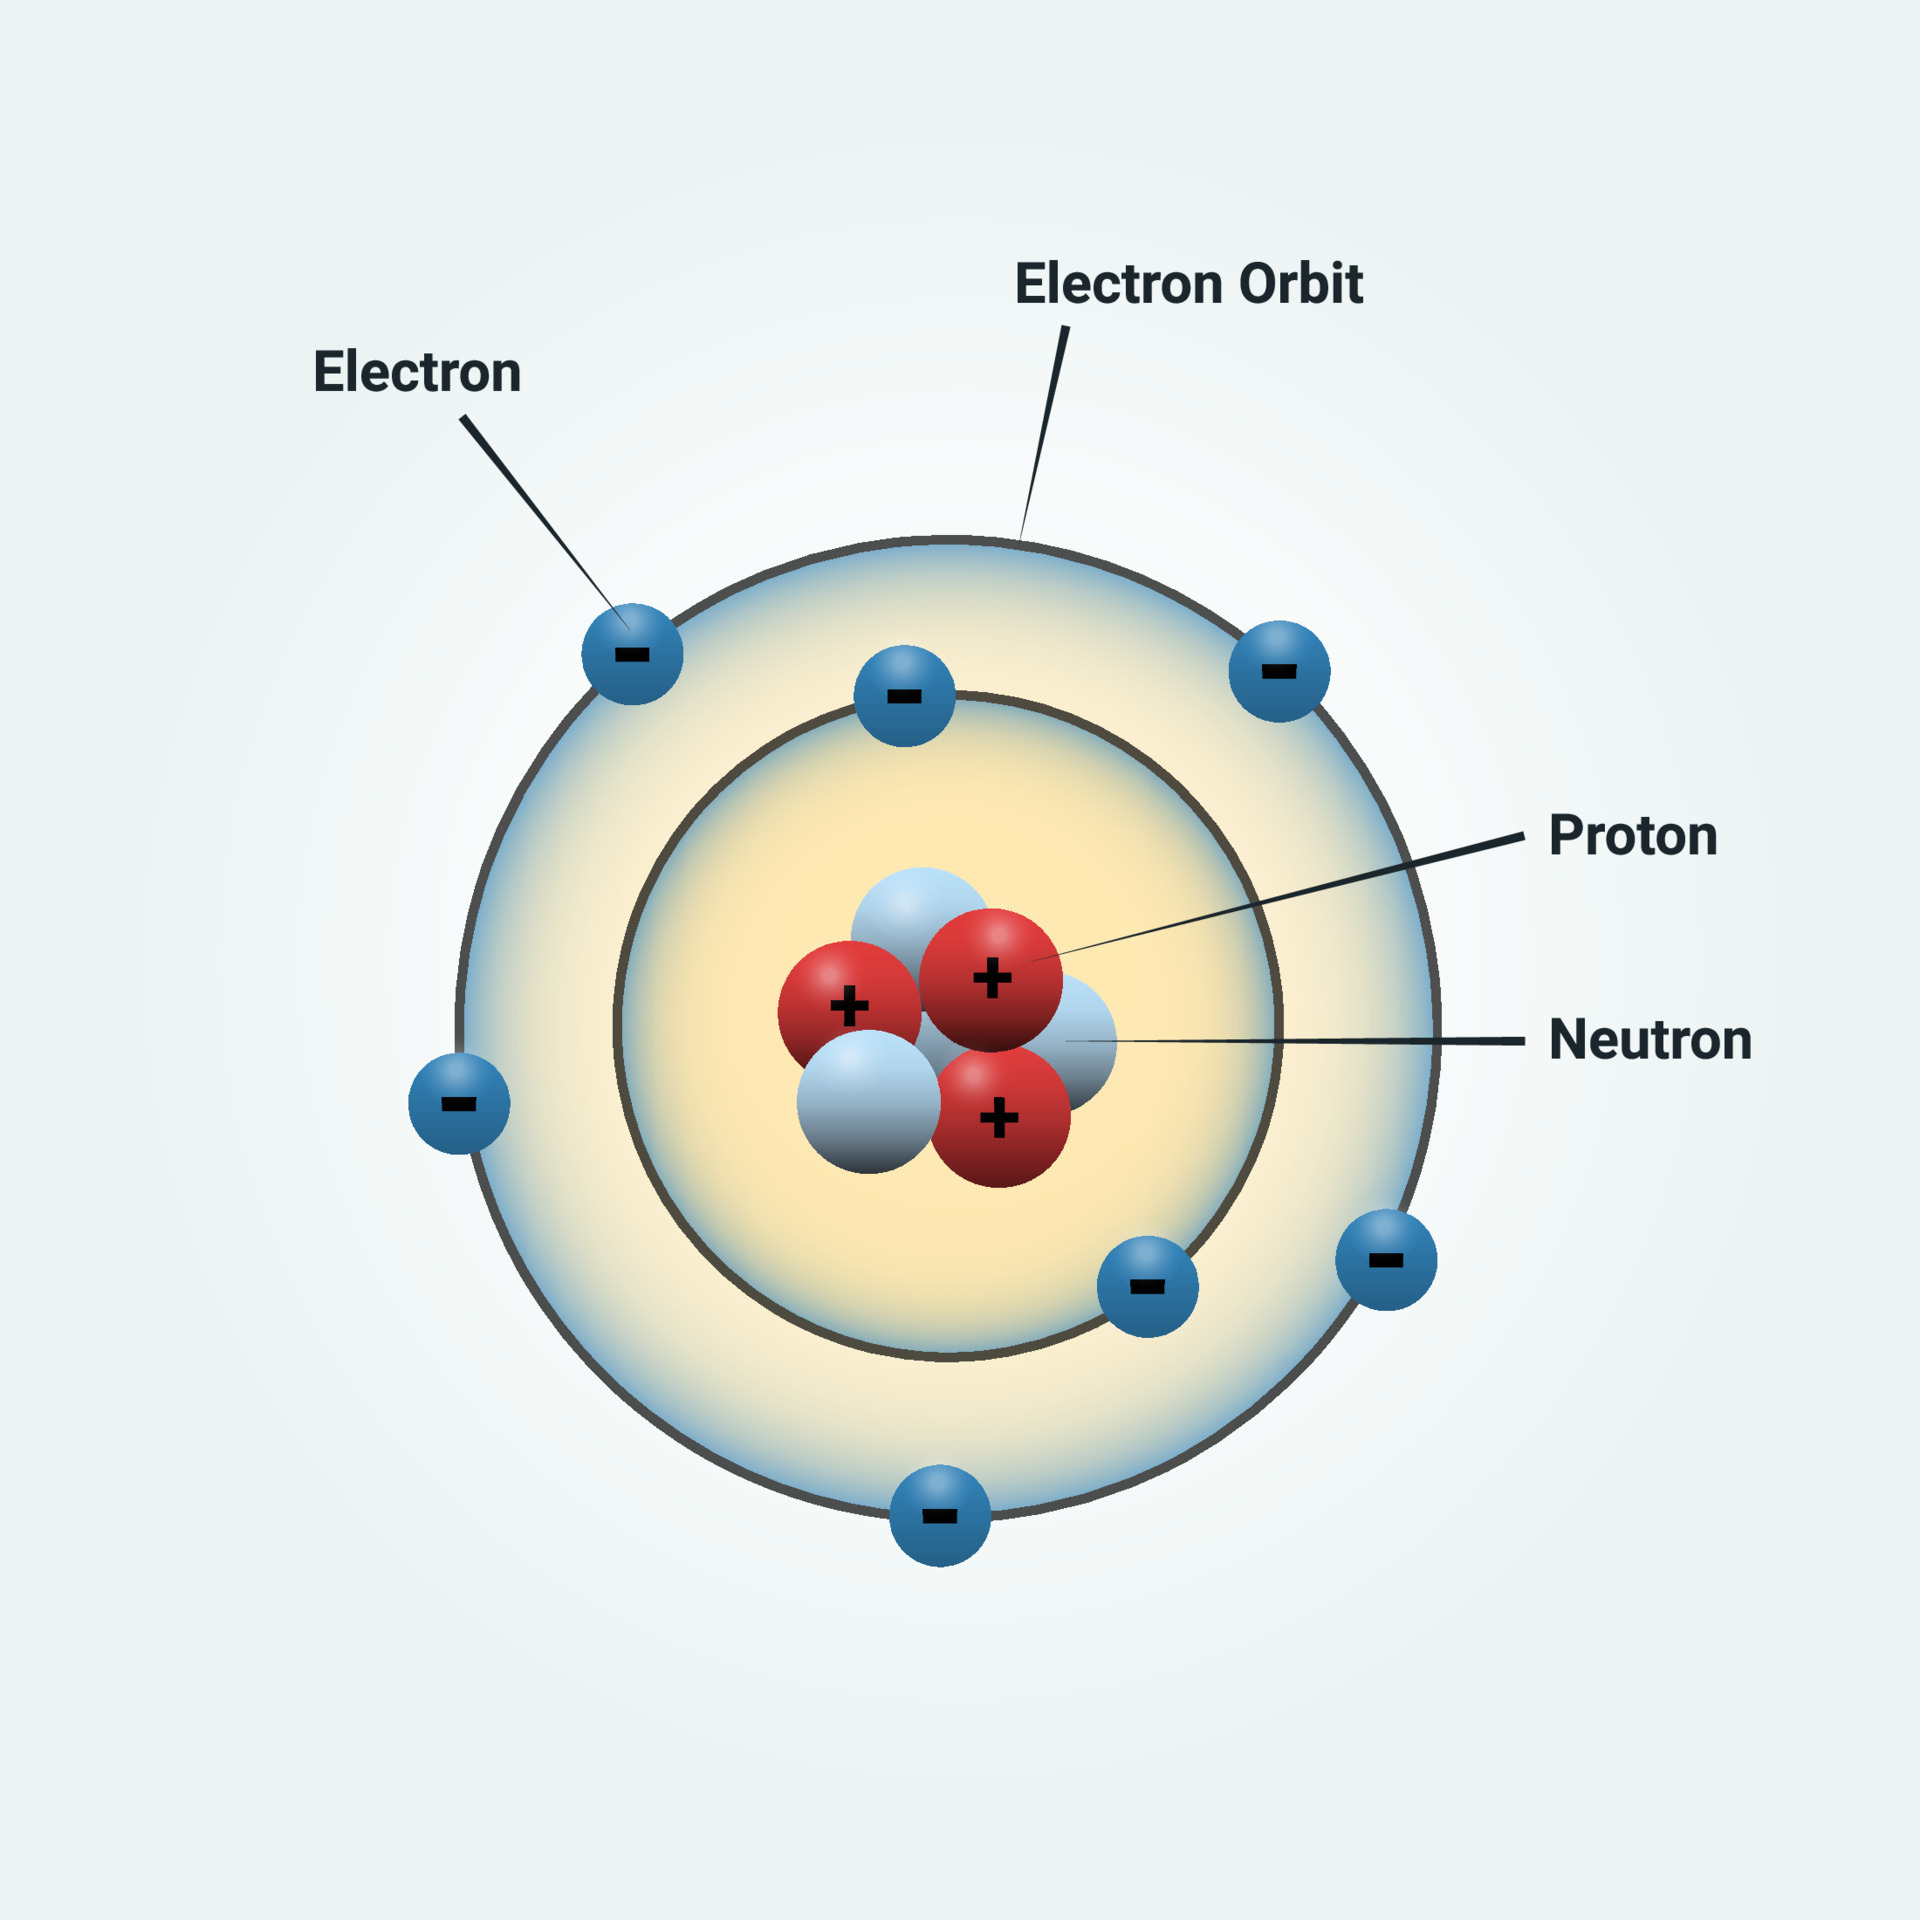
The Model Of An Atom has been rewritten so many times that most people treat it as settled fact when it's actually a stack of approximations built on top of each other like a house of cards. I learned this the hard way when I was trying to explain quantum tunneling to a group of undergrads and someone asked why we still teach the Bohr model if it's wrong. I didn't have a good answer for them because the honest answer is we teach it because it works for introductory calculations and nobody wants to deal with wavefunctions on day one. Dalton started it in 1803 with solid spheres. He had no idea what was inside an atom but he needed a way to explain why elements combined in whole number ratios. His model was effectively a billiard ball, nothing more. Thomson found the electron in 1897 and realized atoms weren't indivisible. His plum pudding model put negative electrons floating in a positive soup. It was wrong but it was the first time anyone treated the atom as having internal structure you could actually measure. Rutherford changed everything with the gold foil experiment in 1911. Most alpha particles went straight through the foil. A few bounced back. That meant the atom was almost entirely empty space with a tiny dense nucleus at the center. The math worked out that the nucleus was about 100,000 times smaller than the atom itself. If an atom were the size of a football stadium, the nucleus would be a marble on the 50 yard line. The rest is empty space and electrons zipping around somewhere in between.
Bohr fixed Rutherford's problem in 1913. Classical physics said orbiting electrons should radiate energy and spiral into the nucleus in a fraction of a second. Atoms would be unstable. They aren't. Bohr said electrons occupy fixed energy levels and only emit or absorb energy when jumping between them. It was a quantum rule slapped onto a classical picture. It worked perfectly for hydrogen and failed completely for everything else. But it introduced quantized energy levels which are real and still used today. The quantum mechanical model replaced orbits with orbitals around 1926. Schrödinger's equation doesn't give you a path for an electron. It gives you a probability distribution. An orbital is just a region where you have a 90 percent chance of finding the electron if you look. Heisenberg's uncertainty principle means you can never know both position and momentum precisely at the same time. This isn't a measurement problem. It's a fundamental property of reality.
What Nobody Tells You About Electron Configuration
The Aufbau principle, the way most people memorize it with that diagonal diagram, works for about 80 percent of the elements you'll encounter in practice. The exceptions exist for a reason and they break the simple rules. Chromium and copper are the classic textbook examples. Chromium should be [Ar] 4s2 3d4 but it's actually [Ar] 4s1 3d5. Copper should be [Ar] 4s2 3d9 but it's [Ar] 4s1 3d10. Half-filled and fully-filled d subshells are more stable than the rules predict because of electron-electron repulsion being minimized. These aren't quirks. They're consequences of the actual physics. I ran into this when I was setting up a computational chemistry workflow for a research group trying to model transition metal complexes. We had DFT calculations failing to converge on chromium complexes because the initial guess was using standard Aufbau configuration. The self-consistent field couldn't find the right ground state. The workaround was setting the initial occupation manually to [Ar] 4s1 3d5 instead of letting the code assume 4s2 3d4. Convergence took one iteration instead of failing outright. This kind of thing happens constantly with transition metals and nobody warns you about it until your calculations blow up.
Get the Full Details
The Orbital Shapes You Need To Actually Visualize
s orbitals are spheres. p orbitals are dumbbell shaped along the x, y, and z axes. d orbitals get weird with four of them looking like four-leaf clovers and one looking like a dumbbell with a donut around the middle. f orbitals are beyond visual intuition and you should stop trying to draw them accurately. The shapes come from the angular part of the wavefunction solutions to Schrödinger's equation. They're not physical objects. They're probability clouds. The boundary surface diagrams showing 90 percent probability are conventions, not measurements. Here's something that trips people up constantly. The radial distribution function for a 1s orbital peaks at the Bohr radius of about 53 picometers for hydrogen. That's the most probable distance from the nucleus. But the average distance is actually 1.5 times that. The electron isn't sitting at a fixed radius. It's smeared out in three dimensions and different statistical measures give you different answers. When someone says "the electron is at distance r," they should specify which measure they mean.
Where The Model Breaks Down Completely
The quantum mechanical model is the best we have but it's not exact for multi-electron systems. The Schrödinger equation can only be solved analytically for hydrogen and hydrogen-like ions with a single electron. Everything else requires approximations. Hartree-Fock treats each electron as moving in the average field of all the others. It's a mean-field theory and it misses electron correlation. Post-Hartree-Fock methods like MP2, CCSD(T), and CI add correlation back in but the computational cost scales poorly. CCSD(T) scales as N7 where N is the number of basis functions. You hit a wall pretty fast. DFT is the workhorse because it includes some correlation at much lower cost, typically N3 or N4. But it's not systematically improvable. You pick a functional and hope it's good enough. For main group chemistry, B3LYP or wB97X-D usually works. For transition metals, you need functionals designed for that like M06-2X or omegaHSEA2. Using B3LYP on a cobalt complex will give you qualitatively wrong results about half the time. The error isn't random either. It systematically underestimates spin state splittings and over-stabilizes low spin configurations. I learned this the hard way when a collaborator sent me DFT results for an iron-sulfur cluster using PBE functional and expected me to interpret the magnetic properties. PBE predicted the wrong ground spin state by 15 kcal/mol. The experimental value was clear. We switched to B3LYP* which is a modified version with less exact exchange and got agreement within 2 kcal/mol. This is a known issue in the literature but it doesn't appear in any textbook. The problem is that no single functional works well across all types of systems and you need to know which ones to trust for your specific case.
How To Actually Use Atomic Models In Practice
Start with what you're trying to calculate. If it's a simple organic molecule and you need geometries and energies, DFT with a medium-sized basis set like def2-SVP or 6-31G* is fine. If you need accuracy, go def2-TZVP or cc-pVTZ. Basis set superposition error becomes significant when you're dealing with weak interactions like hydrogen bonds or van der Waals complexes. Counterpoise correction fixes that but it doubles your computational cost. For molecular orbitals, don't trust the default output without checking. Some programs guess the symmetry and occupation. If you're dealing with open shell systems, check that your alpha and beta electron counts are correct and that you're getting the right multiplicity. I've seen people run unrestricted calculations and get alpha-alpha interactions that should have been alpha-beta because the initial guess put two electrons in the same spatial orbital with parallel spins. The SCF converges to a local minimum that's physically meaningless. When you need to explain atomic structure to someone, show them the quantum mechanical model but be honest about what it means. The electron isn't orbiting like a planet. It's a standing wave in three dimensions described by four quantum numbers. n determines the energy level. l determines the subshell shape. ml determines the orientation. ms is spin. Those quantum numbers come directly from solving the Schrödinger equation with boundary conditions. They're not arbitrary. The exclusion principle follows from the antisymmetry requirement of the wavefunction for fermions. This is why you can only put two electrons per orbital with opposite spins.
There's a practical shortcut for remembering orbital filling order that doesn't involve the diagonal diagram. Write out the orbitals in this order: 1s 2s 2p 3s 3p 4s 3d 4p 5s 4d 5p 6s 4f 5d 6p 7s 5f 6d 7p. Each line goes down one principal quantum number and across one azimuthal quantum number. The sum n plus l increases monotonically. That's Madelung's rule and it's derived from the hydrogenic energy levels with shielding effects approximated. It fails for heavier elements where relativistic effects become important. Gold's yellow color and mercury being liquid at room temperature are relativistic phenomena that the non-relativistic model can't explain. If you need reference data, NIST Atomic Spectra Database is the gold standard. It has energy levels, transition wavelengths, and configuration assignments for thousands of elements. The data comes from experimental measurements combined with theoretical calculations. It's freely available and well-curated. For computational work, the Gaussian basis set exchange has standardized basis sets across multiple programs. Don't mix basis sets from different libraries without checking the exponents. Some programs use different contraction schemes and the same name doesn't guarantee the same numbers.