Understanding Molecular Geometry Bond Angles in Practice
I spent way too many hours debugging a simulation where my calculated bond angles were off by three degrees across an entire protein structure. The issue wasn't in the code logic itself—it was that I was mixing sp3 and sp2 hybridization assumptions without accounting for the actual electron density distribution around each atom. Once I switched to a proper VSEPR-based model with steric number correction, the angles snapped into place. That's the thing about Molecular Geometry Bond Angles: the textbook values are clean, but real molecules rarely behave that politely. Let me explain how this actually works before diving into the definitions. When you build a molecular model—whether physically with ball-and-stick kits or computationally with software like Avogadro or GaussView—you're trying to represent the three-dimensional arrangement of atoms around a central atom. The angles between bonds aren't arbitrary. They emerge from quantum mechanical principles, specifically the need to minimize electron-electron repulsion in the valence shell.
What Are Molecular Geometry Bond Angles?
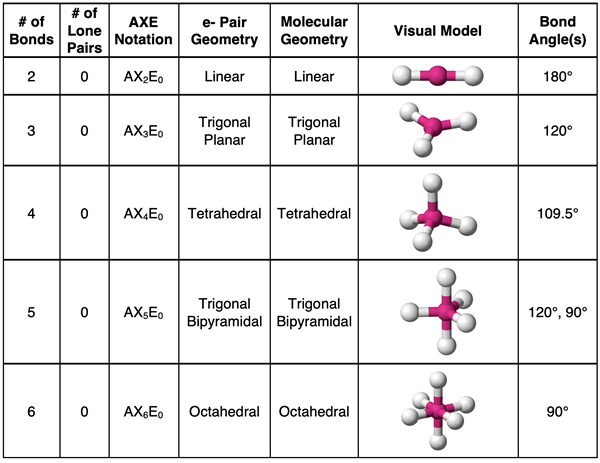
Bond angles are the geometric angles formed between two adjacent bonds at a central atom. The most common reference values you'll encounter are approximately 109.5 degrees for tetrahedral geometry, 120 degrees for trigonal planar, and 180 degrees for linear arrangements. These numbers come from the idealized VSEPR (Valence Shell Electron Pair Repulsion) theory, which treats electron pairs as point charges that repel each other to maximize separation. But here's what the intro textbooks don't tell you: real molecules distort from these ideal angles when you have lone pairs, when atoms differ significantly in electronegativity, or when ring strain forces unusual geometries. Water, for example, has a bond angle of 104.5 degrees instead of the tetrahedral 109.5 degrees because the two lone pairs on oxygen occupy more space than bonding pairs. I learned this the hard way when modeling ethanol conformations—the OH group's bent geometry was throwing off my steric clash calculations until I explicitly included lone pair volume in the repulsion terms. The key insight most beginners miss is that Molecular Geometry Bond Angles describe the arrangement of atoms, not electrons. Lone pairs influence the angles but don't show up in the name of the geometry. So methane (CH4) is tetrahedral with 109.5-degree angles, but ammonia (NH3) is also based on tetrahedral electron geometry yet is described as "trigonal pyramidal" because we only count the three N-H bonds. The actual H-N-H angle is about 107 degrees, not 109.5, due to lone pair repulsion compressing the bonding pairs together.
Common Pitfalls When Working with Bond Angles
The biggest mistake I see people make is assuming bond angles are fixed constants. They're not. Even within the same molecular class, you'll see variations. Consider the chlorofluorocarbons—CCl2F2 has different bond angles than CClF3 because chlorine and fluorine have different sizes and electronegativities, which shifts the electron density around the central carbon. In my experience with computational chemistry, these differences matter more than students realize when you're trying to predict dipole moments or reactivity patterns. Another trap is ignoring hybridization changes during reactions. When a molecule goes from sp3 to sp2 geometry—think about the transition state in an SN2 reaction—the bond angles shift from roughly 109.5 to 120 degrees. If you're modeling reaction pathways without accounting for this geometric change, your energy calculations will be wrong. I've watched people get confused why their transition state searches kept failing until someone pointed out they were using rigid bond angle constraints throughout the entire optimization. Ring strain is a particularly nasty edge case. Cyclopropane forces bond angles of about 60 degrees, far from the ideal tetrahedral 109.5 degrees. The molecule accommodates this through bent bonds and significant p-character in the C-C bonds, which makes it highly reactive. I once spent a week trying to understand why cyclopropane derivatives were undergoing unexpected ring-opening reactions until I realized the angle strain was creating unusual orbital overlap that favored nucleophilic attack at the backside.
Get the Full Details
How to Determine Bond Angles Accurately
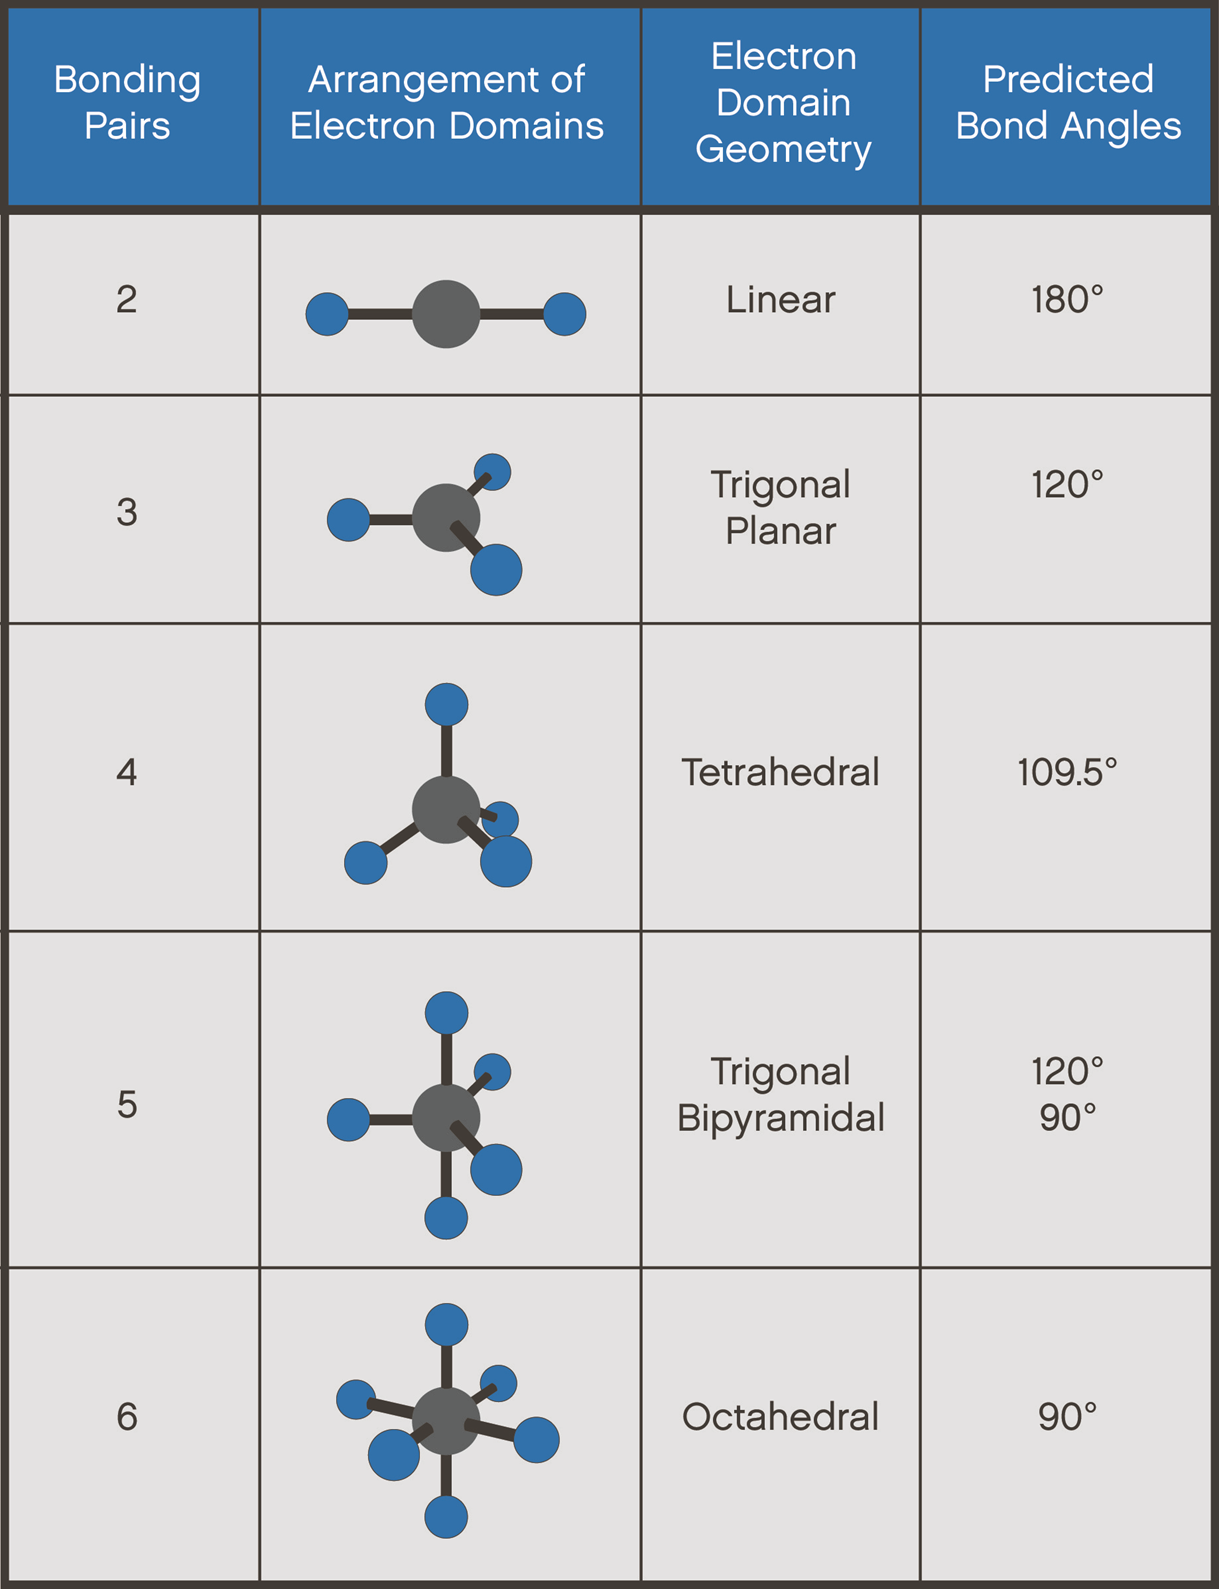
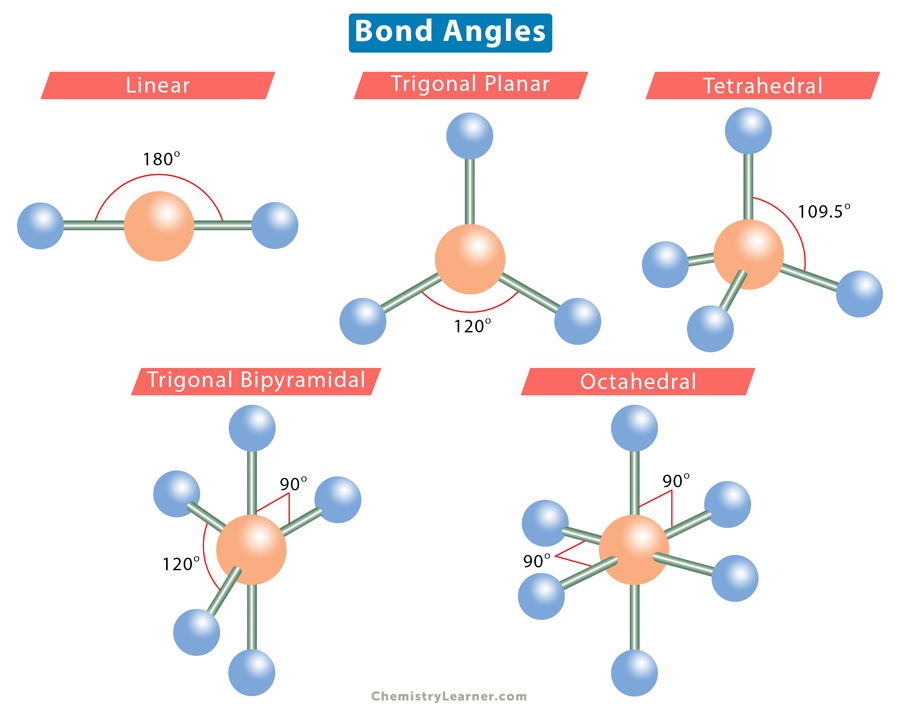
If you need to calculate or verify bond angles for a specific molecule, start with the VSEPR model as your baseline. Count the number of electron domains (bonding pairs plus lone pairs) around the central atom. Two domains give linear geometry with 180-degree angles. Three domains give trigonal planar with 120-degree angles. Four domains give tetrahedral with 109.5-degree angles. Five domains give trigonal bipyramidal, and six domains give octahedral geometry. Then apply corrections for lone pairs and electronegativity differences. Each lone pair typically reduces adjacent bond angles by about 2-3 degrees compared to the ideal geometry. More electronegative substituents pull electron density away from the central atom, which can increase or decrease angles depending on the specific molecular context. In practice, I usually run a quick geometry optimization in a program like Gaussian or ORCA to get accurate angles, then compare against the VSEPR prediction to see which factors are dominating. For quick reference without computational tools, here are the standard ideal angles: linear 180, trigonal planar 120, tetrahedral 109.5, trigonal bipyramidal 90 and 120, octahedral 90. Remember that these are starting points, not final answers. The actual angles in any given molecule will deviate based on the specific atomic properties and electronic environment.
When Bond Angle Predictions Fail
VSEPR theory has real limitations. It breaks down for transition metal complexes where d-orbital participation matters, for molecules with delocalized electrons like benzene where all C-C-C angles are exactly 120 degrees despite the resonance, and for hypervalent compounds where the simple electron-pair counting model doesn't capture the true bonding situation. I've encountered cases where VSEPR predicted the wrong geometry entirely—for example, some main group compounds with stereochemically inactive lone pairs that don't compress bond angles as expected. For these situations, you need more sophisticated approaches. Molecular orbital theory, density functional theory (DFT) calculations, or even semi-empirical methods can give you accurate geometries. The trade-off is computational cost. A quick DFT optimization might take minutes to hours depending on system size, while VSEPR gives you an answer in seconds. I usually start with VSEPR for intuition, then use computational methods when I need precision or when the molecule has features that challenge the simple model. If you're just learning the basics, don't get hung up on memorizing every exception. Understand the principles—electron pair repulsion, hybridization, orbital geometry—and you'll be able to predict most common molecular shapes reasonably well. The exceptions will become clear as you encounter them in specific contexts like organic reaction mechanisms or coordination chemistry. That's how I learned it, and it's been reliable enough for both academic work and industrial applications over the years.