Calculating Molecular Mass in Real Lab Work
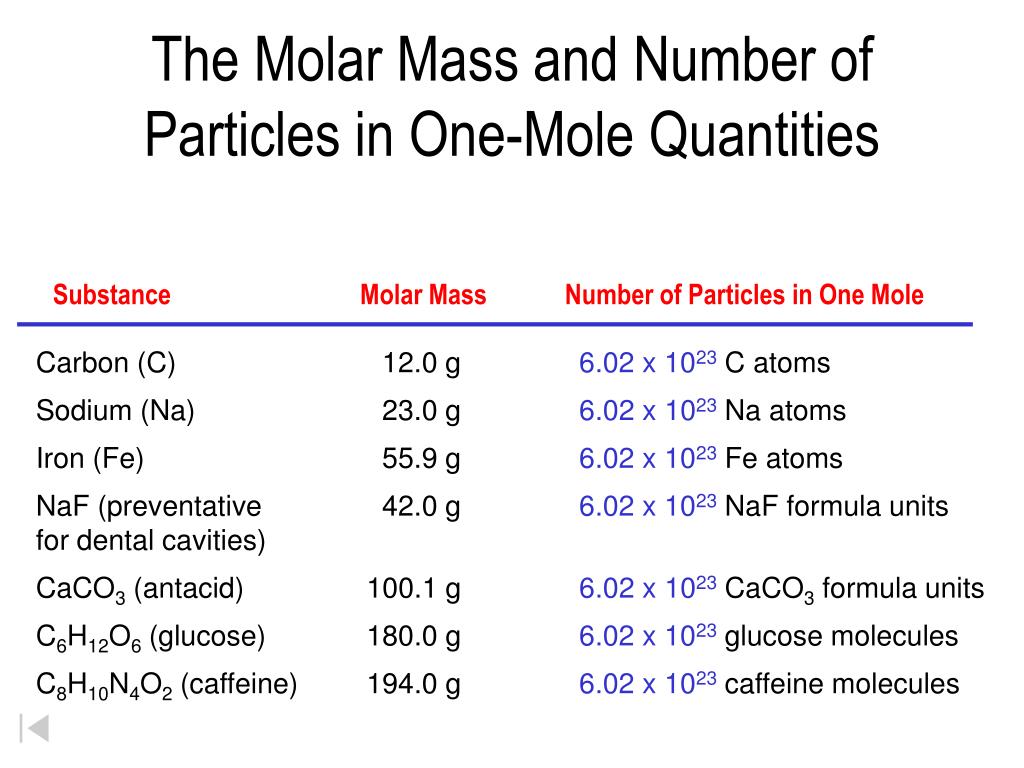
The quick version: you sum the atomic masses of every atom in your formula. Sodium chloride is 22.99 plus 35.45, which gives you 58.44 g/mol. But that simple arithmetic only gets you through undergraduate chemistry labs. The moment you start working with real compounds, real instruments, and real publication requirements, the process gets messier than a textbook suggests. I spent three days last year debugging why my HPLC calibration curves were drifting by 2.3 percent across batches of a particular peptide standard. The culprit wasn't the column, wasn't the mobile phase, wasn't the detector. It was the molecular weight I had been using from a supplier's certificate of analysis. That certificate listed 1247.5 g/mol based on the monoisotopic mass of the sodium adduct form, but my method was quantifying against a free acid reference standard. I had been calculating concentrations off a mass that was 0.04 percent higher than what I actually had in the vial. Over multiple dilution steps, that error compounded into a meaningful bias. The fix was straightforward once I spotted it: I recalculated everything using the average molecular weight of the free acid form, pulled from a proper atomic mass table rather than trusting the cert. Took about twenty minutes.
Understanding the Difference Between Molecular Mass and Molecular Weight
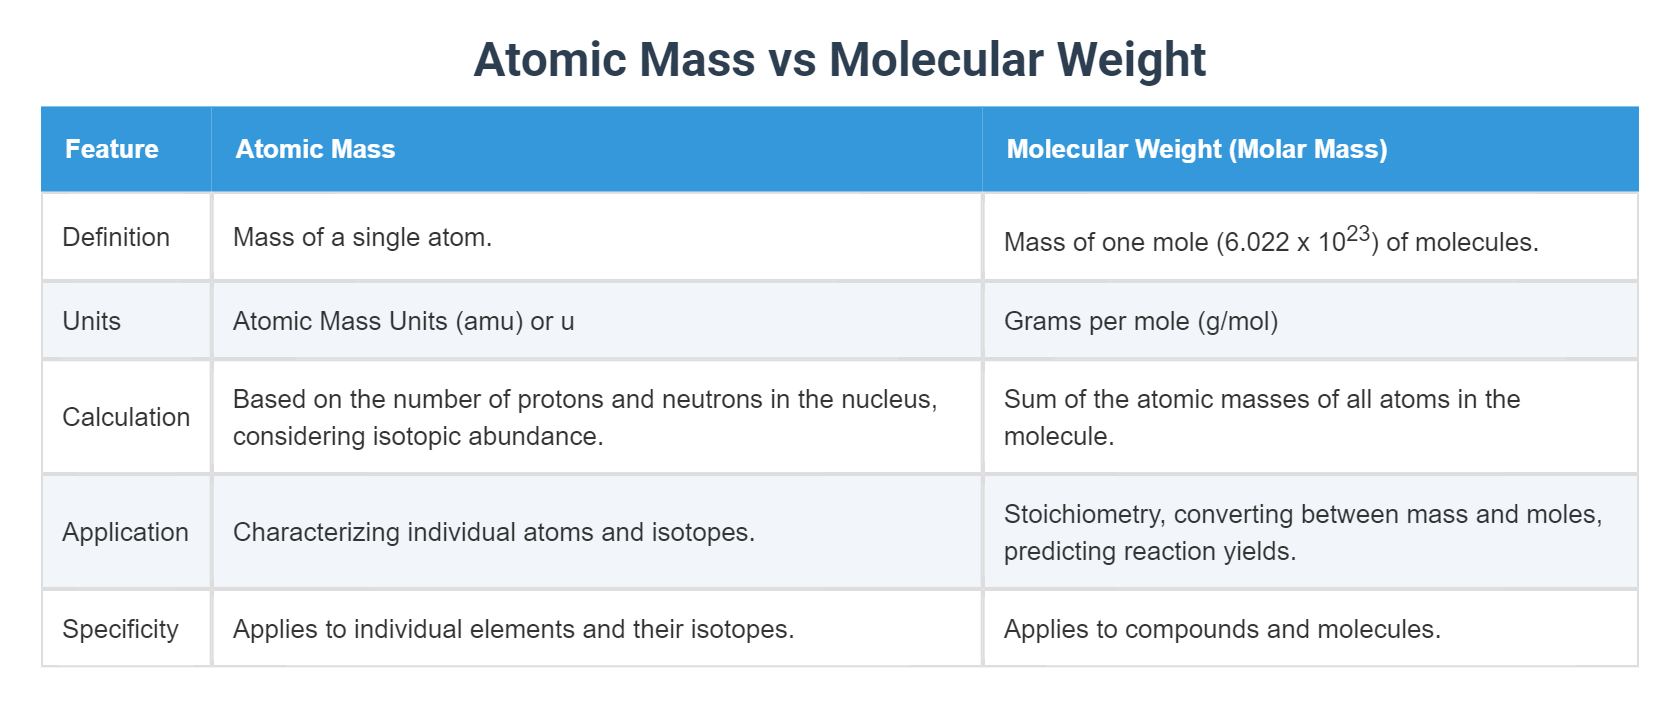
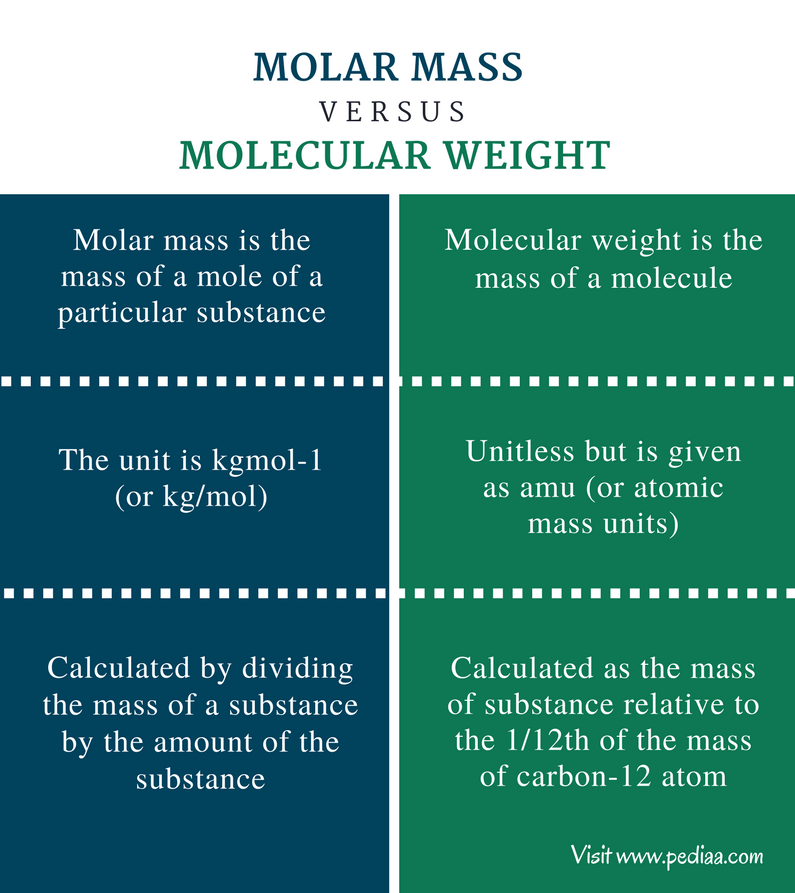
People use these terms interchangeably, and most of the time that works fine. But they are technically different things, and confusing them will bite you if you ever need high precision. Molecular mass refers to the mass of a single molecule, typically expressed in daltons or unified atomic mass units. It is a property of one discrete entity. Molecular weight, or more correctly molar mass, is the mass of one mole of that substance, expressed in grams per mole. For most practical purposes in organic chemistry and biochemistry, the numerical values are identical because one dalton is defined as one gram per mole. The distinction matters when you are dealing with polymers, proteins, or any heterogeneous mixture where there is no single molecular mass to speak of. A protein with fifty different glycoforms doesn't have one molecular mass. It has a distribution. Reporting a single molecular weight for that sample is useful but inherently approximate. Another thing most people gloss over is the difference between monoisotopic mass and average mass. Monoisotopic mass uses the mass of the most abundant isotope for each element. Carbon is 12.00000, hydrogen is 1.00783, nitrogen is 14.00307, oxygen is 15.99491. Average mass uses the weighted average of all naturally occurring isotopes from the periodic table. Carbon becomes 12.011, hydrogen becomes 1.008, and so on. For small molecules under 500 daltons, the difference is usually negligible. For a peptide at 2000 daltons, the gap between monoisotopic and average mass can be several daltons. Matrix-assisted laser desorption ionization time-of-flight instruments resolve monoisotopic peaks for smaller peptides, but for anything over roughly 3000 daltons, you are looking at envelope peaks and average mass becomes the more relevant number. If you feed the wrong mass value into your method, your retention time predictions, your internal standard corrections, and your concentration calculations will all be slightly off.
How to Actually Calculate It Without Making Mistakes
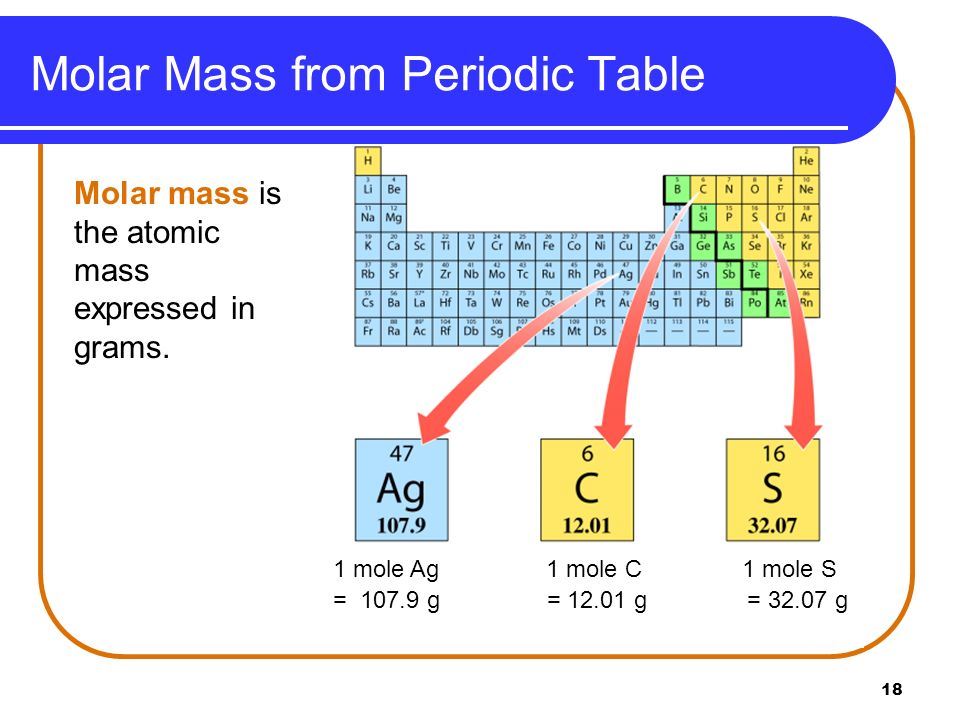
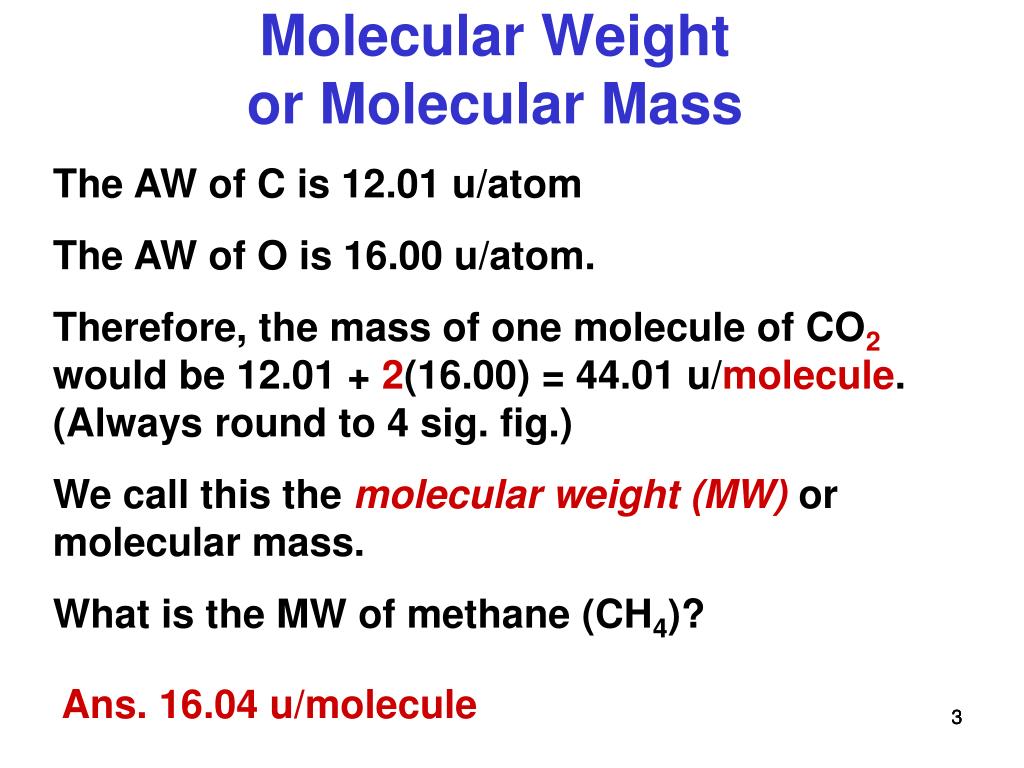
The manual way is tedious and error-prone. The reliable way is to use a proper tool. I use ChemAxon's MarvinSketch for quick lookups because it handles tautomers and protonation states reasonably well, and I use the NIST Chemistry WebBook when I need traceable standard reference data. For calculations across many compounds, I run a small Python script using the rdkit library, which pulls atomic masses directly from its internal periodic table and sums them based on the parsed SMILES string. That cuts a batch of fifty compounds from maybe an hour of manual calculation down to about thirty seconds. Here is the basic procedure whether you are doing it by hand or by software. Write out the full molecular formula. Make sure you have accounted for every atom including hydrogens, which people routinely forget when reading condensed structural formulas. Look up each element's atomic mass from a current periodic table. NIST publishes the most authoritative values and they update them periodically, so check the date on your source. Multiply each atomic mass by the number of atoms of that element. Sum everything. If you need monoisotopic mass, use the monoisotopic values. If you need average molar mass, use the standard atomic weights. Do not mix the two within a single calculation. I should mention one common failure mode that trips up everyone at least once. When you are working with salts, hydrates, or counterions, the molecular weight you report depends entirely on what form the compound is actually in. If your reagent bottle says the compound is a hydrochloride salt monohydrate, you need to include the HCl and the water in your molar mass calculation unless your protocol explicitly calls for the free base. I saw a methods section in a journal once where the authors stated they dissolved their compound to make a 10 mM stock solution, but they used the free base molecular weight while the actual reagent was the sulfate salt dihydrate. Their stock was 13 percent more concentrated than they thought, and every downstream concentration in the paper was wrong by the same margin. The reviewers missed it. The error propagated into every calculated IC50 value. It is the kind of mistake that is almost impossible to catch after the fact unless you go back to the original reagent certificate and compare it against what they reported.
Get the Full Details
When Simple Calculation Fails Completely
The biggest limitation anyone should understand is that molecular weight calculation assumes you know the exact molecular formula. That sounds obvious until you are dealing with natural product extracts, polymer samples, or biological lysates. A synthetic polymer like polyethylene glycol does not have a single molecular weight. It has a number-average molecular weight, a weight-average molecular weight, and a z-average molecular weight, each measured differently and each telling you something different about the sample. Gel permeation chromatography gives you a distribution, not a number. Reporting a single molecular weight for PEG-4000 is conventional shorthand, but it is shorthand nonetheless. The actual sample contains chains ranging from maybe 2000 to 6000 daltons depending on the manufacturer and the batch. Ions and adducts are another area where the simple calculation goes sideways. If you are doing mass spectrometry and your analyte shows up as a sodium adduct [M+Na]+, the observed m/z is not the molecular weight of your compound. It is the molecular weight plus 22.989. If you forget to subtract the adduct mass before using the value for anything else, everything downstream breaks. I have seen people in metabolomics papers use the observed m/z directly as the compound mass in their quantification calculations. It produces systematically inflated molecular weights across the entire dataset. Isotopic labeling compounds introduce yet another layer. Deuterated internal standards do not have the same molecular weight as their non-deuterated counterparts, obviously, but the difference is not always the obvious integer you might expect. Each deuterium substitution adds approximately 1.0063 daltons rather than exactly 1.0 because deuterium is 2.01410 rather than 2.00000. For a standard with six deuterium labels, the mass shift is about 6.038 daltons, not 6.0. That seems trivial until you are calibrating a high-resolution instrument and the shift puts your internal standard peak in the wrong bin relative to your analyte.
Practical Guidelines for Molecular Mass Molecular Weight Determination
Match the mass type to your application. Use monoisotopic mass for small molecule mass spectrometry and peak assignment. Use average molar mass for stoichiometric calculations, solution preparation, and anything involving gravimetric measurements. There is a small but real advantage to using average mass for solution work because balance measurements integrate over the natural isotopic distribution present in your reagent. A balance cannot distinguish between carbon-12 and carbon-13 atoms in a solid sample, so the average mass is the physically correct value to use when you weigh out a compound. Always verify your molecular weight against at least two independent sources when precision matters. I cross-reference whatever a vendor provides against the PubChem compound summary and against a calculation I run myself. Discrepancies between these three sources almost always point to a salt form, hydrate, or tautomer issue rather than a calculation error. If all three disagree on a value that should be a simple sum, check the chemical form before you check your arithmetic. Document the exact form you used for the calculation. State whether it is monoisotopic or average, whether you included counterions or solvent molecules, and which atomic mass standard you referenced. This is trivially easy to do and it prevents confusion when someone else tries to reproduce your work or when you return to the data six months later and cannot remember whether you had accounted for the water of hydration.
For routine lab work, the whole process from formula to final molecular weight takes roughly five minutes per compound if you are doing it carefully by hand, or under a second per compound if you are using an automated script. The time investment is minimal compared to the cost of catching an error after you have already prepared standards, run experiments, and collected data. The only situation where calculation-based molecular weight is insufficient is when you are working with truly undefined or heterogeneous materials, in which case you need empirical measurement methods like osmometry, light scattering, or SEC-MALS rather than any formula you can write down on paper.