Why VSEPR Theory Isn't Actually Useful the Way People Think
Most people learn molecular shape and geometry through VSEPR — valence shell electron pair repulsion — which works fine for simple textbook molecules but falls apart fast when you run into transition metals, hypervalent species, or anything with delocalized electrons. I spent years building models by hand, then moved into computational chemistry, and the gap between what the theory predicts and what actually exists is where most students get lost.
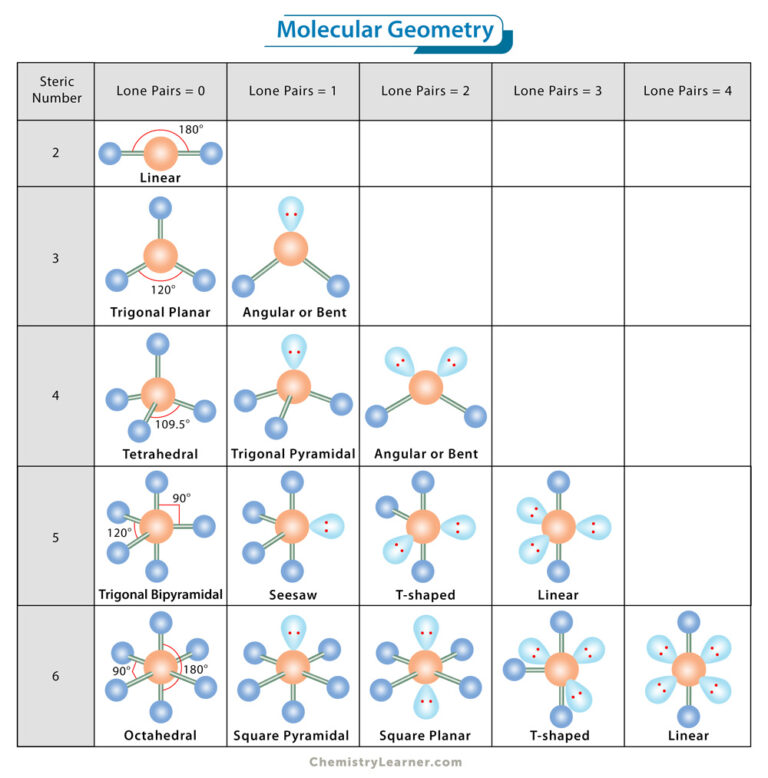
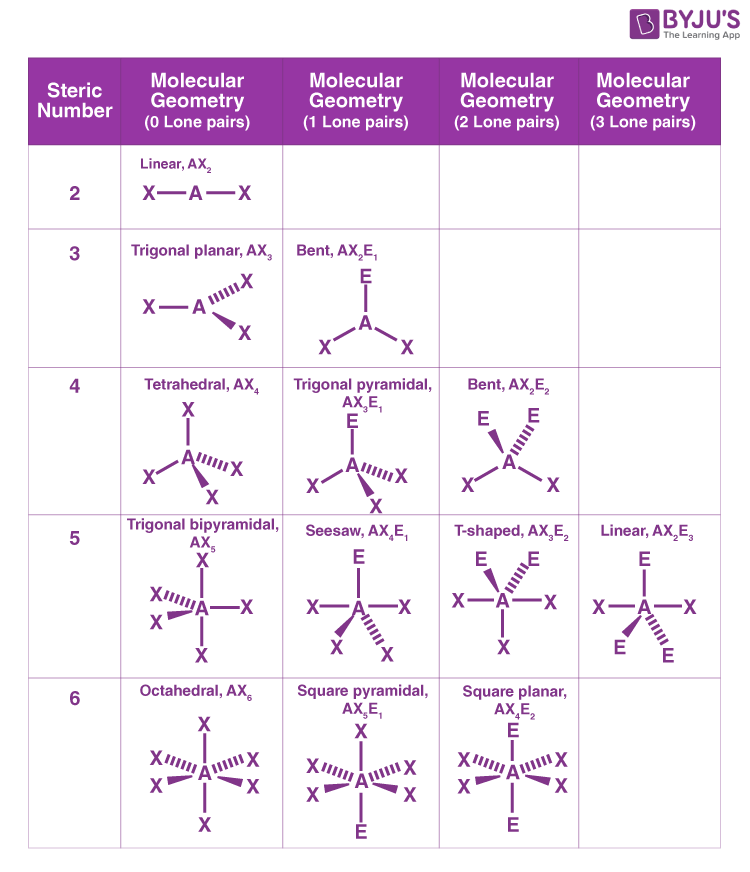
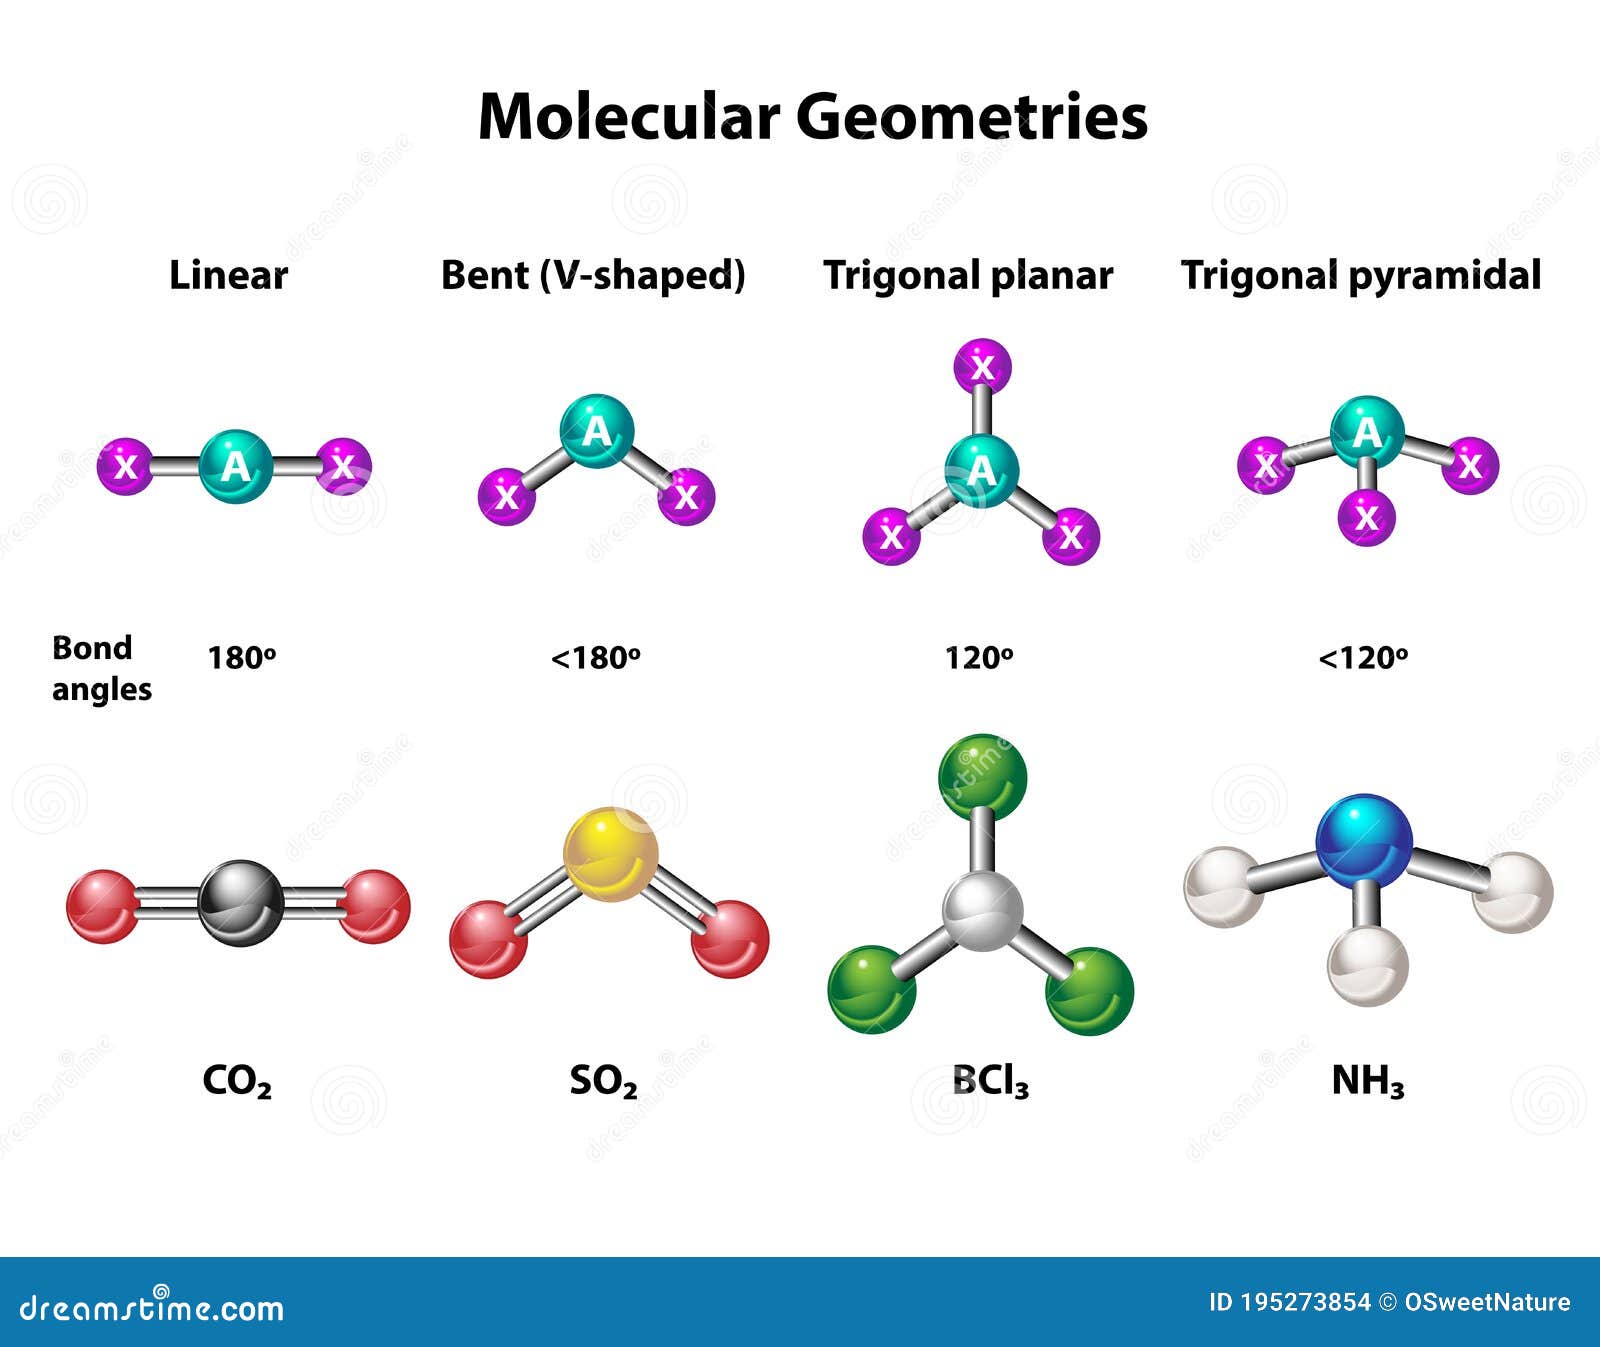
The basic idea is still worth knowing. Electron domains around a central atom arrange themselves to minimize repulsion. Two domains give linear, three give trigonal planar, four give tetrahedral, and so on. Lone pairs take up more space than bonding pairs, which is why water is bent instead of linear even though it has four electron domains. This is standard curriculum and there's no point pretending it's wrong — it's just incomplete.
Practical Molecular Shape And Geometry Beyond the Textbook
When you're actually working with molecules, the geometry you care about comes from two sources: experimental data and computational prediction. X-ray crystallography gives you bond angles and distances directly for solid samples. Gas-phase electron diffraction works for volatile compounds. NMR coupling constants can infer dihedral angles through the Karplus relationship. Computational chemistry — specifically DFT calculations at the B3LYP/6-31G* level or better — gives you optimized geometries that are usually within a few degrees of experimental values for organic molecules.
The computational route is what I use most often now. I run a geometry optimization, check for imaginary frequencies to confirm it's a true minimum, and then read off the bond angles from the output. It takes about ten minutes on a decent workstation for a medium-sized molecule, compared to hours of manual model-building I used to do. The catch is that DFT geometries can drift by two to three degrees from reality depending on the functional and basis set, which matters if you're trying to predict something sensitive like a reaction barrier or an NMR coupling constant.
Common Mistakes That Cost Me Hours of Troubleshooting
The biggest issue I see people make is treating molecular shape and geometry as interchangeable terms. They're related but not the same. Geometry refers to the positions of all atoms in space. Shape, in the VSEPR sense, usually means the arrangement you describe when you ignore lone pairs and just look at the atomic positions. For ammonia, the geometry is tetrahedral around nitrogen because of the four electron domains, but the molecular shape is trigonal pyramidal because the lone pair isn't an atom. Students mix this up constantly and then get confused when their professor marks them wrong.
Another problem is assuming that steric number determines everything. With d-block complexes, crystal field effects and ligand field stabilization energies dominate over simple repulsion arguments. A d8 metal like Pt(II) will almost always adopt square planar geometry regardless of what VSEPR might suggest, and trying to force it into a tetrahedral framework just creates nonsense. I once spent an afternoon trying to rationalize the geometry of a cobalt complex using electron domain counting before someone pointed out that I was dealing with a strong-field low-spin case where the d-orbital splitting pattern made square planar far more stable than tetrahedral. That one mistake cost me a day of work.
How to Determine Geometry When You Actually Need To
If you're doing this by hand without software, start by drawing the Lewis structure correctly. Count all valence electrons, place the central atom, distribute bonds, and put remaining electrons on atoms as lone pairs. Make sure formal charges are minimized — a structure with a +2 on carbon and a -2 on oxygen isn't wrong in terms of electron counting but it's almost certainly not the right one.
Then count electron domains around the central atom. Single bonds count as one domain. Double and triple bonds also count as one domain each for geometry purposes, though they do exert slightly more repulsion than single bonds. Lone pairs count as one domain. The total gives you the steric number, which maps to a base geometry. Then remove the lone pairs from your description to get the molecular shape.
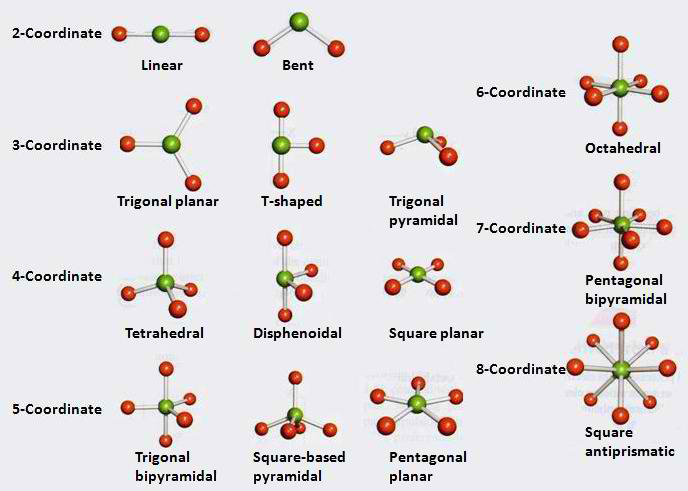
For transition metals, skip the VSEPR shortcut. Look up the coordination number and common geometries for that oxidation state and d-electron count. Four-coordinate d8 metals are square planar. Four-coordinate d10 metals are tetrahedral. Six-coordinate is almost always octahedral unless there's a strong Jahn-Teller distortion, in which case you'll see elongated or compressed octahedra with noticeably different axial and equatorial bond lengths.
When Computational Methods Fail and What to Do Instead
DFT geometry optimization doesn't always converge. Sometimes it oscillates between conformations, sometimes it finds a local minimum that looks reasonable but isn't the global minimum, and sometimes it just refuses to converge no matter what you do. I've had cases where the optimizer kept drifting toward a symmetric structure that had a tiny imaginary frequency — a transition state, not a minimum. Running a frequency calculation immediately after optimization catches this, and it should always be the second step after any geometry optimization.
For very large molecules, semiempirical methods like PM6 or GFN2-xTB give geometries in seconds that are usually good enough for qualitative purposes. They're not accurate for quantitative predictions but they're useful for getting a starting structure before running a fuller DFT calculation. I typically use xTB for initial geometry generation on anything larger than thirty heavy atoms, then refine with DFT if I need precision.
For molecules with strong multireference character — things like singlet oxygen complexes, certain transition states, or diradicals — single-reference DFT gives garbage geometries. In those cases you need CASSCF or at least a broken-symmetry DFT approach, which is significantly more work and requires more expertise to set up correctly. If you're encountering weird bond lengths or angles that don't make chemical sense, multireference character is worth considering.
A Specific Case Where Standard Approaches Broke Down
I was modeling a phosphorus-containing compound with a P=O bond and a nearby positively charged nitrogen. The standard DFT optimization kept producing a structure where the P=O bond was 1.48 angstroms instead of the expected 1.43, and the O-P-N angle was compressed to about 100 degrees instead of the tetrahedral norm. The frequency calculation showed a small imaginary mode corresponding to oxygen motion toward the nitrogen. The issue was that the implicit solvation model I was using didn't account for the strong electrostatic interaction between the phosphoryl oxygen and the cationic nitrogen. Switching to an explicit solvent molecule positioned between the two charges resolved the problem immediately, and the geometry came out clean. This is the kind of thing that doesn't show up in any textbook — you just learn it by hitting the wall.
Recommended Tools and Where to Get Them
For quick geometry visualization and basic VSEPR prediction, Avogadro is free and runs on all platforms. It has a built-in builder that shows electron domains and predicted shapes. For actual computational geometry work, Gaussian is the standard but it requires a license. ORCA is a good free alternative that handles DFT geometry optimizations well and has decent documentation. For semiempirical and tight-binding calculations, the xTB package from Grimme's group is free and fast — you can download it from github or grab a precompiled binary. WebMO provides a web interface for multiple quantum chemistry packages if you want a GUI wrapper.
The practical workflow I recommend is: build the structure in Avogadro, optimize it with xTB for a quick check, then run a DFT geometry optimization in ORCA with a frequency calculation to confirm it's a minimum. This usually takes under thirty minutes for a moderate-sized organic molecule on a laptop and gives you reliable bond lengths and angles for most purposes.
Gallery Molecular Shape And Geometry
Molecular Geometry: Definition, Chart, Shapes, and Examples
Mastering Electron Pair Geometry: A Step-by-Step Guide to VSEPR Shapes and Molecular Structure ...
PPT - Molecular Geometry and Bonding Theories PowerPoint Presentation, free download - ID:2953178
Molecular Geometry Structure of Elements Stock Vector - Illustration of infographic, molecule ...
Molecular Geometry - Chemistry