Of Glass Analysis
Most people approaching this subject treat it like a clean textbook exercise. It isn't. The core idea is straightforward enough: you take a piece of glass, figure out what it's made of, and try to make sense of the result. Where things get messy is everything that happens between those two points. I've spent years working with glass samples across archaeology, materials science, and industrial QA, and the short version is that every lab or field setup handles it differently. There is no single universal method. What follows is a practical overview of how people actually do Of Glass Analysis in 2024-2025, what goes wrong, and how to avoid the obvious traps.
Getting Started With Of Glass Analysis
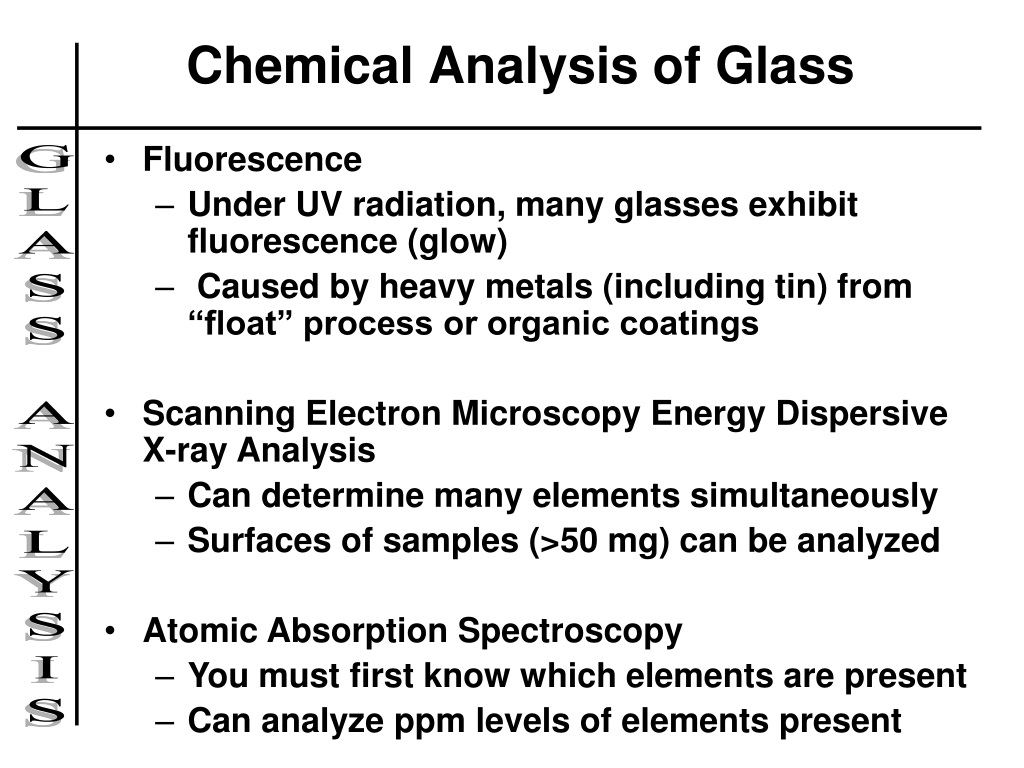
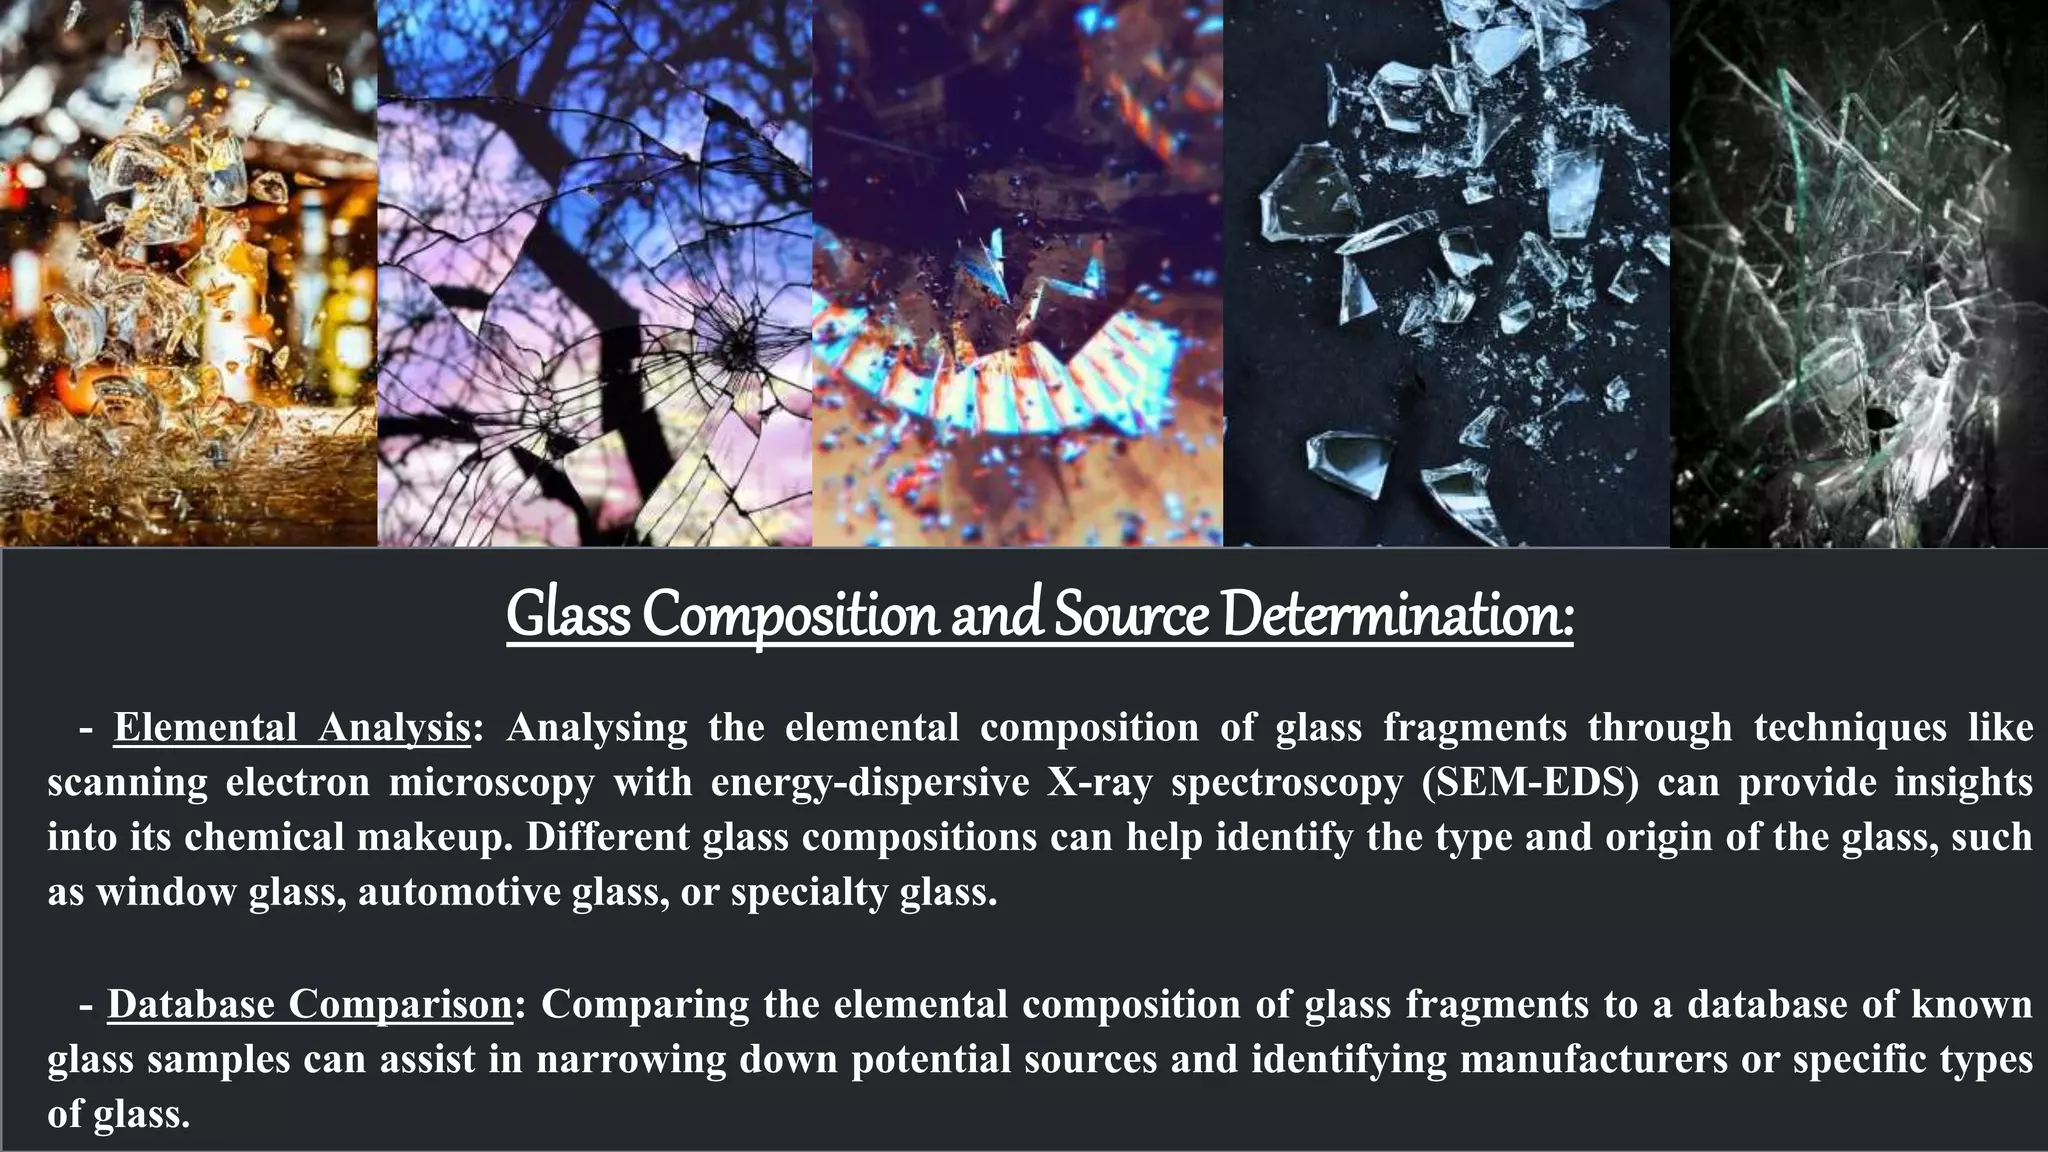
The first decision is analytical technique. In practice, most labs use one or a combination of these: Choose your technique based on what you're actually trying to answer. If you need bulk composition down to the oxide percentage, XRF is usually the workhorse. If you're looking at weathering rinds or surface alteration, SEM-EDS gives you spatial resolution. If you need parts-per-million trace elements for provenance studies, ICP-MS is the right call but the sample prep is significantly more involved. A lot of beginners skip this decision and just send everything to the same instrument. That wastes money and produces data you can't properly interpret later. Take ten minutes to define your question before you touch a sample.
The Practical Side of Of Glass Analysis
Sample preparation is where most projects stall out. Glass is deceptively simple. A clean fragment goes into an XRF and you get numbers. That's the textbook version. The real version involves deciding whether to grind, polish, mount, or analyze in situ, and each choice introduces different kinds of error. For powder XRF, you typically dry the sample, crush it to below 50 microns, and press a pellet with a binder like boric acid or cellulose. The particle size matters because incomplete grinding leaves coarse grains that scatter X-rays differently and skew your results. I've seen entire datasets thrown out because someone skipped the mesh sieve and assumed "fine powder" meant whatever came out of the jaw crusher. For thin sections under SEM-EDS, you mount the fragment in epoxy, grind it flat, and polish it to a 1-micron finish. The polishing step is critical. Scratches and rounding on edges create charging artifacts and false elemental readings, especially on sodium and lighter elements that XRF struggles with anyway.
Get the Full Details
A Problem I Actually Faced
Here's a specific case that cost me about three weeks of work and probably taught me more than any textbook chapter. We were analyzing Roman-period glass fragments with significant surface corrosion. The external rind was enriched in alkalis and depleted in silica compared to the fresh core. Standard XRF on an unpolished surface read the rind, not the glass. Our compositional groupings were all over the place because half the spectra were contaminated by corrosion products. The workaround was to do cross-sectional analysis. We mounted entire fragments in epoxy, ground through from the surface into the fresh core, and took point analyses at regular intervals along the transect. That let us map the corrosion gradient and identify where the unaltered composition actually sat. We then used only those core readings for the compositional grouping. It took longer and required more sample destruction, but the alternative was publishing nonsense. If you're working with ancient or environmentally exposed glass, always check for surface alteration before you assume your bulk composition is accurate. A quick visual inspection under magnification and a few spot checks with EDS can save you from going down the wrong analytical path.
Common Pitfalls and How to Avoid Them
Instrument calibration drifts. Everyone knows this, but the practical implication is that you should run a certified reference material at the start of every session and at least once more halfway through. If your NIST glass standard (like NIST 610 or 612 for silicate glasses) reads outside acceptable tolerance, your entire batch is suspect until you figure out why. Don't assume the machine is fine just because the last calibration was last week. Heterogeneity is another big one. Glass is amorphous, which makes it homogeneous at the atomic scale, but that doesn't mean it's homogeneous at the macro scale. Bubbles, unmelts, crystal inclusions, and phase separation are all common, especially in archaeological and artisanal samples. A single point analysis can be misleading. Take multiple readings across the sample and report the range, not just the average. Data presentation matters more than people admit. A table of oxide percentages means nothing without context. Group your data using statistical methods appropriate to your question—principal components analysis for provenance studies, hierarchical clustering for typological grouping, discriminant function analysis for classification. Raw numbers don't tell the story. The statistics do.
When Of Glass Analysis Fails You
No technique answers every question. XRF has poor sensitivity for light elements like sodium and boron, which are critical components in many glass types. If your research question depends on accurate Na2O or B2O3 measurements, XRF alone won't cut it. You'd need ICP-OES after acid digestion, though that destroys the sample entirely. Portability is another limitation. Handheld XRF units are convenient for field work, but they have thinner detection limits and less accuracy than lab-based instruments. The trade-off is real. If you need publication-quality data, handheld readings are a screening tool, not a final answer. Use them to triage samples, then send the interesting ones to the lab. Statistical overinterpretation is perhaps the most common error in the literature. Small sample sizes, inconsistent methodologies across studies, and poorly documented preparation protocols make cross-study comparisons unreliable. I've seen papers claim strong provenance links based on five samples analyzed with different instruments and different preparation methods. That's not evidence. It's a hypothesis that needs proper testing.

A Note on Tools and Resources
There isn't a single downloadable software package called "Of Glass Analysis" because the term describes a general approach, not a product. What you'll actually need depends on your workflow. For data processing, people commonly use programs like PyMca for XRF analysis, GUPIX for quantification, and R packages like vegan or FactoMineR for multivariate statistics. For instrument control, you're working with whatever the manufacturer provides—Malvern Panalytical, Thermo Fisher, Bruker, and similar vendors all ship their own software suites. If you're looking for reference databases, the Glass Data Bank and the Research Space database from the Getty Conservation Institute are useful starting points. They don't replace good measurement practices, but they help you contextualize your results against known compositions.
Practical Recommendations
Document everything. Sample ID, preparation method, instrument settings, calibration standards, and raw data files should all be recorded in a way that someone else could reproduce your work three years from now. This sounds obvious until you're the one trying to figure out why a dataset from six months ago looks wrong and you have no record of what you actually did. Start small. Run a few known standards before you commit to a full batch. See how your instrument behaves, how your preparation goes, and whether your results make sense. Then scale up. Rushing into a large project without pilot data is the fastest way to waste time and money. Learn the limitations of your technique before you trust the output. XRF is excellent for major and minor elements in silicate glasses but weak on light elements and trace concentrations below a few ppm. ICP-MS covers those gaps but requires acid digestion and introduces contamination risk from reagents and vessels. SEM-EDS gives you spatial context but Quantification is semi-quantitative at best without proper standards. Pick the right tool for the question, not the one that's easiest to use.