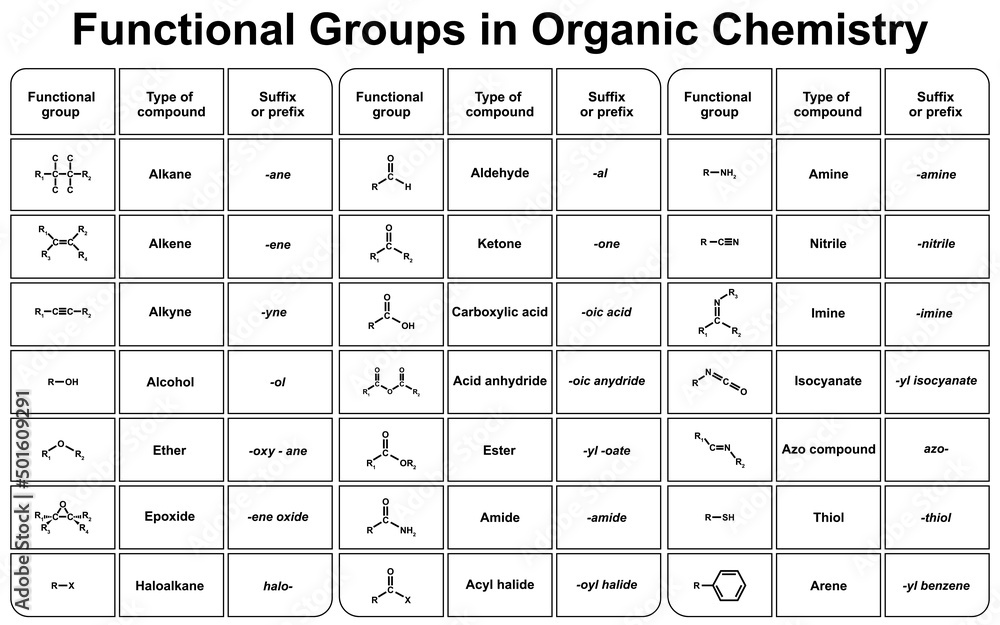
How to Actually Work With Functional Groups Without Losing Your Mind
Functional groups are the reactive parts of molecules. That is about all you need to know at the surface level. The thing nobody tells you when you first open a textbook is that memorizing lists of groups does not prepare you for actually using them in synthesis or analysis. I spent three semesters trying to draw mechanisms by rote before something clicked. Here is what I wish someone had said earlier. The core problem with Organic Chem Functional Groups is that students treat each one as an isolated island. Hydroxyl groups, carbonyls, amines — they get their own chapter with color-coded diagrams and a summary table. But real molecules have multiple groups interacting with each other. A carboxylic acid next to a hydroxyl on the same carbon is a gem-diol and it falls apart in water. An amine near a carbonyl on a five-membered ring forms a stable lactam. These are not exceptions to the rules. They are the rules wearing disguises. The moment you learn to see groups in context instead of in isolation, the subject stops being a memory exercise and starts being predictive.
Learning Organic Chem Functional Groups Through Reactions, Not Definitions
The fastest way to internalize functional groups is to map them onto reactions you already know, not to study them in a vacuum. Take the carbonyl group. It appears in aldehydes, ketones, carboxylic acids, esters, amides, and acid chlorides. The group itself looks the same on paper — a C=O double bond — but its reactivity shifts dramatically depending on what is attached to that carbon. Acid chlorides react with water at room temperature. Amides do not, unless you boil them in strong acid or base for hours. Esters sit somewhere in between. This hierarchy is not arbitrary. It comes down to leaving group ability and resonance stabilization, which are the same concepts repeating across every chapter of organic chemistry. When I was learning this, I built a single reference sheet organized by reaction type instead of by functional group. Nucleophilic acyl substitution, for example, covers acid chlorides, anhydrides, esters, and amides all in one place. Seeing them together made the leaving group trend obvious in a way that four separate textbook sections never did. I spent maybe two hours making that sheet. It saved me dozens more over the rest of the year.
The Spectroscopy Angle Nobody Talks About
In practice, identifying functional groups mostly happens through IR and NMR, not by looking at a structural formula on a page. IR tells you what bonds are present. NMR tells you what environment those atoms sit in. The gap between knowing that a carbonyl absorbs around 1700 cm¹ and actually using that number to distinguish an ester from a ketone from an amide is where most students get stuck. Here is the practical detail that is easy to miss: the exact wavenumber of a C=O stretch depends on conjugation, ring strain, and hydrogen bonding. An unconjugated ketone sits near 1715 cm¹. Put it next to an aromatic ring and it drops to about 1690. Put it in a four-membered ring and it jumps to 1780. An amide C=O shows up lower, around 1650, because nitrogen donates electron density into the carbonyl through resonance. If you memorize one number per group you will be wrong half the time. Memorize the ranges and the reasons instead. For NMR, the proton on a carbonyl carbon — the aldehyde proton — shows up as a distinct triplet or singlet around 9 to 10 ppm. That is almost diagnostic. Carboxylic acid protons are even further downfield, usually 10 to 13 ppm, and they are broad because of hydrogen bonding and exchange. These are the peaks you check first when you are trying to figure out what you actually made in the lab.
Get the Full Details

A Real Problem I Ran Into With Protected Groups
I once ran a Grignard reaction expecting to form a secondary alcohol from an ester, but the product was mostly unreacted starting material. The substrate had a free hydroxyl group elsewhere on the molecule that I had missed during the analysis phase. Grignard reagents are brutally basic. They deprotonate that OH before they ever touch the carbonyl, and the resulting alkoxide kills the reagent through an unwanted side reaction. I spent two days troubleshooting before I realized the hydroxyl was the culprit. The fix was straightforward but it cost me a day I did not have: protect the hydroxyl as a TBS ether using TBSCl and imidazole in DMF, run the Grignard, then remove the protecting group with TBAF. The TBS group is stable to basic conditions and to the Grignard reagent, and it comes off cleanly under mild fluoride conditions. This is exactly the kind of problem that never shows up in the practice problems because the textbook molecules are clean. Real molecules are messy.Pitfalls That Waste More Time Than Anything Else
There are a few recurring mistakes that show up no matter how many times students have seen the material. The first is confusing nucleophilicity with basicity. They correlate but they are not the same thing. A bulky strong base like LDA is a poor nucleophile. Methoxide is both a strong base and a good nucleophile. Picking the right reagent depends on which property matters for your transformation. The second is assuming that a functional group behaves the same way in every solvent. Epoxide opening with a nucleophile proceeds through SN2-like backside attack, so you expect inversion of configuration. That is true in aprotic solvents. In protic solvents, hydrogen bonding can slow the nucleophile enough that the reaction takes a different path or requires heat. Solvent choice is not just about solubility. It changes the mechanism.The third mistake is over-relying on functional group interconversion tables without checking whether the reaction conditions are compatible with other groups on the molecule. Sodium borohydride reduces aldehydes and ketones but leaves esters mostly alone. Lithium aluminum hydride reduces everything. If your molecule has both a ketone and an ester and you only want to reduce the ketone, NaBH in methanol at 0°C is your choice. LAH would wipe out both and you would need a separate reduction strategy to differentiate them later.
What the Standard References Get Wrong
The most commonly used undergraduate textbook, McMurry, handles functional groups accurately but presents them in a way that reinforces the isolation problem. Each chapter treats a single group family as if the rest of the molecule does not exist. Clayden is better on the mechanistic connections between groups, but it assumes a level of mathematical maturity that most students have not yet developed. The answer is to use McMurry for the baseline definitions and Clayden for the why, but you have to actively look for the cross-references. The book will not do it for you.
For spectroscopy, Silverstein is the reference most people point to. It is thorough. It is also dense enough to be intimidating if you are still learning to read an IR spectrum. A more approachable alternative for beginners is Pavia, which walks through interpretation step by step instead of just presenting reference tables. I used Pavia for the first two semesters and switched to Silverstein when I needed to interpret more complex spectra for a research project.
A Practical Workflow for Identifying Functional Groups in Unknown Samples
Here is the sequence I use now, and it cuts down the analysis time significantly compared to how I used to do it. Start with the molecular formula if you have it. Calculate the degree of unsaturation. Two degrees or more usually means either rings plus double bonds or a benzene ring. Then look at the IR. A broad O-H stretch around 3300 cm¹ tells you something is hydroxyl-containing. A sharp C=O around 1700 narrows it down to carbonyl types. Then use the ¹H NMR to distinguish between them. Aldehyde proton at 9 to 10 ppm. Carboxylic acid proton past 10 ppm. Alpha protons next to a carbonyl show up around 2 to 2.5 ppm. The ¹³C NMR confirms the carbonyl carbon — ketones and aldehydes sit around 190 to 210 ppm, esters and acids around 160 to 180 ppm.That sequence usually gets you to a solid identification in under 30 minutes if the spectra are clean. Rough spectra or overlapping signals can push it to an hour or more. The bottleneck is almost always the NMR, not the IR. Modern FTIR instruments give you a decent spectrum in under a minute. ¹H NMR at 400 MHz on a crowded sample can take 15 to 30 minutes for a useable spectrum, and you may need a ²D experiment if the protons are overlapping badly.
Functional Groups in Computational Chemistry
If you are doing any computational work, the way you parameterize functional groups matters more than most people realize. Force fields like GAFF assign partial charges based on atom types, and an oxygen in a hydroxyl group is parameterized differently from an oxygen in an ether, even though both are sp³ hybridized. If you mix up the atom types during prep, your docking scores or MD trajectories will drift. Always verify your ligand parameterization with a known analogue before running production simulations. A five-minute check can save you a week of correcting bad results.When Functional Group Analysis Breaks Down Completely
The standard approaches fail in a few predictable scenarios. Conjugated polyenes do not behave like isolated alkenes. The pi system is delocalized across multiple carbons, and reactivity patterns from simple alkene chemistry do not apply. A Diels-Alder reaction on a conjugated diene follows pericyclic rules, not electrophilic addition rules. Trying to force the latter framework onto the former produces wrong predictions about regiochemistry and stereochemistry. Another failure mode is heteroaromatic systems like pyrrole, furan, and thiophene. These contain heteroatoms with lone pairs that participate in the aromatic pi system. Their reactivity is dominated by aromatic stability, not by the typical behavior of alcohols or ethers. Pyrrole, for instance, undergoes electrophilic substitution at the 2-position, not addition. The oxygen in furan does not behave like a typical ether oxygen because its lone pair is part of the six-electron aromatic system. Students who treat these groups as standalone heteroatoms instead of integral parts of the aromatic framework make consistent mistakes on exam problems and in the lab.