Working With Redox: A Practical Guide
Most people learn oxidation and reduction as two separate definitions that happen to occur simultaneously. In practice, that framing gets you into trouble fast. The moment you stop treating them as independent concepts and start tracking electron flow directly, everything becomes much easier to balance.Oxidation Reduction Reactions Chemistry
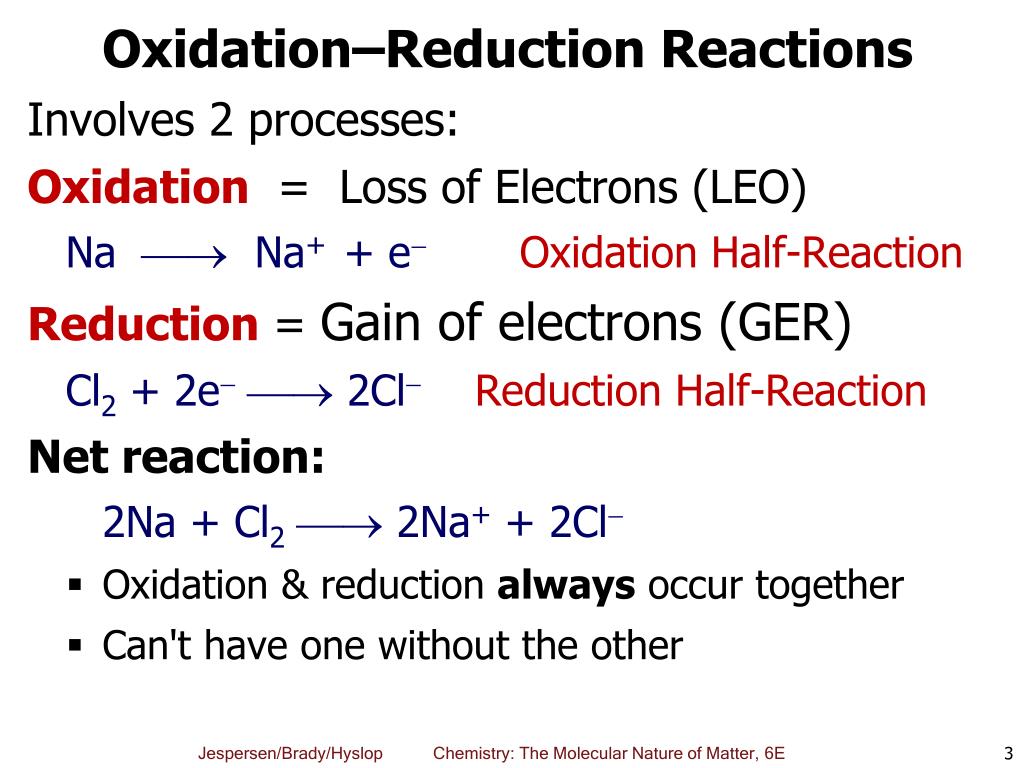
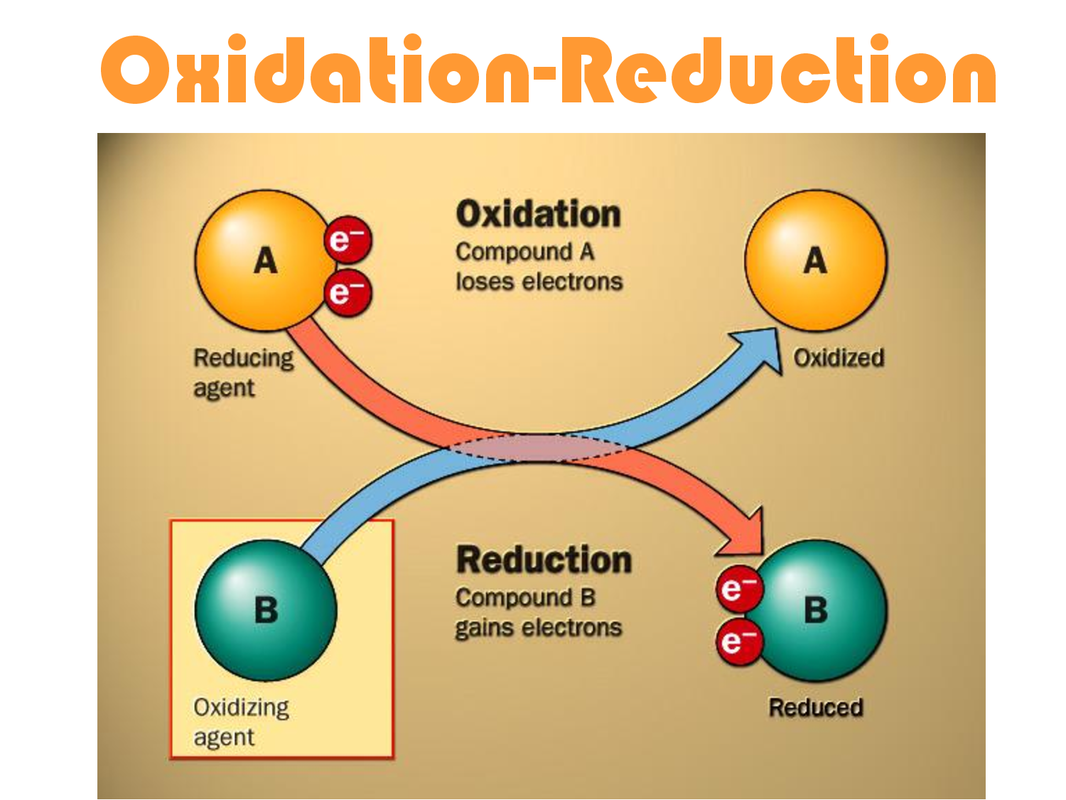
At the core, these reactions involve the transfer of electrons between species. Oxidation is the loss of electrons, reduction is the gain. But here is where students regularly make mistakes: you do not need to memorize the "OIL RIG" acronym to actually work these problems. What matters is learning to assign oxidation states correctly and then following where those numbers change. The standard method for balancing redox reactions in acidic solution goes like this: First, split the overall equation into two half-reactions—one for oxidation, one for reduction. Identify which species is losing electrons and which is gaining. Then balance all atoms except oxygen and hydrogen. After that, add water molecules to balance oxygen atoms. Add H ions to balance hydrogen atoms. Finally, add electrons to each half-reaction so the charge balances on both sides. Multiply each half-reaction by whatever factor is necessary to equalize the electron count, then add them back together and simplify.
I spent three semesters watching students lose points because they skipped the oxidation state verification step and just guessed which side electrons went to. One time, I was grading a particularly stubborn problem involving permanganate reacting with iron(II) in acidic solution. The student had written the correct products but assigned the wrong number of electrons in the half-reaction. It should have been five electrons transferred. They wrote three. The balanced equation came out wrong even though their logic for splitting the reaction was sound. I made them reassign every oxidation state from scratch before they could proceed. That habit alone would have prevented the error entirely. For basic solutions, the process is similar but with an extra step after you finish balancing in acid. Once you have your balanced acidic equation, add OH ions to both sides in the same quantity as the H ions present. Then combine H and OH to form water on whichever side has both. Cancel out any water molecules that appear on both sides. That is it. It is one extra step but it catches people who try to just "switch to base" without adjusting the hydrogen balance.
Common Pitfalls and What Actually Works
The biggest issue I see is people treating redox balancing as a purely algebraic exercise. It is not. You need to understand what is actually happening chemically. Permanganate in acidic solution always reduces to Mn². In basic or neutral solution, it typically reduces to MnO. If you are given a problem with KMnO but no pH information specified, assume acidic unless the problem explicitly says otherwise. That assumption saves you from a whole category of errors. Another counter-intuitive point: the activity series you memorize in high school chemistry only works for single displacement reactions in aqueous solution. It breaks down completely for concentrated acids, for non-aqueous solvents, and for reactions involving oxides or complex ions. When I was running my own lab work with dichromate oxidations in sulfuric acid, I kept trying to predict the direction of electron transfer using the activity series. It failed every time because chromium is not a metal in that context—it exists as an oxoanion. You have to look up standard reduction potentials directly from a table instead. Standard reduction potentials are your actual toolkit. The table lists half-reactions ordered by their E° values. A higher (more positive) E° means that species has a stronger tendency to be reduced. If you pair a reduction half-reaction with an oxidation half-reaction that has a lower E° value, the overall cell potential is positive and the reaction is spontaneous. That is the entire logic behind galvanic cells. You do not need to derive it from first principles during an exam. Just subtract the anode potential from the cathode potential and you get your cell voltage.
Get the Full Details

Here is something most textbooks gloss over: the Nernst equation. Under standard conditions, everything behaves predictably. The moment concentrations deviate from 1 M, or temperature shifts away from 25°C, your standard potentials no longer apply directly. The Nernst equation adjusts the potential based on actual conditions. In my experience, problems that mention non-standard concentrations and then ask for the cell potential are testing whether you know this equation, not whether you can memorize a table. E = E° - (RT/nF) ln Q. At 298 K, that simplifies to E = E° - (0.0592/n) log Q. Use the simplified version on exams. Keep the full form for lab reports where temperature might not be exactly 25°C. I once had a student who could balance every redox equation perfectly but could not explain why a particular reaction was spontaneous. They knew the math but not the chemistry. That is a real problem because when you encounter an unfamiliar reaction on an exam—something not covered in lecture—you need conceptual understanding to fall back on. Memorization gets you through predictable problems. Understanding gets you through the ones designed to trip you up.
What This Approach Does Not Cover Well
The half-reaction method described here works reliably for reactions in aqueous acidic or basic solution. It becomes much less straightforward for reactions in non-aqueous solvents, for solid-state redox processes, and for electrochemical cells with liquid junctions or membrane potentials. In those cases, you need a deeper thermodynamic framework involving Gibbs free energy and chemical potentials. For introductory and intermediate chemistry courses, the half-reaction method is sufficient. But if you encounter problems involving molten salts or industrial electrolysis, the approach has real limitations. You would need to consult specialized references on electrochemistry beyond the standard general chemistry curriculum. Another limitation: the standard reduction potential table assumes all species are at unit activity. In real solutions, especially at higher concentrations, activity coefficients deviate from 1. The potentials shift. For dilute solutions this is negligible. For concentrated electrolytes in industrial settings, it matters significantly. If you are working in a research or engineering context rather than a classroom, you will need to account for this.
A Quick Reference for Common Half-Reactions
MnO + 8H + 5e Mn² + 4HO (E° = +1.51 V) CrO² + 14H + 6e 2Cr³ + 7HO (E° = +1.33 V) Fe³ + e Fe² (E° = +0.77 V)

Zn² + 2e Zn (E° = -0.76 V) Cu² + 2e Cu (E° = +0.34 V) Memorizing these exact equations is unnecessary if you can balance them from first principles. But knowing the standard potentials lets you quickly predict whether a given combination of reactants will actually react. That skill separates people who can balance equations from people who understand what those equations mean.
When I grade papers, the ones that earn full credit are the ones where the student shows every step clearly and checks their work by confirming that both mass and charge balance in the final equation. Skipping steps might get you the right answer sometimes, but it also means you have no way to verify you got there honestly. Take the time to write it out. It pays off consistently.