Understanding How We Represent 3D Molecules on Flat Paper
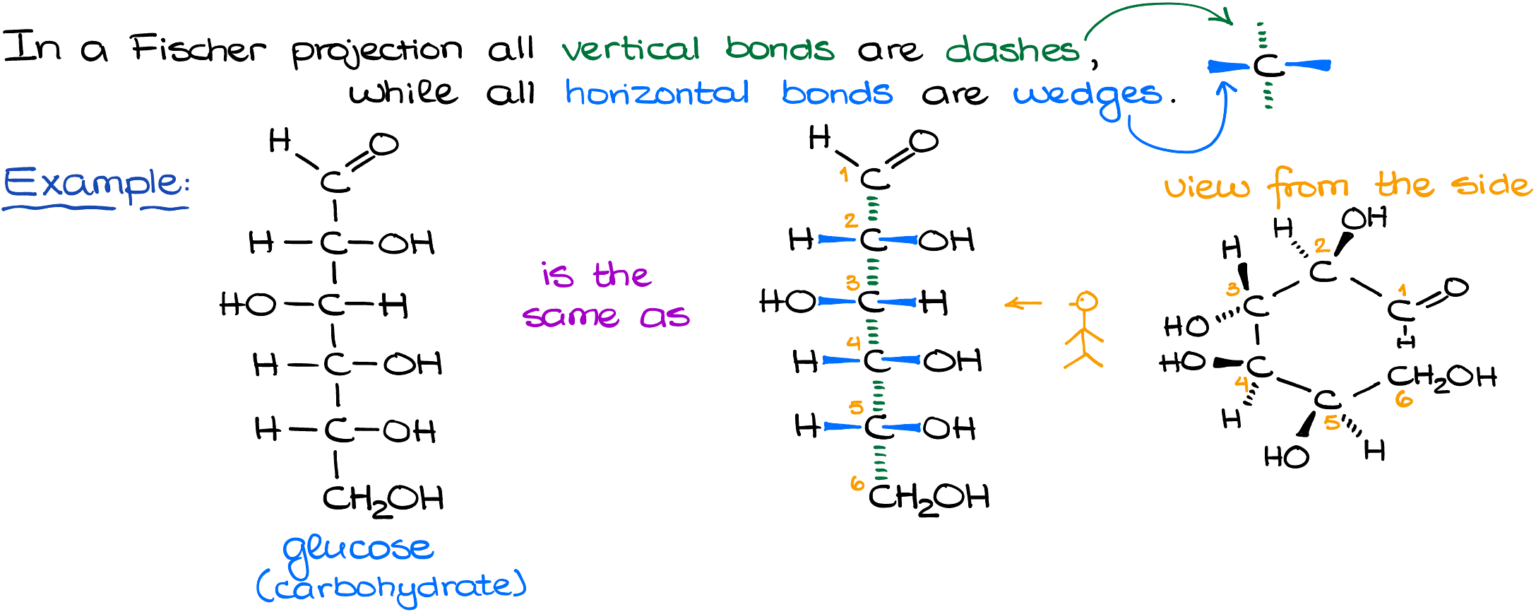
You will see projections of organic molecules in literally every upper-level chemistry course and many industrial publications. They are not decorative. They are a translation problem — taking a three-dimensional structure that exists in space and rendering it on a two-dimensional surface so another chemist can understand stereochemistry, conformation, and reactivity without having to build a model. The main types you need to know are Fischer projections, Newman projections, Haworth projections, sawhorse projections, and wedge-dash notation, which is technically a projection even though it does not always get called that. Each one solves a different problem. Fischer is for sugars and amino acids drawn vertically with the carbon chain running top to bottom. Newman looks down a bond axis to show staggered and eclipsed conformations. Haworth flattens cyclic sugars into rings. Sawhorse shows bonds at an angle rather than directly end-on like Newman.
Projections Organic Chemistry: Getting the Basics Right
The first thing people mess up is confusing representation with reality. A Fischer projection is not a molecule standing upright. It is a convention where horizontal bonds come toward you and vertical bonds go away. If you rotate a Fischer projection by 90 degrees, you have inverted every stereocenter. Rotate it by 180 degrees in the plane of the paper and you get the same molecule. This distinction matters because exam questions and peer reviewers both catch people who treat Fischer as a flexible drawing tool rather than a strict convention. Newman projections require you to pick the correct bond to look down. Beginners will often choose the wrong bond and produce a conformation that is technically valid but irrelevant to the question being asked. If the problem involves rotation around the C2-C3 bond in butane, you look down C2-C3. Not C1-C2. Not C3-C4. Just the bond named in the question. I lost points on this exact mistake during my second semester and stopped doing it after the third time. Wedge-dash is the most commonly used projection in modern literature because it is the fastest to draw and read. Solid wedge means coming out of the plane. Dashed wedge means going into the plane. Plain lines stay in the plane. The convention is standard but you will still see people draw wedges and dashes inconsistently across a single diagram, which makes the stereochemistry impossible to parse.
Working Through a Conformational Problem Step by Step
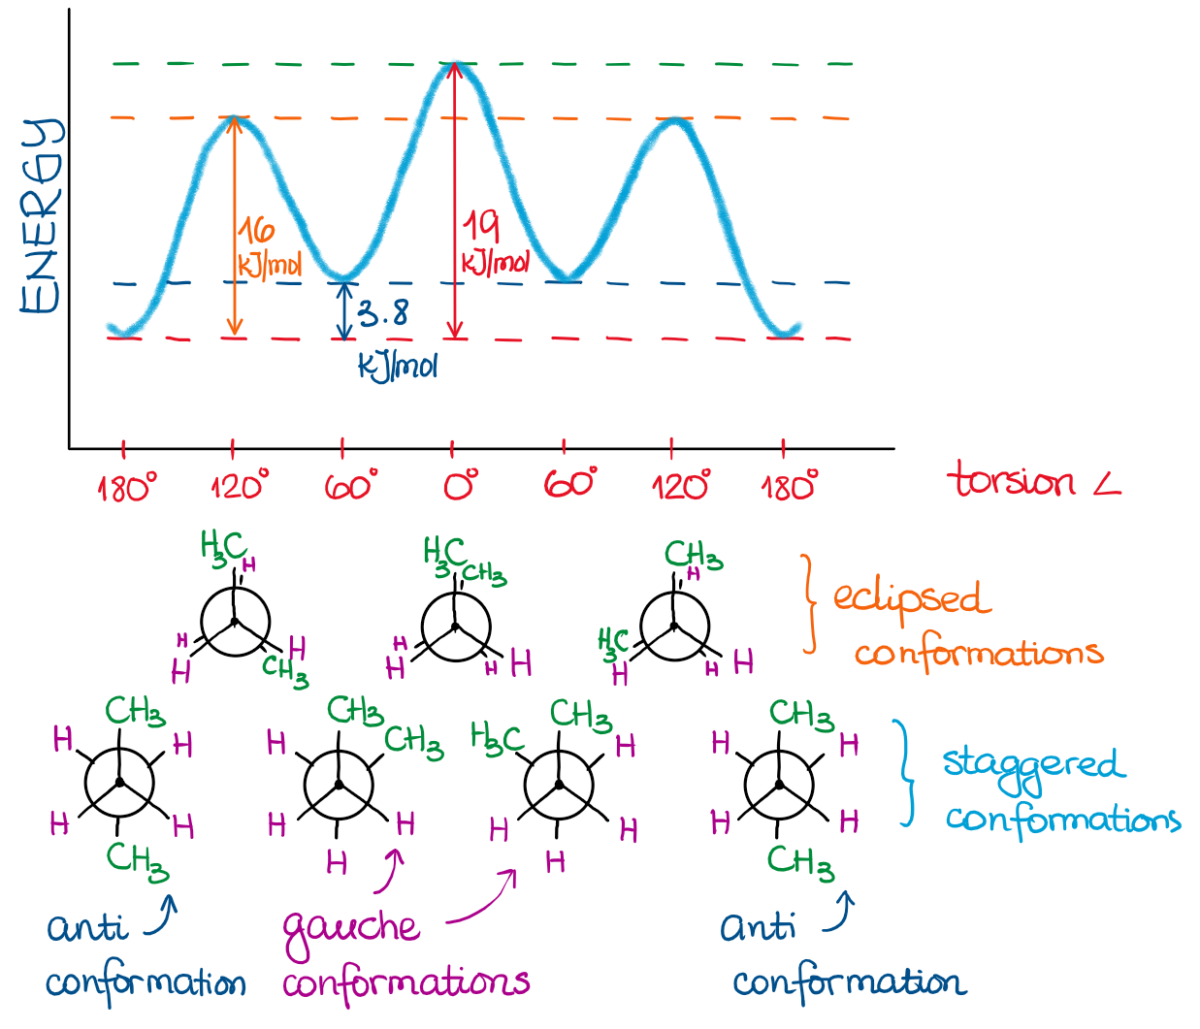
Here is how I actually approach these problems when I need to determine the lowest energy conformer of a substituted cyclohexane or figure out whether a reaction goes through an axial or equatorial pathway. Step one is drawing the base structure with clear stereochemistry. Get the wedges and dashes right before you do anything else. Step two is converting that into a chair conformation if you are dealing with a six-membered ring. I draw the chair first, label each carbon as axial or equatorial, then place substituents one at a time. Step three is evaluating 1,3-diaxial interactions and A-values. A methyl group costs about 1.7 kilocalories per mole in the axial position. A tert-butyl group is so large it essentially locks the ring into the conformation where it is equatorial. That is not a suggestion. That is a near-absolute rule. For acyclic systems, I convert the structure into a Newman projection looking down the bond of interest. I draw the front carbon as a point and the back carbon as a circle. I place the highest priority groups anti to each other in the staggered conformation because that is the lowest energy arrangement. Then I rotate in 60 degree increments to map out all three staggered and three eclipsed conformers. The energy differences between them tell you what the population distribution looks like at room temperature.
Get the Full Details

A Real Problem I Ran Into and How I Fixed It
I was working with a disubstituted cyclohexane where one substituent was a methoxy group and the other was a bulky silyl protecting group on a neighboring carbon. The problem was that the standard chair flip gave two conformers that looked almost equal in energy on paper, but the reaction clearly favored one pathway exclusively. I spent about two hours trying to figure out what I had drawn wrong before I realized I had placed the methoxy group axial in both chairs instead of flipping it properly. The A-value of an OMe group is roughly 0.6 kcal/mol, which is small enough that both conformers exist in significant amounts, but not small enough to ignore. Once I redrew the chair correctly with the methoxy equatorial in the major conformer, everything matched the experimental outcome. The workaround was to build a physical model kit and actually manipulate the ring. Paper drawings had been lying to me because I was not tracking the bond rotations carefully enough during the flip. R.S. configuration assignment is where most people lose track. When you are looking at a Fischer projection and need to assign R or S, remember that the horizontal bonds are coming toward you. That changes how you apply the Cahn-Ingold-Prelog rules compared to a standard wedge-dash drawing. I have seen people assign the wrong configuration simply because they applied the rules as if all bonds were in the plane. Another issue is over-relying on Newman projections for cyclic systems. You can draw a Newman projection for a cyclohexane ring, but it is usually not helpful. The ring constraints make the visualization messy and error-prone. Use a chair conformation instead. It tells you everything you need to know about axial and equatorial positioning in a single drawing.
Haworth projections are straightforward but they hide important conformational detail. A pyranose ring drawn as a flat hexagon with substituents pointing up or down gives you stereochemistry at each carbon, but it does not show you that the actual ring is in a chair conformation with its own axial and equatorial positions. If you need to predict reactivity or steric effects, go from Haworth to chair. It takes about thirty seconds and prevents a lot of wrong predictions.
When Projections Fail You
There are cases where 2D projections simply cannot convey the relevant information. Locked ring systems with multiple fused rings, atropisomers where rotation is restricted but not absent, and molecules with conformational flexibility that interconverts rapidly on the NMR timescale all resist clean projection. In those situations, computational modeling software or crystallographic data from X-ray diffraction is the only reliable way to determine the actual three-dimensional structure. I have used both in my work and both are worth learning. The computational route takes longer to set up but gives you energy profiles across multiple conformations in one run. If you are just starting out, focus on mastering Fischer, Newman, and wedge-dash. Chair conformations come next. Haworth and sawhorse are useful but less frequently needed in daily practice. The skill that makes the biggest difference is not memorizing every projection type but developing the mental rotation ability to move between them quickly. That comes from doing problems, not from reading explanations.