Getting Pure Crystals Without the Headache
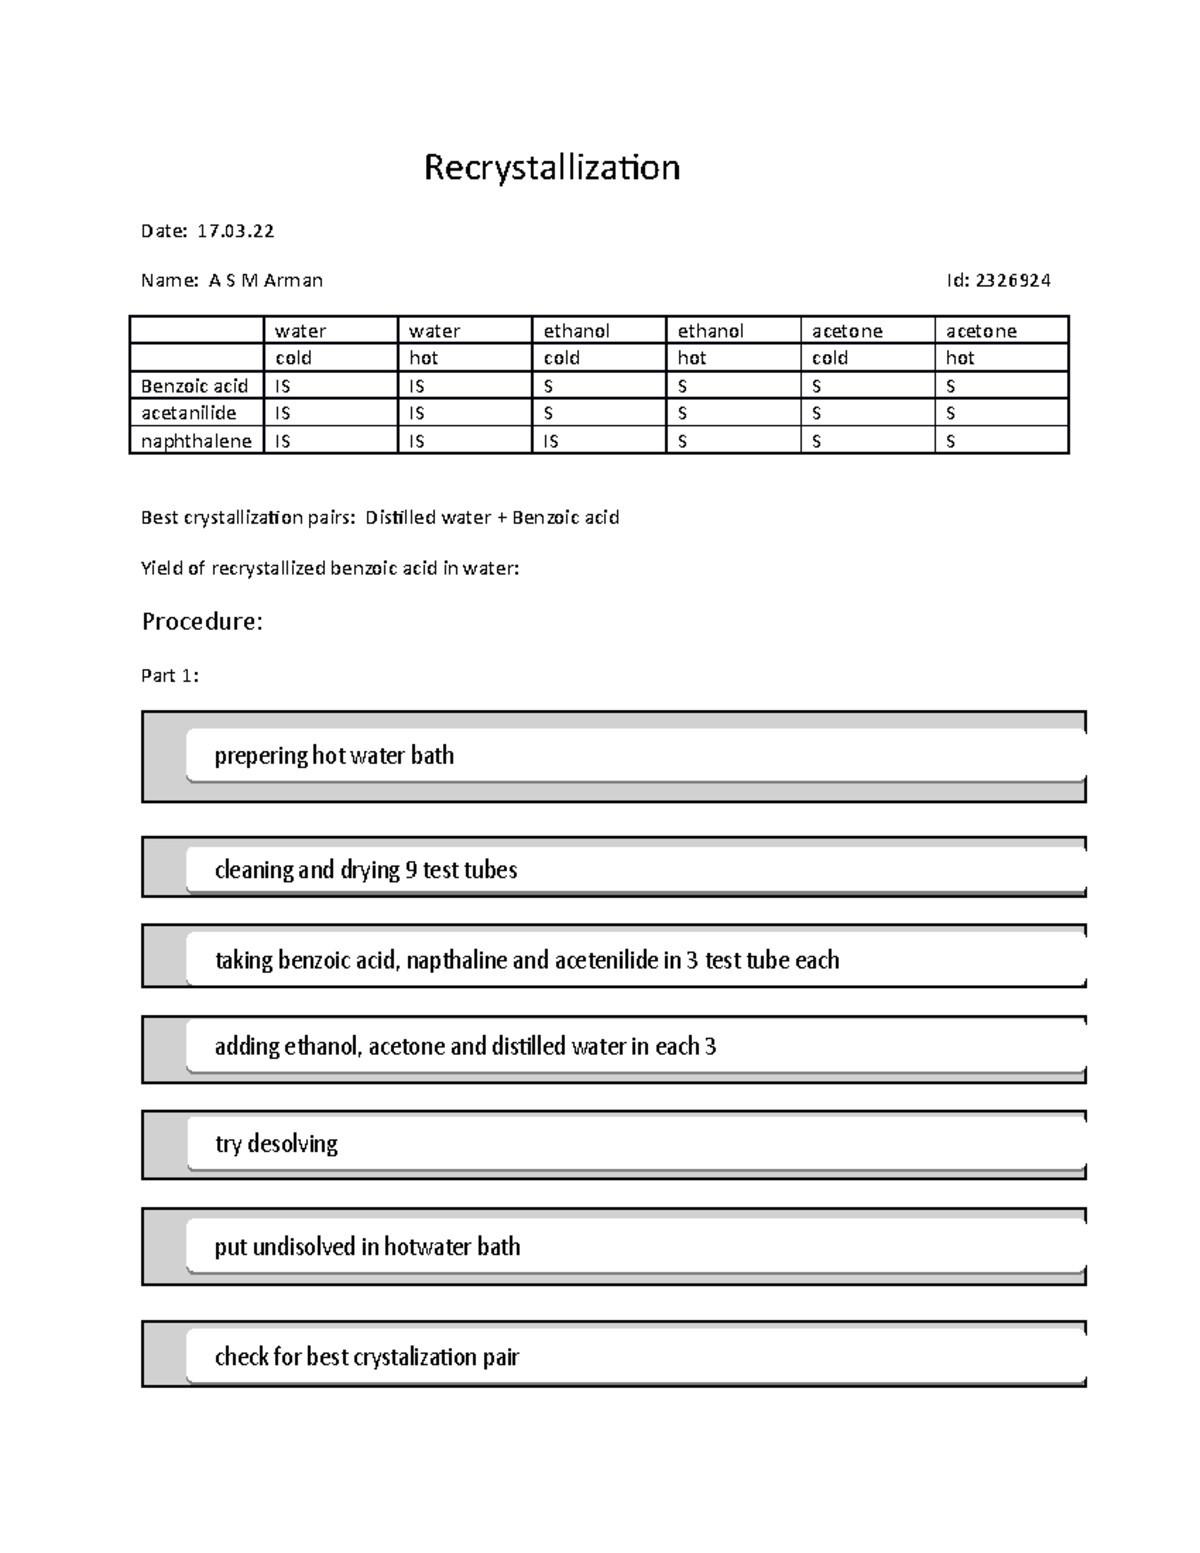
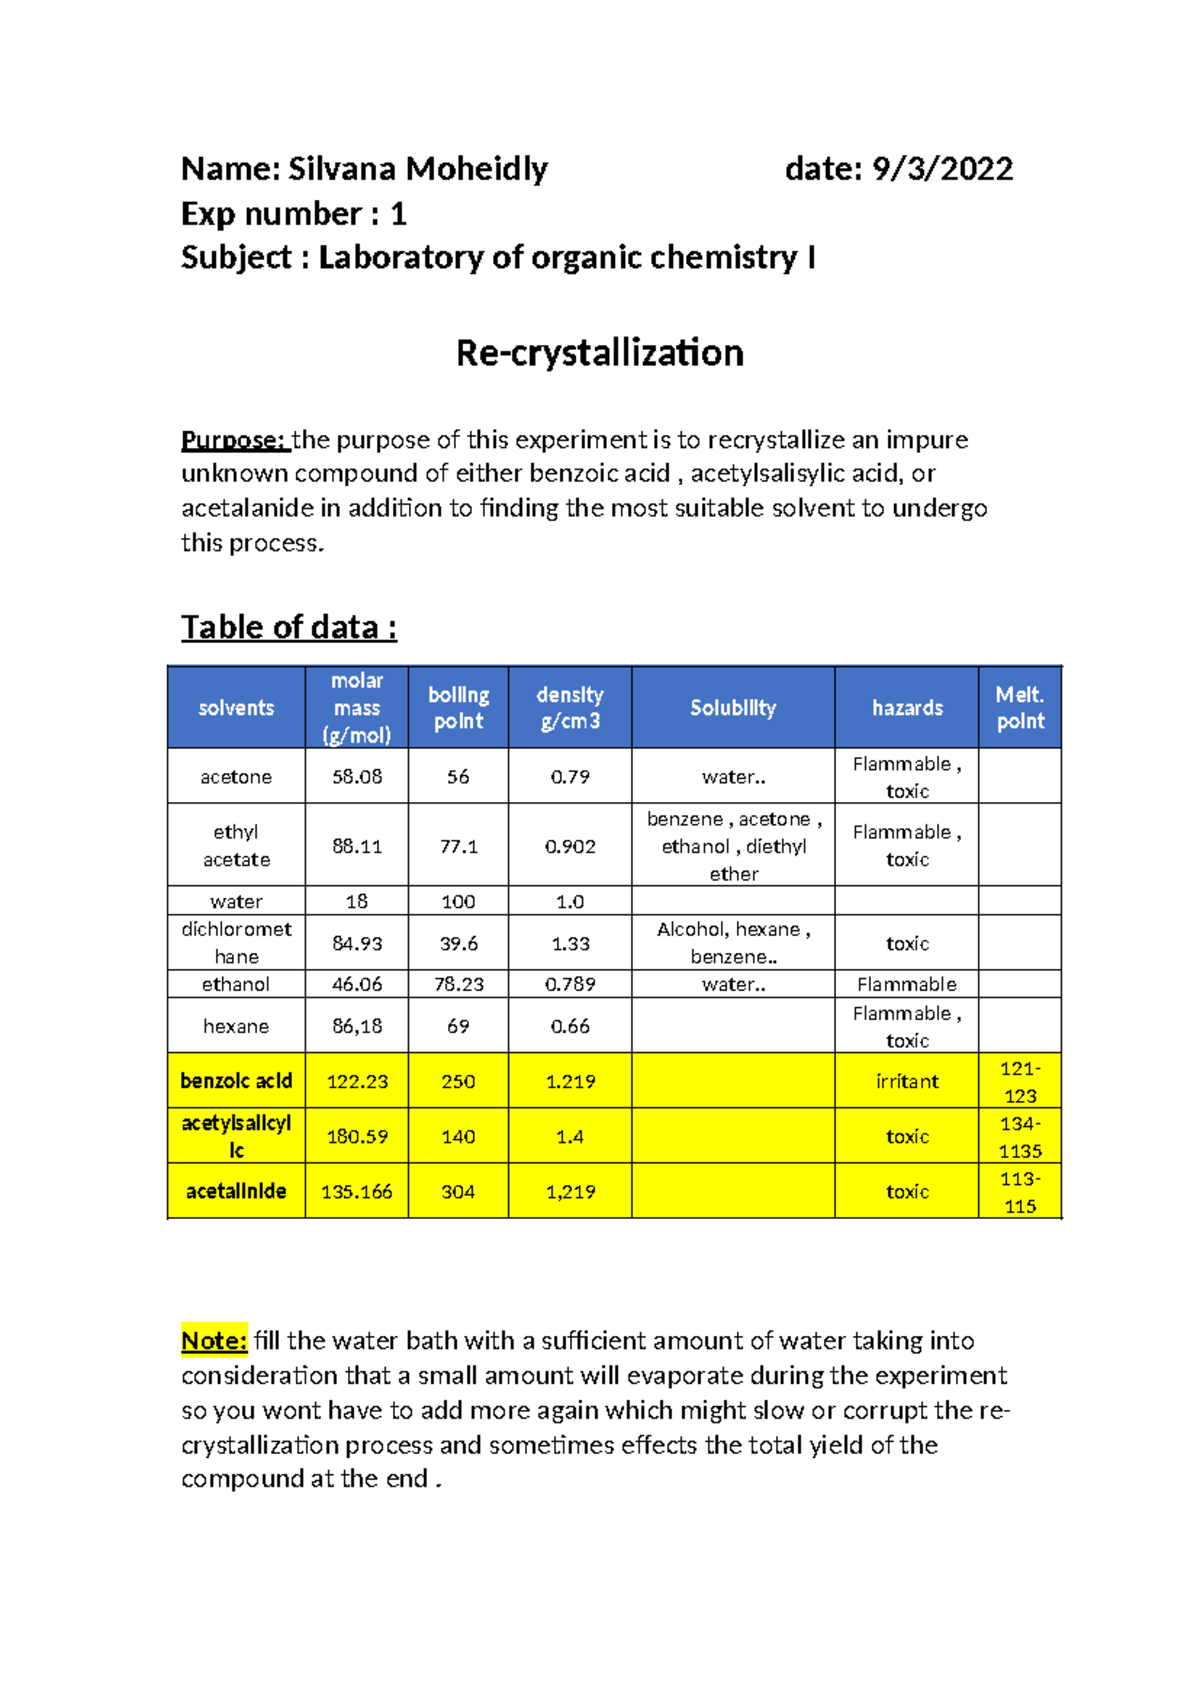
Recrystallization is one of those techniques that looks simple on paper but tends to ruin your day if you get it wrong. I have spent more mornings watching undergraduates lose half their product because they cooled the flask too fast or picked a solvent that formed an oil instead of crystals. The principle is straightforward enough, but the execution has more pitfalls than most students realize before they stand over a hot plate with a beaker of murky solution. The core issue most people face is solvent selection. You need a solvent where your compound is sparingly soluble at room temperature but dissolves readily when hot. Common mistake is picking ethanol or water for everything because those are safe solvents in the stockroom. Sometimes that works fine, but more often you end up with either no crystals forming or a gum that never solidifies. I once had a student trying to purify an unknown solid for three hours using methanol, only to realize the compound was essentially insoluble even at the boiling point. We switched to a hot hexane-ethyl acetate mixture and got clean needles in twenty minutes. The ideal solvent should give you about one gram of solubility per ten milliliters at room temperature and ten grams or more when hot. You can test this by adding a small amount of solvent to a pinch of your solid in a test tube, heating it, and seeing if it clears up. If the solid dissolves at room temperature, the solvent is too good and you will lose product during cooling. If it never dissolves even when boiling, the solvent is too weak.
The Actual Procedure
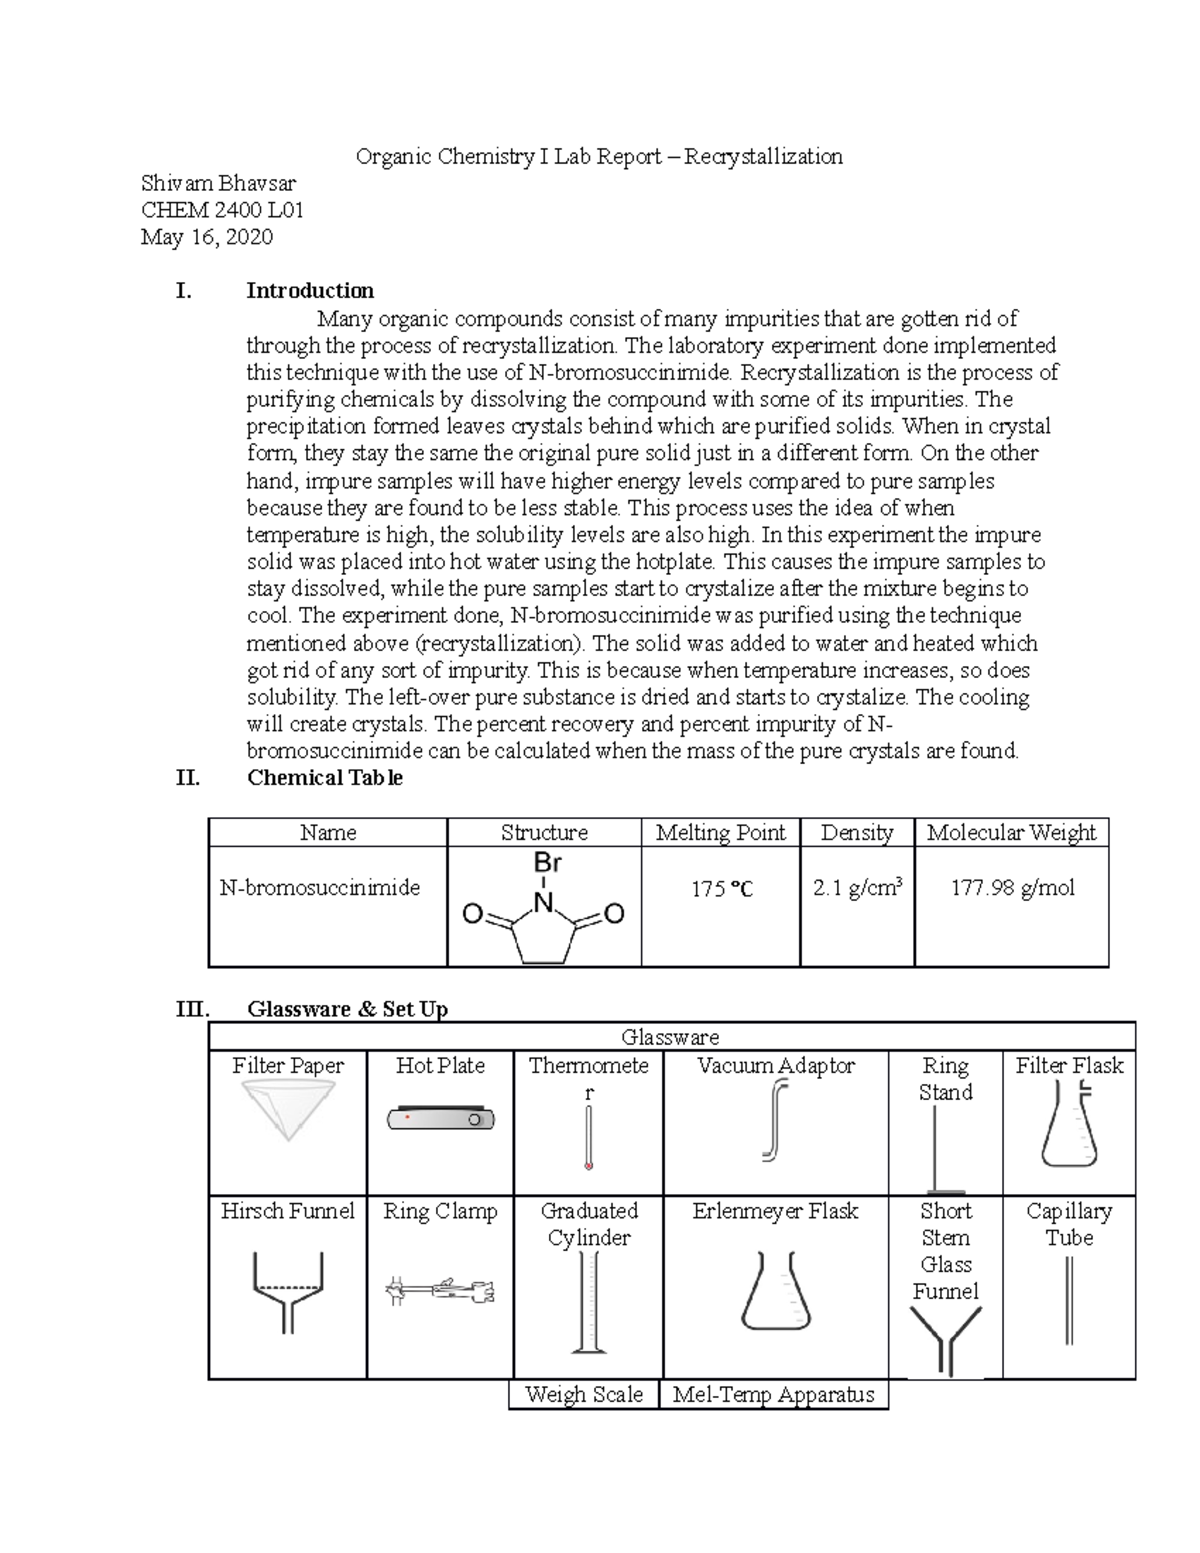
Start by finding the minimum amount of hot solvent needed to dissolve your crude solid. This usually means heating the solvent separately in a flask while you weigh out your compound. Adding solvent to the solid in a large flask and then trying to heat everything together tends to cause premature crystallization on the walls, which means you have to start over. I prefer using an Erlenmeyer flask with a loose stopper or watch glass to prevent excessive evaporation while allowing some airflow. Once the solid dissolves, remove the heat and let it cool slowly to room temperature. Do not put it in an ice bath immediately unless your procedure specifically calls for it. Rapid cooling produces small crystals that trap impurities inside their lattice. Slow cooling allows the molecules to arrange themselves properly and exclude foreign compounds. I usually set the flask on the bench and walk away for twenty minutes. When it reaches room temperature, I check if crystals have formed. If not, I seed the solution with a tiny crystal or scratch the flask interior with a stir bar to provide nucleation sites. After crystals form, cool the mixture in an ice bath for another ten minutes to maximize recovery. Then collect the solid by vacuum filtration using a Büchner funnel. Wash the crystals with a small amount of ice-cold solvent to remove surface impurities. Each wash should be no more than one or two milliliters per gram of solid. Excess washing solvent dissolves your product and lowers your yield significantly.
What Your Lab Report Should Actually Show
Many students write reports that just repeat the procedure without analyzing what went right or wrong. A good report needs the melting point range of both the crude and purified product. The purified sample should melt within one or two degrees of the literature value and over a narrower range. If your melting point is depressed or broad, either the compound is still wet with solvent or impurities remain trapped in the crystal lattice. Yield calculations matter, but so does explaining losses. If you recovered only sixty percent of your starting material, state whether you think product was lost during transfer, left in the mother liquor, or dissolved in the washing solvent. A yield over eighty percent is suspicious unless you started with very pure material. It usually indicates incomplete drying or inclusion of solvent in the crystal structure. I see too many reports that claim perfect results without any discussion of problems encountered. Mention if your crystals formed as an oil initially. Describe how you fixed it, whether by adding more solvent, seeding, or changing the cooling rate. This shows you understand the technique rather than just following instructions mechanically.
Get the Full Details
When Recrystallization Fails Completely
Sometimes no amount of solvent switching will give you clean crystals. This happens most often with compounds that have complex molecular structures or multiple functional groups with different solubilities. If your solid consistently forms oils instead of crystals, try a different approach. You might need to convert the compound to a derivative with better crystallization properties, use column chromatography instead, or attempt zone refining for high-purity requirements. Drawing oil instead of crystals usually means the compound is decomposing or has a eutectic mixture with its impurities. I once worked with a student who kept getting oily residues from a reduction reaction. We discovered the product was forming a stable hydrate that melted below its decomposition point. Switching to a dry solvent system and using anhydrous conditions solved the problem entirely. This took about an hour of troubleshooting but saved weeks of wasted attempts. If recrystallization fails after three or four solvent trials, stop and consider alternative purification methods. Each attempt costs time, solvent, and usually some product. Thin-layer chromatography can help determine whether you actually have a single compound or a mixture that needs separation by other means.
Common Mistakes That Tank Your Grade
Not reporting the volume of solvent used makes your procedure impossible to reproduce. State exactly how many milliliters you added and how many times you had to repeat dissolution. Missing the melting point range of the purified product is an automatic point deduction in most courses. Recording only the initial melting temperature instead of the full range shows you did not observe carefully. Claiming one hundred percent recovery is a red flag that your data is fabricated. Even with optimal technique, you typically recover seventy to eighty-five percent unless you started with nearly pure material. Writing a conclusion that says everything went perfectly without mentioning any problems suggests you did not actually perform the experiment yourself. Forgetting to dry your crystals completely before weighing introduces significant error. Solvent trapped in the crystal lattice or adsorbed on the surface adds mass that is not your product. I usually leave samples in a desiccator overnight or use a warm oven at fifty degrees Celsius for two hours. Weighing wet crystals can inflate your yield by five to ten percent, which professors notice immediately.
Advanced Techniques for Difficult Compounds
Some compounds benefit from controlled cooling rates rather than natural cooling. Using a heating mantle set to forty degrees and programming it to decrease by one degree per minute gives more consistent crystal growth than ambient cooling. This method takes longer, about two hours instead of twenty minutes, but produces larger, purer crystals suitable for X-ray diffraction or precise melting point determination. Sequential recrystallization from different solvents can improve purity when a single solvent gives mediocre results. Crystallize from solvent A, collect and dry the solid, then recrystallize from solvent B. Each step removes different impurities based on solubility differences. This approach usually increases purity by five to ten percent per cycle but reduces overall yield by fifteen to twenty percent. Temperature-controlled nucleation using a syringe pump to add antisolvent slowly gives better results than rapid precipitation for sensitive compounds. Adding acetone to a THF solution dropwise over thirty minutes while stirring produces uniform crystal size distribution. This technique requires more equipment but is standard in process chemistry laboratories for producing consistent particle sizes.