What Actually Happens When a Reducing Agent Gets Used Up
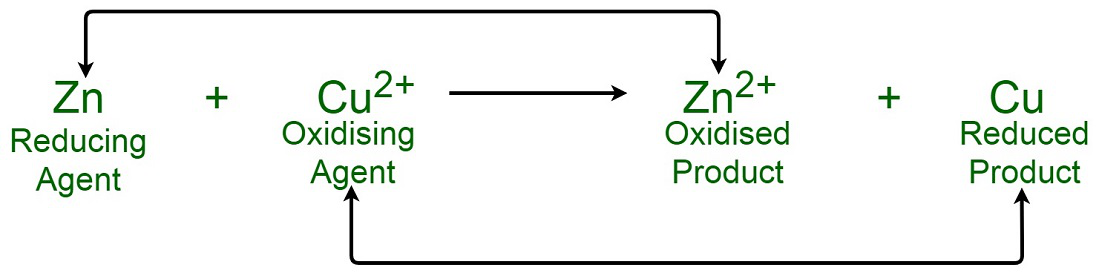
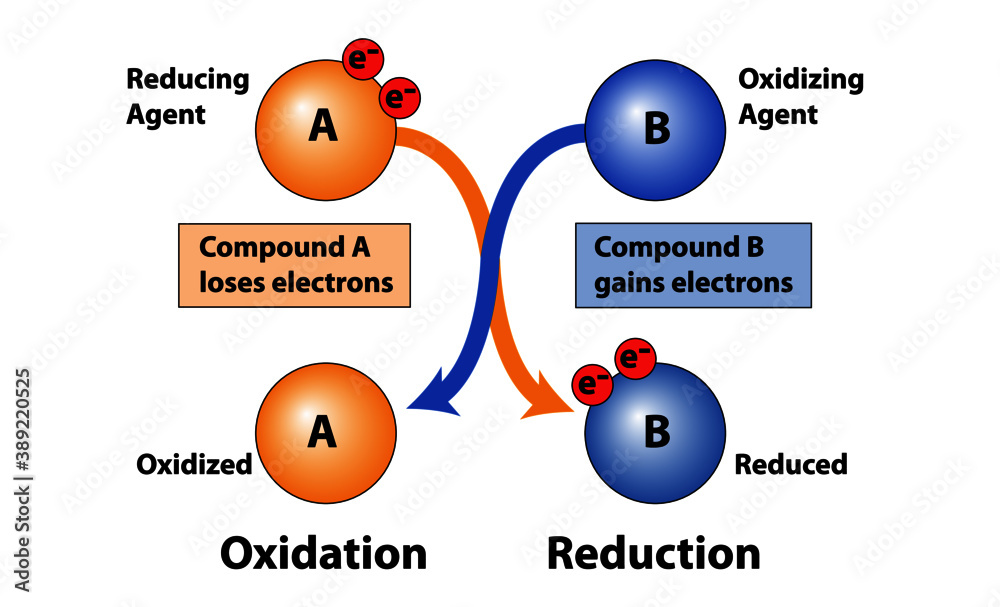
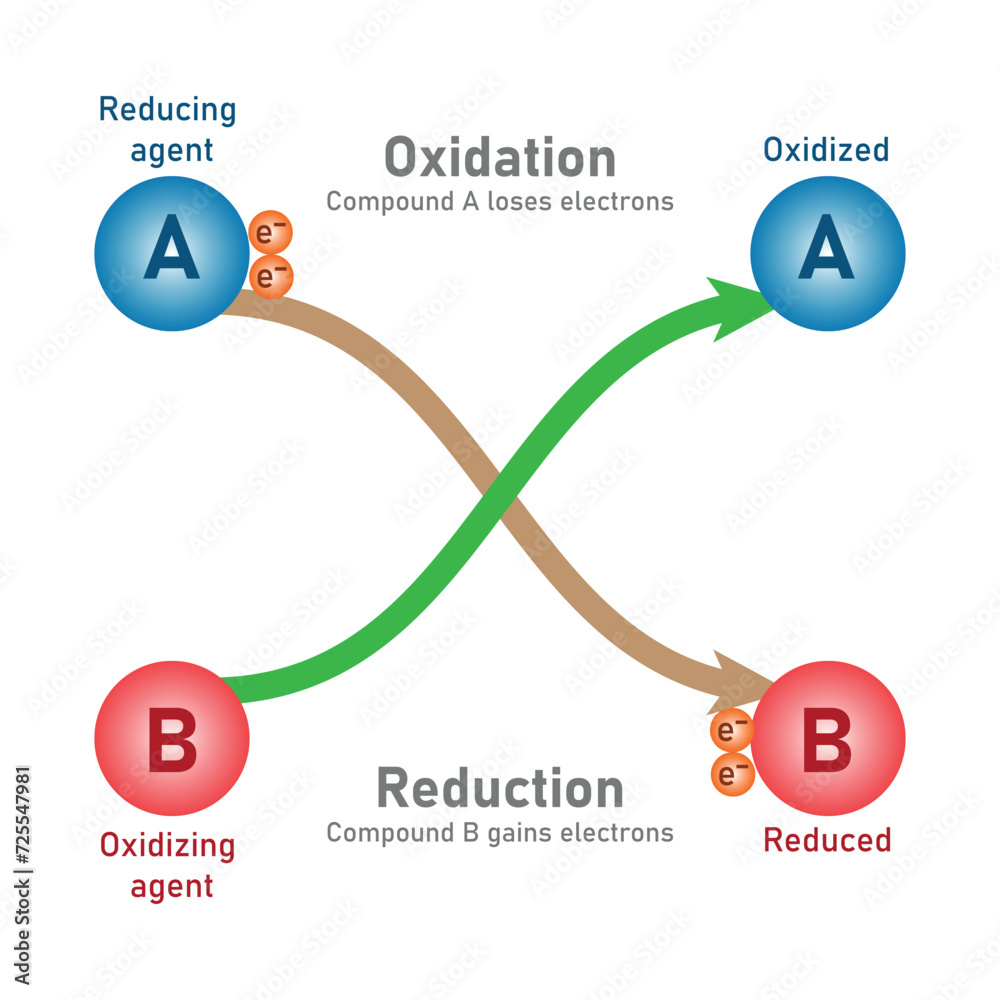
I spent three semesters grading introductory chemistry labs and the same misunderstanding showed up in basically every single report. Students could balance a redox equation on paper but when you asked them what was happening to the electrons in their hands-on experiment, they froze. The gap between written theory and physical observation is where most people stumble, so I am going to walk through this the way I wish someone had explained it to me before I started working with these reactions daily. A reducing agent donates electrons to another species and in doing so becomes oxidized itself. That second part always catches people off guard. The substance that drives the reduction process is the one getting destroyed in the bargain, not preserved. Think of it like paying for something with cash. You hand over the money, the transaction completes, and your wallet is lighter. The reducing agent pays the electron debt and takes the hit on its own oxidation state. In practice, the most common reducing agents you will encounter are things like zinc metal, sodium borohydride, lithium aluminum hydride, and molecular hydrogen over a palladium catalyst. Each one has a different electron-donating strength and a different tolerance for water, air, and temperature. Pick the wrong one for your conditions and you waste reagents, create side products, or sometimes just sit there watching nothing happen for forty-five minutes while your procedure says it should have finished in ten.
I once ran a reduction on a substrate that looked straightforward on the reaction scheme. The literature called for sodium borohydride in methanol at room temperature, which should have been fine. My compound had a chloro group sitting right next to the ketone I wanted to reduce. The borohydride attacked the carbonyl just as intended, but it also started nibbling at the carbon-chlorine bond as a secondary pathway. I ended up with a mixture of the desired alcohol and a dechlorinated byproduct in roughly equal amounts. The yield on paper looked great until I ran the NMR and saw the extra peaks. The workaround was switching to a milder system. I used L-Selectride instead, which is bulkier and more selective for the carbonyl over the alkyl halide. The reaction took longer, about twenty minutes instead of five, but the crude product came out nearly clean. Running the silica column after that took maybe eight minutes instead of the hour I would have spent purifying the messy borohydride result. You trade a bit of time upfront to save a lot downstream. Here is something people rarely emphasize in textbooks. The strength of a reducing agent is not an absolute number you can look up once and forget. It shifts with solvent, temperature, concentration, and what else is floating around in the pot. Standard reduction potentials from a table assume aqueous conditions at one molar and twenty-five degrees Celsius. Your reaction is probably none of those things. A reagent that looks borderline in the tables might work brilliantly in your organic solvent, or it might sit there inert because the kinetics are sluggish even though the thermodynamics look favorable.
When I teach students to approach this, I tell them to think in terms of two separate questions. First, is the reaction thermodynamically possible? Check the potentials or the general reactivity pattern. Second, is it kinetically accessible under your actual conditions? That second question is the one that separates a successful lab day from a frustrating one, and it is also the one nobody tests for until they are already in the middle of it. The mechanism matters more than you might expect. Some reducing agents transfer hydride directly. Others operate through single-electron transfer, especially when you are working with metals like zinc or iron in acidic media. The pathway determines the stereochemical outcome, the regioselectivity, and the kinds of side reactions you should watch for. Hydride transfer tends to give predictable stereochemistry based on sterics and chelation control. Single-electron transfer can generate radical intermediates that do things like cyclize, fragment, or abstract hydrogen from unexpected places. I keep a small notebook of conditions that worked for specific substrates, not because I have a good memory, but because the same functional group arrangement can behave completely differently depending on the order of addition, the solvent grade, or how quickly you cool the reaction after the reducing agent is introduced. A protocol that calls for adding the borohydride in one portion at zero degrees might need to be dosed over fifteen minutes if your substrate is particularly reactive or if you are scaling up beyond a few milligrams.
Get the Full Details
Quantification is another area where things get messy. You do not always get a clean stoichiometric relationship between the reducing agent and the product. Side reactions consume some of the reagent. Incomplete conversion leaves starting material behind. Over-reduction can push the product further than intended, especially with strong agents like lithium aluminum hydride, which will reduce esters, amides, and nitriles almost indiscriminately if you let it warm up too much. I usually run a quick TLC or an in situ NMR sample before quenching anything to check conversion. It saves a lot of wasted time compared to assuming the reaction is done because the colored indicator disappeared or because the procedure said thirty minutes. Thirty minutes is a suggestion, not a law. Some reactions finish in ten. Others need an hour or more, especially with steric hindrance around the reactive site. Workup is where I see the most avoidable mistakes. Quenching a lithium aluminum hydride reaction requires care because the excess reagent reacts violently with water and alcohols, releasing hydrogen gas. I have seen people add water directly to the reaction flask without first letting the hydride consume itself or using a proper sequential quench with ethyl acetate and then dilute acid. That is how you lose product to aerosolization and sometimes damage the glassware if the exotherm is severe enough.
The safer approach is to let the initial reaction finish, cool it down, and then add the quenching reagent slowly, preferably behind a shield. For smaller scale work in teaching labs, I have students use a saturated aqueous Rochelle salt solution to decompose the aluminum complexes cleanly, which gives a much clearer organic layer to separate. It is a small detail that makes the purification step noticeably easier. There are cases where a reducing agent simply does not work no matter how long you wait or how much you increase the temperature. Bulky ketones near a quaternary center resist hydride attack. Conjugated systems sometimes prefer 1,4 reduction over 1,2 depending on the reagent and the solvent. If you push hard enough with a very strong reducing agent, you might force the reaction, but you also increase the chance of reducing other sensitive groups in the molecule. That is a trade-off you have to evaluate based on what you actually need. I recommend keeping a mental checklist before you start any redox procedure. Identify the functional group you want to reduce. Identify the ones you want to leave alone. Choose a reagent that differentiates between them. Plan your quench. Have a way to monitor progress that does not rely solely on time. And be ready to adjust if the reaction behaves differently than the textbook says it should, because it often does.
The chemistry is not harder than it needs to be. The difficulty comes from treating every reaction as if it were the same generic process instead of a specific set of conditions applied to a specific molecule. Once you start thinking in those terms, the whole topic becomes much more manageable and your lab results reflect that shift almost immediately.