
How Retrosynthesis Actually Works When You're Staring at a Target Molecule at 2am
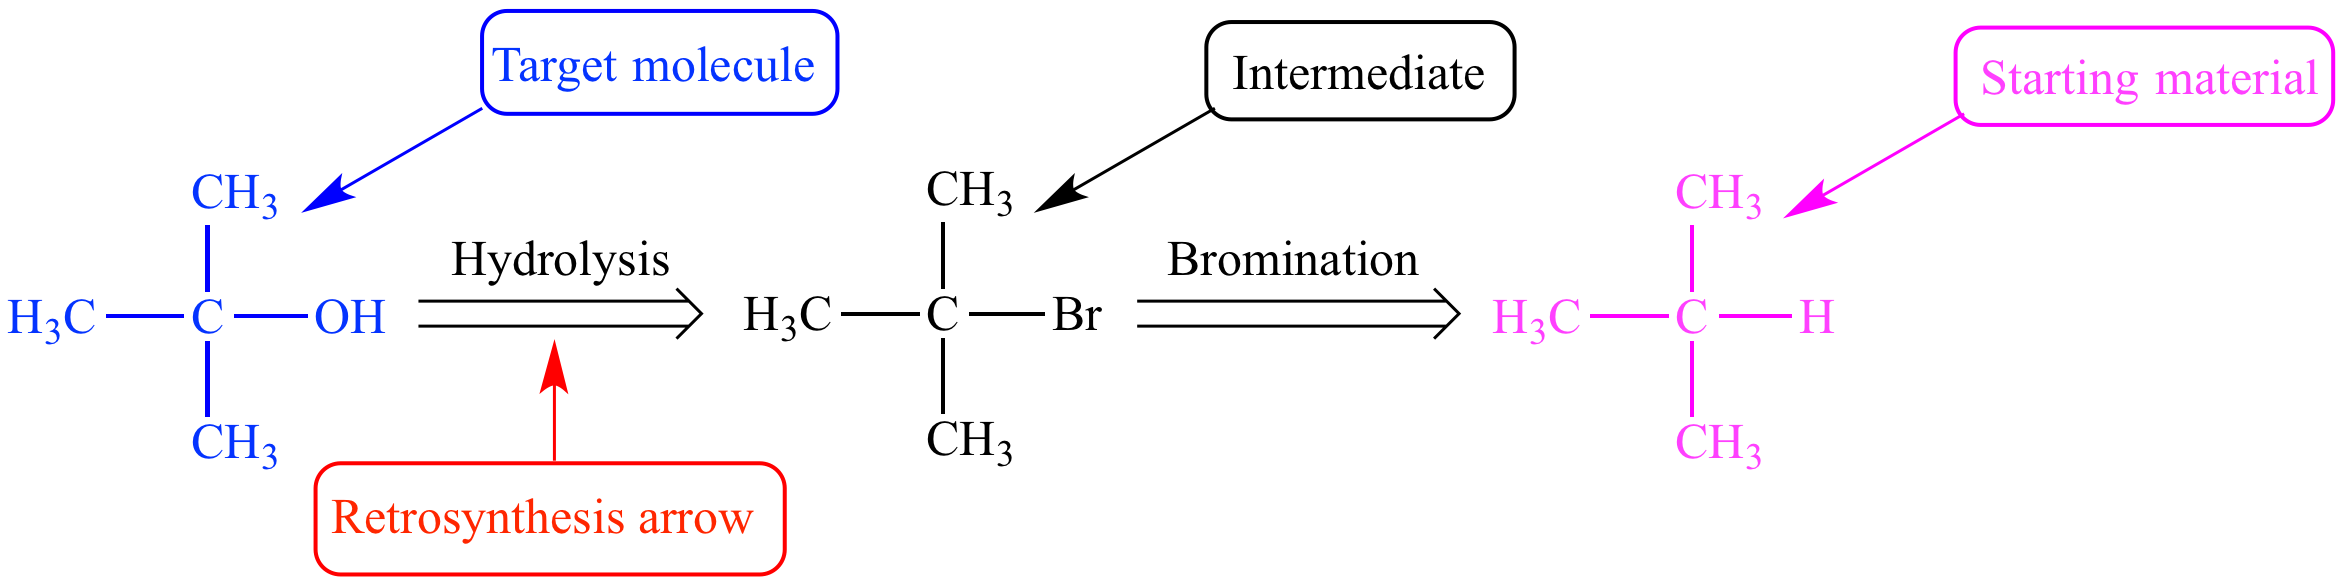
Retrosynthesis is just the practice of breaking a target molecule down into simpler pieces by mentally cutting bonds, then finding real reactions that would forge those bonds in the forward direction. You are not predicting a synthesis. You are reversing one. The whole method was formalized by E.J. Corey in the late 1960s, and it remains the standard mental framework for planning complex organic routes. The basic operation is straightforward. Look at your target. Identify a bond that, if broken, gives you fragments that are commercially available or clearly simpler. Call that a disconnection. Replace it with a synthetic equivalent, which is often a functional group that indicates a known reaction type. Then repeat on each fragment until you reach starting materials. That is the entire algorithm.
Retrosynthesis In Organic Chemistry: The Practical Workflow
Here is how I actually run through a problem, not how it appears in the textbook. I start by mapping the functional groups and noting any stereocenters. The stereochemistry matters early because certain disconnections simply cannot set stereochemistry reliably. If I see a 1,3-diols, diols, amino alcohols, or -hydroxy carbonyls, I immediately flag possible aldol, aldol-type, or epoxide-opening precursors. If I see a 1,5-dicarbonyl, I am thinking about Stork enamine chemistry or Michael additions. If I see an allylic alcohol adjacent to a quaternary center, I consider a Evans aldol or a Julia-Kocienski variant before even drawing a single arrow. The next step is ranking disconnections by reliability, not by how elegant they look on paper. I use retrosynthetic equivalents, also called syntheses, to translate each imagined cut into a real reagent pair. A ketone disconnection near a carbon-carbon bond often maps back to an organometallic addition or an enamine alkylation. An ether might suggest a Williamson ether synthesis from an alkoxide and an alkyl halide, but I also have to consider whether the halide will undergo elimination under the basic conditions. That is a common student blind spot. They draw the disconnection and forget that the synthetic equivalent has to survive the actual reaction conditions. From there I evaluate each branch for availability, cost, and step count. I prefer routes that converge rather than those that are purely linear. A convergent route halves the impact of a failed late-stage step. If I have three fragments that couple in the final two steps instead of assembling the molecule one bond at a time over seven steps, the overall yield improves dramatically even if the individual steps are identical.
Modern software helps with the brute enumeration part. Tools like Chematica, now part of SynQuest, and IBM RXN for retrosynthetic prediction can propose dozens of routes in minutes. I still use them, but I treat the output as a first draft, not an answer. The programs optimize for literature precedent and reagent availability, not for what actually works on a gram scale in a fume hood. They also struggle with reactions that depend heavily on substrate scope or with transformations that require very specific catalysts.
Get the Full Details
A Real Case Where the Textbook Approach Failed Me
I was planning a route for a fused tricyclic intermediate with two adjacent quaternary centers and three stereocenters. The obvious retrosynthesis pointed to a Diels-Alder disconnection. It produced clean fragments on paper. When I actually executed it, the diene was unstable, the reaction gave a messy ratio of endo and exo products, and the subsequent functional group manipulations degraded the core. I spent roughly three days on that path before abandoning it. The workaround was to step back and disconnect the molecule using a ring-closing metathesis (RCM) strategy instead. I opened one of the rings to a diene precursor, built the two quaternary centers through a conjugate addition sequence, and then closed the ring with a Grubbs II catalyst. The RCM step gave clean conversion, the stereochemistry was set earlier in the sequence where I had more control, and the overall route dropped from nine linear steps to six. The lesson was not that Diels-Alder is bad. It is that the most obvious disconnection is not always the one with the highest probability of working in practice. I now always run a quick literature search on the specific substrate class before committing to a named reaction, because scope matters more than the reaction name itself.
Counter-Intuitive Things Beginners Miss
First, more disconnections do not always mean a better route. Each additional disconnection adds a step. The goal is the shortest reliable route to commercially available material, not the route with the most elegant cuts. I have seen students produce five-disconnection pathways for molecules that could be accessed in two disconnections using a convergent coupling strategy. Second, functional group interconversion (FGI) is not a fallback. It is often the most important tool in the sequence. Changing an alcohol to a leaving group, reducing a ketone, or oxidizing an alkene can enable disconnections that are otherwise impossible. The mistake is treating FGI as busywork instead of as a strategic move that unlocks higher-level disconnections. Third, chemoselectivity drives disconnection choice. If a molecule contains both a ketone and an ester, a Grignard reagent will attack the ketone first. A disconnection that relies on adding to the ester while preserving the ketone requires either protection or a different reagent like an organozinc or a lithiated diisopropylamide for deprotonation. I see this error constantly in student proposals. They propose a disconnection that assumes selective reactivity without verifying that the selectivity actually exists for their specific substrate.
Common Pitfalls and When Retrosynthesis Breaks Down
Retrosynthesis is not a perfect method. It fails or becomes unreliable in several clear scenarios. Steric complexity is the first limitation. Targets with multiple contiguous quaternary centers, bridgeheads in small rings, or constrained macrocycles often resist standard disconnection strategies. The theoretical route exists, but the forward chemistry is unpredictable. I have spent weeks on paper plans for strained bicyclic systems only to find that the required cyclization either does not proceed or proceeds with unacceptable erosion of enantiopurity. Reagent availability and cost is a practical bottleneck. A route that looks great on paper might require a catalyst that costs thousands of dollars per gram, a reagent that is unstable at room temperature, or a solvent that is regulated. In medicinal chemistry workflows, this is often the deciding factor. A slightly longer route using cheap, stable reagents beats a shorter route that relies on a specialty catalyst every time in a production setting.
Stereochemical control is another area where retrosynthesis oversimplifies. The method tells you which bonds to form. It does not guarantee that the stereochemistry will be set correctly. A disconnection that implies an asymmetric aldol reaction requires a chiral auxiliary, a chiral catalyst, or an enzymatic resolution. If none of those are feasible for your substrate, the route needs revision. I always check stereochemical precedent before finalizing a proposal. A single incorrect stereocenter can invalidate an entire synthesis plan. Scalability is frequently ignored in academic retrosynthetic exercises. A reaction that works at 10 milligrams on silica may fail completely at 100 grams due to heat transfer, mixing, or workup issues. Exothermic reactions, reactions with gas evolution, and steps requiring cryogenic temperatures become problematic fast. I learned this the hard way when a Pd-catalyzed cross-coupling that ran cleanly on a small scale caused a runaway temperature excursion at scale. The retrosynthetic plan was sound. The process chemistry was not.
How to Actually Learn This
The most effective approach is to work through real targets backwards. Start with moderately complex molecules from the literature. Draw the target. Make one disconnection. Check whether the resulting fragments are available or easily made. Repeat. Time yourself. A practiced chemist can generate a viable retrosynthetic analysis for a medium-complexity target in about 15 to 30 minutes. A beginner might take two hours or more. The difference is pattern recognition, not intelligence. The more disconnections you have seen in published syntheses, the faster you can map new targets onto known reaction types. Keep a personal database of disconnections. When you encounter a new reaction in the literature, record the bond formed, the functional group transformation, and the substrate scope. A simple spreadsheet or a note-taking app works fine. I have been maintaining mine for over a decade. When I face a new target, I can search it in minutes instead of spending hours re-deriving known chemistry. Use computational tools as a supplement, not a replacement. Run your target through RXN or a similar platform. Compare the proposed routes to your own analysis. The discrepancies are where the learning happens. If the software proposes a route you did not consider, investigate why. You will often find a reaction type or a strategic disconnection you were not aware of.
Practice with targets that have increasing complexity. Start with a molecule that has one stereocenter and one disconnection. Move to molecules with two stereocenters and two disconnections. Then tackle targets with multiple stereocenters, convergent coupling points, and protecting group strategies. The progression matters. Jumping straight to complex natural products without building pattern recognition first leads to frustration and incorrect plans. The method is simple in principle. The application requires experience, judgment, and an honest assessment of what actually works in the lab. Most textbooks present retrosynthesis as a clean logical exercise. It is not. It is a conversation between your knowledge of reactivity, your awareness of practical constraints, and your willingness to abandon a plan that looks good on paper but will not function in reality.


