Why the Phase Curve Matters More Than You Think
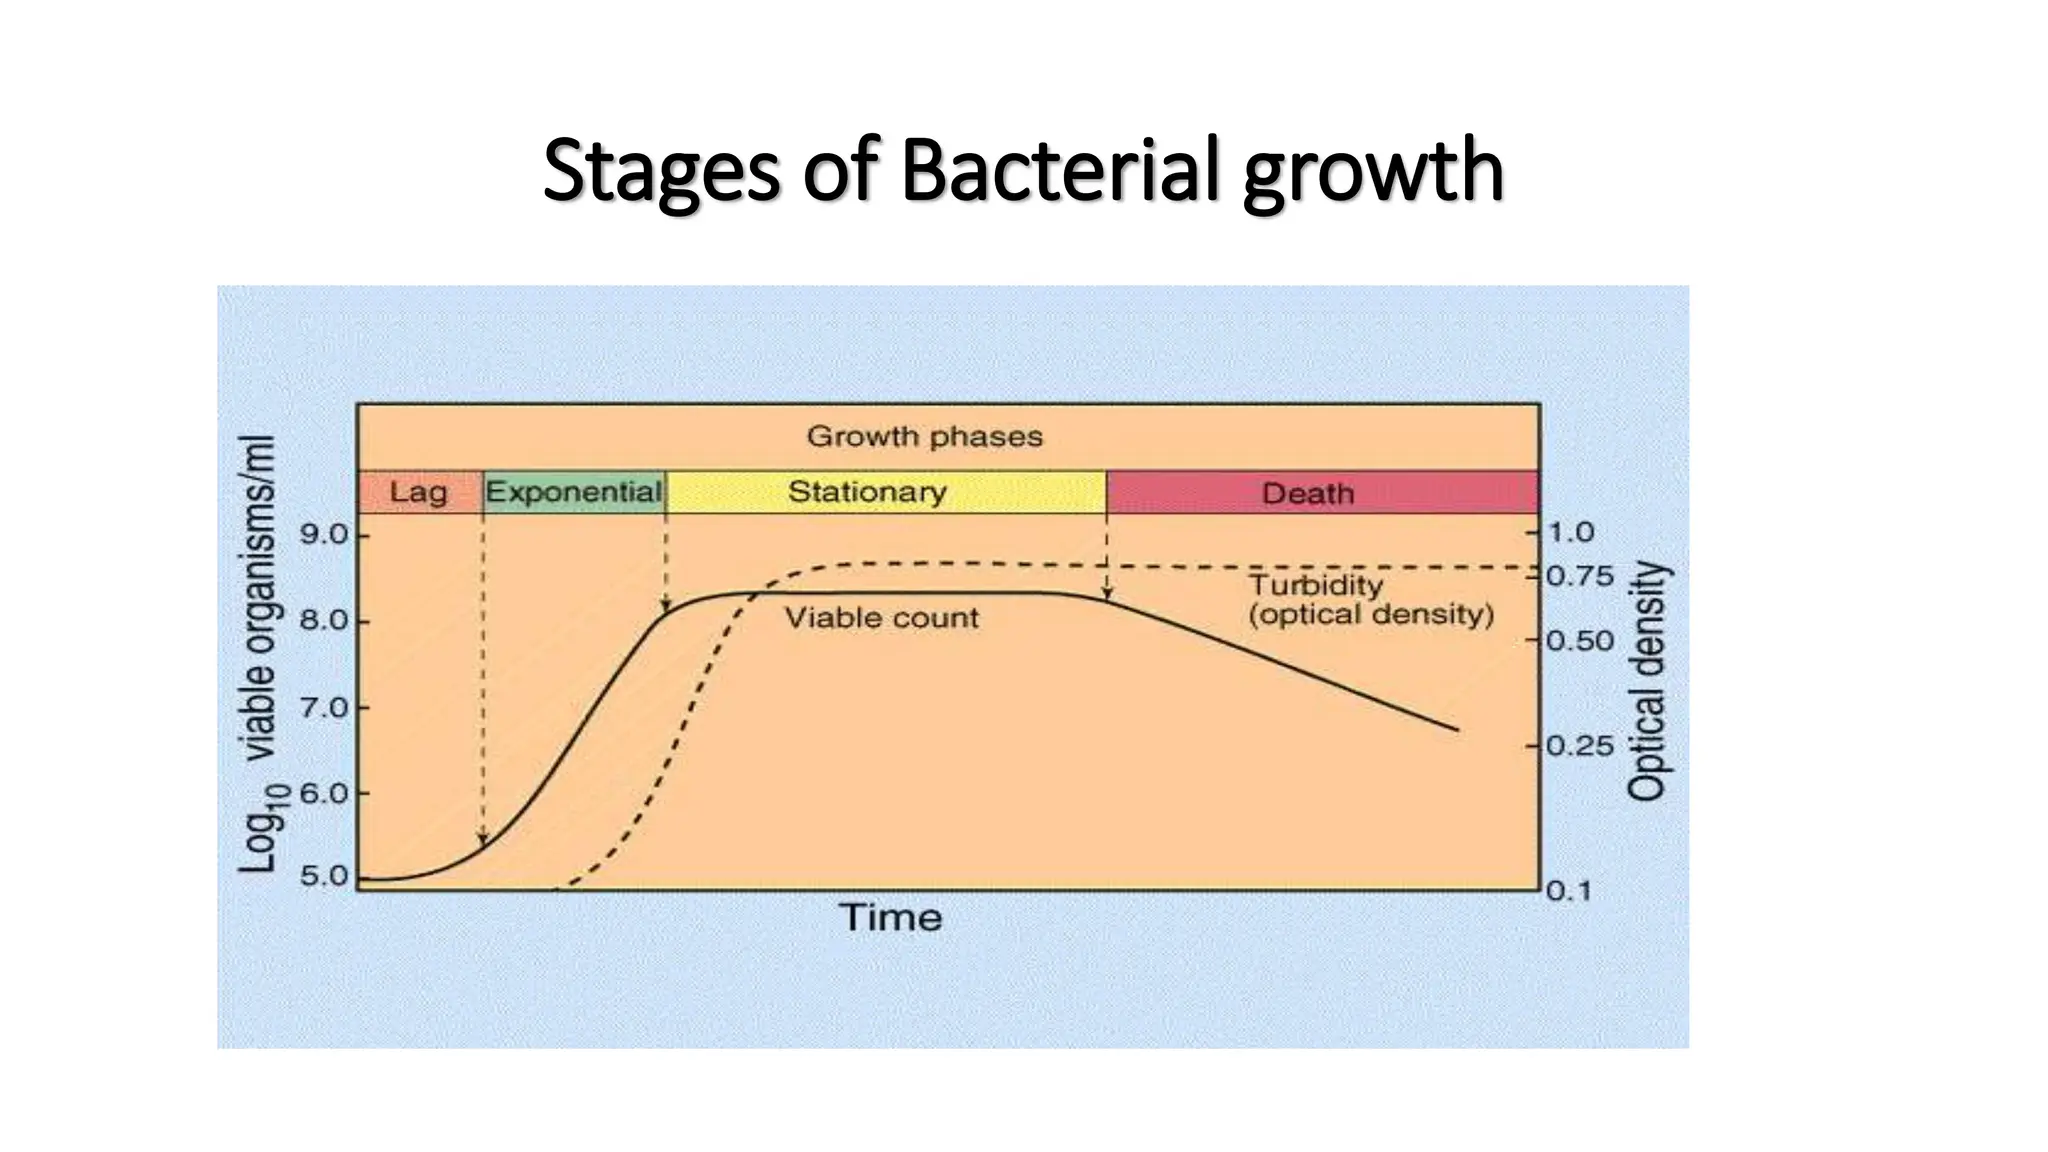
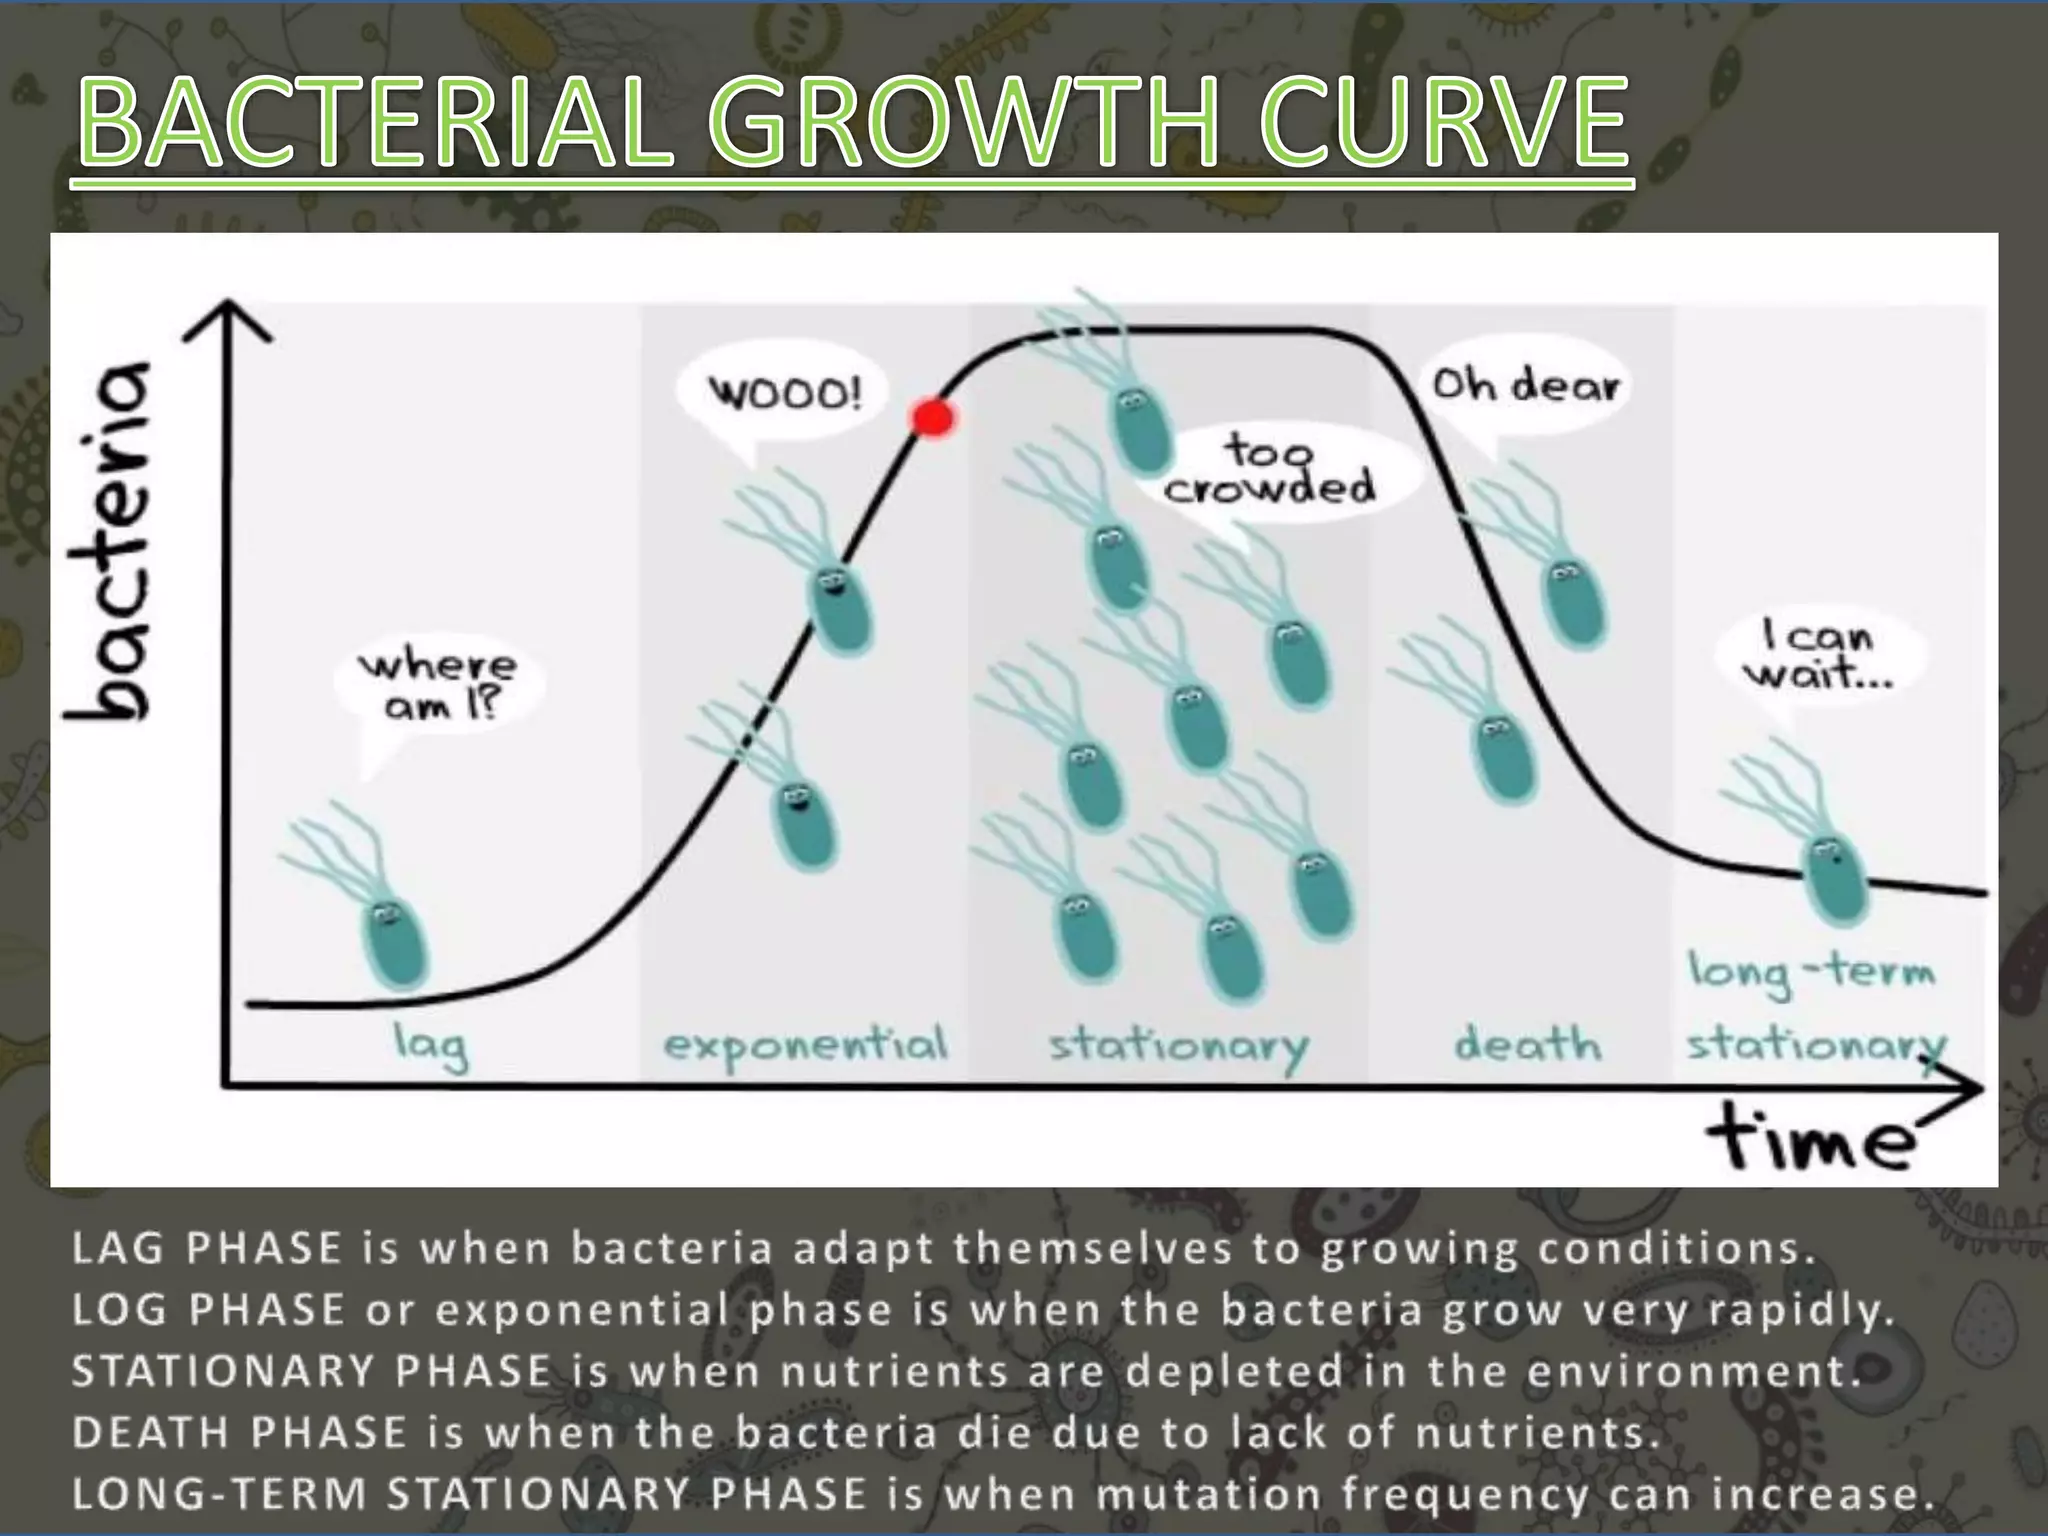
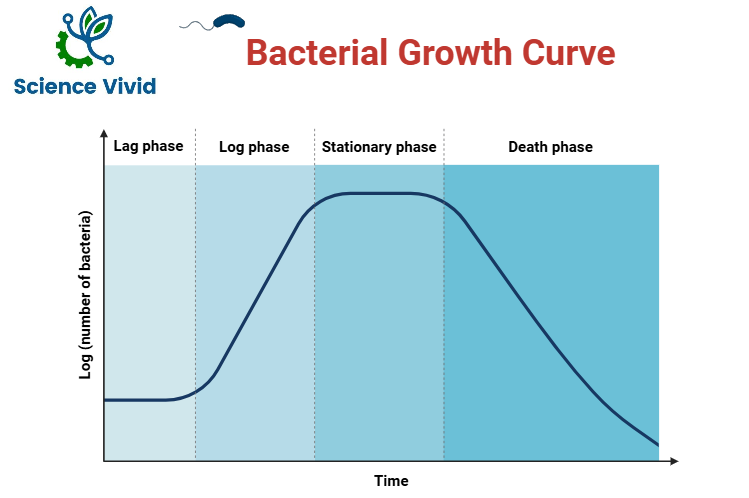
The Stages Of Bacterial Growth aren't something you learn once and remember forever. They're the backbone of everything you do in a microbiology lab, whether you're running cultures for plasmid prep, checking contamination, or troubleshooting why your transformants refused to grow. The standard model has four phases: lag, exponential (log), stationary, and death. That's what the textbooks say. What they don't tell you is how messy it gets when you're actually working with real organisms under non-ideal conditions. Lag phase is when bacteria are adjusting to a new environment. They're metabolically active but not dividing yet. You might see this last 30 minutes or several hours depending on the inoculum health, medium composition, and temperature shift. If you're subculturing from a frozen glycerol stock into fresh LB, that lag can be longer than if you're moving from an overnight culture. It's one of the first things people get wrong—they assume growth starts immediately and time their experiments accordingly. Exponential phase is where cells divide at their maximum rate. In E. coli at 37 degrees Celsius in rich medium, that's roughly one division every 20 minutes. This is the phase you want for most applications—plasmid prep, protein expression, metabolic labeling. The optical density increases linearly on a log scale. If your doubling time is slower than expected, check your temperature, aeration, and whether your starter culture was actually in log phase when you transferred it.
Stationary phase hits when nutrients deplete or waste products accumulate. Growth rate equals death rate. Cells change their metabolism, express stress response genes, and some species start forming spores. This phase is relevant if you're doing long-term survival studies or secondary metabolite production. For routine work, stationary phase cultures are problematic—cells are fragile, plasmids can be lost, and protein yields drop off sharply. Death phase follows stationary. Viable cell counts decline. Not all cells die at the same rate. Some persisters survive for days or weeks. This matters for antibiotic resistance studies and biofilm research. The curve flattens out but doesn't reach zero, which surprises people who expect total clearance.
Working With Real Cultures
I've spent years growing bacteria, and the biggest mistake I see beginners make is ignoring the inoculum quality. A healthy, mid-log culture will hit exponential phase fast. A stationary phase culture dragged into fresh media will have an extended lag. People try to compensate by increasing incubation time, but that just pushes everything downstream. The fix is straightforward—always seed from an active culture, never from old plates or frozen stocks unless you're doing a revival protocol. Here's something that caught me off guard early on. When I was running a large-scale protein expression experiment, the OD readings looked perfect throughout the day. Everything matched the growth curve from my initial trial. Then I plated samples at different time points and the colony counts told a different story. The spectrophotometer was reading dead cell debris as biomass. The actual viable count had dropped significantly by the time I thought I was harvesting. I learned to cross-validate OD measurements with plating, especially when cultures were approaching stationary phase or had been sitting for a while. It added about ten minutes per sample but saved me from wasting an entire day's work. Another common issue is aeration. Shaking flasks at 200 rpm sounds like plenty, but in a 500 ml flask with 100 ml of medium, you're barely getting adequate oxygen transfer. The cells look fine at low density but starve once they hit mid-log. I switched to larger flask volumes or reduced the medium volume and the growth rates improved noticeably. Small changes like that matter more than people give them credit for.
Get the Full Details
Edge Cases and Exceptions
Biofilm-forming organisms don't follow the standard growth curve neatly. They attach to surfaces, produce extracellular matrix, and the planktonic fraction in the superantant can show weird kinetics. If you're working with Pseudomonas or Staphylococcus species, expect deviations from the textbook model. Sampling from the middle of the culture won't represent what's happening at the bottom or on the walls. Slow-growing organisms like Mycobacterium tuberculosis or some environmental isolates have lag phases that stretch into days. Standard incubation times are meaningless for these. I worked with a soil isolate once that took nine days to show any increase in OD. Everyone assumed it wasn't growing and discarded it. A simple plating revealed viable cells were present the whole time, just dividing slowly. The takeaway is that growth curves should be monitored over appropriate time scales for the organism you're working with, not copied from protocols designed for E. coli. There's also the issue of phase locking when you maintain serial transfers. If you always passage cultures at the same optical density, you can create a steady state where the bacteria never fully enter stationary phase. This is useful for some experiments but problematic if you're trying to study stress responses or stationary phase gene expression. The cells adapt to the rhythm and their physiology shifts accordingly.
Tracking Growth Accurately
Optical density at 600 nanometers is the standard measurement. It's quick and non-destructive. But it measures turbidity, not cell count or viability. At high densities above about 0.8 OD the readings become unreliable because light scattering saturates. Dilute the culture and re-measure. At very low densities below 0.05 the signal-to-noise ratio is poor. Use a larger path length or concentrate the sample. For accurate viable counts, plate serial dilutions on agar and count colonies after overnight incubation. This gives you colony forming units per milliliter. It's labor-intensive but it's the ground truth. Flow cytometry with viability dyes is faster but requires equipment most teaching labs don't have. When building a growth curve, take readings every 15 to 30 minutes during exponential phase and every hour during lag and stationary. More frequent measurements during the log phase capture the true doubling time. Fewer readings during lag can miss the transition point entirely. Automated systems like plate readers with kinetic mode handle this well but you need to confirm they're agitating the plates properly between readings.
When Growth Curves Fail You
Not every culture behaves predictably. Contamination can mask the growth curve of your target organism. A fast-growing contaminant might dominate the OD readings while your intended culture sits at low density. Always verify identity through plating or PCR when something looks off. Mixed cultures produce curved growth patterns that don't match any standard phase profile. Antibiotic selection pressure changes growth dynamics. Cells carrying resistance plasmids grow slower than untransformed cells due to the metabolic burden. This is most noticeable during exponential phase where you'll see a clear difference in doubling time. If you're doing growth comparison experiments, make sure you're comparing like with like—same strain background, same plasmid backbone, same selection. Temperature shifts during incubation create phase transitions that look artificial. A refrigerator door opening too often, a shaking incubator with a failing heater, or even moving flasks to a bench for sampling can throw off your curve. Monitor temperature continuously if precision matters for your experiment.
Media composition matters more than people realize. Rich media like LB supports fast growth but produces more acidic byproducts. Minimal media forces slower metabolism and different growth kinetics. Switching between media types without recalibrating your timing is a reliable way to get wrong results. If you're following a published protocol, use the exact medium specified. Substitutions change the curve.
Bottom Line
The Stages Of Bacterial Growth are a framework, not a law. Real cultures deviate based on conditions, organism, and history. The four phases describe the general pattern but the boundaries are fuzzy. Lag isn't always clean. Exponential phase isn't always perfectly straight on a log plot. Stationary and death overlap in ways that matter for your experimental readout. Pay attention to your specific setup, validate your measurements, and don't trust a single data point or an uncritical OD reading. The curve is a tool, not a certificate of correctness.