Titration Isn't as Clean as the Textbooks Make It Look
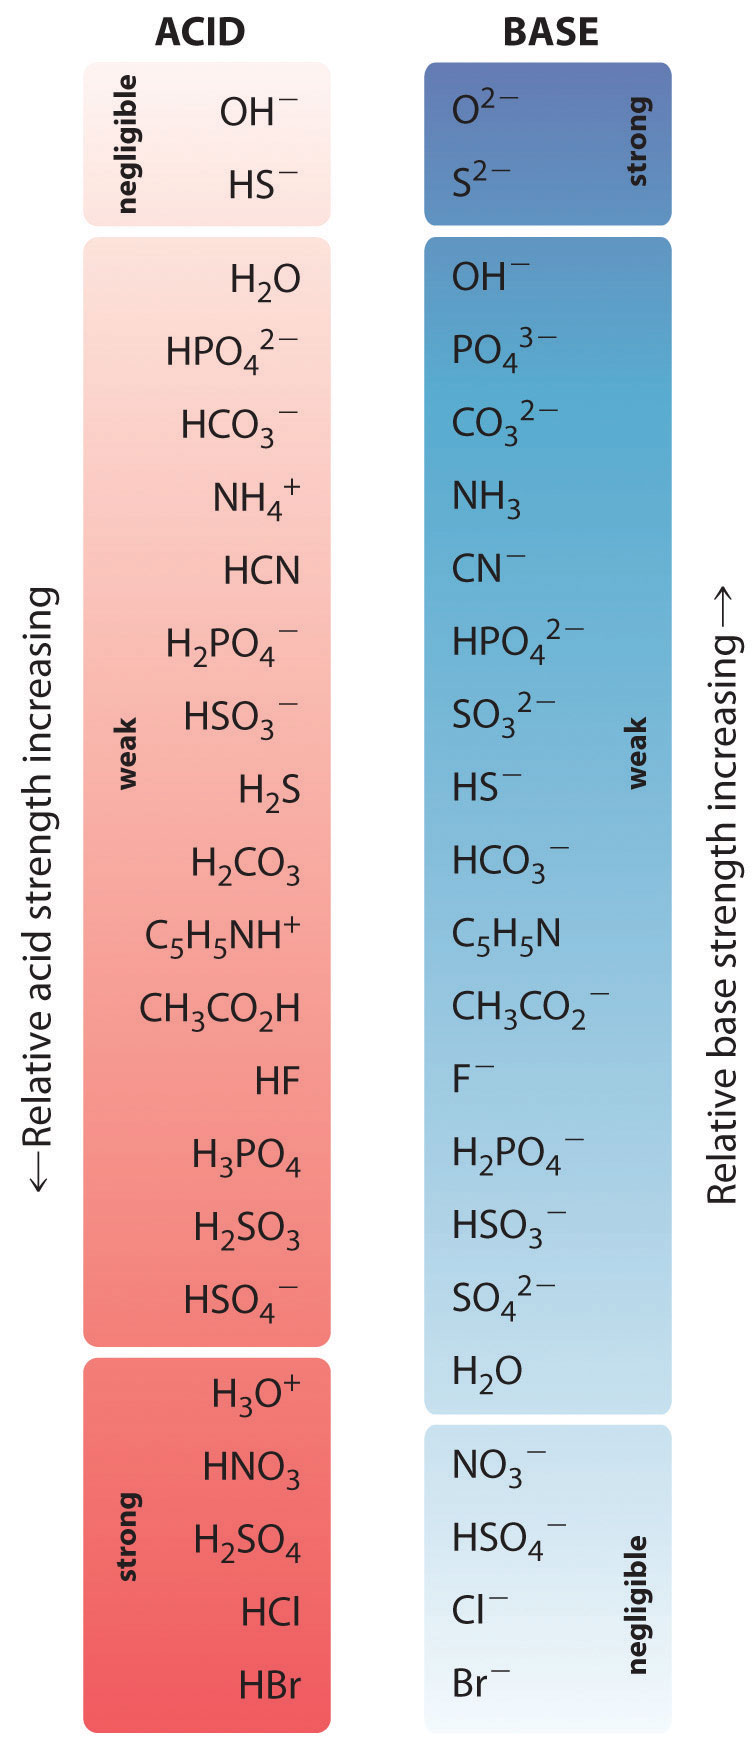
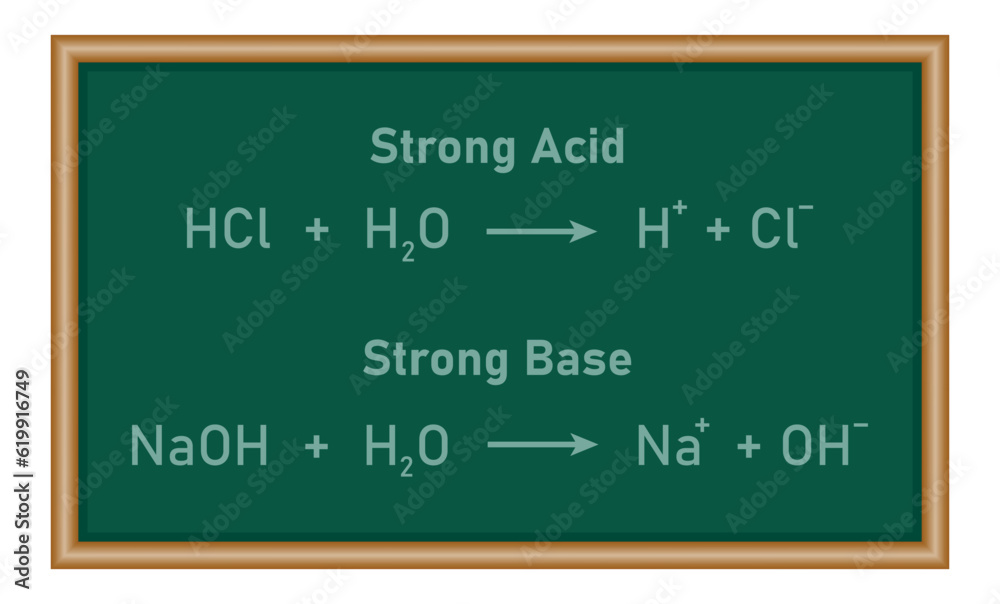
I spent three years as a lab tech running acid-base titrations before I stopped trusting the endpoint indicators. The standard protocol says you add a couple drops of phenolphthalein, swirl the flask, and watch for a faint pink that persists for thirty seconds. That works fine when you're in an undergrad lab with reagent-grade chemicals and distilled water that hasn't been sitting open for a week. Real work is messier. The core concept is straightforward enough. A strong acid dissociates completely in water. Hydrochloric acid, nitric acid, sulfuric acid at reasonable concentrations — they give up their protons without hesitation. A strong base does the same on the other side. Sodium hydroxide, potassium hydroxide, barium hydroxide. They strip protons from water or donate hydroxide ions directly. When you mix them, the H+ and OH- combine to form water. The salt that remains is spectator noise. The pH at the equivalence point sits exactly at 7.0 for a monoprotic strong acid and strong base system at 25 degrees Celsius. That clean symmetry is what makes the titration math work.
Strong Acid And Strong Base Calculations You Actually Need
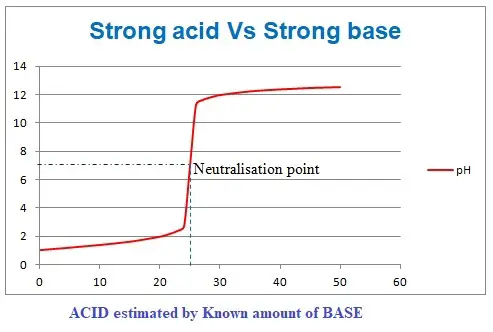
Before you set up any titration, you need to understand what the pH curve looks like across four distinct regions. The starting point is just the pH of your strong acid solution. If you have 0.1 M HCl, the pH is 1.0. No activity coefficient corrections needed at that concentration for rough work. As you add base, the pH rises slowly at first, then shoots up dramatically near the equivalence point, then flattens out again once you've overshot into excess base territory. Here's where most people mess up. They calculate the equivalence point volume using M1V1 = M2V2 and call it done. That gets you through homework. It doesn't get you accurate results in practice. The concentration of your NaOH solution changes over time because it absorbs CO2 from the air. I once spent two weeks troubleshooting why my titration results were drifting downward by about 0.5 percent each day before I remembered that my sodium hydroxide stock had been open on the shelf. The solution was replacing it with a freshly standardized batch and storing it in a soda lime trap. That one issue accounted for more error than anything else in my workflow. For the actual calculation of pH during the titration, you divide the problem into zones. Before the equivalence point, you have excess acid. Calculate the remaining moles of H+, divide by total volume, take the negative log. At the equivalence point, you're just looking at water autoionization, so pH equals 7. Past the equivalence point, you have excess hydroxide. Calculate the remaining moles of OH-, divide by total volume, find pOH, subtract from 14. The tricky region is within about one milliliter of the equivalence point. A single drop of titrant can shift the pH by three or four units. That's why the slope is so steep and why indicator choice matters more than most people realize.
I learned this the hard way with a batch of unknown strong acid samples. The procedure called for phenolphthalein, which changes color around pH 8.2 to 10. For a strong acid-strong base titration, the equivalence point is at pH 7, so phenolphthalein gives you a slight overshoot. Not huge, maybe 0.1 to 0.2 mL of extra titrant depending on concentration. But when you're doing quality control on a production line and your spec tolerance is tight, that matters. I switched to bromothymol blue, which transitions around pH 6.0 to 7.6. The endpoint landed much closer to the true equivalence point. Not perfect, but better. Still not as good as a pH meter, which is what I ended up using anyway. There's a common misconception that strong acid-strong base titrations are the easiest type and therefore you can rush them. They're easy in theory. In practice, the steep pH change near the equivalence point means small errors in volume measurement become large errors in calculated concentration. A 0.05 mL burette reading error on a 25 mL titration is a 0.2 percent uncertainty. That sounds small until you're working with tight specs or when you're standardizing a primary standard and need that uncertainty to propagate correctly. Temperature is another thing nobody accounts for enough. The neutral point of water shifts with temperature. At 50 degrees Celsius, neutral pH is about 6.63, not 7.0. If your lab runs hot or your reactions are exothermic and you're titrating warm solutions, your equivalence point is not at pH 7. Most people ignore this and accept the error. If you need precision, measure the temperature of your solution and adjust. The pKw value is well tabulated at different temperatures.
Get the Full Details
For concentrated strong acids above about 1 M, activity coefficients start to matter. The simple pH equals negative log of concentration formula underestimates the actual hydrogen ion activity. I worked with 6 M HCl once and the measured pH was about 0.6, not the 0.78 the simple calculation would give. Dealing with activity requires the Davies equation or similar models. Most routine titrations don't need this level of correction, but it's worth knowing it exists if your concentrations push past the ideal range. When you're doing the actual lab work, the procedure that consistently gives reliable results is standardizing your base first. Use potassium hydrogen phthalate as a primary standard. It's stable, has a high molecular weight which minimizes weighing error, and gives a sharp endpoint with phenolphthalein. Weigh about 0.4 to 0.5 grams into an Erlenmeyer flask, dissolve in about 50 mL of CO2-free distilled water, add two drops of phenolphthalein, and titrate. Do this at least three times. If your results vary by more than 0.3 percent relative standard deviation, something is wrong with your technique or your reagents. For the acid sample itself, if it's a strong acid of unknown concentration, take an aliquot, dilute if necessary, add your indicator, and titrate against your standardized base. Record the initial and final burette readings to 0.01 mL if your burette allows it. The volume difference is your titrant volume. Calculate the moles of base used, which equals the moles of acid at the equivalence point for a monoprotic acid. Divide by the aliquot volume to get the acid concentration.
The biggest practical limitation of strong acid-strong base titration is that it only works when both the acid and the base are strong. If either is weak, the equivalence point pH shifts away from 7, the buffer region creates a flat portion on the curve that makes endpoint detection ambiguous, and your indicator choice becomes critical. Phenolphthalein fails for weak acid-strong base titrations if you're not paying attention to the equivalence point pH. But that's a different problem. For pure strong acid and strong base systems, the method is robust, fast, and requires minimal equipment beyond a burette, some glassware, and a standardized titrant.