PCR, Gel Electrophoresis, and Everything Between
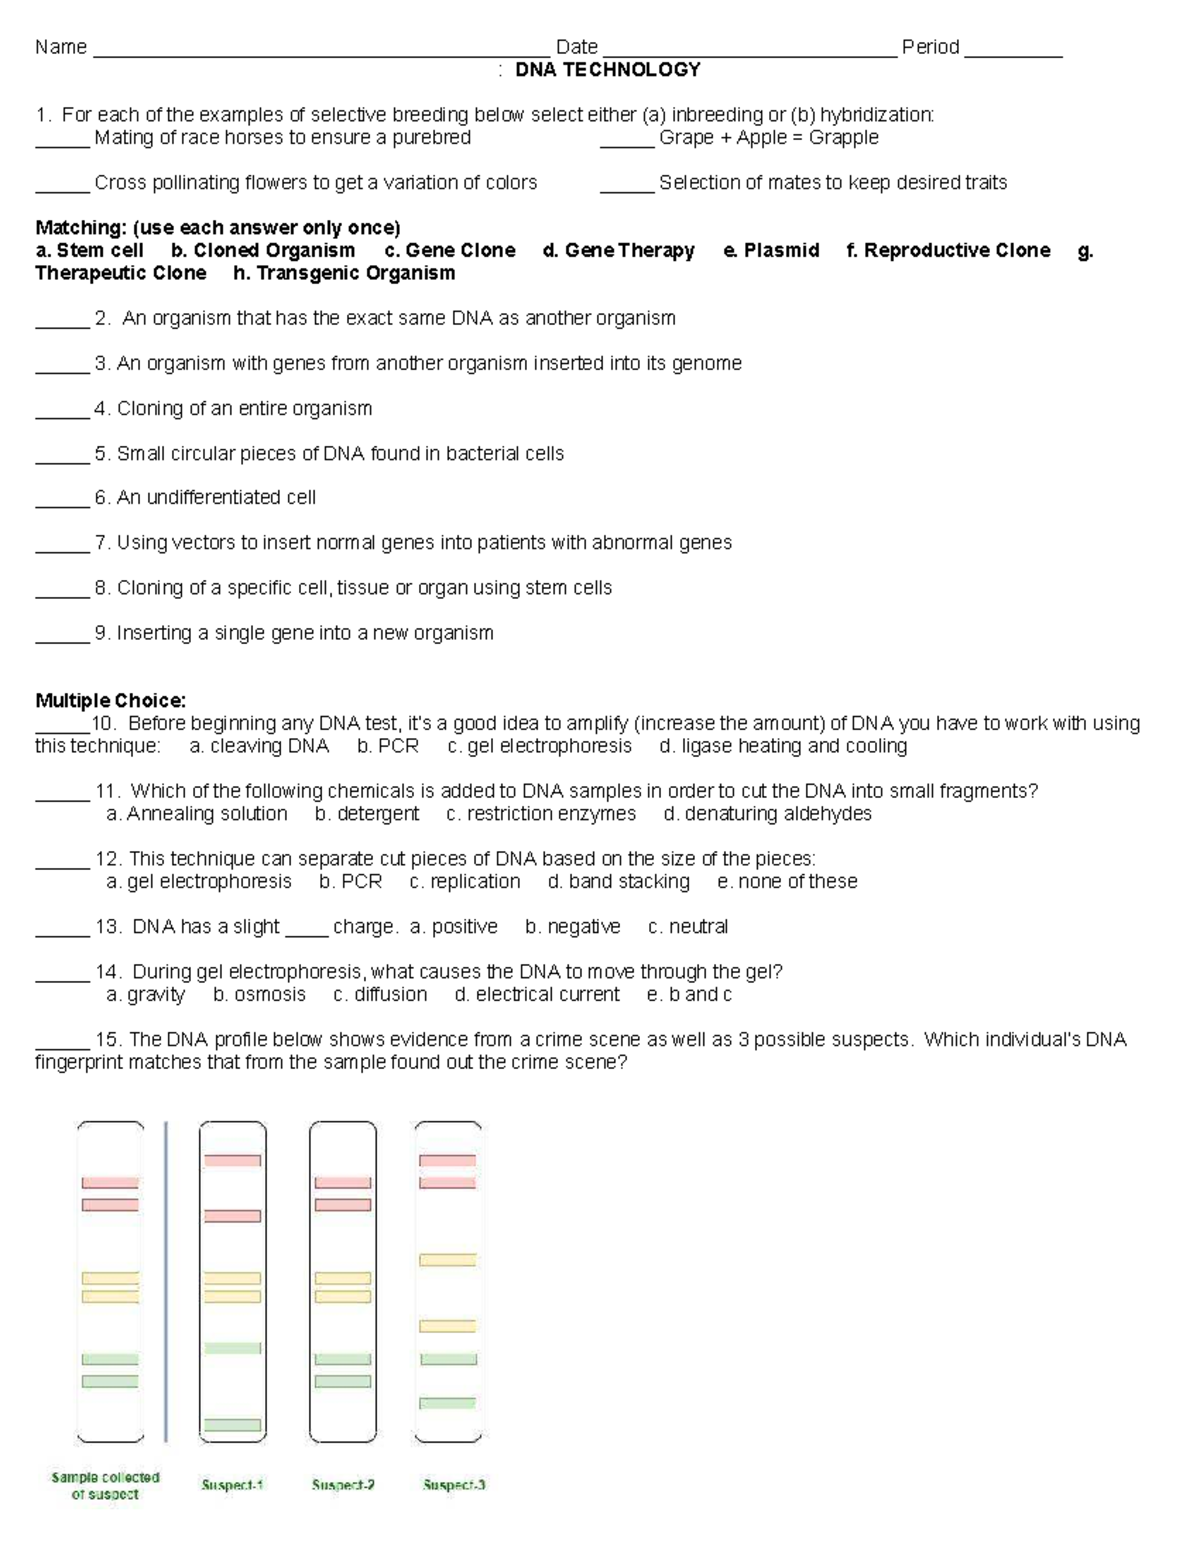
I keep running into people who treat Study Guide Section 3 Dna Technology Continued like a memorization checklist. That approach falls apart fast because the concepts build on each other. If you skip understanding why restriction enzymes cut where they cut, gel electrophoresis looks like magic instead of a straightforward separation method. I learned that the hard way during my first undergraduate lab. The section usually covers recombinant DNA technology, PCR, gel electrophoresis, Southern blotting, DNA sequencing methods, and sometimes CRISPR depending on the curriculum. Each topic is practical. You need to know both the mechanism and the real-world constraints. Take PCR first. The standard protocol involves denaturation at 94-98°C, annealing at 50-65°C depending on primer Tm, and extension at 72°C. That's textbook stuff. What the study guide rarely emphasizes is that your annealing temperature can make or break your reaction if your primers have secondary structures. I once wasted an entire afternoon troubleshooting a reaction where the forward primer was forming a hairpin because I hadn't checked the fold-back structure before ordering. Running a quick mfold or even just scanning for complementarity in the primer sequence itself would have saved hours. The workaround was redesigning the primer with a few mismatched bases at the 3' end to disrupt the secondary structure while keeping the Tm acceptable.
Restriction enzymes come next and they deserve more attention than students give them. Not all enzymes follow the same cut pattern. Some produce sticky ends, others produce blunt ends. The overhang length matters enormously when you're ligating fragments. A 4-base overhang ligates much more efficiently than a 2-base overhang. I remember a lab where we chose EcoRI for its nice six-base cutter and 4-base sticky end, but our plasmid had an internal EcoRI site we hadn't mapped. The vector self-ligated instead of taking up our insert. Solution was a double digestion with two different enzymes that created non-compatible ends, which forces directional cloning and eliminates the self-ligation problem entirely. Gel electrophoresis separates DNA by size using an electric field through an agarose matrix. The smaller fragments migrate faster. That's basic. But here's where people lose marks: the voltage matters. Running a gel at too high a voltage causes heating, which distorts the bands and reduces resolution. For a standard 1% agarose gel, 80-100 volts gives you clean bands in about 45 minutes. If you crank it to 150 to save time, your bands smear and you lose the ability to distinguish fragments that differ by only 50 base pairs. Also worth noting is that ethidium bromide is effective but hazardous. Many labs have switched to SYBR Safe or similar dyes that are less mutagenic while giving comparable sensitivity. Southern blotting is another area where the textbook simplification hides the actual difficulty. Transferring DNA from a gel to a nylon membrane sounds mechanical. In practice, capillary transfer takes several hours and if your gel dries out even slightly during the setup, the transfer fails completely. The alternative is electroblotting, which is faster but requires specialized equipment that not every teaching lab has. A common pitfall is insufficient UV crosslinking afterward, which means your DNA washes off during hybridization and you get no signal. Ten minutes of UV at the right wavelength fixes this, and you should always include a positive control probe so you know your transfer worked.
DNA sequencing has shifted dramatically since the study guides were written. Sanger sequencing is still the gold standard for confirming clones, but the cost and throughput have changed everything. You can send a plasmid prep for Sanger sequencing for about 8-10 dollars per reaction now, and you get results back in 24 hours. The main limitation is read length. You're generally looking at 800-1000 clean bases per reaction, which is fine for confirming a clone but useless for de novo assembly of large regions. Next-generation sequencing solves that but introduces its own problems: you need bioinformatics infrastructure to process the data, and the error rate per read is higher, requiring deeper coverage to call variants confidently. CRISPR-Cas9 appears in updated versions of this section and it's important to understand both the mechanism and the off-target risk. The guide RNA directs Cas9 to a specific PAM-adjacent sequence, and the double-strand break gets repaired through NHEJ or HDR. NHEJ is error-prone and creates indels, which is useful for knockouts. HDR is much less efficient and requires a donor template. I've seen students assume that designing a guide RNA is sufficient. It isn't. You need to check the guide against the whole genome for similar sequences using tools like Benchling or CRISPOR. Off-target cuts can produce phenotypes that have nothing to do with your intended edit, and reviewers will catch that in any publication attempt. Here's something most Study Guide Section 3 Dna Technology Continued materials don't stress enough: transformation efficiency varies wildly between cell lines and preparation methods. Competent cells made by the calcium chloride method typically give you 10^6 to 10^7 colonies per microgram of plasmid. Electroporation-competent cells can reach 10^9 to 10^10. If you're working with a difficult construct or a large library, using electrocompetent cells isn't optional, it's necessary. I learned this when my ligation produced nothing on calcium chloride plates but gave hundreds of colonies the moment I switched to electroporation.
Get the Full Details

The practical takeaway is that DNA technology isn't a series of isolated facts. Restriction digestion feeds into ligation, which feeds into transformation, which feeds into screening, which may feed into sequencing. Understanding the flow and the failure points at each step is what actually helps you on an exam and in a lab. Memorizing the steps without that context just means you'll forget them under pressure or be lost when a protocol doesn't work as expected, which is always.