Reading Reaction Energy Diagrams Without Losing Your Mind
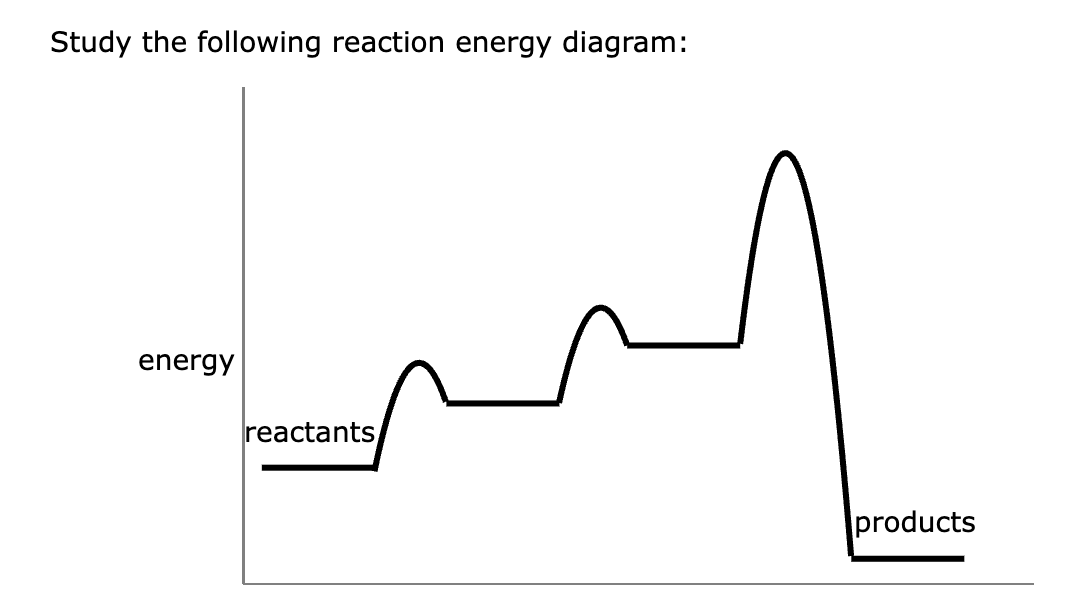
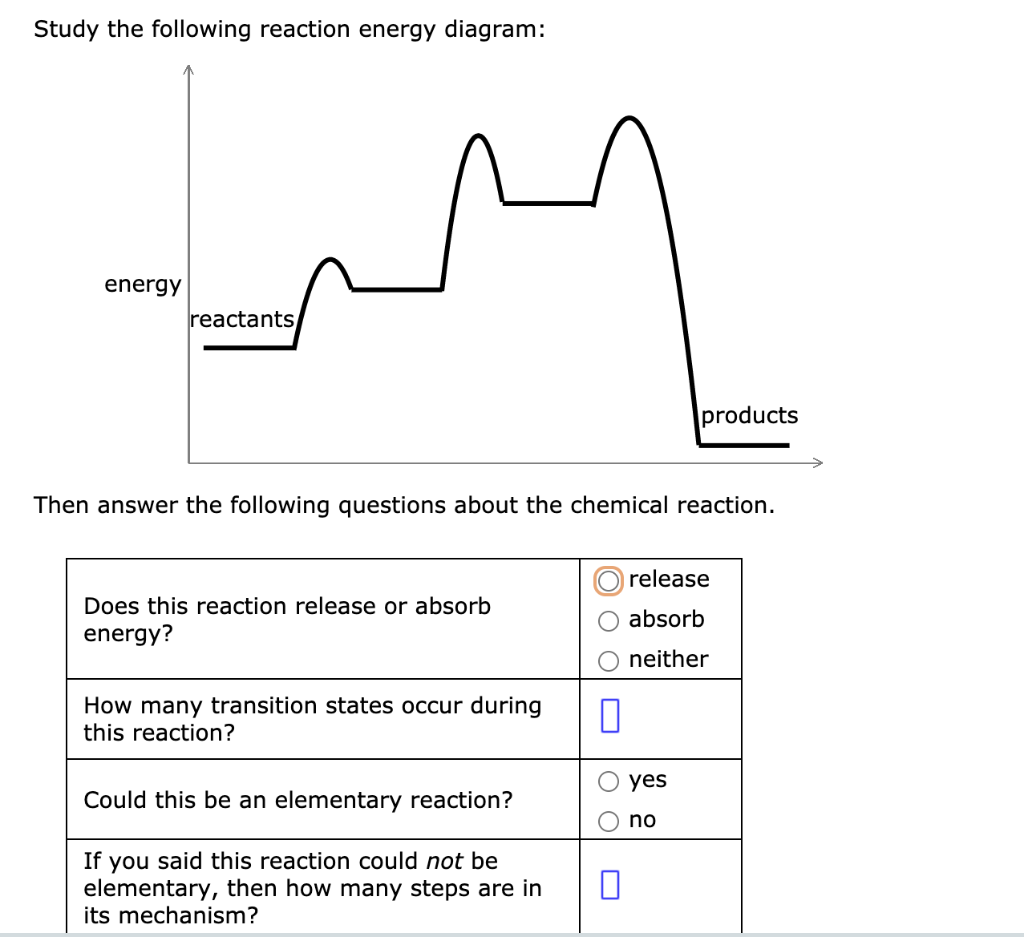
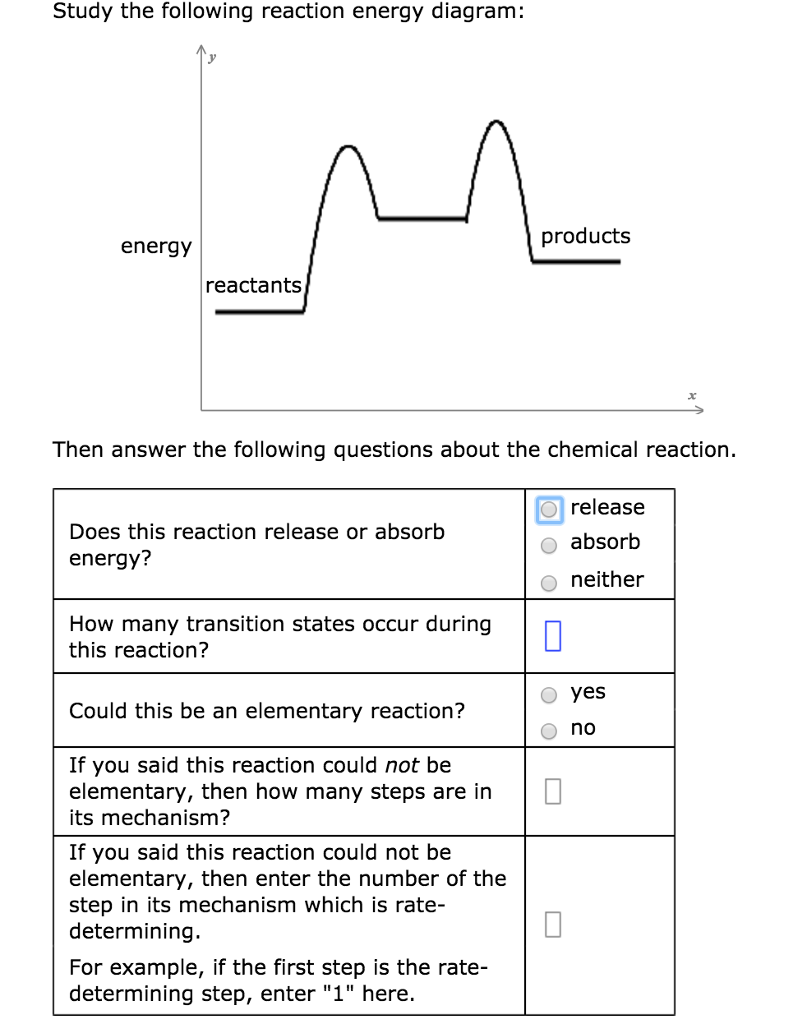
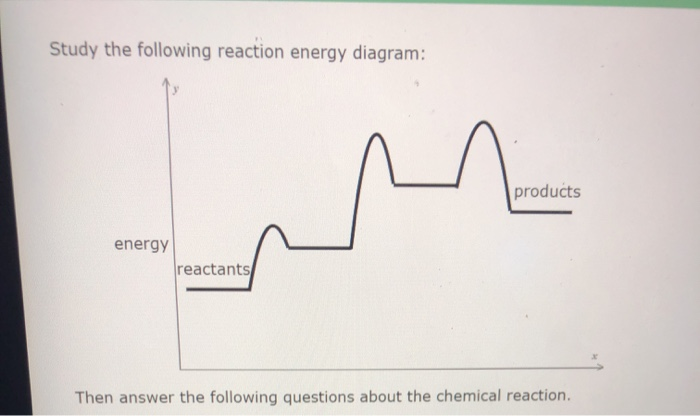
Most people treat reaction energy diagrams as if they were simple line graphs from a middle school science class. They are not. The diagram on your screen or paper is a two-dimensional projection of a potential energy surface that exists in 3N-6 dimensions for a molecule with N atoms. What you are really looking at is a slice through a much higher-dimensional landscape, and understanding that fact alone will save you from half the mistakes students make on exams. When you Study The Following Reaction Energy Diagram, the first thing to check is the axis labels. I cannot count how many times I have seen a graph where the x-axis was labeled "Reaction Coordinate" with no indication of what that coordinate actually represents. Sometimes it is an imaginary frequency eigenvector from a transition state calculation. Sometimes it is a interpolated bond length or an angle. Sometimes it is literally nothing physical — just a mathematical parameter that connects reactants to products. The shape of the curve depends entirely on which coordinate you chose. This matters because the activation energy you read off the diagram is only meaningful relative to that specific coordinate. Change the path and you change the barrier height. This is one of those counter-intuitive points that beginners consistently miss. The barrier is not an intrinsic property of the reaction alone. It is a property of the reaction plus the path you chose to trace through configuration space.
What the Axes Actually Tell You
The y-axis is potential energy, usually in kJ/mol or kcal/mol. In computational chemistry it is often shown in Hartrees. The important detail here is what kind of energy. Is it electronic energy from a DFT calculation? Does it include zero-point vibrational energy corrections? Are enthalpy or Gibbs free energy terms folded in? A diagram can look dramatically different depending on this choice. A reaction that appears endothermic in electronic energy might be exothermic once ZPE is included. I spent a full day once chasing an apparent thermodynamic inconsistency in a catalytic cycle because someone had mixed unscaled electronic energies with zero-point-corrected values from different calculations. The fix was straightforward — recompute everything at the same level of theory with consistent frequency scaling — but spotting the mismatch took far longer than it should have. A standard single-step reaction diagram shows reactants on the left, products on the right, and a hump between them. The height from reactants to the peak is the activation energy. The difference in energy between reactants and products tells you whether the reaction is exothermic or endothermic. That is the textbook version. Here is what the textbooks leave out. Multiple steps do not always mean multiple humps. A reaction can have a detectable intermediate and still show a single barrier if the well is very shallow. Conversely, some diagrams with two obvious humps actually represent a concerted mechanism where the "intermediate" is a shallow plateau, not a true minimum on the potential energy surface. You need to verify by checking the Hessian at the stationary point. One imaginary frequency means a transition state. All real frequencies means a minimum. If someone hands you a diagram and claims a species is an intermediate without showing the frequency analysis, that claim is unverified.
The transition state structure does not always resemble the highest-energy point on the diagram. This is Hammond's postulate, but people apply it incorrectly. Hammond says the transition state structure shifts toward the closer species in energy. It does not say the TS geometry can be approximated by linear interpolation between reactant and product coordinates. I once optimized a TS structure using a linear geometric guess between two conformers and landed on a completely different stationary point — a bicyclic arrangement the starting materials could not have accessed through simple bond stretching. The diagram would have looked wrong because the reaction coordinate I assumed was not the one the molecule actually followed.
Get the Full Details
Practical Pitfalls When Working With These Diagrams
Conformational degeneracy is the silent killer of accuracy. A reactant might have three low-energy conformers, each leading to a different transition state with different barrier heights. If you only model the lowest-energy conformer and ignore the others, your calculated rate constant could be off by orders of magnitude. The fix is to scan the relevant degrees of freedom systematically, map all low-lying conformers, and use Boltzmann weighting when summing their contributions. This can add hours to a calculation but it is the difference between a number that is roughly correct and one that is useless. Solvation effects are another source of error that people underweight. Gas-phase diagrams and solution-phase diagrams can show inverted reactivity. A nucleophilic substitution that is barrierless in the gas phase might develop a significant barrier in polar solvent because the charged nucleophile is stabilized by solvation more than the transition state is. If your diagram comes from a continuum solvation model like PCM or SMD, remember that these are approximations. They work reasonably well for equilibrium geometries but can struggle with charge localization in tight transition states. Here is a specific edge case I ran into last year. We were studying an organometallic oxidative addition and the computed diagram showed a surprisingly low barrier for a reaction that experimental kinetics suggested should be much slower. The issue was that the transition state involved a solvent-separated ion pair, and our implicit solvation model did not capture the discrete solvent coordination around the emerging charges. Adding two explicit solvent molecules to the model and reoptimizing raised the barrier by about 12 kJ/mol, bringing the computed rate into agreement with experiment. Implicit models alone would have given a misleadingly clean diagram.
How to Actually Use a Reaction Energy Diagram
Start by identifying every stationary point: reactants, intermediates, transition states, and products. Verify each one with a frequency calculation. Transition states must have exactly one imaginary frequency, and that mode must correspond to motion along the reaction path connecting the claimed reactant and product. Pull back along that normal mode vector from the TS geometry and confirm you reach the right minima on both sides. This is called intrinsic reaction coordinate (IRC) verification, and skipping it is the most common error I see in student work and in published computations alike. Once the stationary points are verified, the energy profile is constructed by plotting the Gibbs free energies at the temperature of interest. Electronic energies alone are fine for qualitative sketches but insufficient for anything quantitative. Thermal corrections from the frequency calculation give you enthalpy and entropy contributions. At room temperature, the thermal correction to Gibbs energy is typically in the range of -20 to -50 kJ/mol for small organic molecules, which is large enough to flip the sign of a borderline enthalpy change. When you read the activation barrier from the diagram, remember that the relevant quantity for kinetics is the Gibbs free energy of activation, not the electronic energy difference. The Arrhenius equation uses an effective barrier that includes entropic effects. A reaction with a modest enthalpy barrier but a highly ordered transition state can be slower than one with a larger enthalpy barrier but a loose transition state. This is why entropy matters and why diagrams based purely on electronic energy can mislead you about relative reaction rates.
When the Diagram Lie To You
There are situations where a reaction energy diagram is fundamentally inadequate. Tunneling is the first example. For hydrogen transfer reactions at low temperatures, the effective barrier can be significantly lower than what the diagram shows because the particle tunnels through the barrier rather than going over it. A diagram based on classical transition state theory will overestimate the rate. Corrections exist but they require additional calculations and they complicate the simple picture. Dynamics effects are another limitation. The diagram assumes that every trajectory crossing the transition state proceeds to products. In reality, some trajectories recross the barrier and return to reactants. The transmission coefficient accounts for this, and for complex molecules with many vibrational modes, it can deviate substantially from unity. This is particularly relevant for unimolecular reactions in the fall-off regime where collisional energy transfer plays a role. Finally, there is the issue of multi-dimensional barrier crossing. A one-coordinate diagram implies that there is a single dominant reaction path. Many reactions, especially in solution phase and in enzymatic catalysis, proceed through ensemble of paths with different geometries and barriers. The diagram you see is a projection, and important chemistry can be hidden in the dimensions you are not looking at. Conformational sampling and enhanced sampling methods like metadynamics or umbrella sampling are the tools used to explore this, but they require significantly more computational effort than a standard TS optimization.
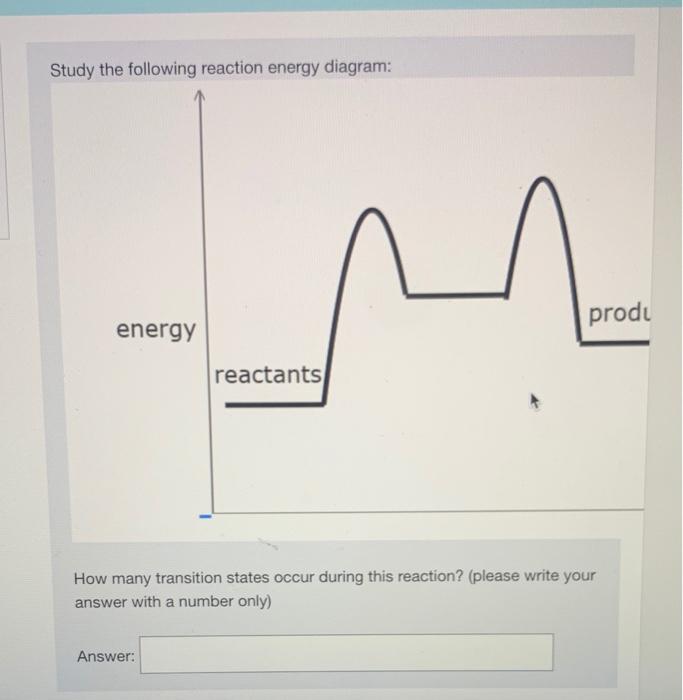
The takeaway is that a reaction energy diagram is a useful abstraction, not a complete description of what a molecule does. It gives you the landscape. It does not tell you how the molecule moves across it. Treat it as a starting point for analysis rather than the final answer, and always verify the stationary points that define the profile before you build any kinetic model on top of it.