The Actual Way You Predict Molecular Geometry
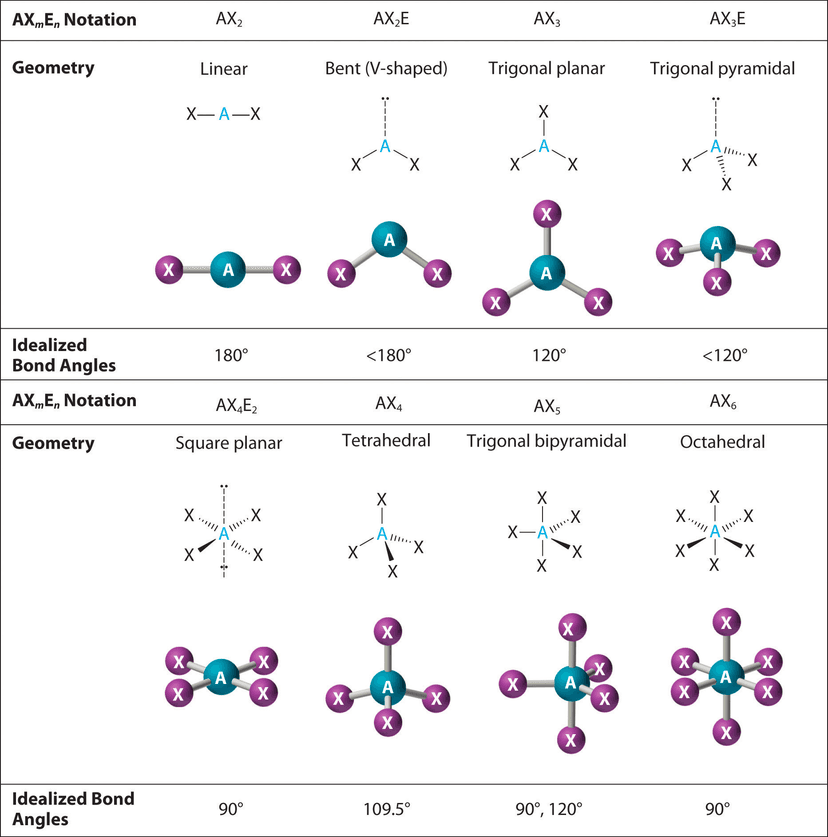
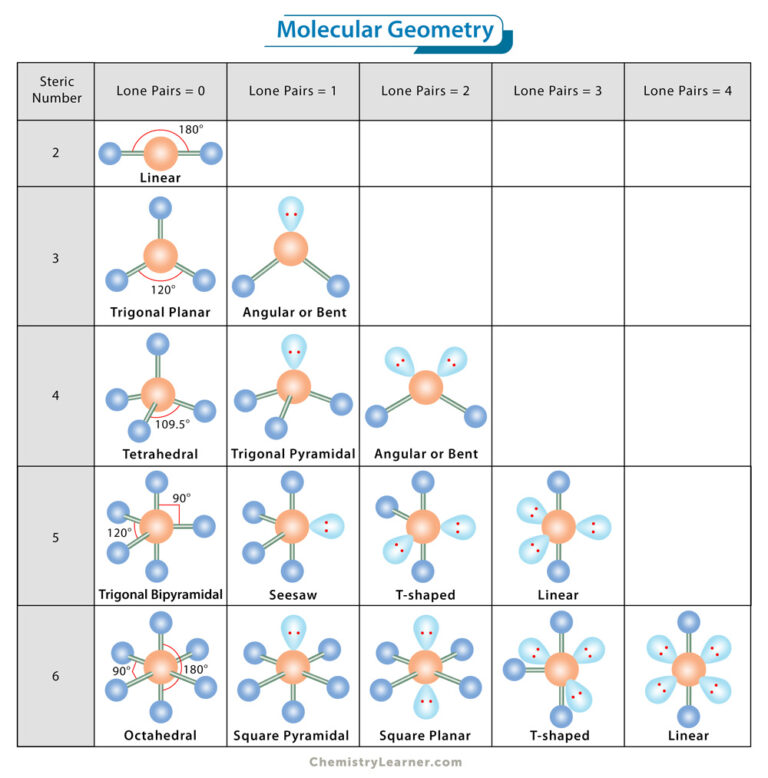
Most people learn Types Of Molecular Geometry as a memorization chore. Count domains, match to a chart, move on. That approach works for quiz questions but falls apart when you actually need to reason through something unfamiliar. I spent three years doing computational chemistry modeling, and the ones who got tripped up weren't the students who didn't know their geometries—they were the ones who treated VSEPR like a rigid rule set instead of a qualitative repulsion model. Here is how I actually approach it now. Start with the Lewis structure. This sounds obvious but it is where most mistakes originate. Get your electron count wrong on the central atom and everything downstream is garbage. Draw single bonds to each substituent, fill octets on the outer atoms, and dump any leftover electrons on the central atom as lone pairs. The total number of electron domains around the central atom—bonding pairs plus lone pairs—is your steric number. That number is the single most important thing in the entire process. Steric number 2 gives linear electron geometry. If both domains are bonding pairs, the molecular geometry is also linear with a 180 degree bond angle. Carbon dioxide is the textbook case. Steric number 3 gives trigonal planar electron geometry at roughly 120 degrees. Two bonding pairs and one lone pair produce a bent molecular geometry, which is why sulfur dioxide has a bond angle closer to 119 degrees rather than a perfect 120. The lone pair takes up slightly more space and compresses the bond angle by a couple of degrees.
Types Of Molecular Geometry: The Core Framework
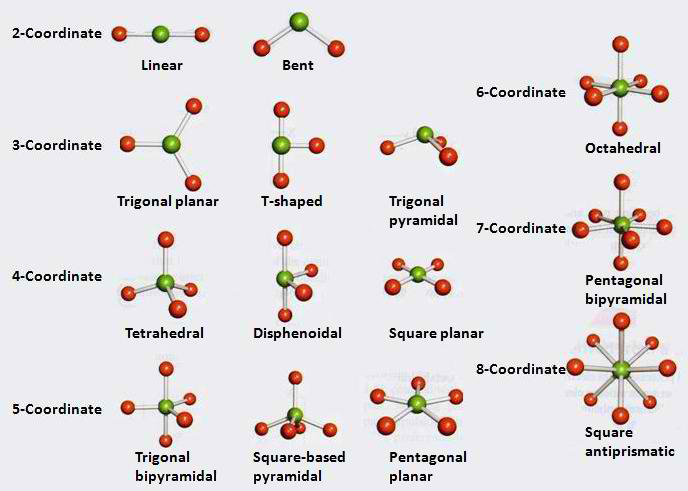
Steric number 4 is where things get crowded. The electron geometry is tetrahedral at 109.5 degrees. Four bonding pairs with no lone pairs keeps that shape intact—methane is the reference point. Three bonding pairs and one lone pair give trigonal pyramidal geometry. Ammonia is the canonical example, and its bond angle drops to about 107 degrees because the lone pair pushes the N-H bonds downward. Two bonding pairs and two lone pairs produce bent geometry. Water sits at roughly 104.5 degrees. The difference between the steric number 3 bent angle and the steric number 4 bent angle matters because students routinely mix them up on exams. Steric number 5 introduces the trigonal bipyramidal electron geometry, and this is where the model gets interesting. The five positions are not equivalent. Three occupy an equatorial plane at 120 degrees from each other, and two sit in axial positions perpendicular to that plane. The axial-equatorial angle is 90 degrees. Lone pairs always prefer the equatorial sites because that placement minimizes the number of 90-degree interactions. One lone pair gives a see-saw shape. Two lone pairs produce T-shaped geometry. Three lone pairs leave you with linear molecular geometry despite the underlying trigonal bipyramidal arrangement. Iodine trifluoride is a case where this matters—you might expect it to be bent or pyramidal, but the three lone pairs force a T-shaped structure. Steric number 6 gives octahedral electron geometry with all positions equivalent at 90 degrees. One lone pair produces square pyramidal geometry. Two lone pairs give square planar geometry, and they go opposite each other to minimize repulsion. Xenon tetrafluoride is the standard example. The key thing to notice here is that square planar and see-saw look superficially similar—four atoms around a central atom—but their origins are completely different. One comes from steric number 6 with two lone pairs, the other from steric number 5 with one lone pair.
I ran into a problem once modeling a bromine trifluoride complex where the bond angles were significantly off from what VSEPR predicted. The model was giving me angles around 86 degrees experimentally, but the basic VSEPR treatment suggested something closer to 90 for the axial positions with the lone pairs pushing inward. The issue was that bromine is a third-row element, and its larger atomic radius means the lone pairs are more diffuse and exert less directional repulsion than the model assumes. I ended up switching to a DFT calculation with a dispersion correction instead of relying on VSEPR alone. It took about twenty minutes to set up the calculation and another ten to parse the output, but it gave me actual geometries that matched the literature values rather than educated guesses.
Get the Full Details
Where VSEPR Actually Breaks Down
The model works reliably for second-row elements and simple p-block species. Once you move to heavier elements like iodine, xenon, or tellurium, the predictions start drifting. Lone pairs on large atoms are stereochemically less active than VSEPR assumes. The inert pair effect becomes relevant, and bond angles can deviate significantly from textbook values. I have seen cases where VSEPR predicted approximately 90-degree angles and the experimental value was closer to 84 or 85. Not catastrophic for a homework problem, but enough to cause real confusion when you are trying to correlate structure with reactivity. Transition metal complexes are another area where VSEPR largely fails. The d-orbital contributions to bonding change the picture entirely, and crystal field theory or ligand field theory is the appropriate framework. Trying to force VSEPR onto an octahedral cobalt complex is an exercise in futility. You will get the right answer by accident sometimes, but you will not understand why. There is also a subtlety with resonance that most resources gloss over. When a molecule has resonance structures, the electron domain count should be based on the actual delocalized system, not any single Lewis structure. Sulfate ion is a common example. All four S-O bonds are equivalent due to resonance, and the geometry is tetrahedral. If you count double bonds as separate domains from single bonds, you will incorrectly predict a different structure. The correct approach is to recognize that resonance hybrids don't have distinct single and double bonds—they have bond orders somewhere in between, and the steric number reflects the actual electron density distribution around the central atom.
Another thing that catches people out is the difference between electron geometry and molecular geometry. Electron geometry describes the arrangement of all electron domains. Molecular geometry describes the arrangement of only the atoms. For steric number 4 with one lone pair, the electron geometry is tetrahedral but the molecular geometry is trigonal pyramidal. These are two different things, and conflating them leads to errors in predicting polarity and reactivity. A trigonal pyramidal molecule like ammonia is polar because the lone pair creates an asymmetric charge distribution. A tetrahedral molecule like methane is nonpolar because all four positions are equivalent. The difference is entirely about whether you are looking at electron domains or atomic positions. X-ray crystallography and gas-phase electron diffraction are the experimental methods you actually use to determine molecular geometry. Computational chemistry packages like Gaussian or ORCA can predict geometries with reasonable accuracy for small to medium molecules, though you need to choose the right functional and basis set. For quick hand calculations, VSEPR is sufficient. For anything requiring quantitative accuracy, you are looking at DFT with at least a triple-zeta basis set, and even then you are dealing with approximations. The whole process for a moderately sized organic molecule usually runs between thirty minutes and two hours depending on convergence behavior. Hybridization is a related concept that many courses conflate with VSEPR. sp, sp2, and sp3 hybridization correlate with steric numbers 2, 3, and 4 respectively, but hybridization is a mathematical construct, not a physical reality. It is useful for organizing your thinking about bonding, but it does not predict geometry on its own. VSEPR does that. The two frameworks are complementary, and using them together gives you a more complete picture than either one alone.