Titration Calculations That Actually Work in Practice
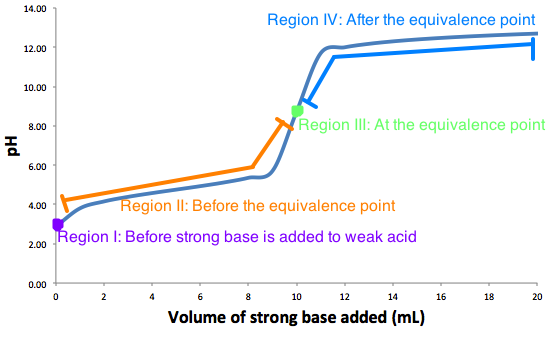
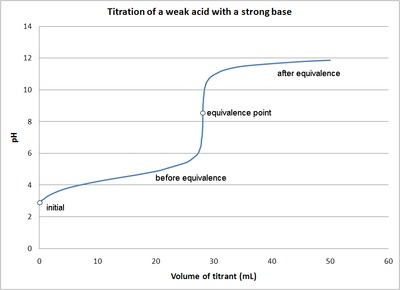
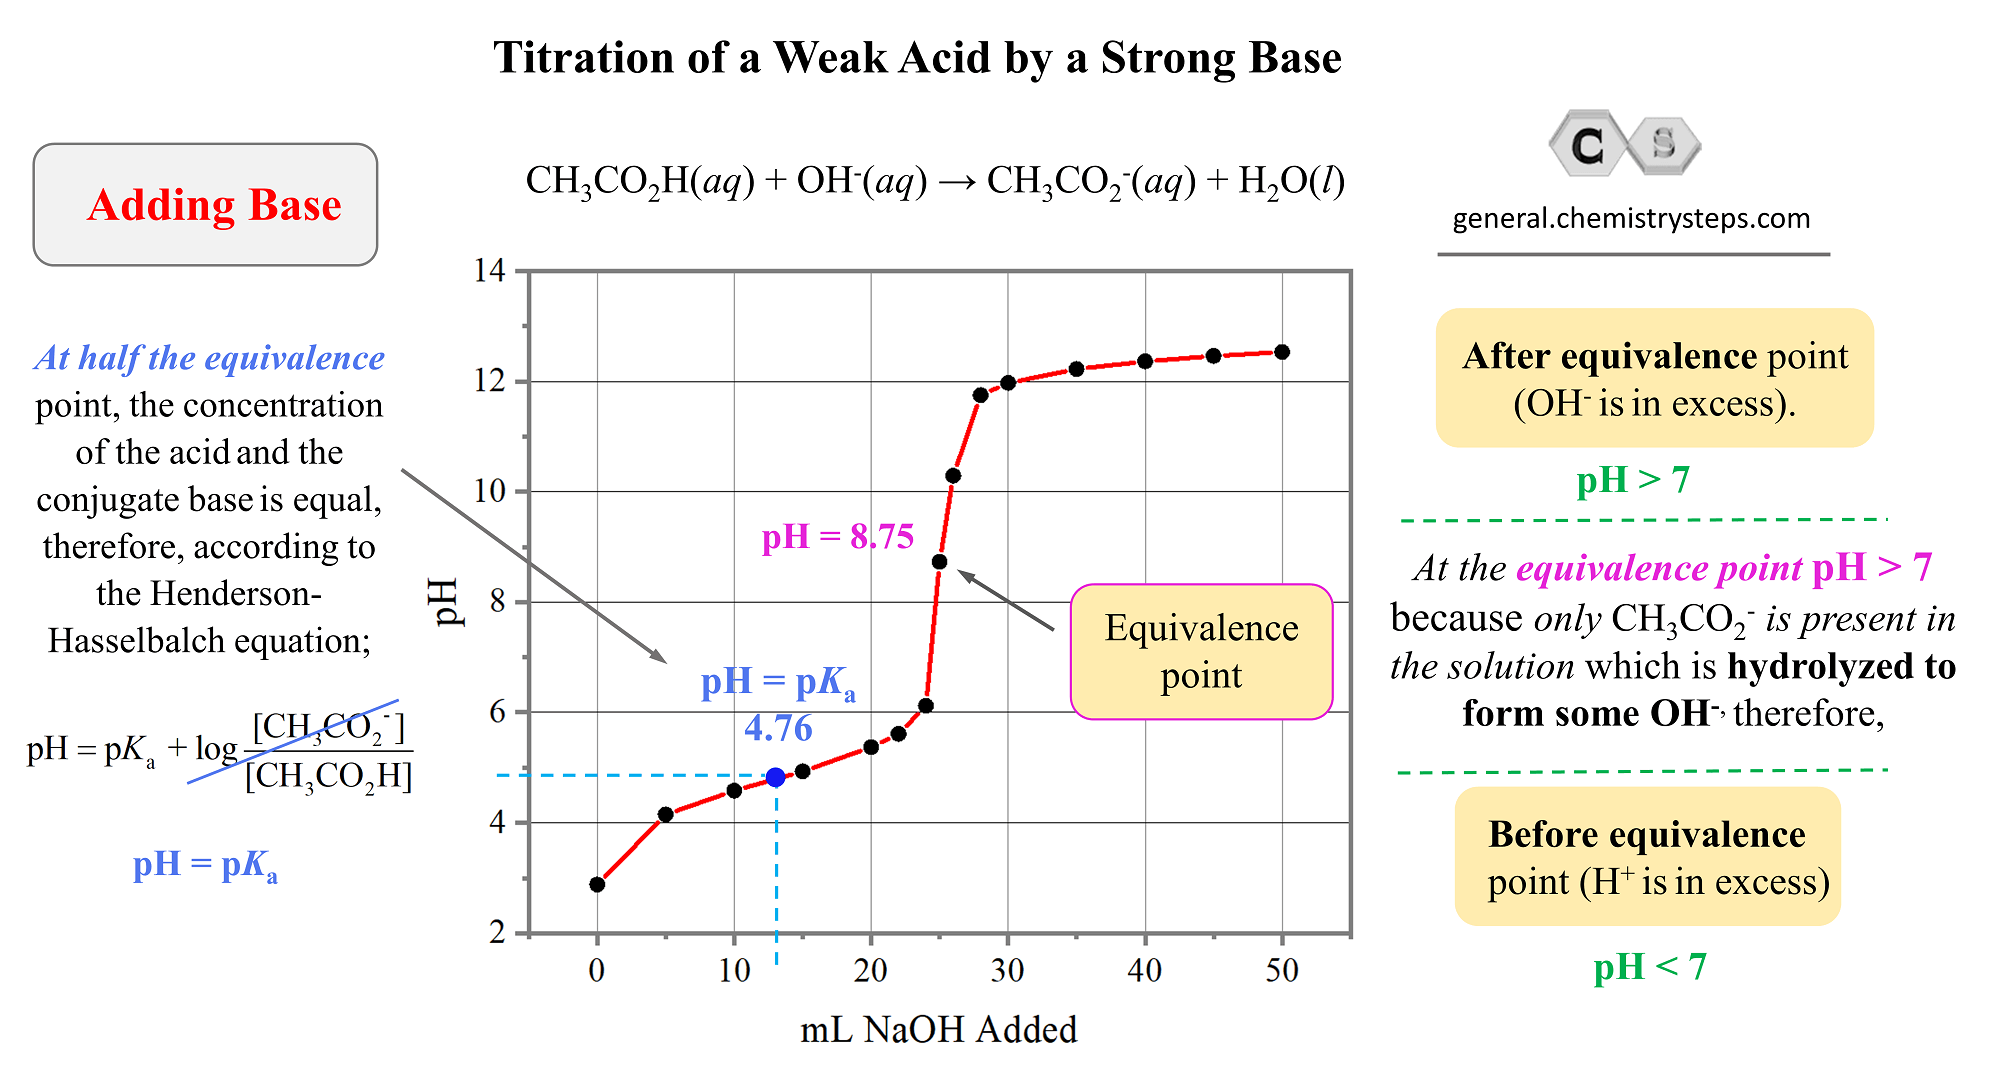
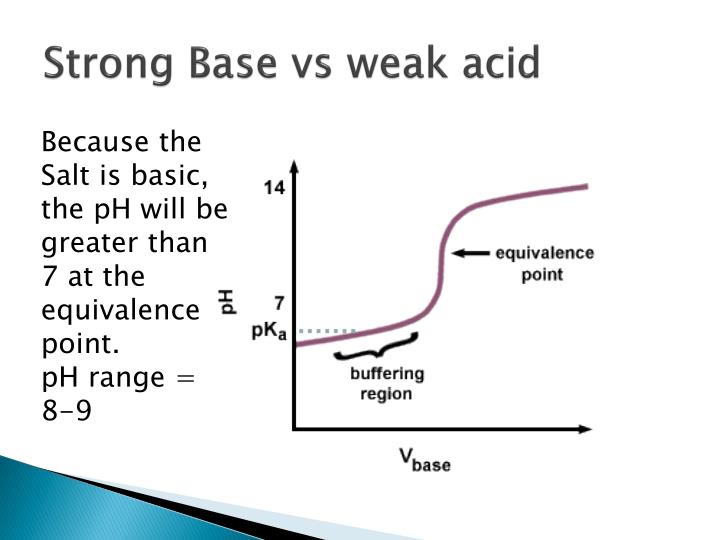
I spent way too long learning that pH curves for weak acid strong base reactions aren't symmetric. The equivalence point sits well above pH 7, and if you assume it's at 7 like you'd do for strong-strong pairs, your whole calculation is wrong. Here's how to handle it properly. Take acetic acid titrated with NaOH. You're neutralizing a proton donor that doesn't fully dissociate, so the math diverges from what you learned in Gen Chem lab before the TA actually explained anything. The key species shifts throughout the curve: you start with HA dominating, move through a buffer region where both HA and A- coexist, hit the equivalence point where only A- remains (and it's hydrolyzing water to make OH-), and finally you're just looking at excess strong base. Most people memorize Henderson-Hasselbalch and think they understand titrations. They don't. Henderson-Hasselbalch works fine for the buffer region only. Before the equivalence point, sure, pH = pKa + log([A-]/[HA]). But that equation means nothing at the equivalence point or beyond, and students write it on exams anyway because they're trying to fill space.
Where the Equivalence Point Actually Lands
At equivalence, you have a solution of the conjugate base A- in water. You need to calculate its Kb using Kb = Kw/Ka, then treat it as a weak base equilibrium problem. For 0.1 M acetic acid titrated with 0.1 M NaOH at equivalence, the acetate concentration is roughly 0.05 M (volume doubled from equal mixing). The resulting pH comes out around 8.7, not 7. This matters because it determines which indicator you should use. Phenolphthalein works here. Bromothymol blue would be useless — it changes color way too early. I ran into this exact issue running a routine acid-base assay on a pharmaceutical batch. Our SOP specified methyl orange as the indicator, and we kept getting endpoint readings about 15 mL too early on every sample. The titrant was 0.1 M NaOH against an acetic acid matrix. After two failed runs wasting about $400 in reagents and three days of analyst time, I recalculated the equivalence point pH on a scrap paper and realized methyl orange (transition range 3.1-4.4) was completely wrong for this system. We switched to phenolphthalein and got consistent results on the third attempt. The method validation was delayed by four days because of it.
Step-by-Step Calculation Method
Before equivalence point: Calculate moles of acid remaining and moles of conjugate base formed. Use Henderson-Hasselbalch. Simple. At the half-equivalence point: [HA] = [A-], so pH = pKa. This is one of the few places where the math is trivial, and it's useful for experimentally determining Ka if you have an unknown acid. I've used this to confirm impurity peaks in HPLC runs by doing small titrations on collected fractions. At the equivalence point: All HA has been converted to A-. Find [A-] accounting for volume change from titrant addition. Calculate Kb from Ka. Set up the equilibrium ICE table for A- + H2O HA + OH-. Solve for [OH-], convert to pOH, then pH = 14 - pOH.
Get the Full Details
After the equivalence point: The conjugate base's contribution to pH becomes negligible compared to excess OH- from the strong base. Just calculate excess moles of NaOH divided by total volume. Ignore the weak base equilibrium entirely. This approximation introduces less than 1% error once you're past equivalence by even 10%.
Common Pitfalls
Not adjusting for dilution. The concentration of every species changes as you add titrant volume. If you started with 25 mL of acid and added 25 mL of base to reach equivalence, everything is halved. People forget this and get pH values that are 0.1 to 0.3 units off. Using concentrations instead of moles when setting up the ratio in Henderson-Hasselbalch. The volume cancels in the log term, so you can use mole ratios directly. This saves you from calculating new molarities at every point, which is where most arithmetic errors creep in. Assuming the weak acid dissociation contributes significantly to pH before the equivalence point. In a 0.1 M acetic acid solution, the initial pH is about 2.87, but if you account for water autoionization or solve the full quadratic from the Ka expression without approximating x as small, you get the same answer to two decimal places. The approximation holds. It usually holds.
When This Approach Breaks Down
If your acid is extremely weak (pKa > 10) and your concentration is very low (below 0.01 M), the pH at equivalence may be so high that the OH- from A- hydrolysis competes with water's contribution. The standard weak base approximation fails. You need to solve the full charge balance equation including [H+] and [OH-] simultaneously. In practice this shows up when you're working with very dilute samples or when analyzing amino acid titrations where you're dealing with polyprotic systems. Polyprotic acids add another layer of complexity that most undergraduate courses barely scratch the surface of. Each equivalence point has its own pH calculation, and the buffer regions overlap if the pKa values are within about 3-4 units of each other. Carbonic acid is the classic example where this gets messy in real samples like blood or river water analysis. The one thing I always check when doing these calculations manually is whether my pH result makes physical sense. If I'm getting a pH below 2 after adding strong base to a weak acid, or above 13 before reaching equivalence, I've almost certainly forgotten to account for volume or mixed up moles and molarity. I've caught my own mistakes this way more times than I'd like to admit.

Practical Shortcut for Quick Estimates
If you need a rough pH at the equivalence point without grinding through the full calculation, remember that for typical weak acid-strong base titrations with concentrations around 0.1 M, the pH will fall in the 8 to 10 range depending on Ka. The weaker the acid, the higher the pH at equivalence. This is backwards from strong acid-strong base where the equivalence is always 7, and it's the single most common conceptual error I see on exams and in lab reports. For actual lab work, the titration curve itself tells you where you are. The steepest vertical section marks the equivalence point regardless of what the pH is. You don't always need to calculate it precisely if your indicator choice covers the steep region. That's what real analysts do rather than computing every point by hand. I still compute them for method documentation and when someone asks why the curve looks different from the textbook, but in day-to-day work the visual inflection point is good enough.