A Practical Look at Core Electrons in Real Chemistry Work
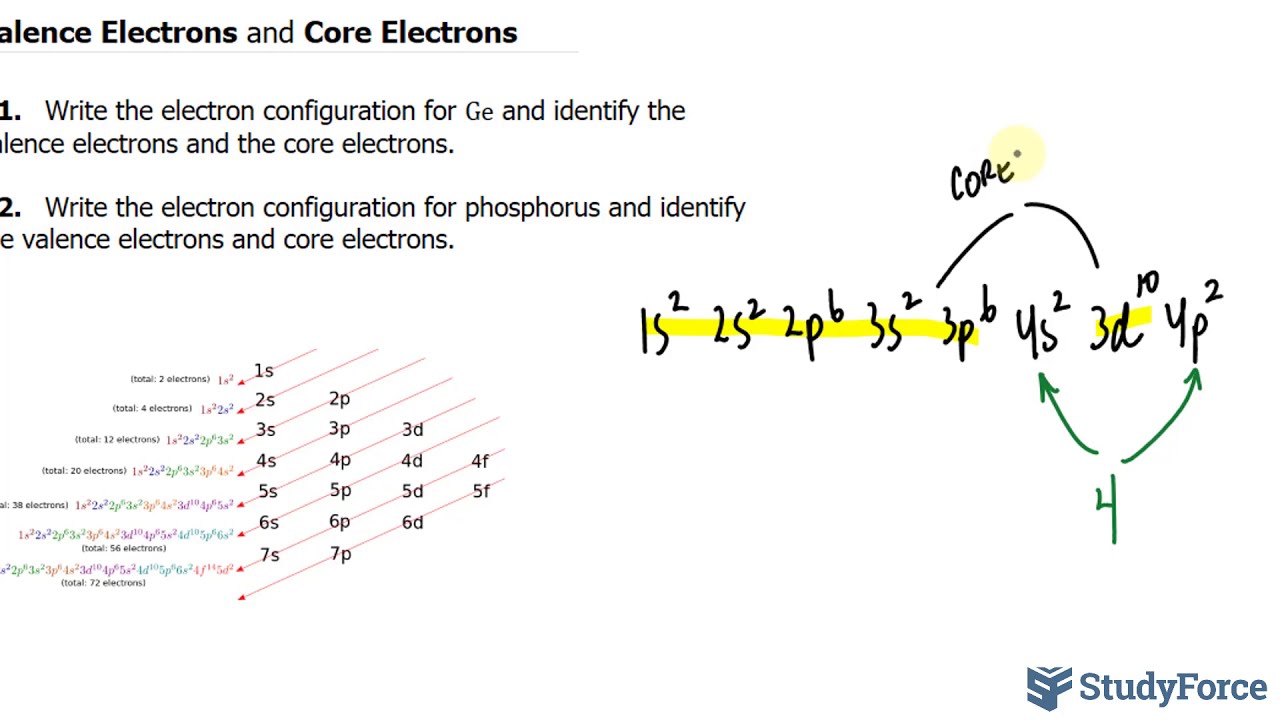
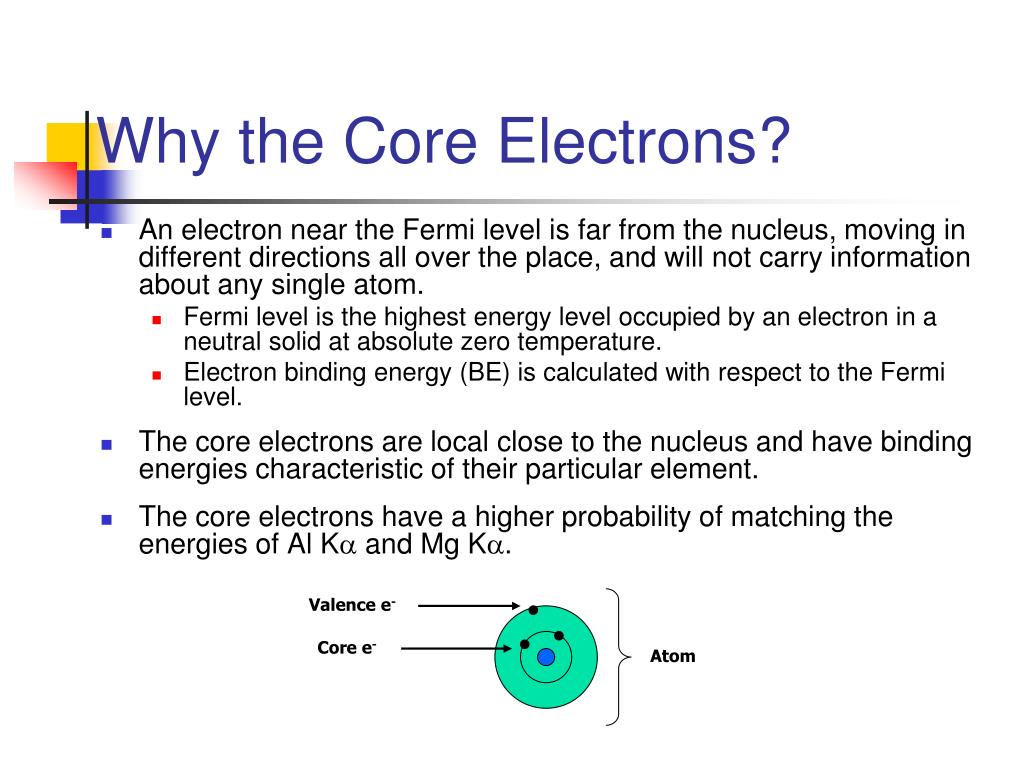
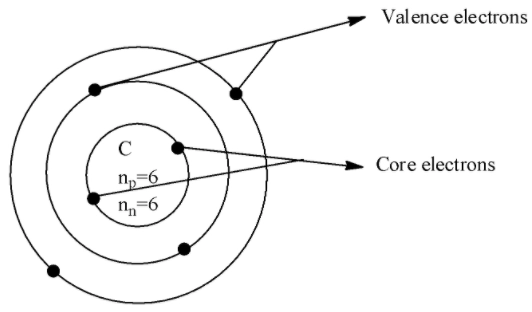
When you're working with computational chemistry or even just trying to understand periodic trends, the distinction between core and valence electrons matters more than most people realize. A core electron is any electron in an atom that sits in an inner shell and doesn't participate directly in bonding. For a sodium atom, that's the ten electrons filling the 1s, 2s, and 2p orbitals. The one valence electron in 3s is what actually does the chemistry. Simple enough on paper, but the complications show up fast when you start working with real data. In practice, core electrons matter because they determine how much computational work you have to do. If you're running a DFT calculation on a transition metal complex, you absolutely do not want to treat every single electron in the system explicitly. That would be computationally brutal and mostly pointless. The core electrons are tightly bound to the nucleus, they're chemically inert, and they don't participate in the reactions you care about. So most people use effective core potentials or pseudopotentials to replace the core region with a mathematical approximation, which reduces your basis set size dramatically. I've seen calculations on moderately sized organometallics go from taking four days to about six hours once we switched from all-electron to an ECP treatment for the heavier atoms. The catch is that not all core electrons are equal. There's a whole gray area where electrons are deep enough to be mostly core-like but close enough to the valence shell that they can still influence chemistry. This is especially true for second-row elements and beyond. Take phosphorus, for example. The standard treatment calls 1s²2s²2p the core and leaves 3s²3p³ as valence. But those 2p electrons aren't entirely inert. When I was working on a project involving phosphine ligands and their interaction with early transition metals, I noticed that using a standard ECP for phosphorus gave slightly off bond angles compared to experimental gas-phase electron diffraction data. The discrepancy was small—about two degrees on the key angles—but it mattered for the publication. Switching to an all-electron treatment for phosphorus and using a larger basis set closed the gap almost entirely. It cost me extra computation time, but it was the right call.
Where People Get Confused About Core Electrons
The biggest misconception I see is assuming that core electrons are static. They're not. Under extreme conditions—high pressure, intense fields, or when you're dealing with heavy elements where relativistic effects become significant—core orbitals actually contract or expand in ways that affect the valence shell. This isn't some fringe edge case. It matters if you're modeling anything involving gold, mercury, or the actinides. Relativistic contraction of the 1s and 2s orbitals in gold, for instance, is part of why gold is yellow and why mercury is liquid at room temperature. Those are bulk properties driven by core electron behavior. Another thing that trips people up is the assumption that the core/valence boundary is fixed. It's not. The choice of what counts as a core electron depends on your method and your basis set. In a typical Gaussian calculation on sulfur, you might treat 1s²2s²2p as core and keep 3s²3p as valence. But if you switch to a different software package with a different pseudopotential, that boundary can shift. Some treatments even include semi-core states—those 2s and 2p electrons in sulfur—in the valence space because they turn out to participate in bonding under certain conditions. This is especially common in solid-state physics codes where you're dealing with periodic systems. The decision has real consequences for accuracy and computational cost, and it's something you should be making consciously rather than accepting the default blindly. Here's a practical tip that isn't widely emphasized: always check what your effective core potential actually includes. When you specify, say, a LANL2DZ basis set for an iron complex, the ECP on iron replaces the 1s through 3p electrons. That's twenty-eight core electrons being approximated. But if you're studying a reaction where iron's oxidation state changes significantly, those replaced electrons aren't as passive as the name suggests. The ECP parameters are fitted to reproduce atomic properties, and while they work well for ground-state geometries, they can drift when you're looking at excited states or unusual oxidation states. I've seen papers where authors used a standard ECP for cerium in a +4 oxidation state compound and got reasonable geometries but completely wrong magnetic properties. Going all-electron with a specialized basis set for the lanthanide fixed it, but it took roughly three times longer to converge.
How to Decide What Counts as Core in Your Calculations
Start by identifying which atoms in your system are heavy enough that all-electron treatment is impractical. That's usually anything past calcium in the periodic table, though even scandium through zinc benefit from ECPs in larger basis sets. Then look at what your method supports. Some functionals and basis sets are specifically designed to work with particular ECPs, and mixing them arbitrarily introduces errors. Stick to published combinations. The Sapporo-DKH tables, for example, list which ECPs pair correctly with which basis sets for relativistic calculations. For lighter elements, run a test. Calculate your property of interest with the standard core/valence split and then recalculate including the semi-core electrons in the valence space. If the result changes by less than your tolerance—say, 0.01 eV for a reaction energy or 0.005 Å for a bond length—then the standard treatment is fine. If it changes more, you need the larger valence space. This typically adds maybe 20 to 40 percent to your computational time, but it saves you from publishing inaccurate results. I lost a week of work once because I trusted the default pseudopotential for selenium without checking, and the Se-O bond lengths were off by almost 0.03 Å compared to what the crystallographers measured. A two-hour test calculation would have caught it immediately.
Get the Full Details

Common Pitfalls That Waste Time
The most expensive mistake is realizing too late that your ECP isn't suitable for the property you're studying. ECPs are generally reliable for geometries and energies, but they struggle with properties that depend on electron density near the nucleus—NMR shielding constants, hyperfine coupling, isomer shifts in Mössbauer spectroscopy. If any of these are in your output requirements, you need an all-electron treatment or a specially designed ECP that includes those effects in its fitting. There's no workaround around that limitation. The physics simply isn't captured by a pseudopotential approximation. A second issue is consistency across your entire system. If you're using ECPs for the heavy atoms and all-electron basis sets for the light ones, make sure the treatment is balanced. I've seen cases where mixing a small-core ECP on bromine with a large all-electron basis set on hydrogen led to artificial polarization effects at the interface. The electron density got pushed around in ways that had nothing to do with the actual chemistry and everything to do with the mismatch in how the cores were treated. Running a convergence test on the basis set size for the hybrid region helped identify this, though it added another round of calculations to the timeline. There's also the matter of core polarization potentials. Standard ECPs don't account for the fact that the valence electrons can polarize the core electrons, creating an induced dipole that feeds back into the valence interaction. For most organic molecules this is negligible. For transition metal complexes with strongly donating or accepting ligands, it can shift energies by a few kilocalories per mole. There are core polarization potentials available for some ECP families, but they're not always the default and they add complexity. You have to decide whether the extra accuracy is worth the extra trouble for your specific system.