The Molecular Building Blocks You Actually Need to Know
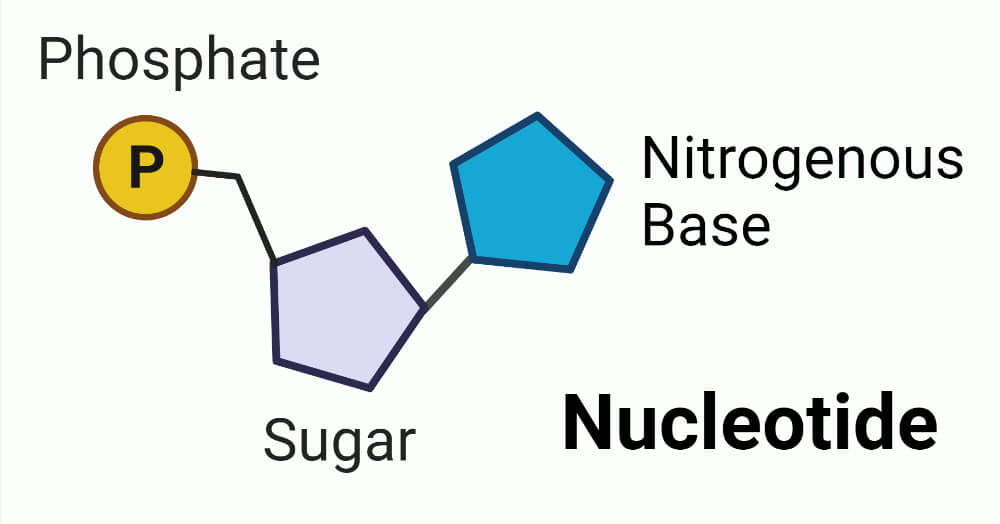
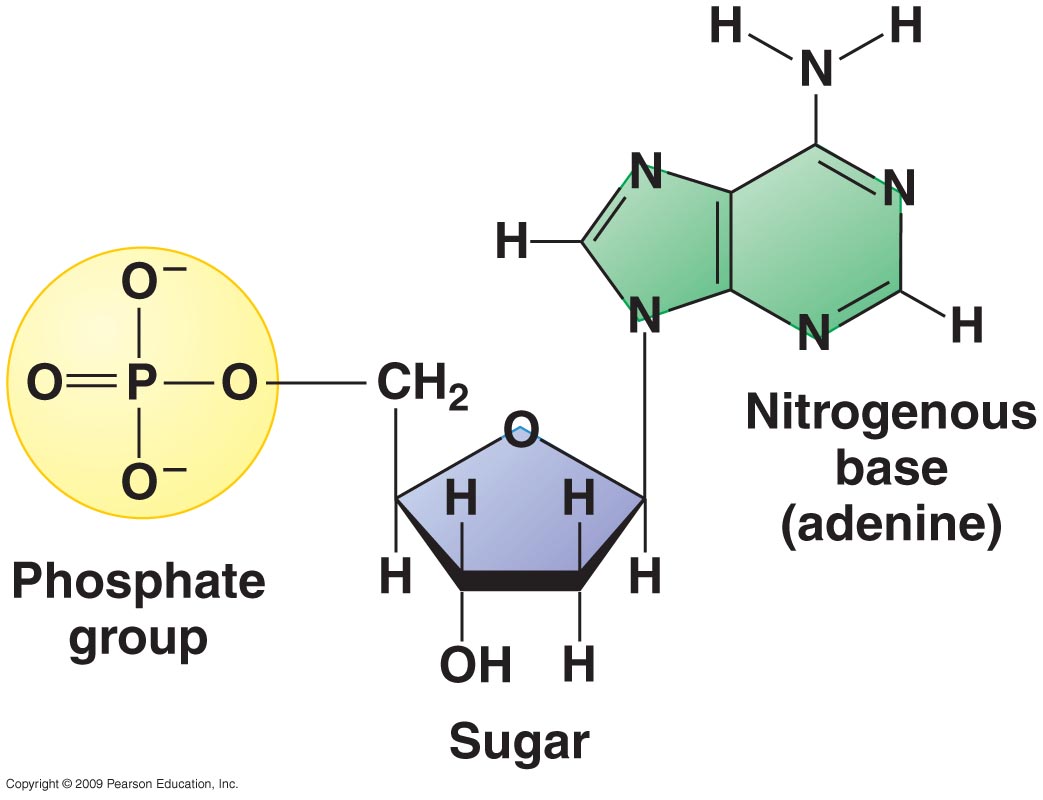
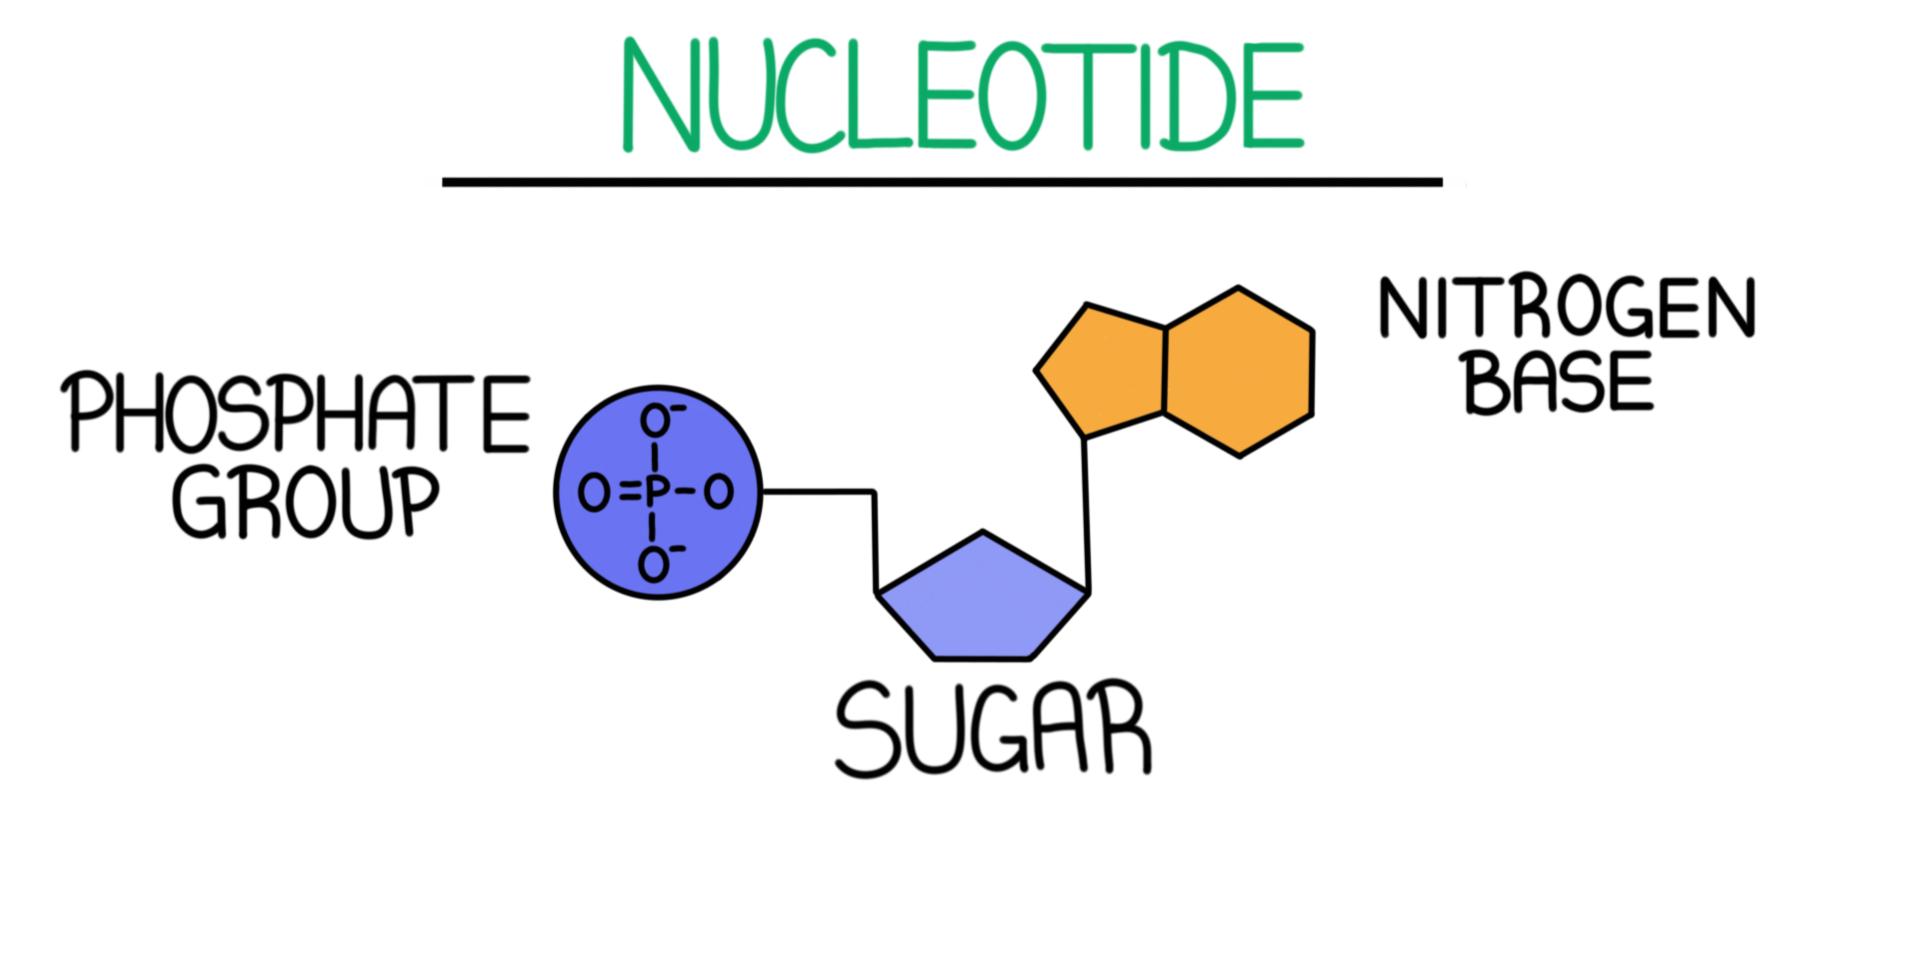
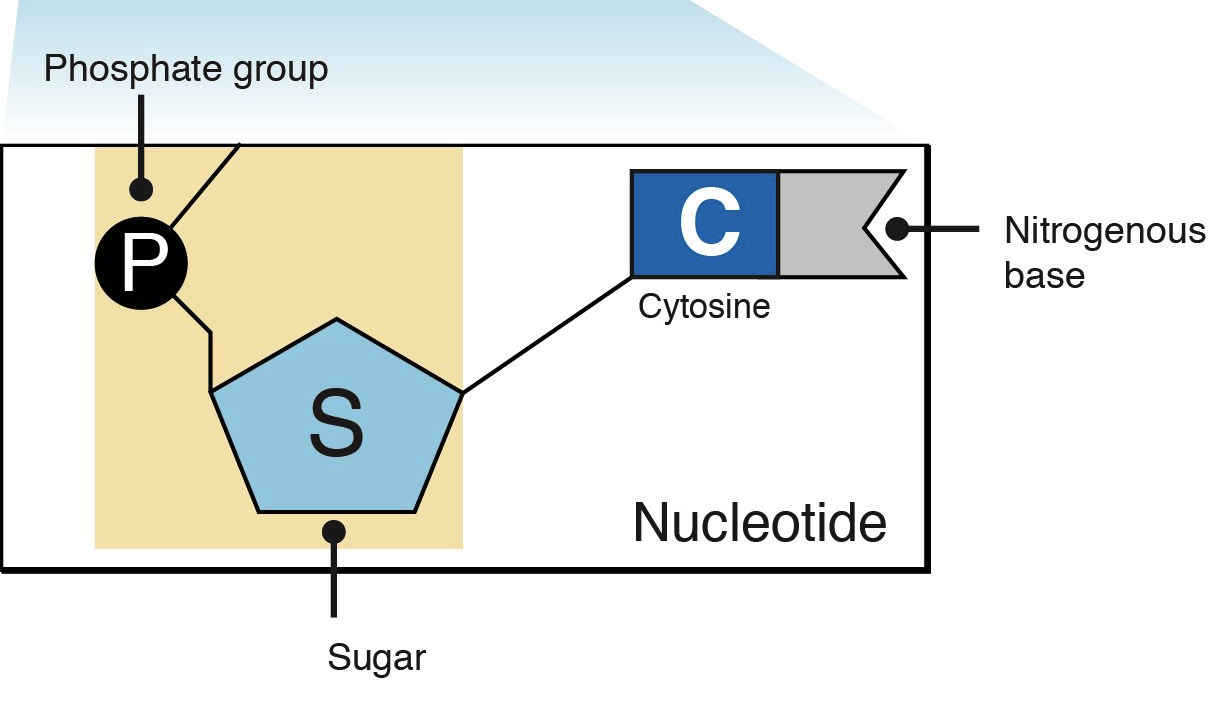
A nucleotide is a single molecular unit made of three parts: a sugar, a phosphate group, and a nitrogenous base. That's it. They link together through phosphodiester bonds to form nucleic acids like DNA and RNA. In a lab setting, you encounter them every day whether you're running a PCR, preparing sequencing libraries, or working with oligonucleotides. I've spent years in molecular biology labs dealing with nucleotide-related workflows, and the thing nobody tells you is that the distinction between nucleotides and nucleosides trips people up constantly. A nucleoside has the sugar and the base but no phosphate. Add one phosphate group and you've got a nucleotide. This matters because protocols specify one or the other and getting them mixed up will invalidate your experiment.
What Is A Nucleotide
The precise definition breaks down into the five standard bases: adenine, guanine, cytosine, thymine, and uracil. DNA uses A, G, C, and T. RNA swaps thymine for uracil. The sugar is deoxyribose in DNA and ribose in RNA. The phosphate backbone is what gives nucleic acids their directional polarity — 5' to 3' — which is fundamental to everything from replication to primer design. Here's a nuance most introductory courses skip over. The nucleotide isn't just a structural unit. It's also an energy currency. ATP is technically a nucleotide. GTP, UTP, and CTP all play roles in cellular metabolism beyond just being RNA building blocks. When you're doing in vitro transcription or working with RNA ligases, you're relying on those energy nucleotides, not just the informational ones. I once wasted two days troubleshooting a failed ligation before I realized the protocol called for NTPs, not dNTPs. The names look similar but they behave completely differently in enzymatic reactions.
How Nucleotides Function in Practice
In a typical molecular biology workflow, nucleotides show up in several forms. Free nucleotides come as powder and need to be reconstituted in the right buffer at specific concentrations. For PCR, you're working with dNTP mixtures usually at 10 millimolar each. The concentration matters because too much dNTP chelates magnesium and kills polymerase activity. Too little and you get incomplete extension or premature termination. Modified nucleotides are another layer. Fluorescently labeled dUTPs, biotinylated nucleotides, phosphorothioate bonds for antisense work — these are standard in many applications now. The catch is that polymerases handle modified nucleotides differently. Some work fine at low substitution rates. Others stall or drop off entirely. I ran into this with a custom sequencing probe design where the label position caused the polymerase to slip and generate a messy read. Moving the modification two bases down the chain fixed it completely.
Get the Full Details

Common Pitfalls with Nucleotide Work
The most practical issue people face is degradation. Nucleotides are relatively stable when stored properly — minus twenty degrees Celsius, protected from light, in the right buffer — but they degrade fast if you're careless. Freeze-thaw cycles are the main enemy. Each time you thaw a dNTP master mix, you lose quality. Aliquoting into single-use volumes at the start saves a enormous amount of headaches later. Another issue is concentration accuracy. Buying a 100 millimolar stock and diluting it down to working concentrations introduces error at every step. I found that preparing fresh dilutions from the lyophilized powder and verifying concentration spectrophotometrically before each major experiment cut down my failed reactions by about sixty percent. The extra twenty minutes per prep is worth it when you consider how much time you waste repeating bad runs.
Why This Matters Beyond the Textbook
Understanding nucleotides at a practical level changes how you approach experiments. It's not enough to know they're the letters of genetic code. You need to understand how modifications affect enzyme kinetics, how concentration impacts fidelity, and how storage conditions determine whether your reagents are still viable. The difference between a clean band on a gel and a smear often traces back to nucleotide quality or concentration. I've seen protocols that treat dNTPs as a commodity — add the standard volume and move on. That approach works until it doesn't, and then you're spending weeks troubleshooting something that came down to degraded reagents or an incorrect final concentration. Pay attention to the nucleotide details in your protocols. They're usually there for a reason.