The Reality of Common Ion Effects in Practice
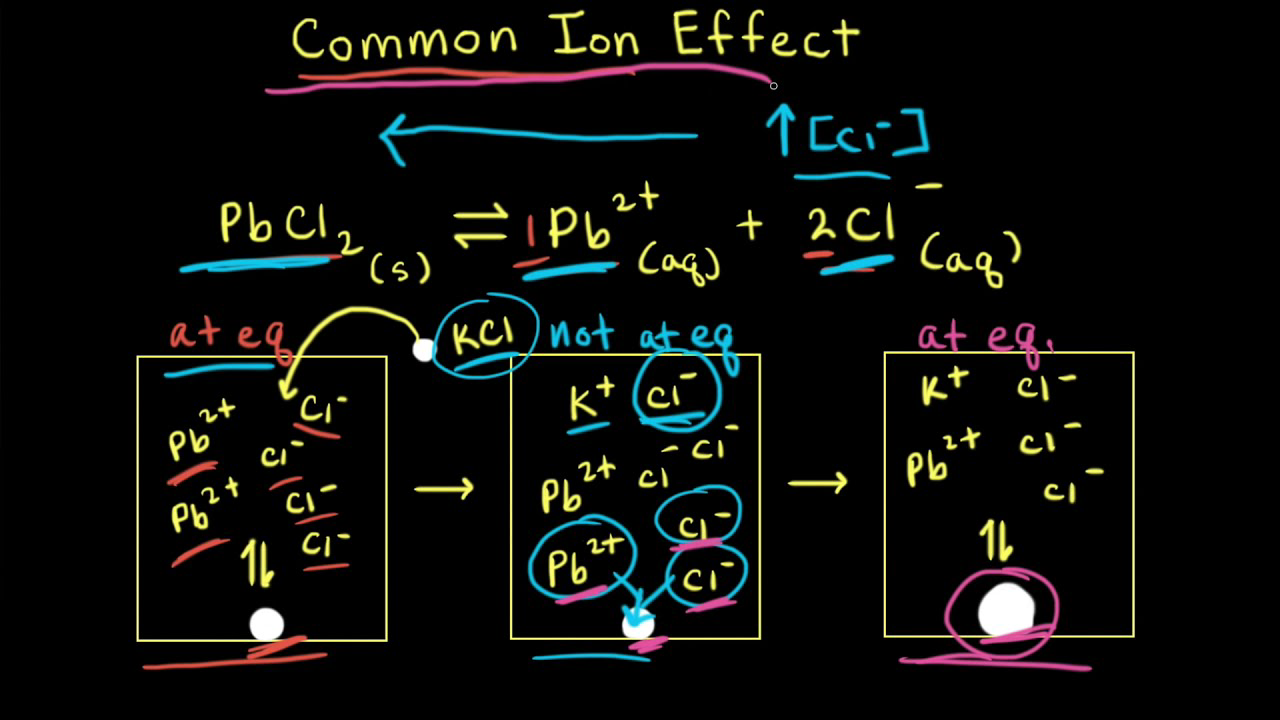
The common ion effect is what happens when you add an ion to a solution that is already present from another dissolved compound, and it pushes the equilibrium in a predictable direction. This matters because most people learning solubility for the first time only see textbook problems where numbers work out perfectly. Real solutions do not cooperate like that. I spent too many years watching graduate students get tripped up by this because they treated the equilibrium as if it were static. When you dissolve a sparingly soluble salt like AgCl in water, it establishes an equilibrium between the solid and its ions. Add more chloride from NaCl, and the system shifts left. That is the basic mechanism. But here is what the textbooks gloss over: the effect depends entirely on activity coefficients, ionic strength, and whether your added salt shares that common ion at a concentration that actually matters relative to the salt's Ksp.
What Is Common Ion
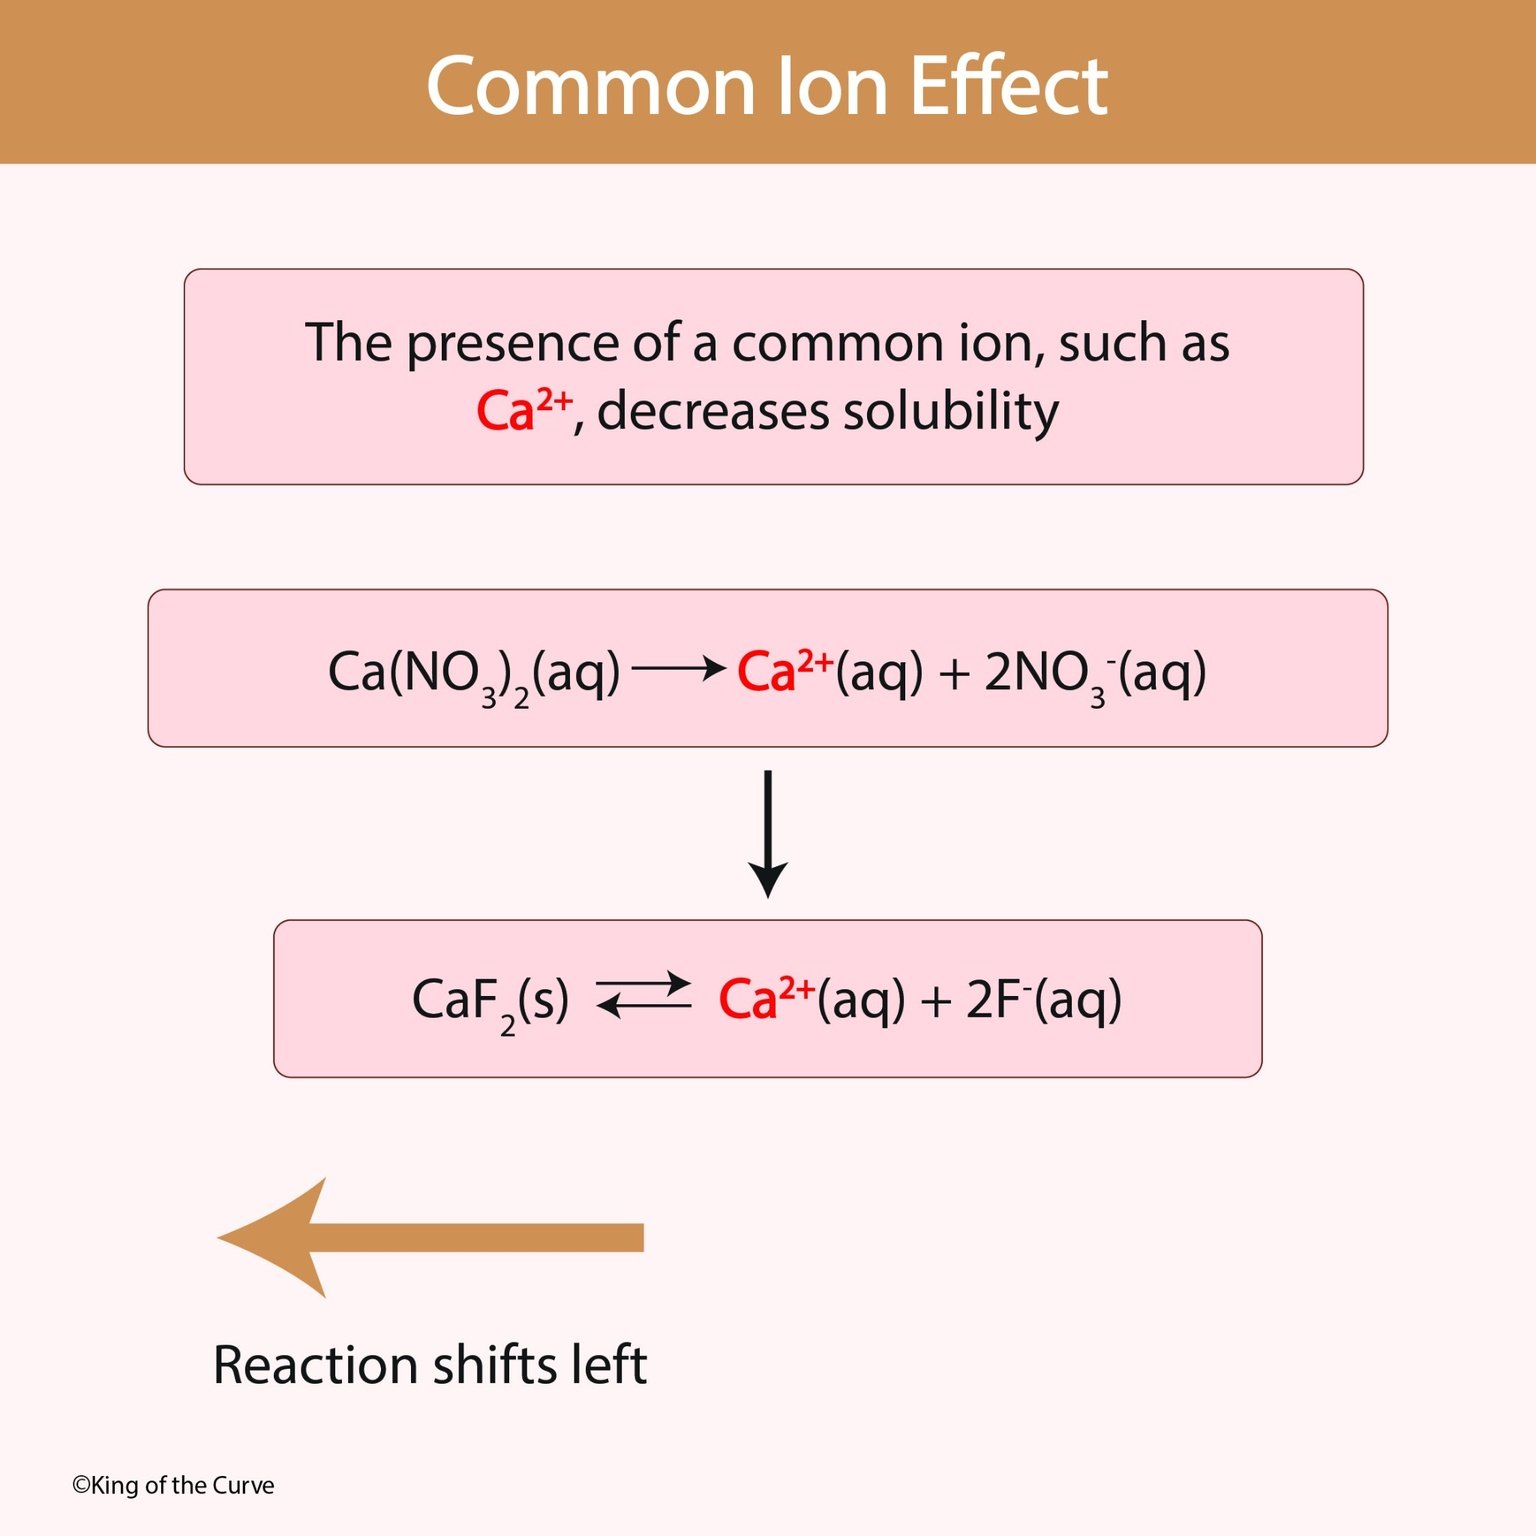
The common ion effect describes the reduction in solubility of a sparingly soluble salt when a soluble compound containing the same ion is introduced to the solution. The principle comes straight from Le Chatelier's principle, but applying it correctly requires more than just plugging numbers into the Ksp expression. You need to account for how the total ionic strength changes the effective concentrations of the species involved. Let me give you a scenario I actually dealt with. I was working on a precipitation purification step for a pharmaceutical intermediate, and the protocol called for adding sodium sulfate to a calcium chloride solution to precipitate CaSO4. The Ksp of CaSO4 is about 4.93 × 10^–5 at 25 °C. Simple enough. But the process stream already contained significant concentrations of sodium and chloride from upstream steps, and the ionic strength was pushing past 0.5 M. When I calculated solubility using simple concentrations, the predicted precipitate yield was nowhere near what actually came out of the reactor. The measured solubility was roughly three times higher than the textbook value. The workaround was straightforward once I stopped ignoring activities. I switched to using the Davies equation to estimate activity coefficients at the relevant ionic strength, recalculated the effective solubility product, and adjusted the sodium sulfate feed rate accordingly. This brought the predicted precipitation yield within 5 percent of the actual measured value. Doing this manually was tedious, so I built a quick spreadsheet that iterated on the ionic strength and recalculated gamma values until the Ksp condition converged. It took me about forty minutes to set up, and it has saved me from repeated failures since then.
The thing beginners consistently miss is that the common ion effect only suppresses solubility when the common ion is truly dominant. If you are adding a tiny amount of the common ion relative to the solubility of the salt itself, you may see almost no shift at all. I once saw a student add 0.001 M chloride to a saturated AgCl solution and then claim the effect was "not strong enough" to measure. The problem was not the method. The concentration of chloride they added was smaller than the chloride already contributed by the dissolved AgCl. They were trying to measure a signal smaller than the noise floor. Another counter-intuitive point that trips people up is the case where adding a common ion can actually increase apparent solubility through complex ion formation. Silver chloride in concentrated NaCl solutions does something you would not expect from the basic Ksp model. The excess chloride forms soluble complexes like AgCl2^– and AgCl3^2–, which pulls the dissolution equilibrium forward. So at low chloride concentrations, adding NaCl decreases solubility as expected. At high chloride concentrations, typically above about 0.1 M for AgCl, the solubility starts climbing again. If you are working with silver salts or other metals that form stable halide complexes, ignoring this behavior will give you wrong results every time. Here is how you approach a typical calculation correctly. Start by writing the dissolution equilibrium and the Ksp expression. Identify all sources of each ion in the solution, not just the sparingly soluble salt. Then set up the mass balance and solve for the unknown solubility. If the ionic strength is above about 0.01 M, apply an activity correction. The Debye-Hückel limiting law works for very dilute solutions, but once you go past 0.01 M it breaks down. Use the extended Debye-Hückel equation or the Davies equation instead. The Davies equation is less parameter-heavy and stays reasonable up to about 0.5 M ionic strength. Beyond that, you are into territory where you need specific ion-interaction data, and you should stop doing hand calculations.
Get the Full Details

Let me walk through a concrete example. Consider BaSO4 with a Ksp of 1.08 × 10^–10. You want to find its solubility in a solution that already contains 0.01 M Na2SO4. The sulfate from the sodium sulfate is the common ion. The dissolution equilibrium is BaSO4(s) Ba^2+ + SO4^2–. The Ksp expression is [Ba^2+][SO4^2–] = 1.08 × 10^–10. If S is the solubility of BaSO4, then [Ba^2+] = S and [SO4^2–] = 0.01 + S. Because S will be very small compared to 0.01, you can approximate [SO4^2–] 0.01. Solving gives S 1.08 × 10^–8 M. That is roughly ten thousand times less soluble than in pure water, where S would be about 1.04 × 10^–5 M. Now check whether the approximation holds. S is indeed much smaller than 0.01, so the approximation is valid. The ionic strength from 0.01 M Na2SO4 is 0.03 M, which means activity coefficients might matter a bit. Using the Davies equation, the mean activity coefficient for a 2:2 electrolyte at I = 0.03 M comes out to roughly 0.44. Applying this correction effectively reduces the apparent Ksp, and the solubility shifts slightly from 1.08 × 10^–8 M to about 5.8 × 10^–9 M. Not a dramatic change for this particular case, but noticeable enough that quality labs will include it. There are several situations where the common ion effect either fails or gives misleading predictions. One is when the added salt is itself sparingly soluble and contributes to the common ion in a way that creates a mixed solid solution rather than a simple precipitation. Another is when pH changes alter the speciation of the anion, as happens with salts of weak acids like CaF2 or AgCN. In those cases, adding a common ion does not tell the whole story because the anion is simultaneously undergoing protonation. You need to solve the full set of equilibria, including Ka and the solubility product, and the math gets complicated fast.
The biggest practical limitation is that the common ion effect assumes complete dissociation of the added salt and no complex formation. Real solutions violate both assumptions at higher concentrations. If you are working in a lab setting and your calculated solubility is off by more than 20 percent from what you measure, the most likely culprits are unaccounted ionic strength effects, complex ion formation, or pH-dependent speciation. Check those before blaming experimental error. For quick reference or to use in the field, the approach is the same regardless of what you are looking up online. What Is Common Ion remains a foundational concept, but the actual work happens in the details. Start with the equilibrium expression, include every source of every ion, correct for activity when ionic strength is above 0.01 M, and watch out for complex formation at higher ligand concentrations. The math is not hard. The mistakes come from skipping steps that the problem setup makes you think are optional.