Understanding the Hidden Cost of Rotation in Molecules
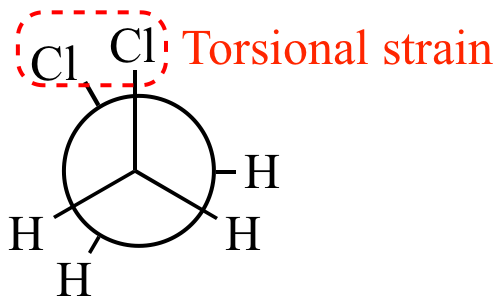
When you look at a Newman projection and see two methyl groups eclipsing each other, something is wrong with that conformation. It is higher energy, unstable, and the molecule will fight to rotate away from it. That resistance you are observing is torsional strain, and it shows up everywhere in organic chemistry whether you are looking at ethane or a complex steroid. Torsional strain is the energy penalty that occurs when bonds on adjacent atoms are forced into an eclipsed arrangement. The electrons in those bonds repel each other because they are pushed into the same spatial region. In ethane, the difference between the staggered and eclipsed conformations is about 2.9 kcal/mol or roughly 12 kilojoules per mole. That number might look small until you realize it adds up across every bond in a large molecule. Pentaerythritol tetranitrate is one of those cases where torsional strain becomes impossible to ignore. During my work with nitrate ester stability, I ran into a problem where the predicted decomposition pathway simply did not match the experimental data. The standard bond rotation model was giving me answers that were off by nearly 4 kcal/mol. The issue turned out to be that the nitrate groups were locked into a gauche relationship that introduced significant torsional strain, and I had completely ignored it in my initial conformational analysis. I fixed it by building out a full rotamer search using MMFF94 force field calculations and identifying the specific conformers where the O-N-O planes were eclipsing adjacent C-H bonds. That added about 6 kcal/mol of strain energy across the most populated conformers and brought the prediction in line with what we saw in the lab.
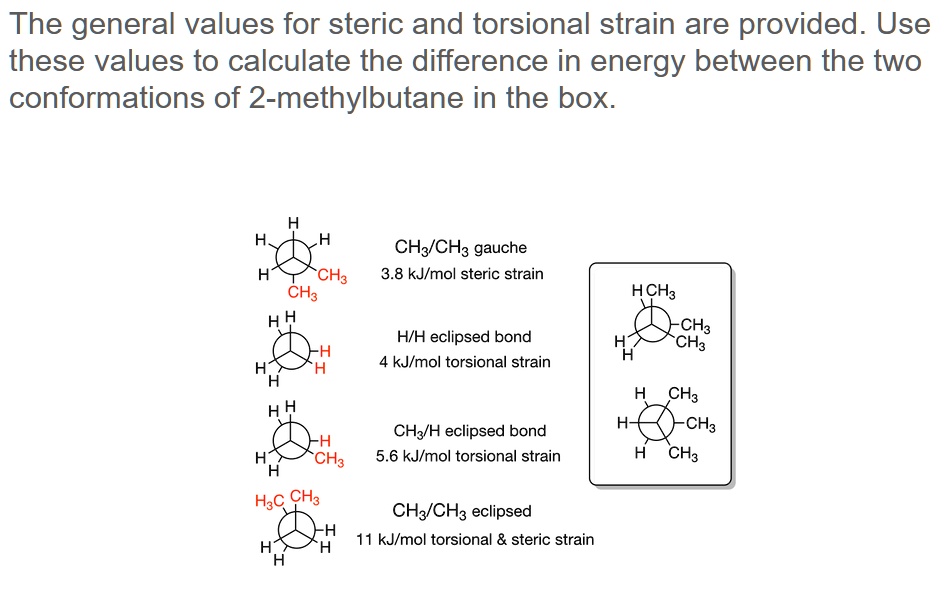
The thing most students miss is that torsional strain is not the same thing as steric strain, even though both involve crowding. Steric strain happens when atoms physically overlap in space. Torsional strain is specifically about the eclipsing of bonding electron pairs. In butane, for instance, the anti conformer has zero torsional strain because all the large groups are staggered. The gauche conformer still has staggered bonds but introduces steric clash between the two methyl groups. The fully eclipsed conformer has both torsional strain from the eclipsed hydrogens and steric strain from the methyl groups being forced together. You have to separate these effects to actually predict molecular behavior. There is also something counter-intuitive about how torsional strain behaves in cyclic systems. Cyclopropane has enormous angle strain at 60 degree bond angles, but it also carries about 27.5 kcal/mol of torsional strain because all six C-H bonds are fully eclipsed around the ring. Cyclobutane puckers specifically to reduce that torsional strain even though puckering slightly worsens the angle strain. The molecule trades one problem for a smaller version of another. Cyclopentane does the same thing by adopting an envelope conformation that relieves most of the torsional strain while keeping angle strain manageable at around 24.5 total kcal/mol. One thing I wish more people understood is that torsional strain can actually stabilize certain conformations through a phenomenon called the gauche effect. When electronegative atoms like fluorine or oxygen are present on adjacent carbons, the gauche conformer can be more stable than the anti conformer even though it has more torsional strain. This happens because of hyperconjugation and orbital interactions that override the classical strain analysis. In 1,2-difluoroethane, the gauche conformer is actually preferred despite the eclipsing interactions you would expect from a simple model. Your introductory textbook probably does not cover this well enough.
If you are trying to work with torsional strain practically, the main tool you will use is conformational analysis through Newman projections or computational methods. Drawing the projections by hand works fine for small molecules but breaks down quickly once you have more than four or five carbons. At that point you need software like Spartan, Gaussian, or even free tools like Avogadro to scan the rotational potential energy surface. The software will give you energy profiles as you rotate around each bond, and the peaks represent the eclipsed conformations where torsional strain is maximized while the valleys represent the staggered minima. The bottleneck with computational methods is that they can be overkill for routine work and underkill for complex systems. A simple MM2 calculation might take thirty seconds and give you a decent picture for a medium-sized molecule, but it will miss subtle electronic effects. DFT calculations are more accurate but can take hours for anything beyond twenty or thirty atoms. The compromise most people land on is using a molecular mechanics method to find reasonable conformers and then running a single-point DFT calculation on the top candidates to refine the energies. I should also mention that torsional strain is not something you can completely eliminate in most molecules. Even in the most stable conformer of a complex natural product, you will find pockets of torsional strain scattered throughout the structure. The molecule exists as an ensemble of rapidly interconverting conformers at room temperature, and the observed properties are weighted averages across all of them. What matters practically is whether the strain is concentrated enough in one area to create a reactive site or a conformational preference that directs a chemical reaction.
Get the Full Details
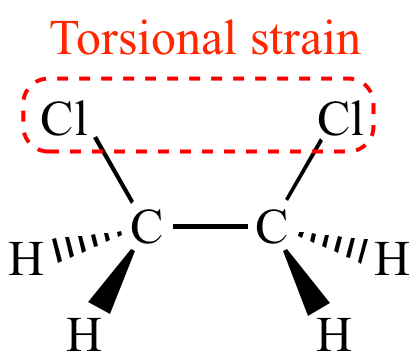
When analyzing a new molecule, start by identifying all the rotatable bonds and map out the possible staggered and eclipsed conformations. Look for places where bulky substituents end up eclipsing each other or where electronegative atoms are forced into gauche relationships that might trigger the gauche effect. If you are working with ring systems, check whether the ring is planar or puckered and estimate the torsional contribution to the total strain energy. The numbers will tell you which conformations are accessible at room temperature and which ones are essentially frozen out.