How Viral Replication Actually Works
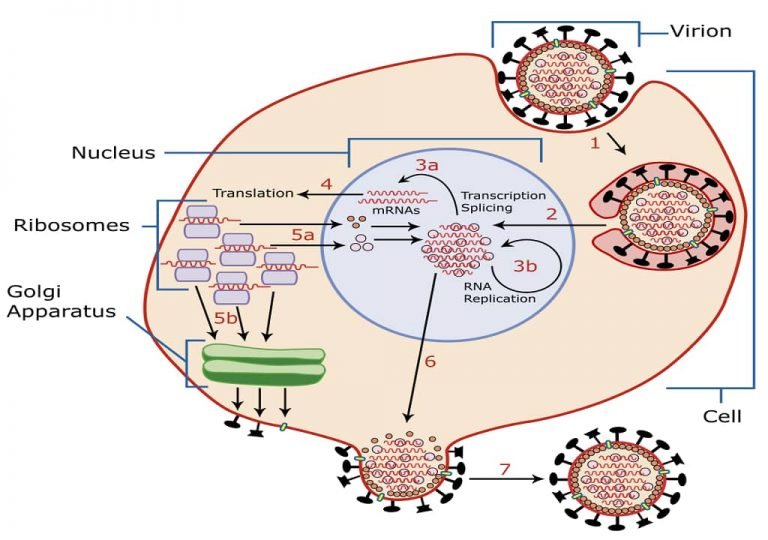
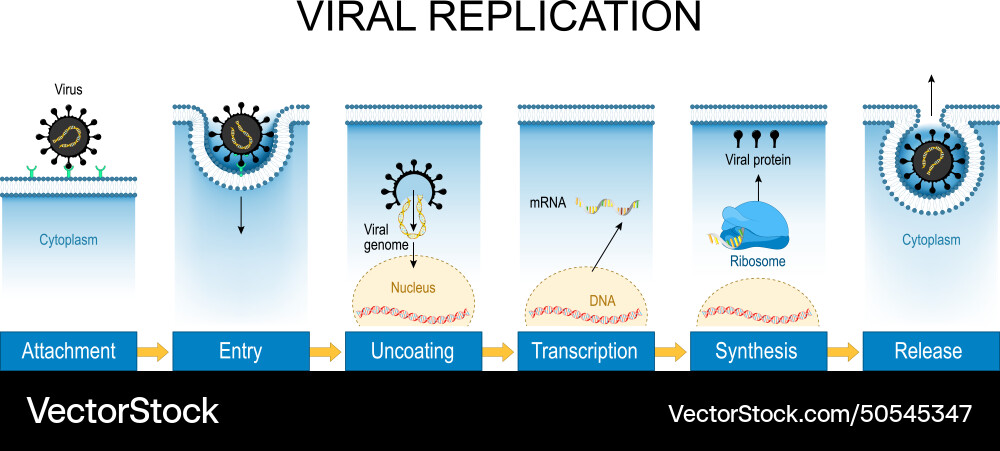
Viruses don't reproduce the way bacteria or any living cell does. They can't split in two, build proteins on their own ribosomes, or generate energy. A virus is essentially a genetic payload wrapped in protein, and sometimes a lipid shell. It exists in two states: outside a host cell as an inert particle (a virion), and inside a host cell where it hijacks cellular machinery to make copies of itself. Understanding this distinction matters because most people conflate viral replication with biological reproduction. The basic replication cycle runs through a few stages: attachment, entry, uncoating, replication, assembly, and release. Attachment means a viral surface protein binds to a specific receptor on the host cell membrane. Entry gets the virion inside. Uncoating releases the viral genome. Replication copies the genetic material and produces viral proteins. Assembly puts new virions together. Release lets them exit the cell, usually by budding or lysing it. The specifics vary dramatically depending on what kind of virus you are dealing with. DNA viruses and RNA viruses take fundamentally different paths once they get inside the cell. Retroviruses reverse transcribe their RNA into DNA before integrating into the host genome. This is where people commonly get confused and assume all viruses replicate the same way.
I spent weeks troubleshooting a persistent lab contamination issue years ago. We kept finding unexpected RNA virus particles in cell culture lines that were supposedly virus-free. The problem traced back to a specific strain of vesicular stomatitis virus that was budding from the cell surface at extremely low levels without causing visible cytopathic effect. It was replicating, assembling, and releasing virions continuously but the cells looked healthy enough to keep growing. We solved it by adding a neutralizing antibody directly to the culture medium, which bound free virions before they could re-infect neighboring cells. The takeaway: not every active infection causes obvious cell death, and low-level continuous replication can go unnoticed for a long time if you are not specifically looking for it. Attachment and entry are governed by molecular compatibility. The virus needs the right receptor. HIV targets CD4 receptors on helper T cells. Influenza binds to sialic acid residues on respiratory epithelial cells. Hepatitis C targets the CD81 receptor on hepatocytes. If a cell lacks the receptor, the virus cannot enter regardless of how many virions are floating around. This is also why tissue tropism exists. Some viruses can only infect certain organs or cell types. Once inside, uncoating exposes the viral genome. For many enveloped viruses, this happens during entry when the lipid envelope fuses with the cell membrane or gets degraded in the endosome. For non-enveloped viruses, uncoating often requires proteolytic cleavage or changes in pH inside endosomal compartments. The genome then either goes straight to the nucleus (most DNA viruses) or stays in the cytoplasm (most RNA viruses).
Replication strategy is where things diverge the most. Positive-sense single-stranded RNA viruses like poliovirus or SARS-CoV-2 can essentially function as messenger RNA the moment they are uncoated. Host ribosomes translate the viral genome directly into protein. The viral RNA-dependent RNA polymerase then copies the genome. Negative-sense RNA viruses like influenza carry their own polymerase inside the virion because host ribosomes cannot read negative-sense RNA. They must transcribe their genome into positive-sense mRNA first. DNA viruses generally replicate in the nucleus where host DNA-dependent RNA polymerase and other transcription machinery are available. Herpesviruses are an exception. They bring their own enzymes and replicate their DNA in the nucleus using a rolling circle mechanism that produces long concatemers, which then get cleaved into individual genomes during packaging. I have seen people miss this detail in exams because they blanket-group all DNA viruses together. Retroviruses like HIV add another layer. Their single-stranded RNA genome is reverse transcribed into double-stranded DNA by the viral enzyme reverse transcriptase. This DNA enters the nucleus and gets integrated into the host chromosome by integrase. From there, host RNA polymerase II transcribes the proviral DNA into new viral RNA genomes and mRNA. The integrated provirus can remain dormant for years. This latent reservoir is why HIV is currently incurable with standard antiretroviral therapy alone. The drugs suppress active replication but do not touch integrated proviral DNA sitting quietly in long-lived memory T cells.
Get the Full Details
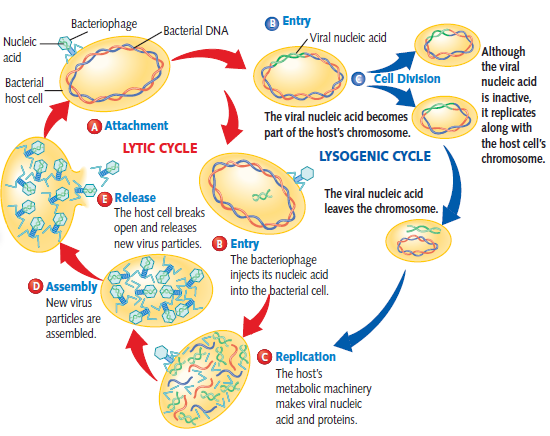
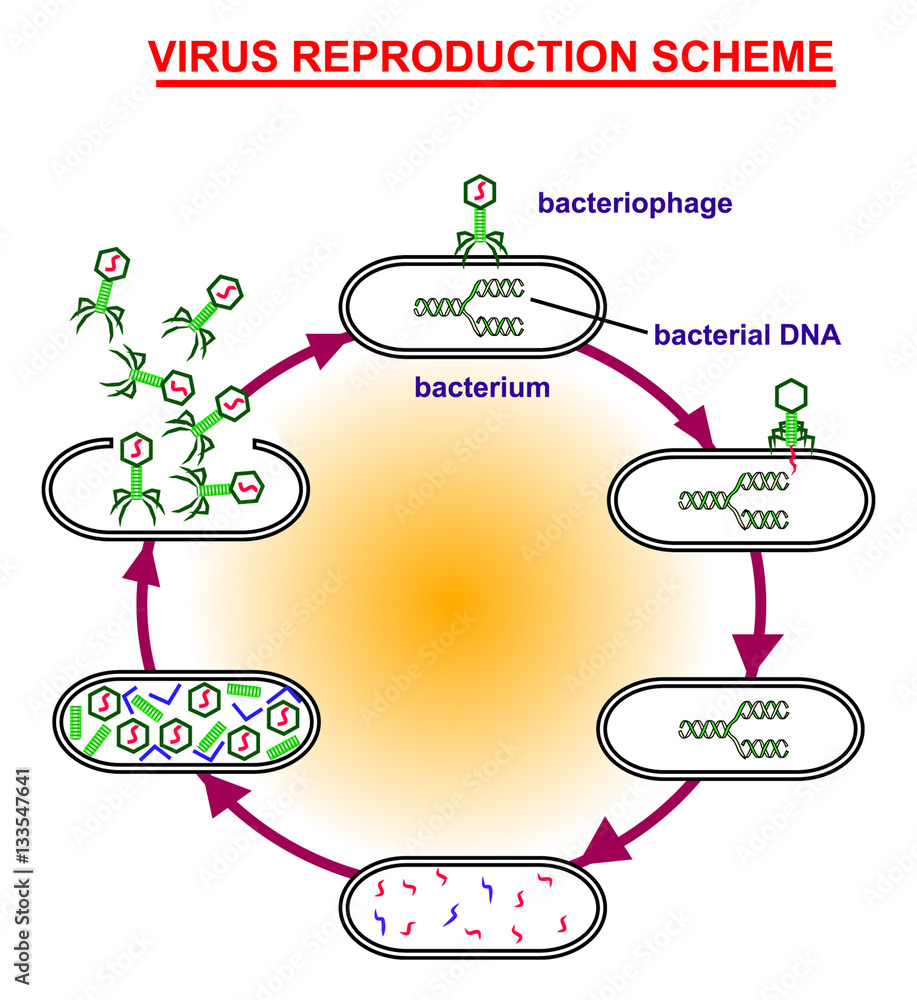
Assembly and release follow replication. New virions assemble from replicated genomes and newly synthesized structural proteins. Enveloped viruses typically acquire their lipid membrane by budding through cellular membranes such as the plasma membrane, Golgi, or endoplasmic reticulum. The viral glycoproteins are embedded in these membranes before budding occurs. Non-enveloped viruses often accumulate in the cytoplasm or nucleus and are released when the cell lyses, though some can exit through non-lytic pathways like exosomes or membrane blebbing. The burst size per cell varies widely. A single influenza-infected cell might produce thousands of virions. A retrovirus might produce hundreds. Bacteriophages, which infect bacteria rather than human cells, can produce tens of thousands per cell through the lytic cycle. These numbers depend heavily on multiplicity of infection, cell health, and whether the host has mounted an effective interferon response. One counter-intuitive point that rarely gets emphasized: not all replication is productive replication. Defective interfering particles are viral genomes with large deletions that remove essential genes. They cannot replicate on their own but can hijack the replication machinery of a co-infecting standard virus. They are shorter and replicate faster, which lets them outcompete the full-length virus over serial passages. This is a real phenomenon observed with influenza and vesicular stomatitis virus in lab settings. It also complicates vaccine production because defective particles can dominate a seed virus stock and reduce yield significantly.
Another common pitfall is assuming a virus is dead after a short time at room temperature. Many enveloped viruses lose infectivity within hours on dry surfaces because the lipid envelope desiccates and the surface proteins denature. Non-enveloped viruses like norovirus and adenovirus remain infectious on surfaces for days. This difference matters enormously for infection control protocols but gets glossed over in most general explanations. The limits of viral replication are set by host range, receptor availability, intracellular antiviral defenses, and immune surveillance. Species barriers exist because receptors differ between animals. Within a single host, restriction factors like APOBEC3G and TRIM5 can degrade or block viral replication at the post-entry stage. Interferons induce hundreds of antiviral genes that collectively create an intracellular environment hostile to replication. No virus has evolved to completely evade all of these mechanisms, which is why most acute infections resolve on their own. Replication kinetics matter clinically. The eclipse period, where no infectious virions can be detected inside the cell, lasts from several hours to over a day depending on the virus. After that, virion production ramps up until cell death or immune clearance stops it. The entire cycle from attachment to release typically takes anywhere from 6 hours for fast-replicating RNA viruses to several days for slower DNA viruses like herpes simplex. Knowing where in this timeline a diagnostic test is most sensitive can mean the difference between a false negative and a confirmed result. Testing during the eclipse period will miss the infection even if the patient is clearly symptomatic shortly after.