Writing the Configuration Without Guessing
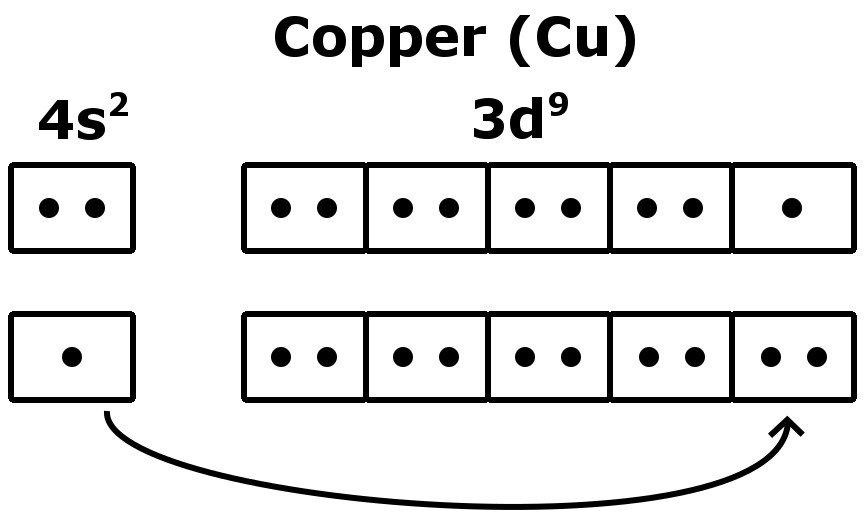
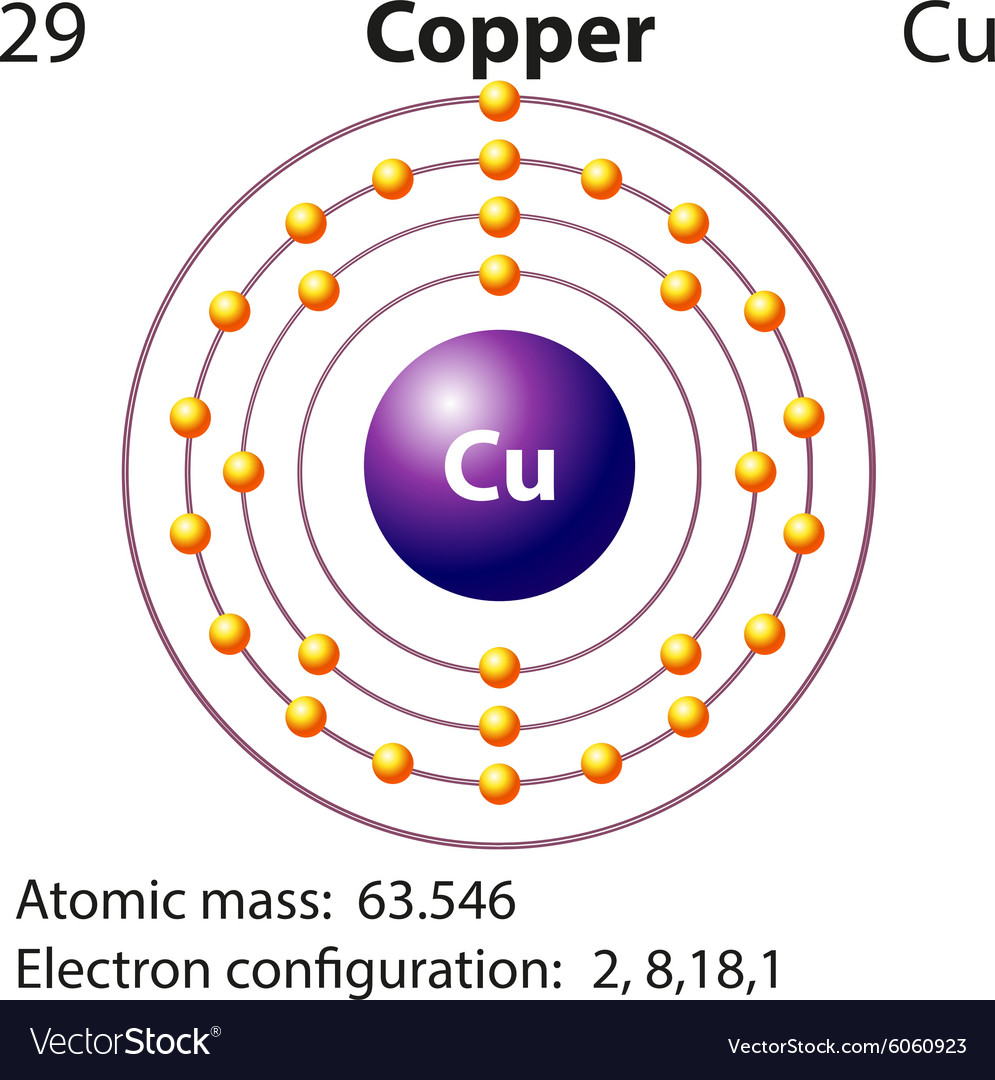
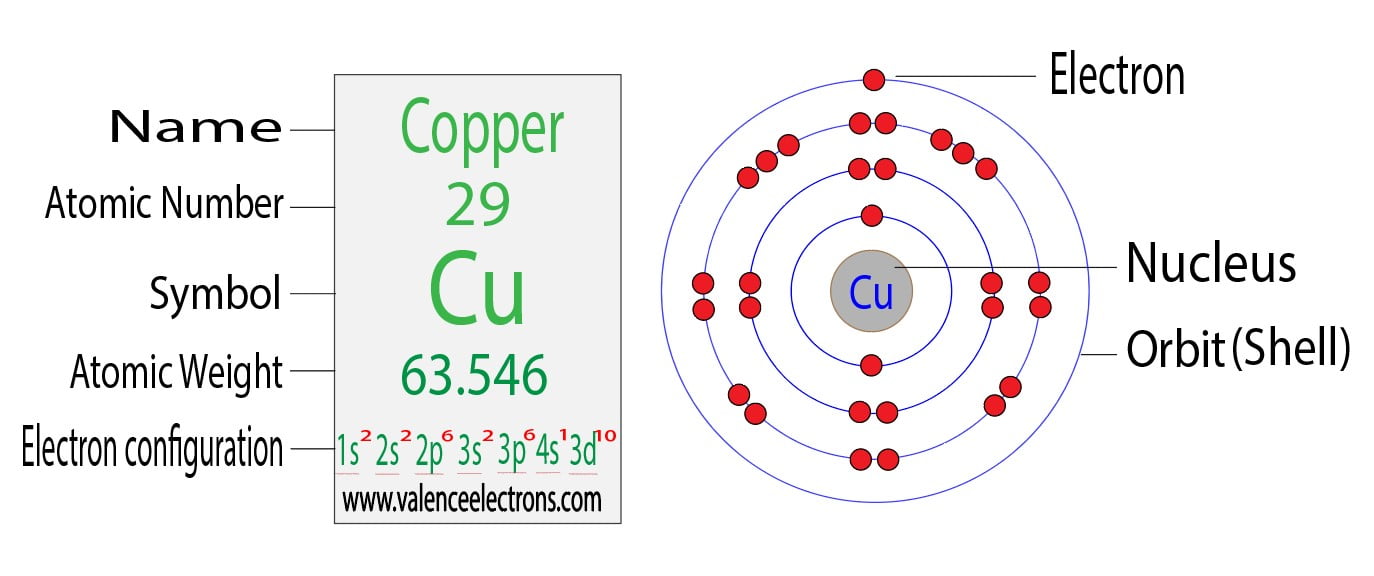
Most people memorize the copper configuration as just another exception to the Aufbau principle, which gets them through homework and then immediately forgets it. I ran into a real problem last year when I was calibrating XPS spectra for a thin-film copper sample on an oxide substrate. The binding energy shifts I was seeing didn't match standard reference data, and it took me about three weeks to realize I had been modeling the wrong ground state in my fitting software. The config matters more than you think when you are actually measuring electronic structure rather than just writing it on a whiteboard.The shorthand is [Ar] 3d10 4s1. The full version is 1s2 2s2 2p6 3s2 3p6 3d10 4s1. That looks wrong if you are following the standard n+l rule strictly, because 4s fills before 3d in most textbook diagrams, so you would expect [Ar] 4s2 3d9. It does not happen that way. The actual ground state rearranges because a completely filled d-subshell is lower in energy than a nearly filled one, even though it means moving an electron out of the 4s orbital. Under normal filling rules, you add electrons in order of increasing n+l value, which puts 4s below 3d. So writing out copper with atomic number 29, you would fill 1s2, 2s2, 2p6, 3s2, 3p6, then 4s2, then 3d9. That gives you [Ar] 4s2 3d9, which is the prediction most students write down and most automated tools default to unless they have exception tables built in. The real energy levels shift once you actually account for electron-electron repulsion and exchange energy within the d-subshell. A filled d10 configuration gains roughly 3 to 4 eV of stabilization from exchange effects alone, and that stabilization outweighs the small energy cost of promoting one electron from 4s to 3d. The 4s orbital also contracts less predictably than the d orbitals, so the energy gap between 4s and 3d narrows significantly by the time you reach copper. I learned this the hard way when a colleague handed me a spectral simulation and I noticed the density of states plot had a peak at the Fermi level that was too broad for metallic copper. I checked the configuration file he had generated, and it was using the predicted [Ar] 4s2 3d9 instead of the actual [Ar] 3d10 4s1. That single electron misplacement changed the entire projected density of states near the d-band edge. We had to regenerate the pseudopotential with the correct occupation, which took about two days on our cluster because we were recalculating the radial part of the wavefunction for each k-point. It is the kind of mistake that does not show up in a multiple choice question but costs real time in practice.
How to Get the Correct Result Without Looking It Up
You can derive the copper configuration without relying on memorization by checking the energy gap between 4s and 3d for elements around Z = 29. Once the d-subshell is more than half filled, the exchange energy stabilization from completing it starts to dominate over the simple Aufbau ordering. This applies to chromium too, which is [Ar] 4s1 3d5 instead of [Ar] 4s2 3d4, and to silver, which follows the same pattern down the group with [Kr] 5s1 4d10. The rule of thumb is that whenever a d-orbital can go from d4 to d5 or d9 to d10, the promotion usually happens. If you are using computational chemistry software and need to set the occupation manually, make sure your input file specifies the correct ground state multiplicity. Copper in its ground state has one unpaired electron in the 4s orbital, so the spin multiplicity is 2, not 1. I have seen this error repeatedly in DFT input files where users set a closed shell singlet because the program defaults to it, and the self-consistent field cycle then converges to an excited state or fails to converge entirely. The workaround is to force an unrestricted calculation or set the initial occupation explicitly in the keyword file.
Pitfalls and Where This Breaks Down
The electron configuration for copper is straightforward on paper but becomes problematic when you move beyond isolated atoms. In a metallic lattice, the 4s and 3d bands overlap significantly, and the concept of a single localized configuration loses meaning. Band structure calculations do not use discrete orbital occupations in the same way, so writing [Ar] 3d10 4s1 for bulk copper is more of a shorthand than a physically rigorous statement. If you are working with bulk properties like conductivity or work function, the atomic configuration is a starting point at best. Another issue comes up with oxidation states. Cu+ has the configuration [Ar] 3d10, and Cu2+ has [Ar] 3d9. People often assume Cu+ is more stable because the d-shell is full, but in aqueous solution Cu2+ is actually the dominant species due to hydration energy compensating for the second ionization potential. The configuration alone does not tell you that, and relying on it to predict chemical behavior will get you wrong answers fairly often. If you need the configuration for spectroscopic fitting or theoretical modeling, the best approach is to use a database like NIST Atomic Spectra instead of deriving it manually. NIST lists the confirmed ground state as 3d10 4s1 with term symbol 2S1/2, and they include the excited configurations if you need those as well. Their data is experimentally verified through laser spectroscopy and discharge lamp measurements, so it is more reliable than any automated generator that does not have a built-in exception table.
Get the Full Details
Common Mistakes in Lab Reports
I grade a lot of undergraduate lab reports, and the recurring errors are surprisingly consistent. Students will write the correct final answer but show the wrong reasoning, usually applying the Aufbau diagram mechanically and then fudging the result because they remember it is an exception. Another common error is flipping the order and writing [Ar] 4s1 3d10, which has the same electron count but implies a different energy ordering than what is actually observed in photoelectron spectroscopy. The 3d electrons are more tightly bound than the 4s electron, so the notation [Ar] 3d10 4s1 reflects the actual binding energy sequence, not just the filling order. A third mistake is confusing the configuration with the oxidation state behavior. Writing the ground state configuration and then predicting that copper should form only +1 ions because of the filled d-shell is a logical leap that does not hold up experimentally. You can note the configuration accurately and still get the chemistry wrong if you treat it as a deterministic rule rather than one factor among many. The electron configuration for copper as an atomic property is well established and not controversial, but the moment you apply it to anything beyond an isolated atom in a vacuum, the simple picture breaks down. That is not a failure of the concept, it is just a reminder that atomic orbitals are a model, not the thing itself. The configuration [Ar] 3d10 4s1 is the best single-determinant description we have, and it is useful as long as you know where its limits are.