Working with Thiol Ene Click Chemistry in Practice
I spent about three years trying to make this work consistently in my lab before I stopped treating it like magic and started treating it like what it actually is: a radical chain reaction that is easy to start, hard to stop, and extremely sensitive to everything in your reaction mixture. Most protocol papers make it look straightforward because they only report the conditions that worked. They don't tell you about the batches that gelled in the syringe or the ones that gave you 40 percent conversion because someone left the nitrogen line connected for too long and scrubbed all the oxygen out. The reaction itself is a photoinitiated or thermally initiated addition of a thiol across a vinyl double bond. You mix a thiol, an alkene, and an initiator, expose it to light or heat, and you get a thioether linkage. That's the whole thing. It's called "click" chemistry because the byproduct profile is essentially zero and the kinetics are fast under the right conditions. But "fast" here means seconds to minutes under UV at high intensity, not instant. Under ambient light or with a cheap LED array, expect ten to twenty minutes depending on your chromophore and concentration.
Thiol Ene Click Chemistry
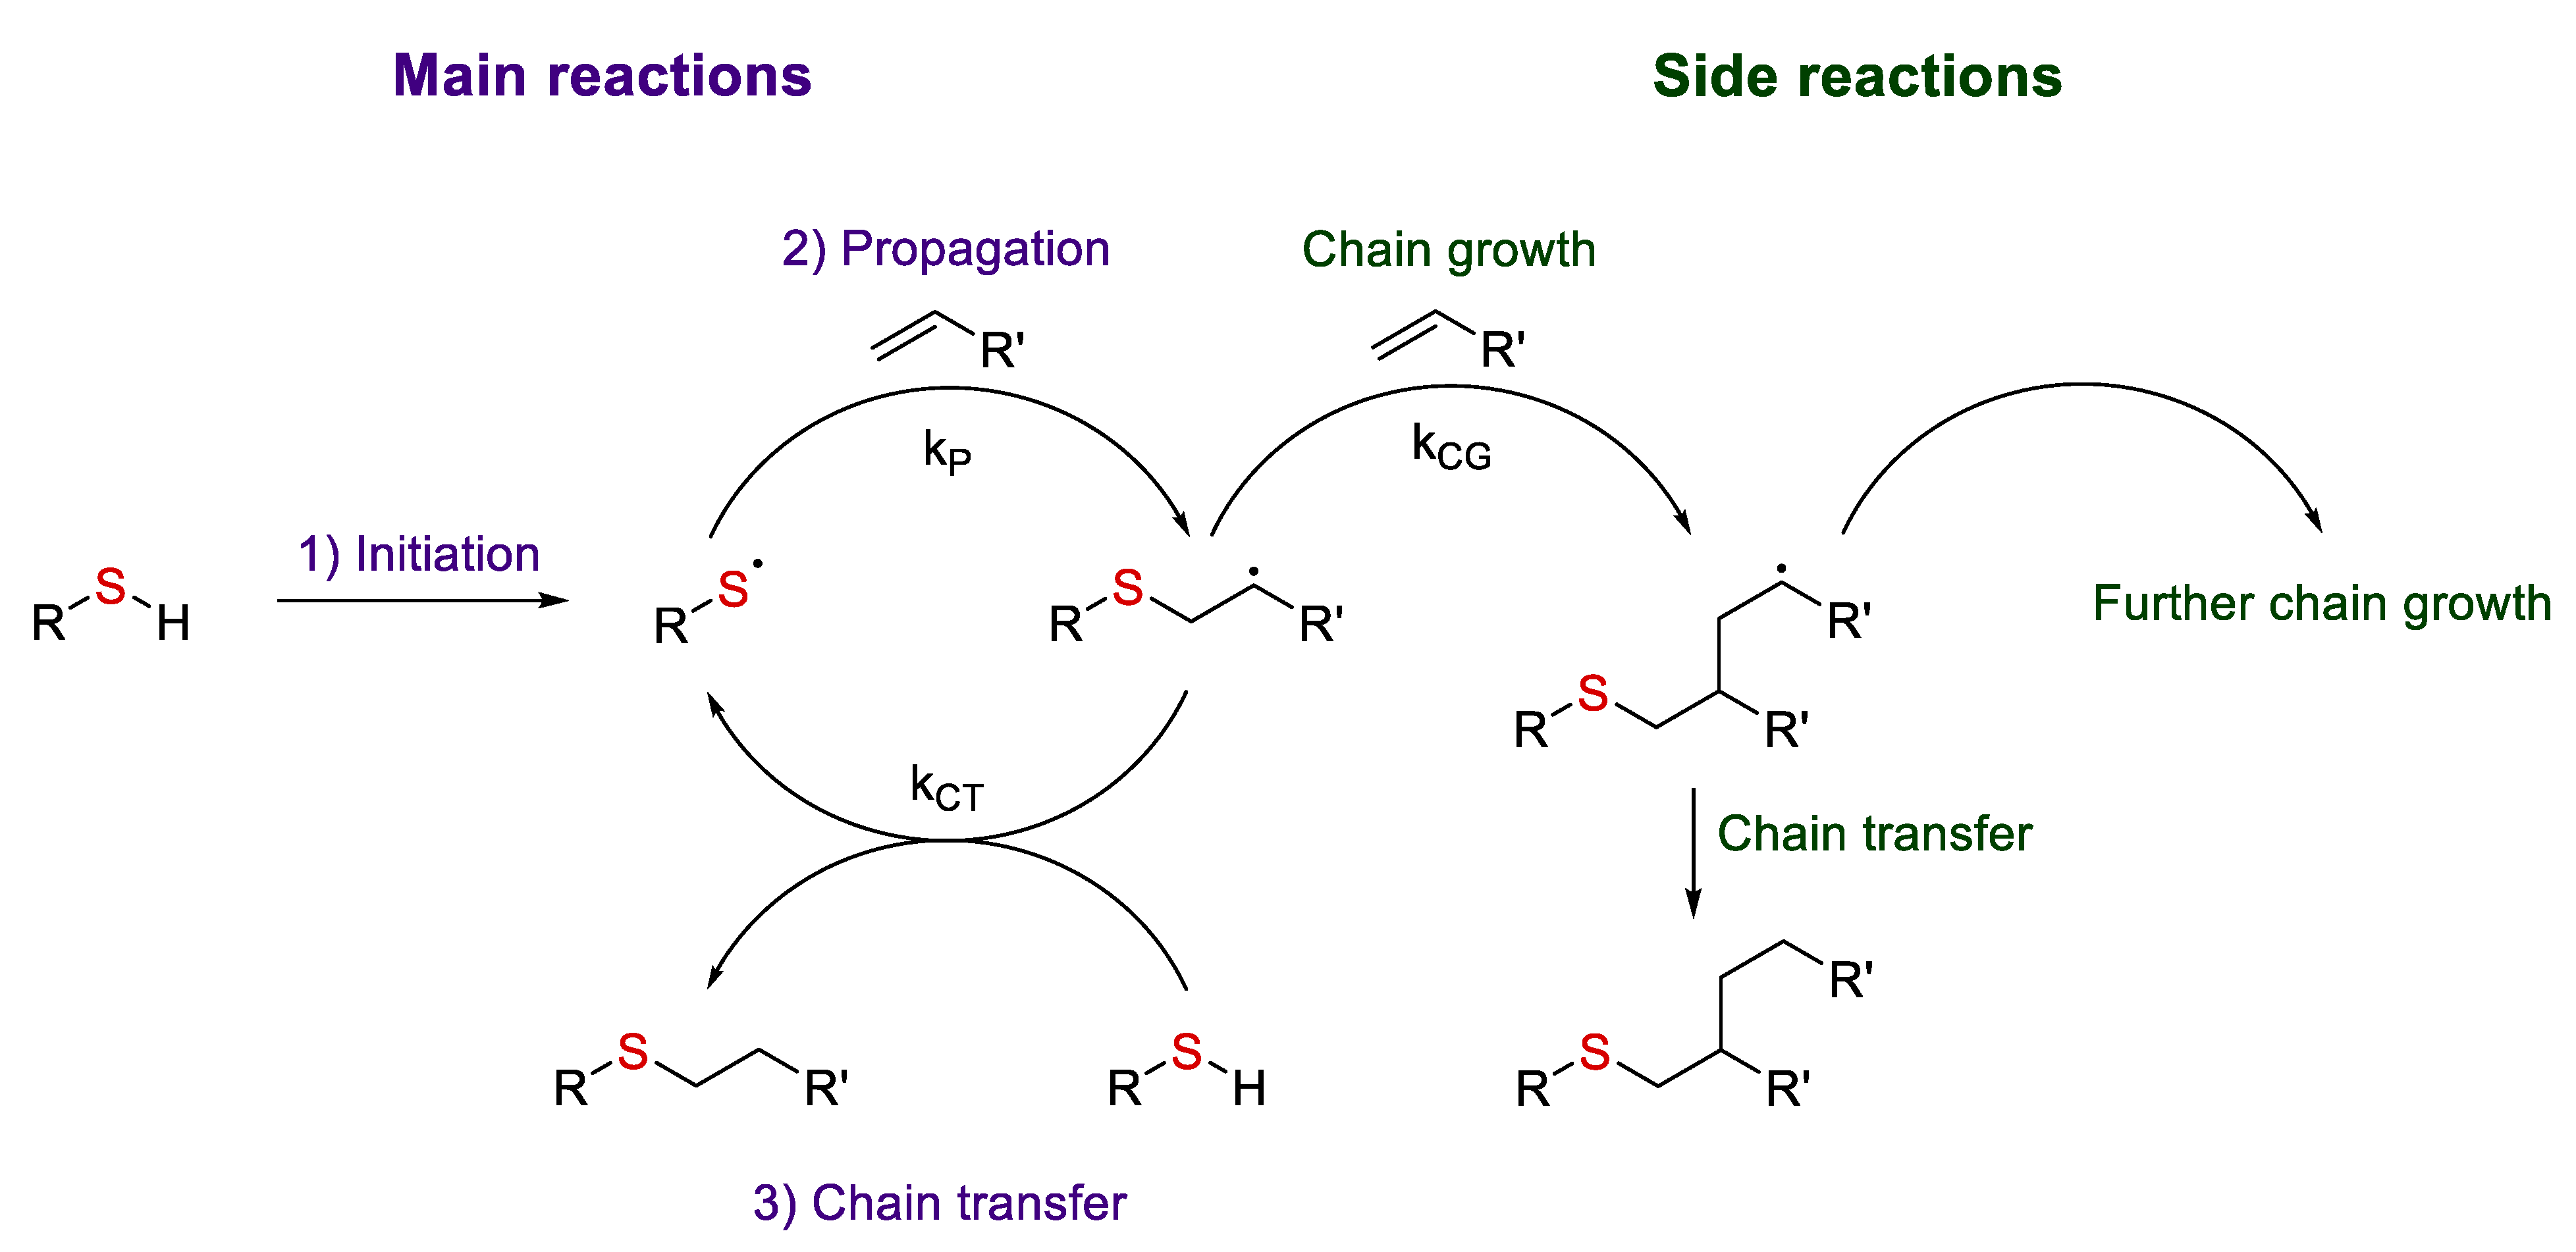
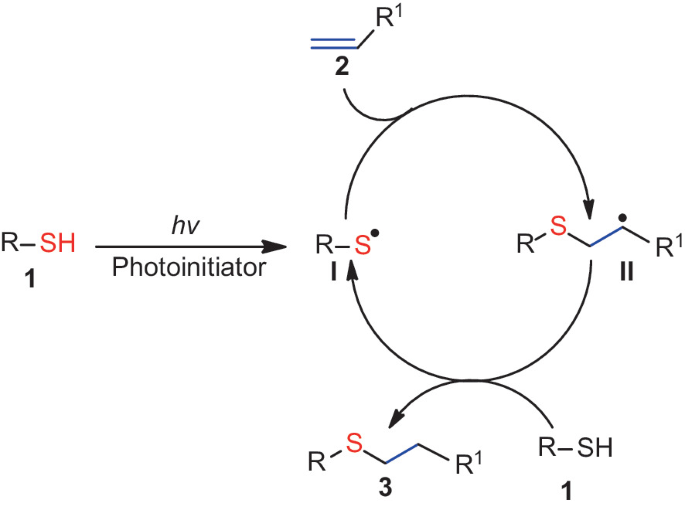
The mechanism runs through a thiyl radical. The initiator generates a radical that abstracts a hydrogen from the thiol, giving you RS·. That thiyl radical adds across the carbon-carbon double bond of the alkene, forming a carbon-centered radical intermediate. That intermediate then abstracts a hydrogen from another thiol molecule, propagating the chain. It's step-growth in the sense that every thiol can react with every alkene, but it's radical in nature, which means inhibition by oxygen is real and immediate. The regioselectivity is anti-Markovnikov. That's one of the things that makes this useful. With a terminal alkene like allyl methacrylate or a simple acrylate, the thiyl radical adds to the terminal carbon, and the hydrogen ends up on the internal carbon. You don't get the Markovnikov product cluttering things up the way you would in a conventional radical addition of HBr or something equally messy. This matters when you're building network structures or functionalizing surfaces because it keeps your topology predictable. I ran into a specific problem last year that took me about two weeks to isolate. I was crosslinking a PEG-diacrylate with a dithiol using TPO as the photoinitiator under a 365 nm LED at 15 mW/cm². The formulations that had higher thiol-to-alkene ratios than 1:1 were giving me incomplete conversion and sticky, tacky surfaces even after extended curing. The residual NMR peaks for the vinyl protons were still there at about twelve percent. I thought it was an oxygen inhibition issue at first and kept increasing the nitrogen purging time. Nothing changed.
The actual culprit was gel-effect autoacceleration working against me in reverse. At higher thiol concentrations relative to the diacrylate, the growing network didn't percolate early enough to trap the radicals. The thiyl radicals just kept finding free thiol hydrogens and terminating rather than propagating into new double bonds. The system stayed fluid longer than it should have, which meant oxygen diffuse in from the surface unchecked during the extended open time. My workaround was switching to a dual-cure system. I added a small amount of camphorquinone with an amine co-initiator for thermal activation, started the reaction thermally at 60 °C for five minutes to build enough molecular weight and viscosity to exclude oxygen, then hit it with UV for final conversion. That dropped residual vinyl to below two percent consistently. If you're starting out, pick your stoichiometry carefully. The classic recommendation is a 1:1 thiol-to-alkene ratio for maximal crosslink density in network formation. But if you're doing surface functionalization or dip-coating where you need to leave unreacted vinyl groups for a secondary reaction, run it thiol-rich. The excess thiol stays in solution and can be washed away, or you can use it for a thiol-Michael addition later if your substrate has maleimide groups. Initiator choice matters more than people admit. TPO is popular because it absorbs at longer wavelengths and has good molar extinction coefficients, but it leaves yellowing issues in clear formulations. IAP absorbs in the visible range and is cleaner optically, but it's significantly more expensive and slower at low loadings. If you're working with translucent or opaque samples, neither of those will penetrate deeply. You'll get a cured skin and a wet core. In those cases, switch to a Type II photoinitiator system like eosin Y with a tertiary amine donor. It works through visible light and the electron transfer mechanism gives you deeper penetration, though the reaction rate drops substantially. I've seen cure depths go from about two millimeters with TPO to over eight millimeters with eosin Y in the same formulation, but the total conversion time increases from roughly three minutes to maybe fifteen.
Get the Full Details
One thing that trips people up is that not all alkenes react at the same rate. Acrylates are fast. Methacrylates are noticeably slower because of the extra methyl group creating steric hindrance at the double bond. Vinyl ethers are in a different category entirely and follow a somewhat different reactivity pattern with thiols. If your protocol says "mix and cure" without specifying which alkene you're using, assume it's optimized for an acrylate and adjust your exposure time accordingly if you're working with a methacrylate. Doubling the cure time is a reasonable starting point, but check your conversion with FTIR because the difference isn't always linear. Sulfur-containing compounds have a notorious odor problem that no one mentions in the methods section. Even at low concentrations, the smell of unreacted thiol lingers in the fume hood and on your gloves. I use 11-mercaptoundecanoic acid sometimes because the carboxylic acid gives me a handle for further modification, and the smell alone will make your eyes water at concentrations above fifty millimolar. Working at lower concentrations helps, but then your reaction rates drop. It's a tradeoff you just have to manage with good ventilation and nitrile gloves changed frequently. The thiol permeates standard latex quickly. For quantitative results, FTIR is your friend. Track the disappearance of the vinyl C-H stretch around 1635 to 1640 cm¹ and the S-H stretch around 2550 to 2570 cm¹. Both should decrease proportionally if the reaction is proceeding cleanly. If the vinyl peak drops but the S-H peak doesn't, you have side reactions happening, probably radical-radical termination without chain propagation. That usually means your initiator concentration is too high relative to your thiol, or you're getting significant oxygen interference.
There are limitations that make this technique unsuitable for certain applications. It doesn't work well in aqueous media unless your thiol and alkene components are sufficiently hydrophilic or you're using a surfactant template. The thiyl radical can be quenched by water in ways that slow propagation, and phase separation will ruin your kinetics. If you need to do thiol-ene chemistry in water, look into using water-soluble initiators like lithium phenyl-2,4,6-trimethylbenzoylphosphinate or switching to a thiol-ene click variant that uses a different mechanism entirely, such as copper-catalyzed azide-alkyne cycloaddition, which is more forgiving in aqueous environments even though it requires a metal catalyst that some biological applications can't tolerate. The other major constraint is that thiols oxidize over time. If you're storing a thiol-containing formulation for more than a few days, especially at room temperature, you'll get disulfide formation and the effective thiol concentration drops. I keep my thiol stocks refrigerated and use them within a month. If you're making a bulk batch of pre-mixed thiol-ene resin, add a small amount of hydroquinone or BHT as a stabilizer, but remember that radical scavengers will interfere with your photoinitiation. You'll need to increase your initiator loading slightly to compensate, and the exact amount depends on how much stabilizer you're using. Ten ppm of BHT might only require a five percent increase in TPO, but fifty ppm could eat half your effective initiator concentration. For most routine lab work, a practical recipe that works reliably is a PEG-diacrylate (MW 575 to 1000) mixed with a dithiol like PEG-dithiol or a simple aliphatic dithiol at a 1:1 thiol-to-alkene ratio, two percent TPO by total weight, cured under a 365 nm LED at 10 to 20 mW/cm² for five to ten minutes in a nitrogen-purged environment. That gives you conversions above ninety percent in most cases. Adjust the components and conditions based on your specific viscosity requirements, mechanical property targets, and whether you need optical clarity or mechanical robustness. They pull in different directions and you'll have to choose which one matters more for your application.