Why Your GMP Compliance Keeps Failing at Inspection
Most people think GMP is just about filling out forms and keeping the facility clean. It's not. GMP is a system of checks that, when done right, catches problems before products reach consumers. When it's done wrong, you're one COI away from getting shut down. I've been working in supplement manufacturing for over a decade, and I've seen companies blow $200,000 on compliance only to fail because they missed something obvious. The core issue isn't knowledge. It's consistency.
Implementing Good Manufacturing Practices Dietary Supplements Correctly
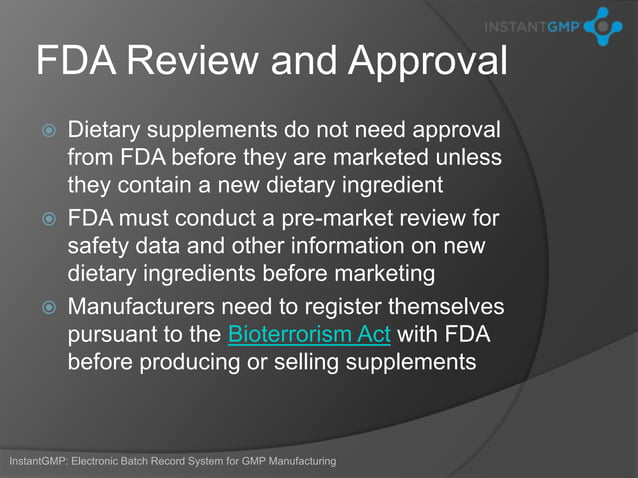
Let's start with the actual regulatory framework. In the US, 21 CFR Part 111 governs GMP for dietary supplements. It's not optional if you want to sell in this country. The FDA enforces it, and they don't joke around. The first thing you need is a proper quality control unit. This isn't just a person who checks labels. The QC unit needs authority to approve or reject all components, packaging, in-process materials, labels, rework, finished products, and any other material used in manufacturing. They need to have the qualifications to do this, and they need documented procedures for everything. Here's what most companies get wrong: they treat the QC unit like an afterthought. They hire someone part-time or give the responsibility to the production manager. This is a compliance violation waiting to happen. The QC unit must operate independently from production. If the same person approves incoming raw materials and also manufactures the final product, you have a conflict of interest the FDA will notice immediately during an inspection.
Component verification testing is where the real work begins. Every supplier's certificate of analysis needs to be validated. I've seen suppliers claim 99% purity on their CoAs, and the incoming test results came back at 87%. The supplier was either lying or their own QC was garbage. Either way, your CoA is not proof that your ingredient is what you think it is. You need to test every batch of every component using appropriate analytical methods. For identity testing, you need at least one method that can confirm the botanical or chemical identity of the component. HPLC, GC-MS, FTIR, or even simple microbiological testing depending on the material. For specifications, you need to set limits for purity, strength, potency, and composition. These specs should be based on your supplier's data, published literature, and your own testing history. Process validation is another area where companies cut corners. If you're blending a powder, you need to demonstrate that your blending process produces a homogeneous mixture. There are standard tests for this. The USP has general chapters on blending validation, and the FDA expects you to reference them. I ran a validation once where we had to take 30 samples from different locations in a 500kg batch, and the variability in active content had to be within a very tight range. This is not something you can skip. It typically takes 2-3 weeks and costs anywhere from $3,000 to $10,000 depending on your formulation complexity.
Get the Full Details

Water activity and moisture content matter more than you'd think. I once had a problem with a probiotic supplement where the final product was passing all microbial tests but was failing shelf-life stability. Turns out the water activity in the capsule shell was higher than expected, and it was affecting the viability of the probiotic strains over time. We had to switch suppliers for the capsules and revalidate the shelf life. That cost us about four months of delayed launch and roughly $45,000 in lost revenue and retesting. Labeling is a minefield. The Supplement Facts panel has very specific formatting requirements. Font sizes, column widths, vitamin and mineral ordering by daily value percentage — all of this is regulated. I've seen companies get warning letters for placing the Supplement Facts panel on the front of the container instead of the side, for using incorrect units of measure, and for making structure/function claims that crossed into disease claim territory without the required disclaimer. The structure/function claim disclaimer is mandatory if you make any claim about the structure or function of the body. It has to say "This statement has not been evaluated by the Food and Drug Administration. This product is not intended to diagnose, treat, cure, or prevent any disease." And it has to appear with "reasonable prominence." The FDA defines reasonable prominence as being near the claim itself, not hidden somewhere on the back panel in font size 4.
Record keeping is the backbone of GMP compliance. You need to maintain records for at least one year past the shelf life of the product, or two years from the date of distribution, whichever is longer. These records include batch production records, quality control records, distribution records, and complaint files. Everything needs to be traceable. If a customer complains about a batch, you should be able to pull the complete production history within hours, not days. Here's a specific problem I encountered with a client who was doing contract manufacturing. They had a contract manufacturer who was storing their raw materials in a warehouse across town, separate from the production facility. The CoAs came from the warehouse, but the actual manufacturing happened at the production site. When the FDA inspector asked to see the storage conditions for a specific batch of rosemary extract, there was a gap in the records. The warehouse logs showed the material arrived but the temperature logs only went back six months, and the production site didn't have its own storage records for that material. The workaround was to implement a dual-documentation system. The contract manufacturer maintained complete records at both locations, and the finished product file included temperature-controlled storage data from point of receipt through production. We also added a clause to the contract requiring the contract manufacturer to provide full environmental monitoring data for any storage area where the company's materials were kept. This added about $500 per year in administrative overhead but eliminated the record gap that would have otherwise resulted in a Form 483 observation.
Another common pitfall: sanitation. You need written procedures for cleaning and sanitizing all equipment that contacts the product. These procedures need to specify the method, the frequency, the responsible person, and how you verify that cleaning was effective. Many companies rely on visual inspection alone. That's insufficient. You need ATP swabbing or other verification methods to confirm that your cleaning actually removed residue. Personnel training is often overlooked. Every employee who handles product or touches equipment needs training in GMP practices relevant to their job. This isn't a one-time thing. It needs to be ongoing, and it needs to be documented. I've seen inspectors cite companies because they couldn't produce training records for a employee who had been there for three years. The employee knew the procedures because they'd been on the job, but without documentation, the lack of training records was a violation. The downside of strict GMP compliance is that it slows things down. Raw material testing alone can add 2-4 weeks to your production cycle depending on lab capacity. Process validation takes months. Stability testing requires you to hold batches for the full intended shelf life before you can fully validate expiration dating. For a small startup, this can be the difference between launching in six months and launching in eighteen months.

If you're a small operation with limited resources, consider outsourcing some functions. Contract QC labs can handle component testing faster than building an in-house lab. Third-party GMP auditors can identify gaps before the FDA does. And consulting firms that specialize in supplement compliance can help you set up systems that actually work rather than creating paperwork that collects dust. But outsourcing doesn't absolve you of responsibility. You are still the legal manufacturer, and the FDA holds you accountable for your contract manufacturers' compliance. So you need to audit your suppliers and contract manufacturers regularly. At minimum, annual on-site audits. If something changes at their facility, you need to know about it. The biggest mistake I see is treating GMP as a checklist instead of a culture. Forms filled out correctly don't matter if the people filling them out don't understand why they're filling them out. Training needs to be thorough, supervision needs to be real, and management needs to prioritize quality over speed. Products that ship without proper testing will come back to haunt you through complaints, recalls, and regulatory action.
One more thing about stability testing. Accelerated stability studies can give you early data, but they don't replace real-time stability. The FDA expects real-time data at the stated expiration date. If you claim 24 months of shelf life, you need to have stability data supporting that claim at the time of distribution. Accelerated data is useful for formulation development, but it's not a substitute for the actual study. I've also noticed that many companies focus heavily on the final product and underinvest in incoming material controls. A bad raw material can't be fixed by better processing. If your ashwagandha root extract has lead contamination, no amount of blending or encapsulation will remove it. Testing at the raw material stage is cheaper than a recall. This should be your highest priority investment. Cross-contamination is another area that gets messy fast. If you're producing multiple products on the same line, you need validation that your cleaning procedures between product changes actually prevent cross-contamination. This includes allergen cross-contact, potent actives carrying over into other products, and microbial contamination from one batch to the next. I worked on a project where a company was switching from a product containing fish oil to one containing a botanical blend, and the residual fish oil in the blender was causing oxidation issues in the new product. The validation study showed that a single wipe-down wasn't sufficient. They needed a full disassembly and wash cycle, which added 45 minutes of downtime between product changes.
Documentation control is also critical. Your procedures need version numbers, effective dates, and approval signatures. Obsolete versions need to be removed from use areas. I've seen inspection teams find superseded procedures still posted on walls in production areas. That's a red flag that your document control system isn't working. If you're looking for guidance documents, the FDA has a GMP compliance guide for dietary supplements on their website. It's not legally binding, but it represents the FDA's current thinking on the subject. Following it will generally keep you in compliance. There's also the Dietary Supplement Current Good Manufacturing Practice (CGMP) regulations themselves, which are the actual legal requirements. The industry self-regulation organizations like the Council for Responsible Nutrition have their own GMP guidelines, but they're voluntary. They can be helpful as a starting point, but they're not a substitute for regulatory compliance. The FDA doesn't recognize industry standards as meeting legal requirements.
Packaging integrity is another area that causes problems. Moisture-sensitive products need appropriate packaging. Oxygen-sensitive products need nitrogen flushing or oxygen absorbers. Light-sensitive products need amber bottles or opaque packaging. If your product degrades during storage or shipping, it's not a manufacturing problem — it's a packaging specification problem. But the FDA will still cite you for it. Receiving and storage conditions for raw materials matter too. Some materials need refrigeration. Some need desiccation. Some need protection from light. Your receiving procedures should include verification that transportation conditions were appropriate. If a shipment arrives that shows evidence of temperature abuse, you reject it or test it more thoroughly. Don't just accept it because the CoA looks good. Final product testing needs to verify identity, strength, purity, composition, and limits on contaminants. Heavy metals, pesticides, microbial pathogens, and adulterants are all things you need to be testing for. The FDA has published maximum limits for some contaminants, and other testing needs to be based on your risk assessment and historical data.
Distribution records should allow you to trace your product to each customer. This is important for recall purposes. If the FDA orders a recall, you need to be able to identify every customer who received the affected batch and where that batch went. Without proper distribution records, you'll be pulling your hair out trying to reconstruct that information after the fact. Complaint files need to be maintained for every consumer or professional complaint you receive. Even complaints that seem insignificant should be documented and reviewed. A pattern of similar complaints can indicate a systemic problem. I once saw a company that received ten complaints about capsules sticking together in hot weather. They dismissed them as isolated incidents. Three months later, a recall was issued because the capsule coating was failing, and those ten complaints were the first warning sign. Return and salvage of product is another area with specific requirements. You need procedures for handling returned product, including quarantine, evaluation, and disposition. Salvaged product — product that is reworked or processed to meet specifications — needs to be handled according to written procedures and approved by the QC unit before it can be released for distribution.
The reality is that GMP compliance is expensive and time-consuming. For a small operation, the cost of compliance can represent a significant portion of operating expenses. But the cost of non-compliance is far higher. Fines, seized product, injunctions, and criminal prosecution are all possibilities. And beyond the legal consequences, there's the reputational damage. One recall or warning letter can destroy a brand that took years to build. The best approach is to build compliance into your processes from the beginning. Don't treat it as an afterthought. Design your facility, your equipment, your procedures, and your training programs with GMP in mind. It's easier to do it right the first time than to retrofit your operations later.


