Why your DFT results are probably garbage
I spent three weeks last year debugging a simulation that turned out to have been wrong from line one of the input file. The geometry looked fine. The output files were beautifully formatted. The energy values were reasonable. And completely wrong because I had forgotten to include spin-orbit coupling for a heavy element system. That took me back six months of planning. This is the thing people don't tell you when they start learning Modelling And Simulation In Materials Science And Engineering. The simulations lie to you convincingly. They produce numbers. They produce pretty visualizations. They don't come with a big red stamp that says "this is nonsense" unless you know exactly what to look for.
Getting Started With Modelling And Simulation In Materials Science And Engineering
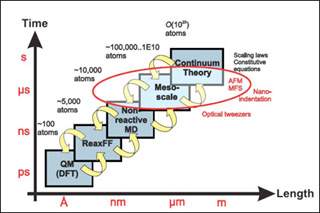
The first step is picking a tool that matches your problem, not the other way around. There are too many people running molecular dynamics on systems where density functional theory would have given them better answers in less time, or vice versa. The hierarchy goes roughly like this: quantum mechanical methods for electronic structure, atomistic simulations for time-dependent behavior at the nanoscale, continuum methods for macroscopic properties. Most real projects sit somewhere in the middle, stitching multiple methods together. For beginners, I recommend starting with an open-source package like VASP if your institution has a license, or Quantum ESPRESSO for a fully free option. Both are well documented. The documentation is actually useful, which is rare. LAMMPS works well for classical molecular dynamics once you have your force field sorted out. GIMP or OVITO for visualization. Don't skip the visualization step. Your atoms are going to look like a mess for the first few runs and you need to see that to understand what's happening. Here's a specific workflow I use when setting up a new project. I start with the crystal structure, preferably from a database like the Materials Project or COD. If the structure isn't available, I build it manually using known bond lengths and angles as a first guess. Then I run a geometry optimization with loose convergence criteria just to clean things up before tightening everything. The typical process goes from initial setup to a converged structure in about four to eight hours on a decent workstation, depending on system size.
I once had a graduate student who ran a full phonon calculation on a 200-atom supercell because I told them to check for imaginary modes. That took three days of CPU time on our cluster. We had already confirmed the structure was stable using a much smaller 12-atom cell with Gamma-point only sampling. The 200-atom run produced the same answer. They learned the lesson though, which is worth more than the saved compute time.
Get the Full Details

The methods and what they actually do
Density functional theory calculates the electronic structure by solving the Kohn-Sham equations self-consistently. You input atomic positions and get out electron densities, band structures, density of states. It's approximate by design. The exchange-correlation functional is the weak point, and no single functional works well for every material type. The PBE functional is the default for a reason, but it underestimates band gaps by roughly forty percent and sometimes overbinds by a significant margin. If you're working with transition metal oxides, you probably need DFT+U or a hybrid functional. If you're working with van der Waals systems, you need dispersion corrections or you'll get the interlayer spacing wrong by a full angstrom. Molecular dynamics uses classical force fields to move atoms through time. The forces come from empirical potentials rather than quantum mechanics. That makes it orders of magnitude faster than ab initio MD, but also much less accurate for bond breaking and electronic phenomena. The most common force fields are EAM for metals, Tersoff and REBO for carbon systems, and COMPASS or UFF for organic and mixed systems. Getting the force field right matters more than most people realize. A bad potential will produce plausible-looking trajectories that are physically meaningless. Finite element analysis handles the continuum side. You mesh a geometry, apply boundary conditions, and solve for stress, strain, thermal distribution, or whatever field you care about. COMSOL and ANSYS are the commercial standards. FEniCS is a solid open-source alternative. The bottleneck here is usually mesh generation, not the solver. A poor mesh introduces numerical artifacts that are hard to distinguish from real physics. Always run a mesh independence study. If your results change by more than five percent when you refine the mesh, you haven't converged yet.
Pitfalls that will waste your time
K-point sampling is where most beginners make their first mistake. Too few k-points and your total energy is meaningless. Too many and your calculation becomes impractical. The rule of thumb is roughly one k-point per two angstroms of reciprocal lattice vector, but that's a starting point, not a guarantee. I always run a convergence test by incrementing the k-point mesh and watching the total energy stabilize. It typically takes three to five test runs to find the sweet spot, and that adds maybe an hour to your setup time but saves you from publishing wrong numbers later. Another common issue is the pseudo-potential choice. You need to match the pseudo-potential to your functional. Some packages ship with default pseudo-potentials that work fine for PBE but perform badly with HSE or other functionals. The pseudo-potential library in Quantum ESPRESSO is better organized than VASP's. If you switch codes mid-project, don't assume your pseudo-potentials will transfer correctly without checking. Boundary conditions matter more than people admit. Periodic boundary conditions are the default in almost every electronic structure code because they're convenient, but they introduce artificial interactions between your periodic images. If your system has a dipole moment or you're simulating a surface, you need dipole corrections or a sufficiently thick vacuum layer. I use at least fifteen angstroms of vacuum for surface calculations. Anything less and the two surfaces interact through the periodic image, which shifts your work function by several tenths of an electronvolt.
I encountered a problem last year where my MD simulation of a grain boundary migration event produced impossible atomic displacements. The time step was set to one femtosecond, which should be fine. The issue turned out to be that I had accidentally created overlapping atoms during the initial relaxation because the energy minimization had stopped prematurely. The atoms were so close that the repulsive part of the potential exploded. The fix was simple: run a longer relaxation with tighter convergence criteria before switching to dynamics. The initial relaxation went from fifteen minutes to about forty-five minutes, but it prevented an hour-long simulation from producing garbage data.
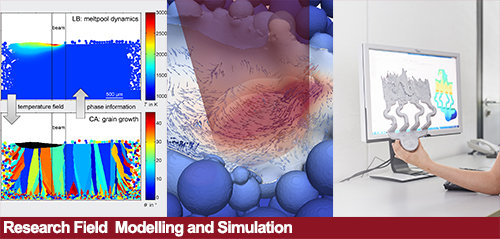
Combining methods for real problems
Most interesting materials problems require more than one method. You might use DFT to parameterize a force field, then run MD simulations with that force field to study temperature effects, then feed the results into a finite element model for macroscopic predictions. This multiscale approach is standard practice but poorly executed more often than not. The handoff between methods is where information gets lost. When I pass DFT results into a force field, I verify the force field reproduces the lattice constants and elastic constants within ten percent before trusting it for anything else. That's a quick check that catches most problems early. When I move from atomistic to continuum, I extract representative volumes large enough to capture the microstructure but small enough to keep the finite element mesh manageable. Usually a few hundred nanometers on each side for polycrystalline materials. Validation against experiment is non-negotiable. No simulation is useful unless it matches real data, even approximately. XRD patterns, SEM images, mechanical testing results, thermal analysis. Pick at least two experimental measurements to compare against your simulation output. If your simulated band gap doesn't match optical absorption data, your model is wrong regardless of how internally consistent it appears.
What this approach cannot do
Simulation fails when the physics you need to capture happens at a length or time scale beyond what your method can reach. Molecular dynamics struggles to simulate anything slower than microseconds for most systems, and even that requires specialized hardware or clever techniques like hyperdynamics. DFT is limited to hundreds of atoms and picoseconds of effective time. Finite element methods can handle anything you can mesh, but they require constitutive relationships that are usually derived from experiments or higher-level simulations. If you're trying to study fracture propagation in a bulk material, atomistic methods won't work because you need millions or billions of atoms. If you're studying exciton dynamics in a quantum dot, continuum methods won't capture the quantum confinement effects. The right tool depends entirely on the question. There's no universal simulation that solves everything. Computational cost remains a real constraint even with modern hardware. A well-converged DFT calculation on a medium-sized system can take days on a single CPU core. Running it on a GPU cluster cuts that to hours, but access to those resources is not guaranteed. Plan your calculations with this in mind. Start small. Converge your parameters. Only then run the full calculation.